Related Experiment Video
Updated: Jun 8, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Improved description of the structure of molecular and layered crystals: ab initio DFT calculations with van der
Tomáš Bučko1, Jürgen Hafner, Sébastien Lebègue
1Department of Physical and Theoretical Chemistry, Faculty of Natural Sciences, Comenius University, Mlynská Dolina, SK-84215 Bratislava, Slovakia. tomas.bucko@univie.ac.at

Abstract:
The implementation of technique for full structural optimizations of complex periodic systems in the DFT-PAW package VASP, including the volume and shape of the unit cell and the internal coordinates of the atoms, together with a correction that allows an appropriate modeling of London dispersion forces, as given by the DFT-D2 approach of Grimme [Grimme, S. J. Comp. Chem. 2006, 27, 1787], is reported. Dispersion corrections are calculated not only for the forces acting on the atoms, but also for the stresses on the unit cell. This permits a simultaneous optimization of all degrees of freedom. Benchmark results on a series of prototype systems are presented and compared to results obtained by other methods and experimental data. It is demonstrated that the computationally inexpensive DFT-D2 scheme yields reasonable predictions for the structure, bulk moduli, and cohesive energies of weakly bonded materials.

Related Concept Videos
 01:29
Determination of Crystal Structures
 01:10
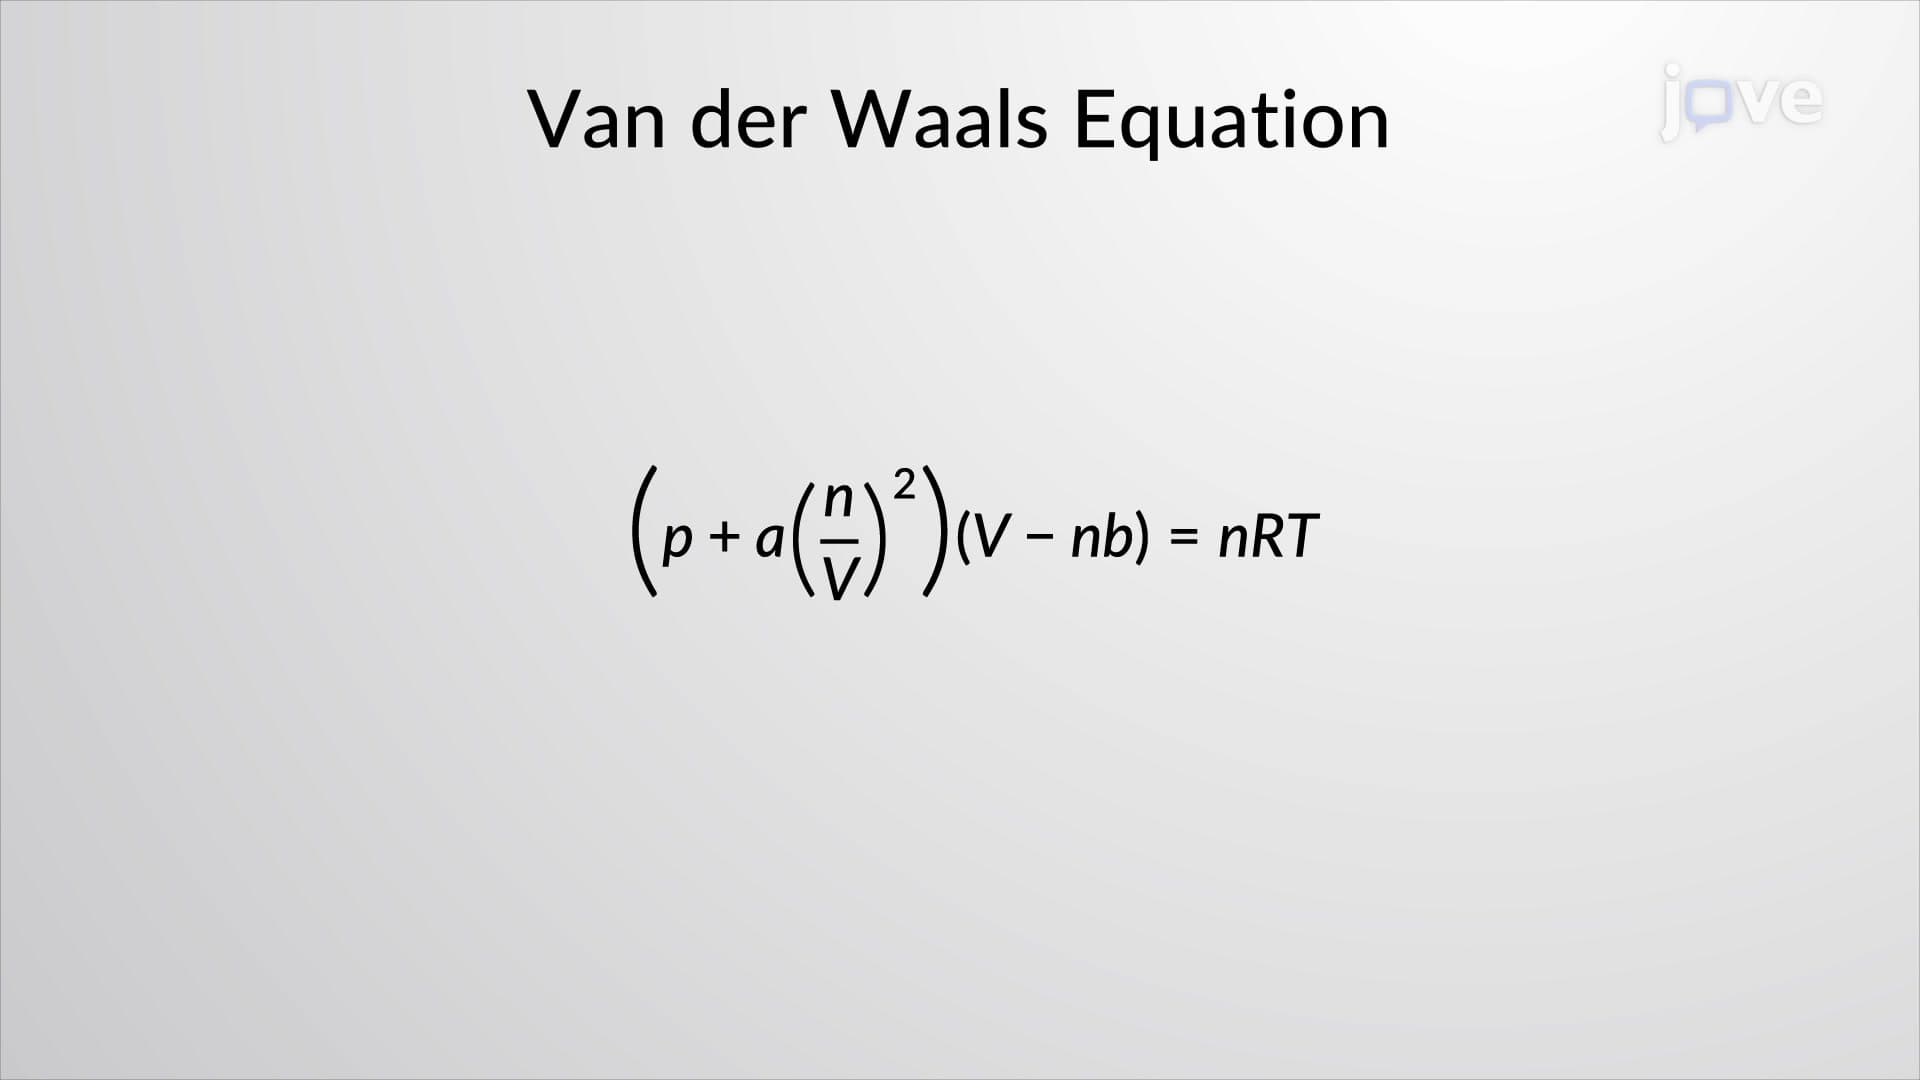
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
Van der Waals Interactions
 02:18
X-ray Crystallography
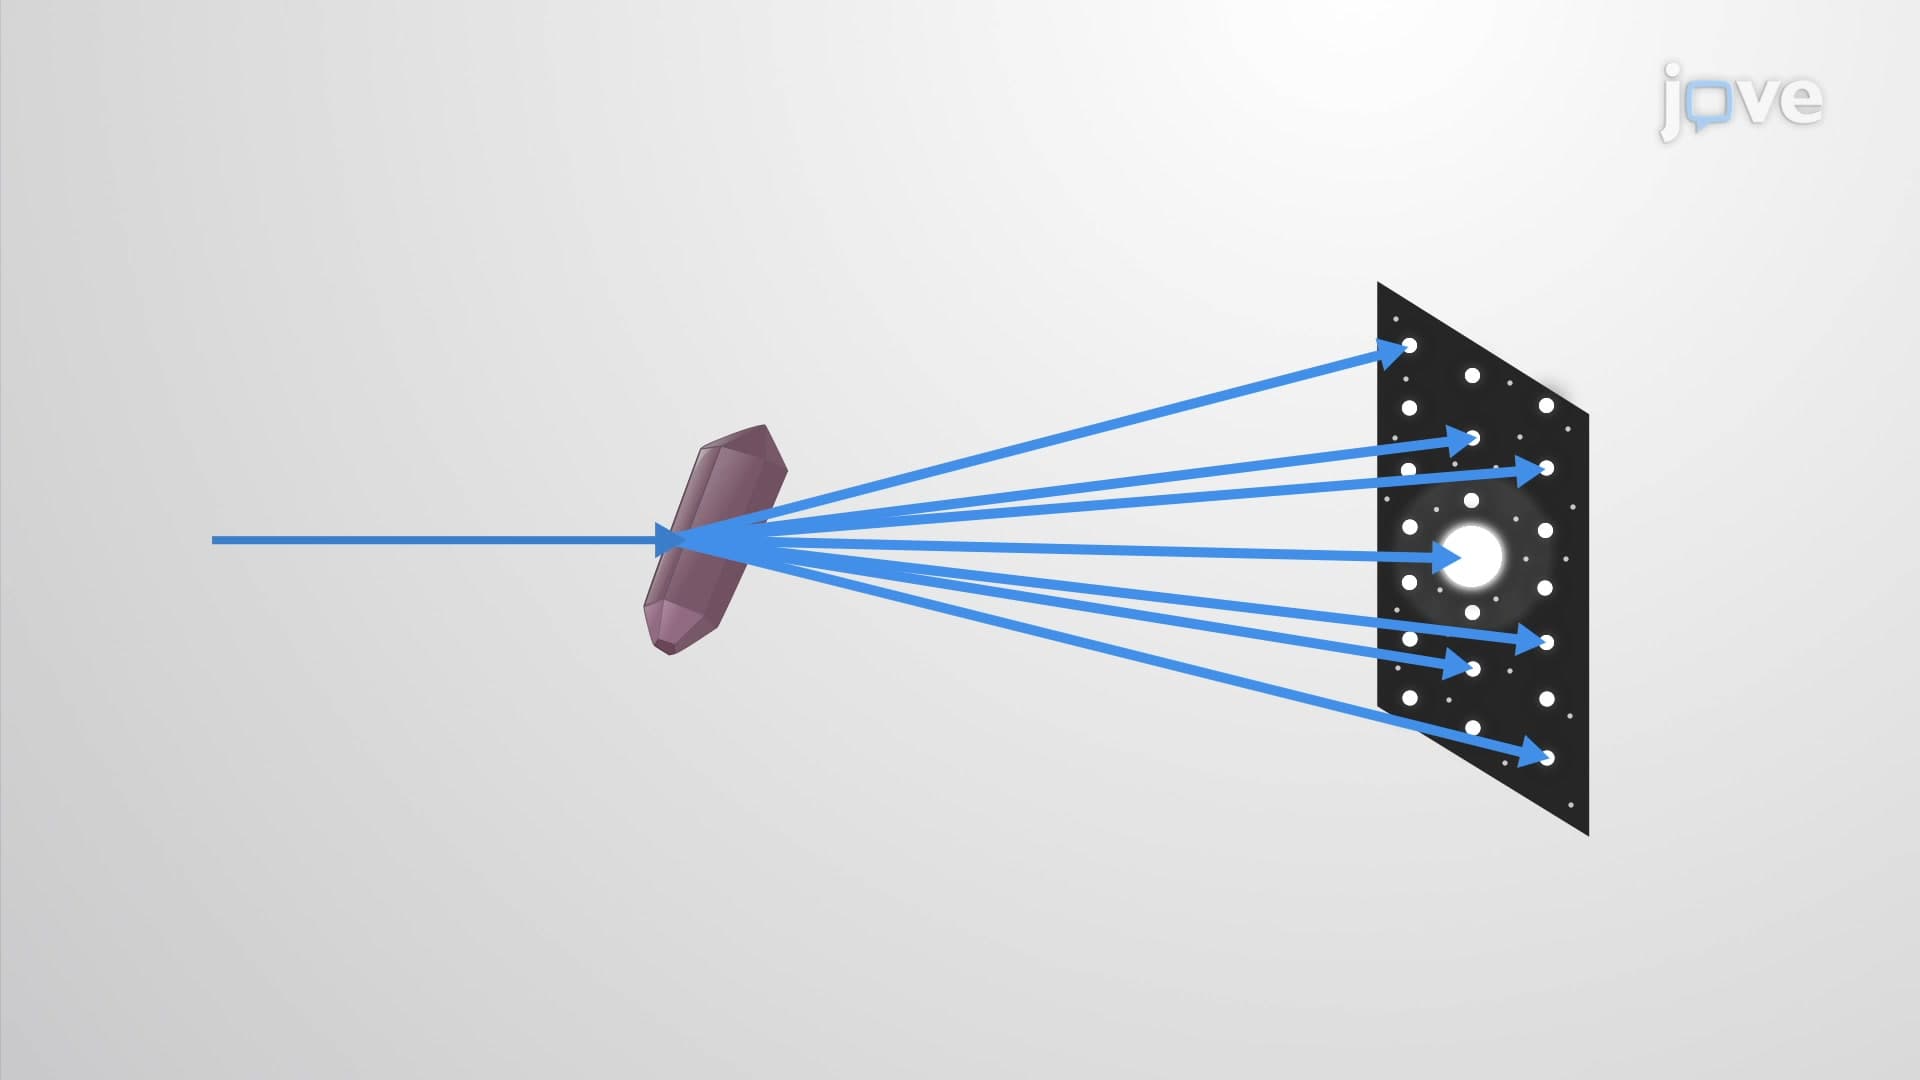
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...