Iron complexes for the catalytic transfer hydrogenation of acetophenone: steric and electronic effects imposed by alkyl substituents at phosphorus
Paraskevi O Lagaditis1, Alan J Lough, Robert H Morris
1Department of Chemistry, University of Toronto, Toronto, Ontario M5S 3H6, Canada.

Abstract:
A series of iron(II) complexes, trans-[Fe(NCMe)(2)(PR(2)CH(2)CH═NCH(2)CH(2)N═CHCH(2)PR(2))][BPh(4)](2) (5, R = Cy; 7, R = iPr; 9, R = Et) were prepared via the template synthesis in one-pot involving air-stable phosphonium dimers, [cyclo-(-PR(2)CH(2)CH(OH)-)(2)](Br)(2) (4, R = Cy; 6, R = iPr; 8, R = Et), KOtBu, [Fe(H(2)O)(6)][BF(4)](2) and ethylenediamine in acetonitrile. In the synthesis of 9, a methanol/acetonitrile solvent mixture was required; otherwise an intermediate iron bis(tridentate) complex, [Fe(PEt(2)CH(2)CH═NCH(2)CH(2)NH(2))(2)](2+), formed as determined by electrospray ionization mass spectrometry (ESI-MS). The crude iron(II) complexes from a template synthesis with ethylenediamine or (S,S)-1,2-diphenylethylenediamine are stirred in acetone under a CO atmosphere (∼2 atm) overnight to displace a NCMe ligand; however, in addition to this, bromide displaces an NCMe ligand as well to form a new class of the iron complexes trans-[Fe(CO)(Br)(PR(2)CH(2)CH═NCHR'CHR'N═CHCH(2)PR(2))](+) (10 R = Cy, R' = H; (S,S)-11, R = Cy, R' = Ph; 12, R = iPr, R' = H; (S,S)-13, R = iPr, R' = Ph; 14, R = Et, R' = H; (S,S)-15, R = Et, R' = Ph). These complexes were isolated in moderate yields (55-84%) as tetraphenylborate salts. Complexes 10-15 were tested for the catalytic transfer hydrogenation of acetophenone in basic iso-propanol at 25 and 50 °C. The complexes 10-13 (where R = Cy or iPr) were inactive while the complexes 14 and (S,S)-15 (where R = Et) were active at 25 °C but had better activity at 50 °C. Complex (S,S)-15 was higher in activity than complex 14, achieving turnover frequencies as high as 4100 h(-1), conversions of acetophenone to (R)-1-phenylethanol as high as 80% and an enantiomeric excess (e.e.) of 50% in the product. As catalysis progressed, the e.e. diminished to as low as 26%.
Related Concept Videos
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 02:13
Reduction of Alkenes: Catalytic Hydrogenation
Metals like palladium, platinum, and nickel are commonly used in their solid forms — fine powder on an inert surface. As these catalysts remain insoluble in the reaction mixture, they are referred to as heterogeneous catalysts.
The hydrogenation process takes place on the surface of...
 01:14
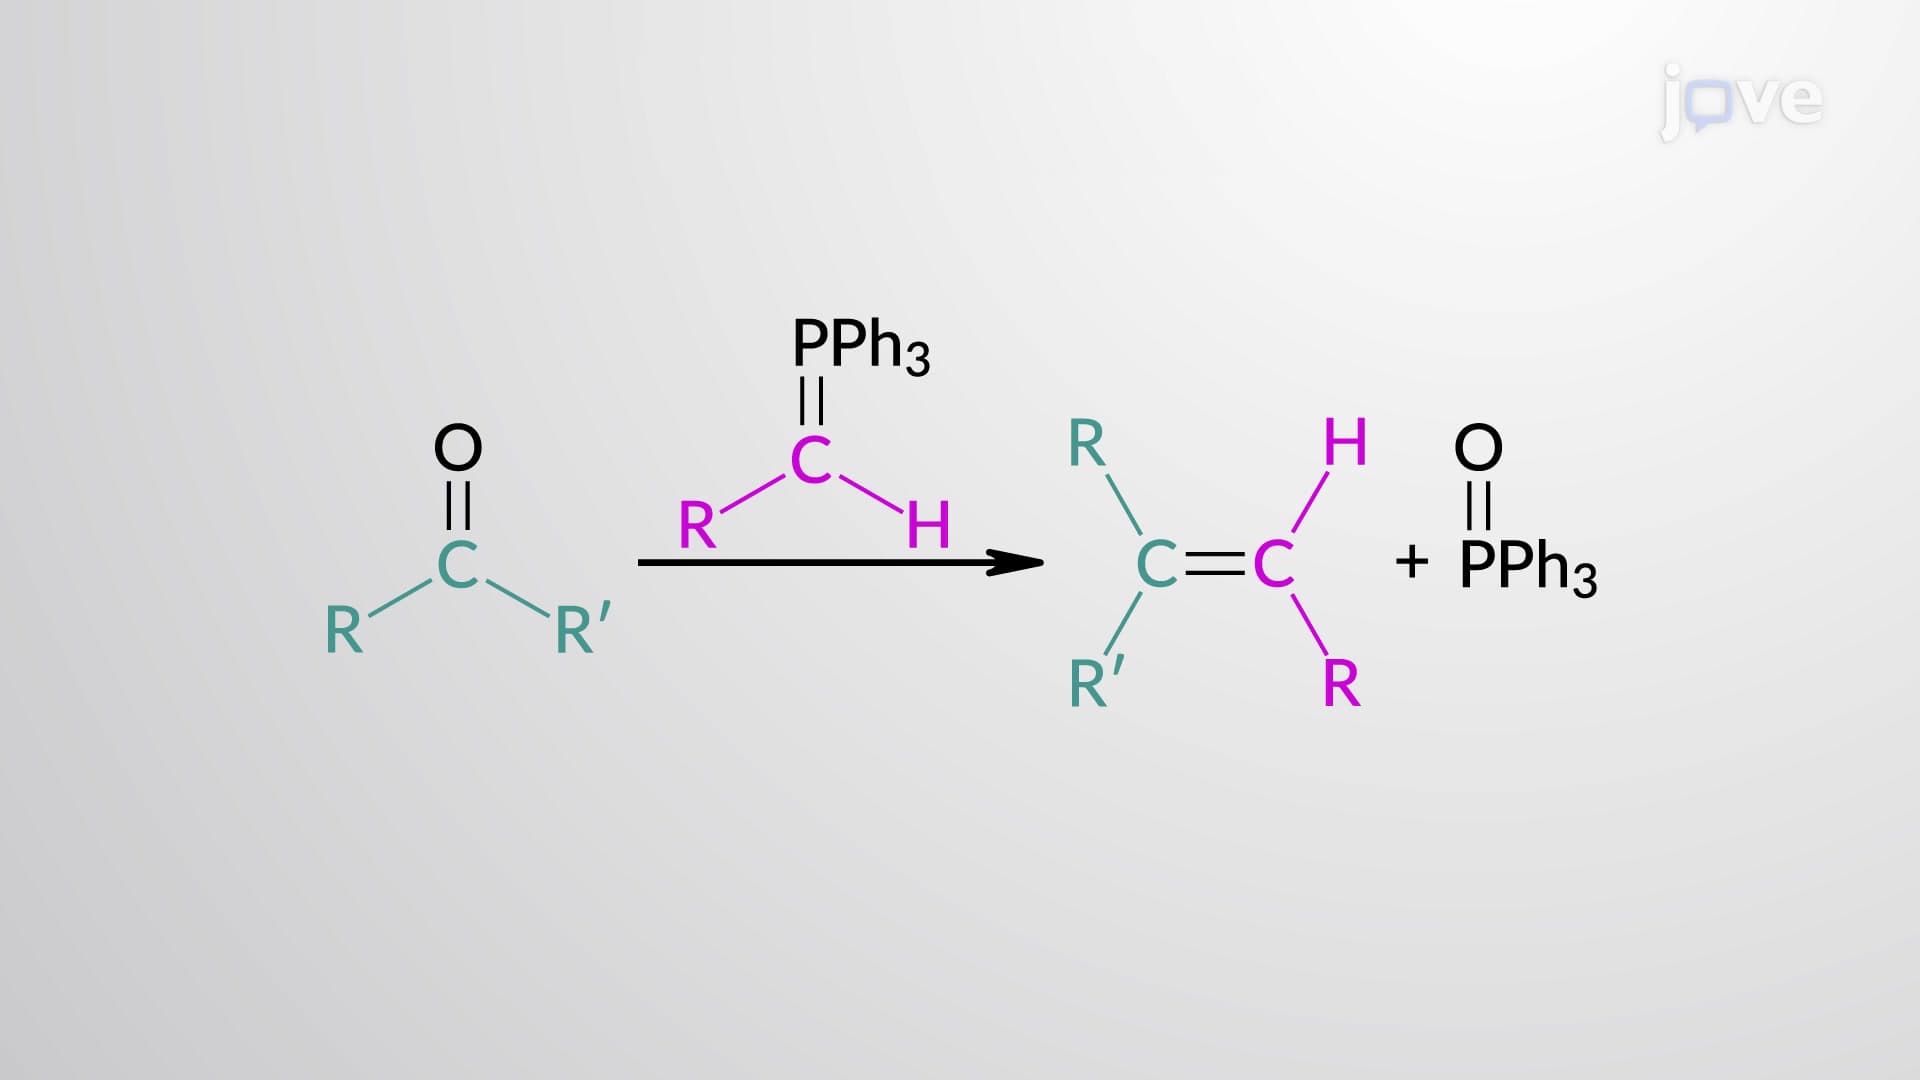
Aldehydes and Ketones to Alkenes: Wittig Reaction Mechanism
The reaction begins with the nucleophilic addition between a phosphorus ylide and the carbonyl compound. Due to its carbanionic character, phosphorus ylide acts as a strong nucleophile and attacks the electrophilic carbonyl group. This generates a charge-separated dipolar intermediate called betaine. The negatively charged oxygen atom and...
 02:47
Alkynes to Aldehydes and Ketones: Hydroboration-Oxidation
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.