
Comparison of aromatic NH···π, OH···π, and CH···π interactions of alanine using MP2, CCSD, and DFT methods
Neetha Mohan1, Kunduchi P Vijayalakshmi, Nobuaki Koga
1Computational Modeling and Simulation Section, National Institute for Interdisciplinary Science and Technology, Trivandrum 695 019, India.

Abstract:
A comparison of the performance of various density functional methods including long-range corrected and dispersion corrected methods [MPW1PW91, B3LYP, B3PW91, B97-D, B1B95, MPWB1K, M06-2X, SVWN5, ωB97XD, long-range correction (LC)-ωPBE, and CAM-B3LYP using 6-31+G(d,p) basis set] in the study of CH···π, OH···π, and NH···π interactions were done using weak complexes of neutral (A) and cationic (A(+)) forms of alanine with benzene by taking the Møller-Plesset (MP2)/6-31+G(d,p) results as the reference. Further, the binding energies of the neutral alanine-benzene complexes were assessed at coupled cluster (CCSD)/6-31G(d,p) method. Analysis of the molecular geometries and interaction energies at density functional theory (DFT), MP2, CCSD methods and CCSD(T) single point level reveal that MP2 is the best overall performer for noncovalent interactions giving accuracy close to CCSD method. MPWB1K fared better in interaction energy calculations than other DFT methods. In the case of M06-2X, SVWN5, and the dispersion corrected B97-D, the interaction energies are significantly overrated for neutral systems compared to other methods. However, for cationic systems, B97-D yields structures and interaction energies similar to MP2 and MPWB1K methods. Among the long-range corrected methods, LC-ωPBE and CAM-B3LYP methods show close agreement with MP2 values while ωB97XD energies are notably higher than MP2 values.

Related Concept Videos
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:18
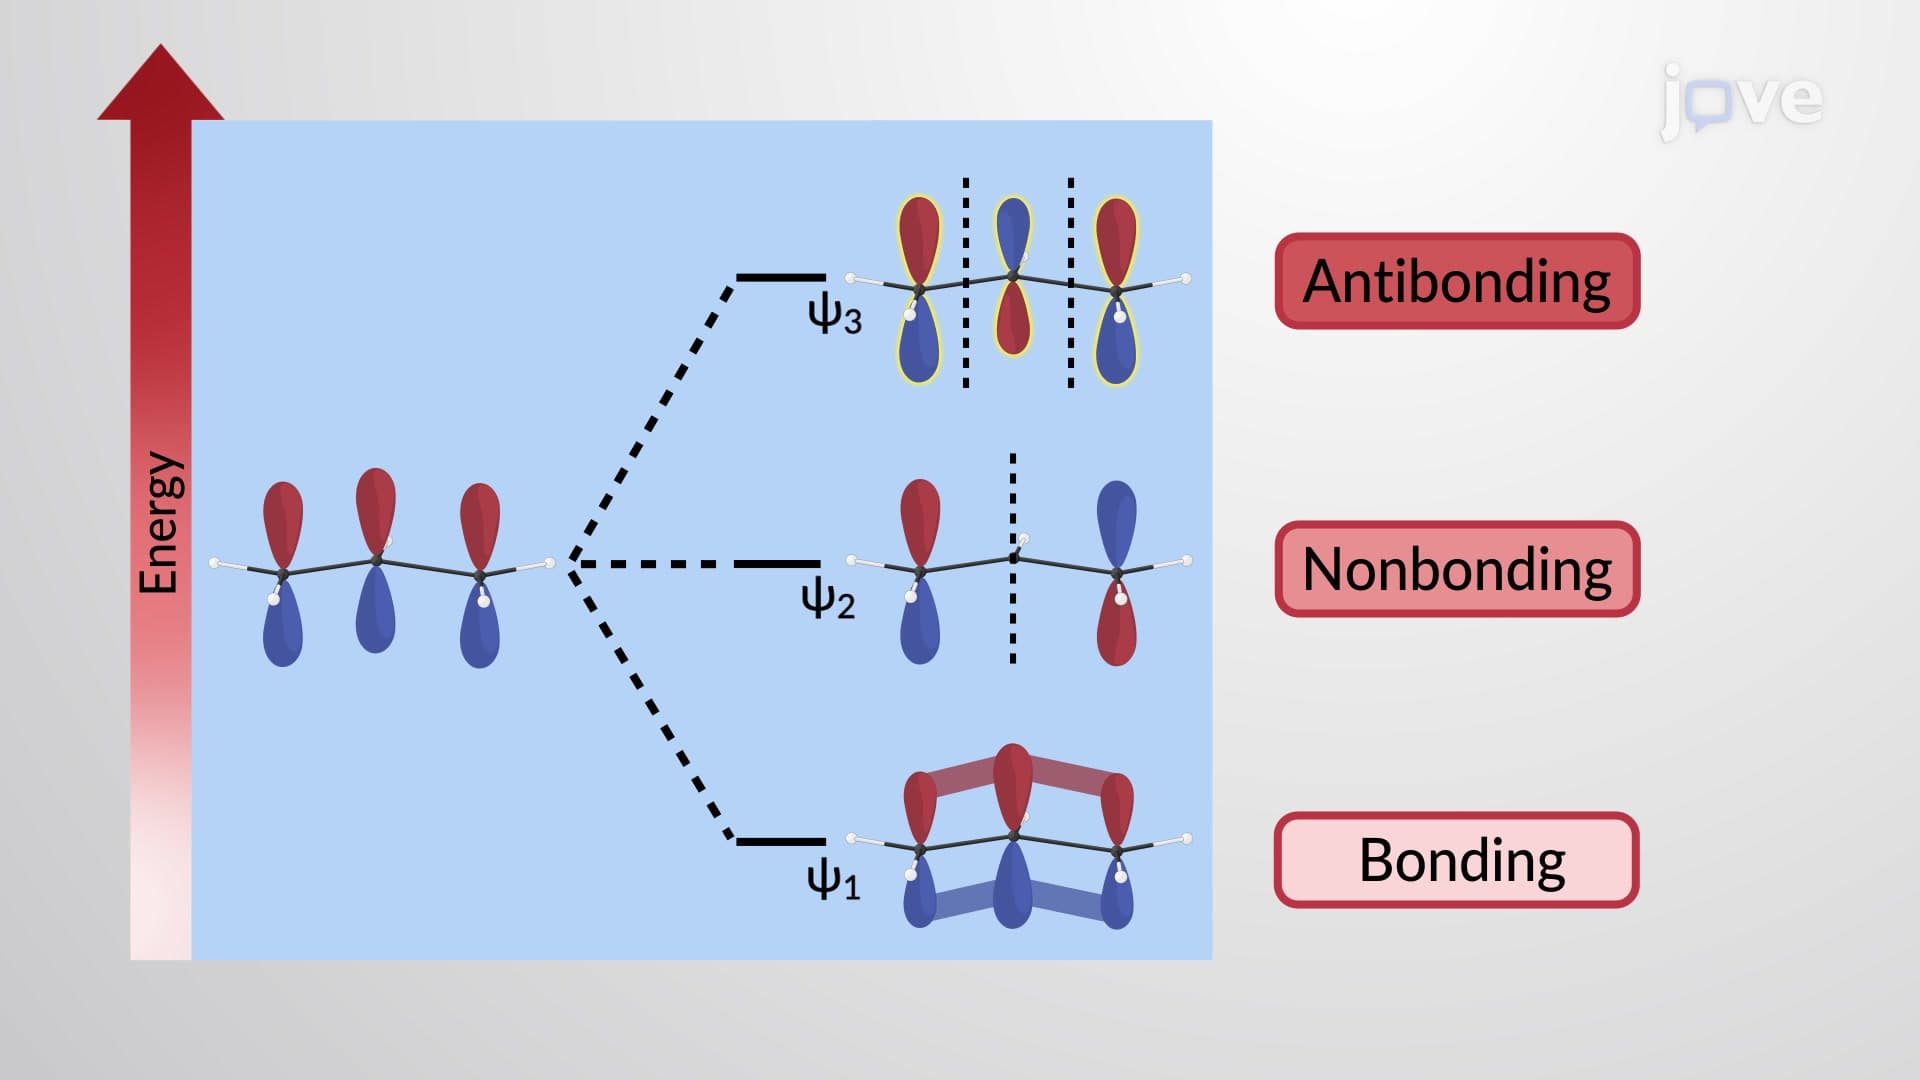
π Molecular Orbitals of the Allyl Cation and Anion
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.
 01:27
π Electron Effects on Chemical Shift: Overview
 01:18
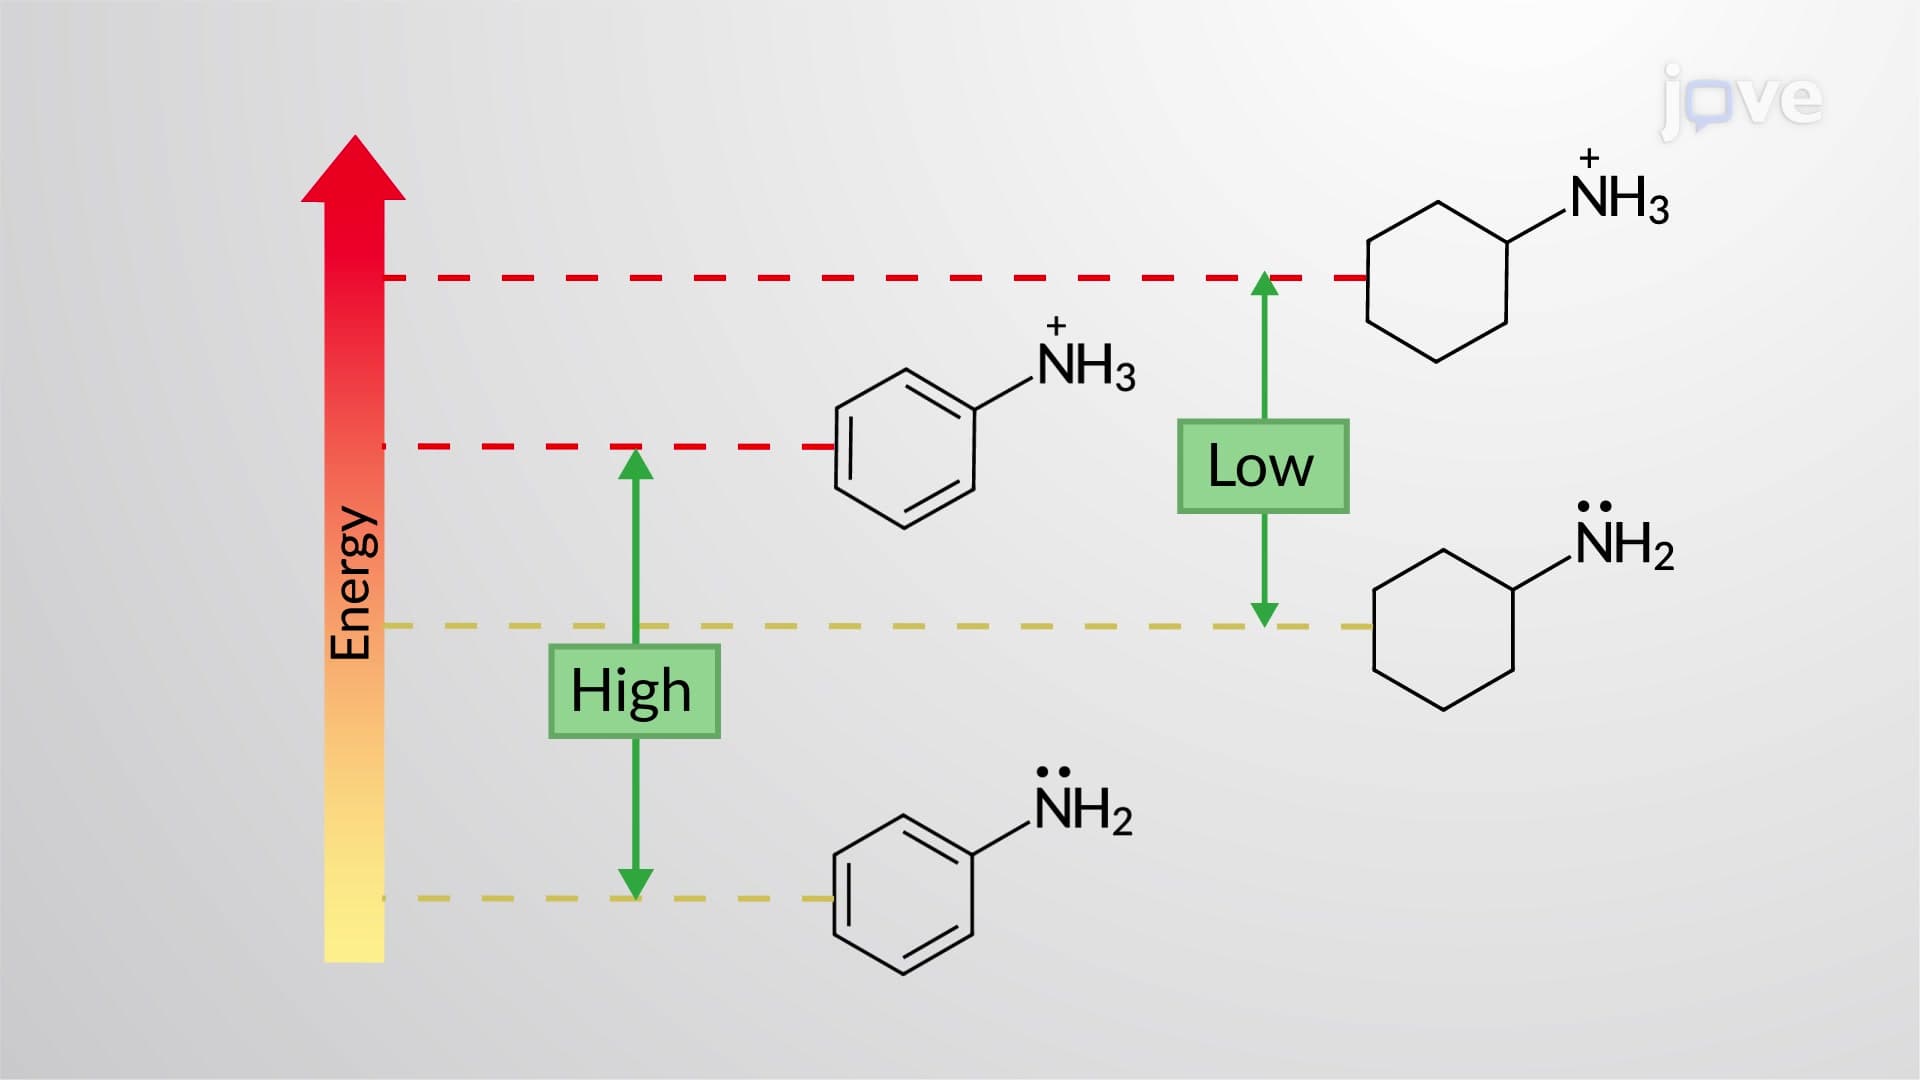
Basicity of Aromatic Amines
 01:19
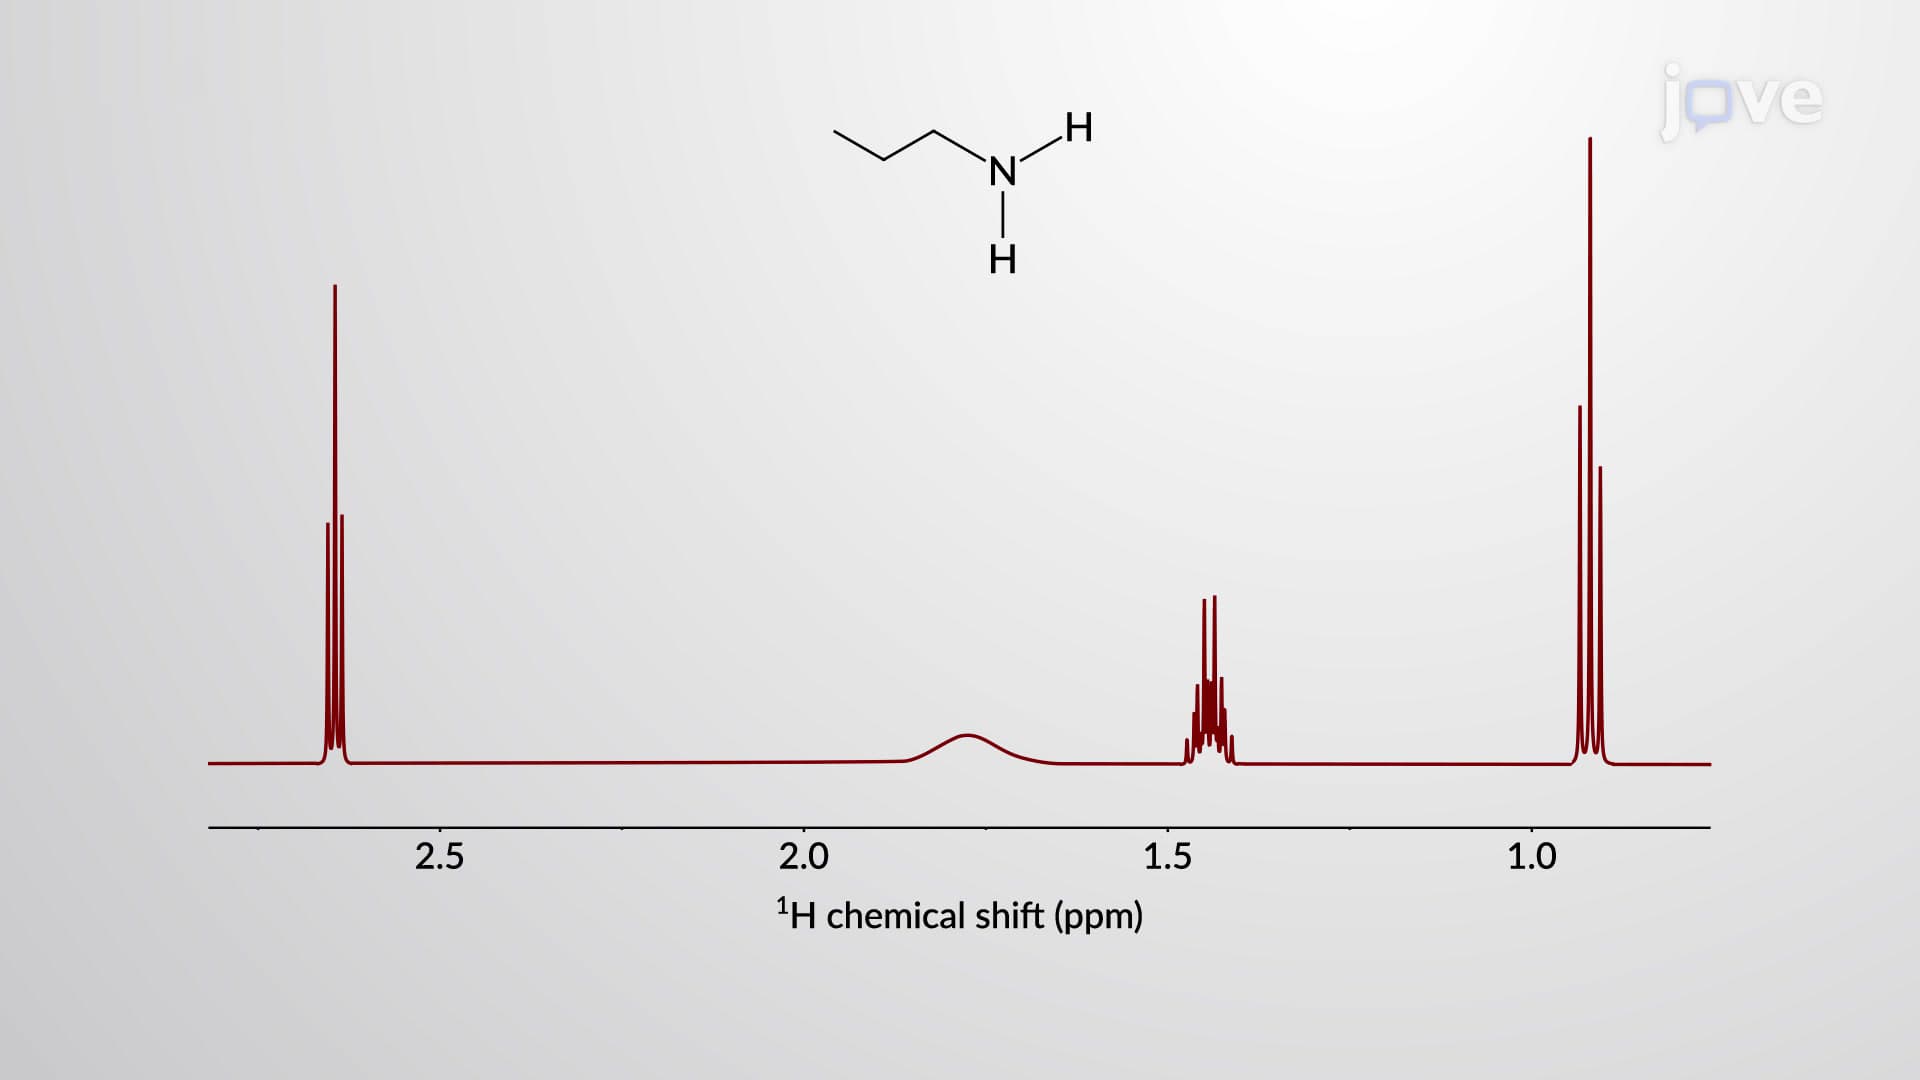
NMR Spectroscopy Of Amines