Related Experiment Video
Updated: Jun 7, 2026
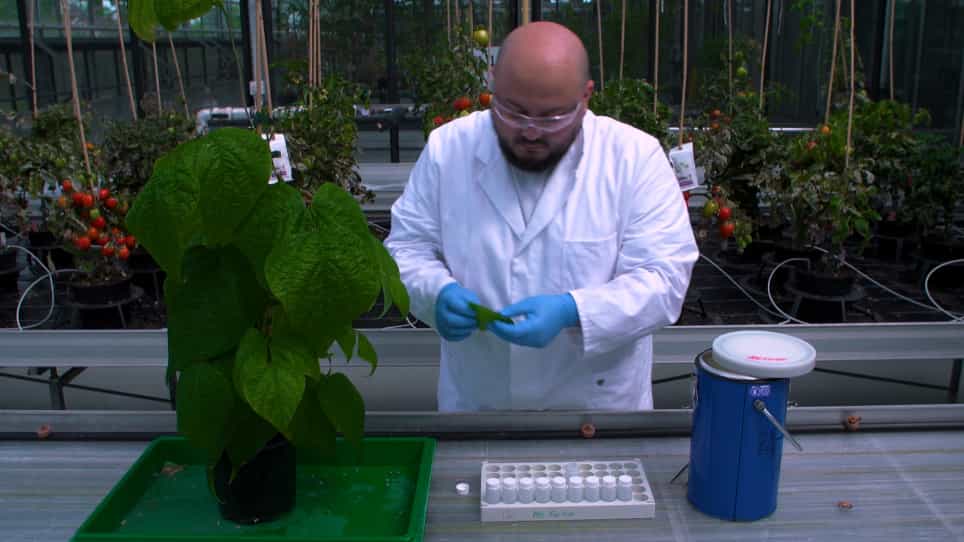
Large-Scale Multi-Omics Genome-Wide Association Studies (Mo-GWAS): Guidelines for Sample Preparation and Normalization
Published on: July 27, 2021
A statistical approach to finding overlooked genetic associations
Andrew K Rider1, Geoffrey Siwo, Nitesh V Chawla
1Department of Computer Science and Engineering, University of Notre Dame, Notre Dame, Indiana, USA.

Background:
Complexity and noise in expression quantitative trait loci (eQTL) studies make it difficult to distinguish potential regulatory relationships among the many interactions. The predominant method of identifying eQTLs finds associations that are significant at a genome-wide level. The vast number of statistical tests carried out on these data make false negatives very likely. Corrections for multiple testing error render genome-wide eQTL techniques unable to detect modest regulatory effects. We propose an alternative method to identify eQTLs that builds on traditional approaches. In contrast to genome-wide techniques, our method determines the significance of an association between an expression trait and a locus with respect to the set of all associations to the expression trait. The use of this specific information facilitates identification of expression traits that have an expression profile that is characterized by a single exceptional association to a locus. Our approach identifies expression traits that have exceptional associations regardless of the genome-wide significance of those associations. This property facilitates the identification of possible false negatives for genome-wide significance. Further, our approach has the property of prioritizing expression traits that are affected by few strong associations. Expression traits identified by this method may warrant additional study because their expression level may be affected by targeting genes near a single locus.
Results:
We demonstrate our method by identifying eQTL hotspots in Plasmodium falciparum (malaria) and Saccharomyces cerevisiae (yeast). We demonstrate the prioritization of traits with few strong genetic effects through Gene Ontology (GO) analysis of Yeast. Our results are strongly consistent with results gathered using genome-wide methods and identify additional hotspots and eQTLs.
Conclusions:
New eQTLs and hotspots found with this method may represent regions of the genome or biological processes that are controlled through few relatively strong genetic interactions. These points of interest warrant experimental investigation.
More Related Videos
 05:53
Candidate Gene Testing in Clinical Cohort Studies with Multiplexed Genotyping and Mass Spectrometry
Published on: June 21, 2018
 11:35
Screening for Functional Non-coding Genetic Variants Using Electrophoretic Mobility Shift Assay (EMSA) and DNA-affinity Precipitation Assay (DAPA)
Published on: August 21, 2016
Related Concept Videos
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:23
Behavioral Genetics and Its Designs
The primary methodologies used in behavior genetics include family studies, twin studies, and adoption studies, each providing unique...
 02:46
Genetic Screens
Forward genetic screens
Forward or “classical” genetic screens involve creating random mutations in an organism’s DNA using radiation, mutagens, or insertion of additional bases, which result in visible changes...
 01:25
Strategies for Assessing and Addressing Confounding
Confounding can be addressed at both the design phase of a study and through analytical methods after data...
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...