

A hybrid method for estimation of molecular dynamics of diazepam-density functional theory combined with NMR and
J Mielcarek1, D M Nowak, A Pajzderska
1Department of Inorganic and Analytical Chemistry, Poznan University of Medical Sciences, Grunwaldzka 6, 60-780 Poznan, Poland. jmielcar@am.poznan.pl

Abstract:
Reorientation of the molecule of diazepam was investigated by calorimetric methods, IR absorption and NMR. The investigation of dynamics was complemented by density functional study (DFT) of vibrational frequencies and infrared intensities, calculations of steric hindrances and Monte Carlo simulations. The results indicated the occurrence of reorientation jumps of the CH(3) group and conformational motion of the benzodiazepine ring. The activation parameters of the methyl group reorientation were determined and the activation barrier obtained was in good agreement with the theoretically estimated value. The FT-IR spectra were assigned using results of DFT calculations.
Related Concept Videos
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 00:58
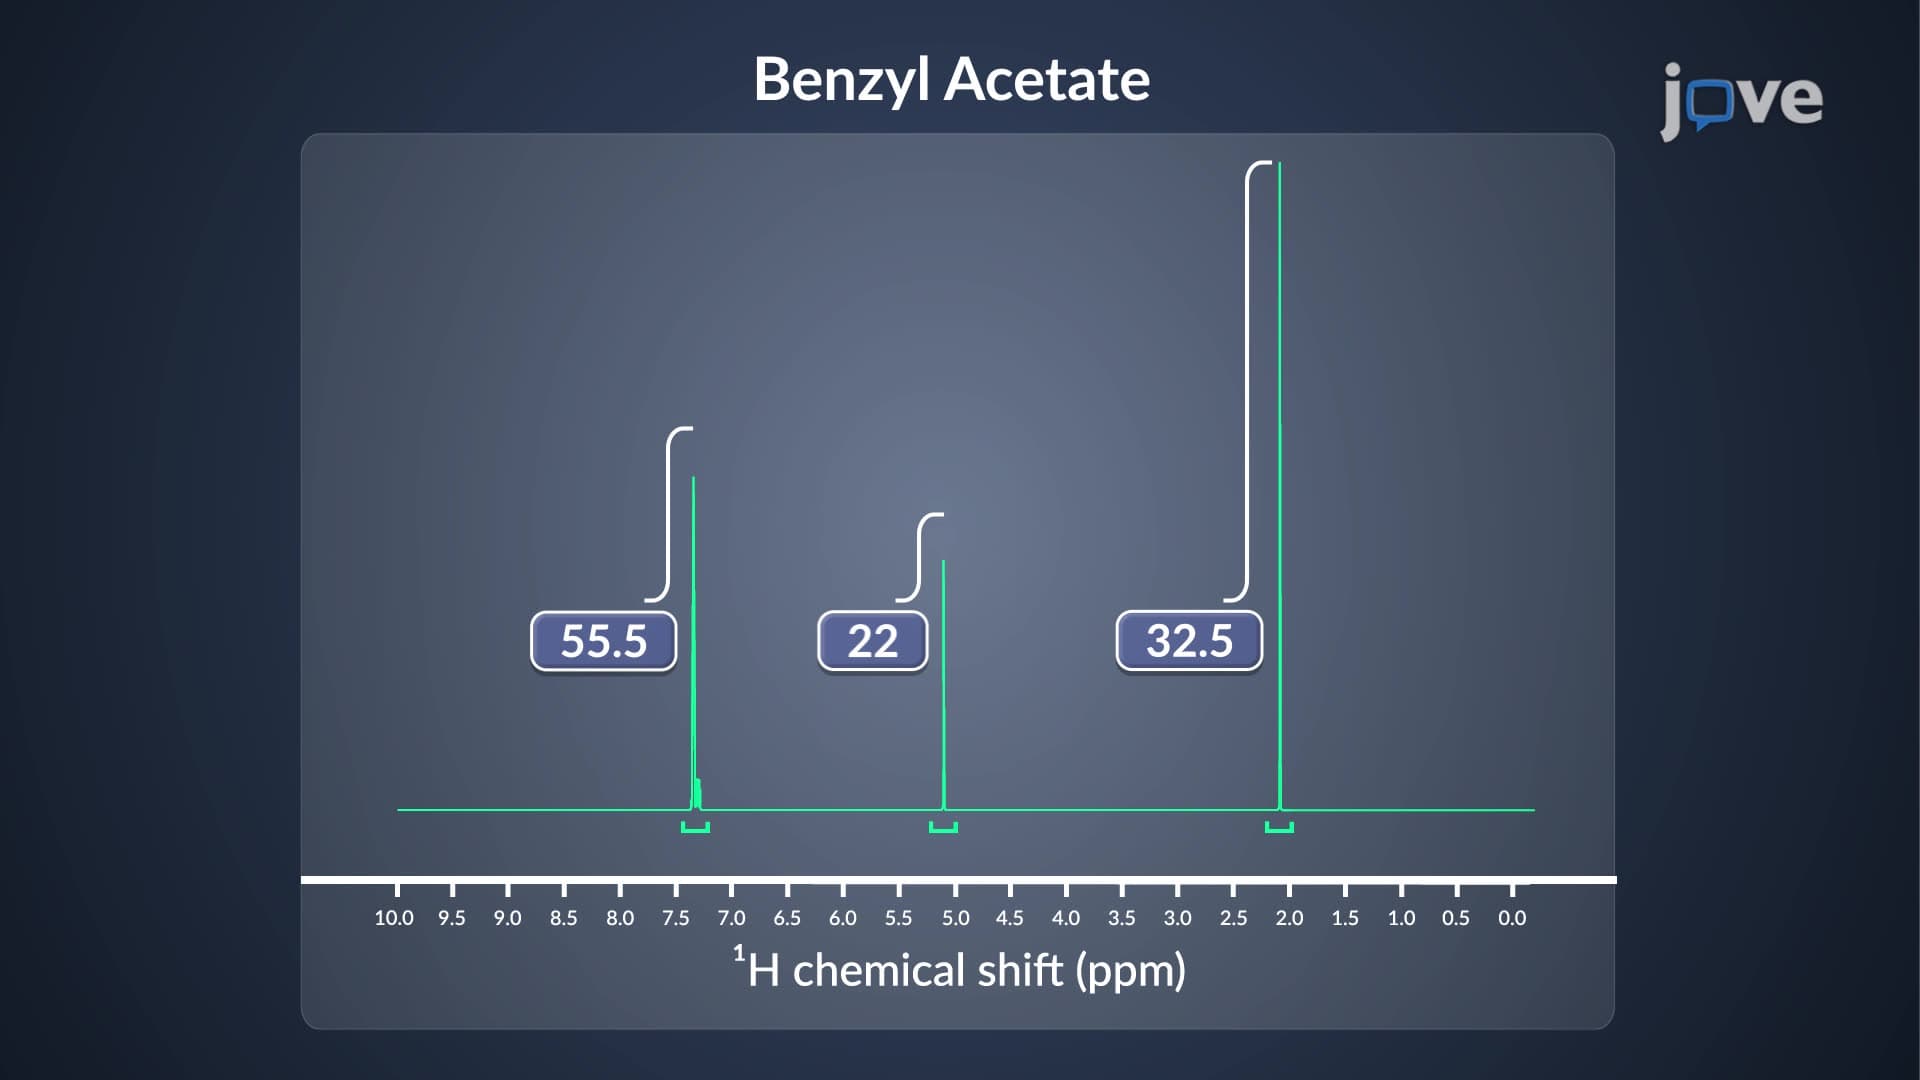
¹H NMR Signal Integration: Overview
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...