Related Experiment Video
Updated: Jun 6, 2026

Targeting Alpha Synuclein Aggregates in Cutaneous Peripheral Nerve Fibers by Free-floating Immunofluorescence Assay
Published on: June 25, 2019
Late form of Pompe disease with glycogen storage in peripheral nerves axons
Anna Fidziańska1, Agnieszka Ługowska, Anna Tylki-Szymańska
1Neuromuscular Unit. MRC. Polish Academy of Science, Warsaw, Poland.

Abstract:
Pompe disease is caused by the deficiency of acid α-glucosidase (GAA), which degrades glycogen into glucose. Its manifestation is characterized by a broad and continuous spectrum of clinical severity ranging from severe infantile to relatively benign adult form. We describe a 12-year-old girl diagnosed at a presymptomatic stage of late form Pompe disease due to fortuitous detection of an elevated level of serum creatine kinase (CK) at the age of 4. Biopsies were taken from the quadriceps muscle and studied with histological and histochemical techniques, as well as in electron microscope. Sporadic muscle cells showed the accumulation of lysosomal glycogen, suggesting Pompe disease. Interestingly, we found lysosomal bound glycogen, located in the axons of intramuscular nerves. The diagnosis was confirmed by deficient GAA activity in leukocytes. Mutation analysis revealed changes IVS1-13T>G and p.C103G in the GAA gene. The patient was able to obtain enzyme replacement therapy in the early asymptomatic stage of the disease.
More Related Videos
 10:16
In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
 06:35
In Vivo Electrophysiological Measurement of Compound Muscle Action Potential from the Forelimbs in Mouse Models of Motor Neuron Degeneration
Published on: June 15, 2018
Related Concept Videos
 01:22
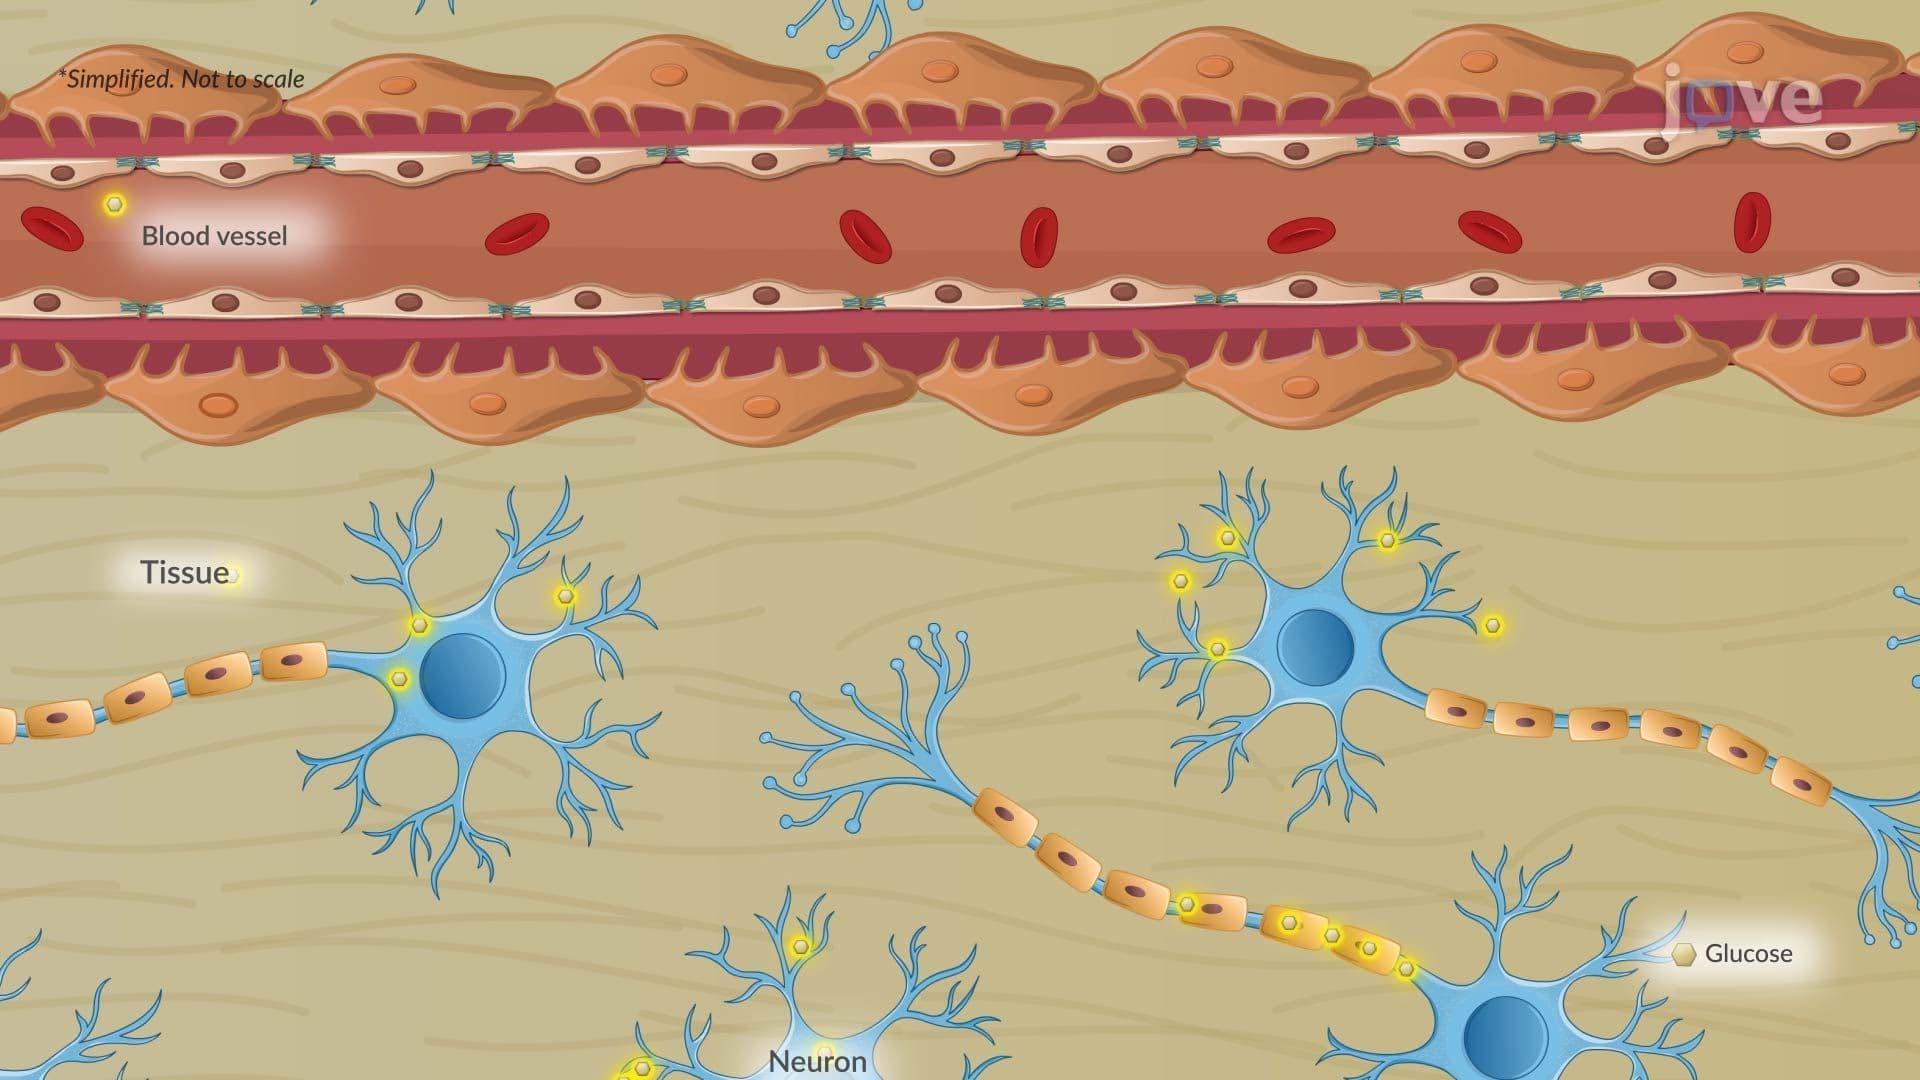
Lysosomal Hydrolases
 01:22
Diabetic Neuropathy
 01:22
Myasthenia Gravis ll: Pathophysiology
 01:15
Parkinson's Disease: Overview
 01:24
Parkinson Disease ll: Pathophysiology
 01:24
Parkinson Disease l: Introduction