Related Experiment Video
Updated: Jun 6, 2026

An In Vitro Model for the Study of Cellular Pathophysiology in Globoid Cell Leukodystrophy
Published on: October 21, 2014
Molecular mechanisms of pathogenesis in a glycosphingolipid and a glycoprotein storage disease
Alessandra d'Azzo1, Erik Bonten
1Department of Genetics, St. Jude Children's Research Hospital, 262 Danny Thomas Place, Memphis, TN 38105-3678, USA. Alessandra.dazzo@stjude.org

Abstract:
The lysosomal system comprises a specialized network of organelles crucial for the sorting, digestion, recycling and secretion of cellular components. With their content of hydrolytic enzymes, lysosomes regulate the degradation of a multitude of substrates that reach these organelles via the biosynthetic or the endocytic route. Gene defects that affect one or more of these hydrolases lead to LSDs (lysosomal storage diseases). This underscores the apparent lack of redundancy of these enzymes and the importance of the lysosomal system in cell and tissue homoeostasis. Some of the lysosomal enzymes may form multiprotein complexes, which usually work synergistically on substrates and, in this configuration, may respond more efficiently to changes in substrate load and composition. A well-characterized lysosomal multienzyme complex is the one comprising the glycosidases β-gal (β-galactosidase) and NEU1 (neuramidase-1), and of the serine carboxypeptidase PPCA (protective protein/cathepsin A). Three neurodegenerative LSDs are caused by either single or combined deficiency of these lysosomal enzymes. Sialidosis (NEU1 deficiency) and galactosialidosis (combined NEU1 and β-gal deficiency, secondary to a primary defect of PPCA) belong to the glycoprotein storage diseases, whereas GM1-gangliosidosis (β-gal deficiency) is a glycosphingolipid storage disease. Identification of novel molecular pathways that are deregulated because of loss of enzyme activity and/or accumulation of specific metabolites in various cell types has shed light on mechanisms of disease pathogenesis and may pave the way for future development of new therapies for these LSDs.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:05
Proteoglycans
 01:24
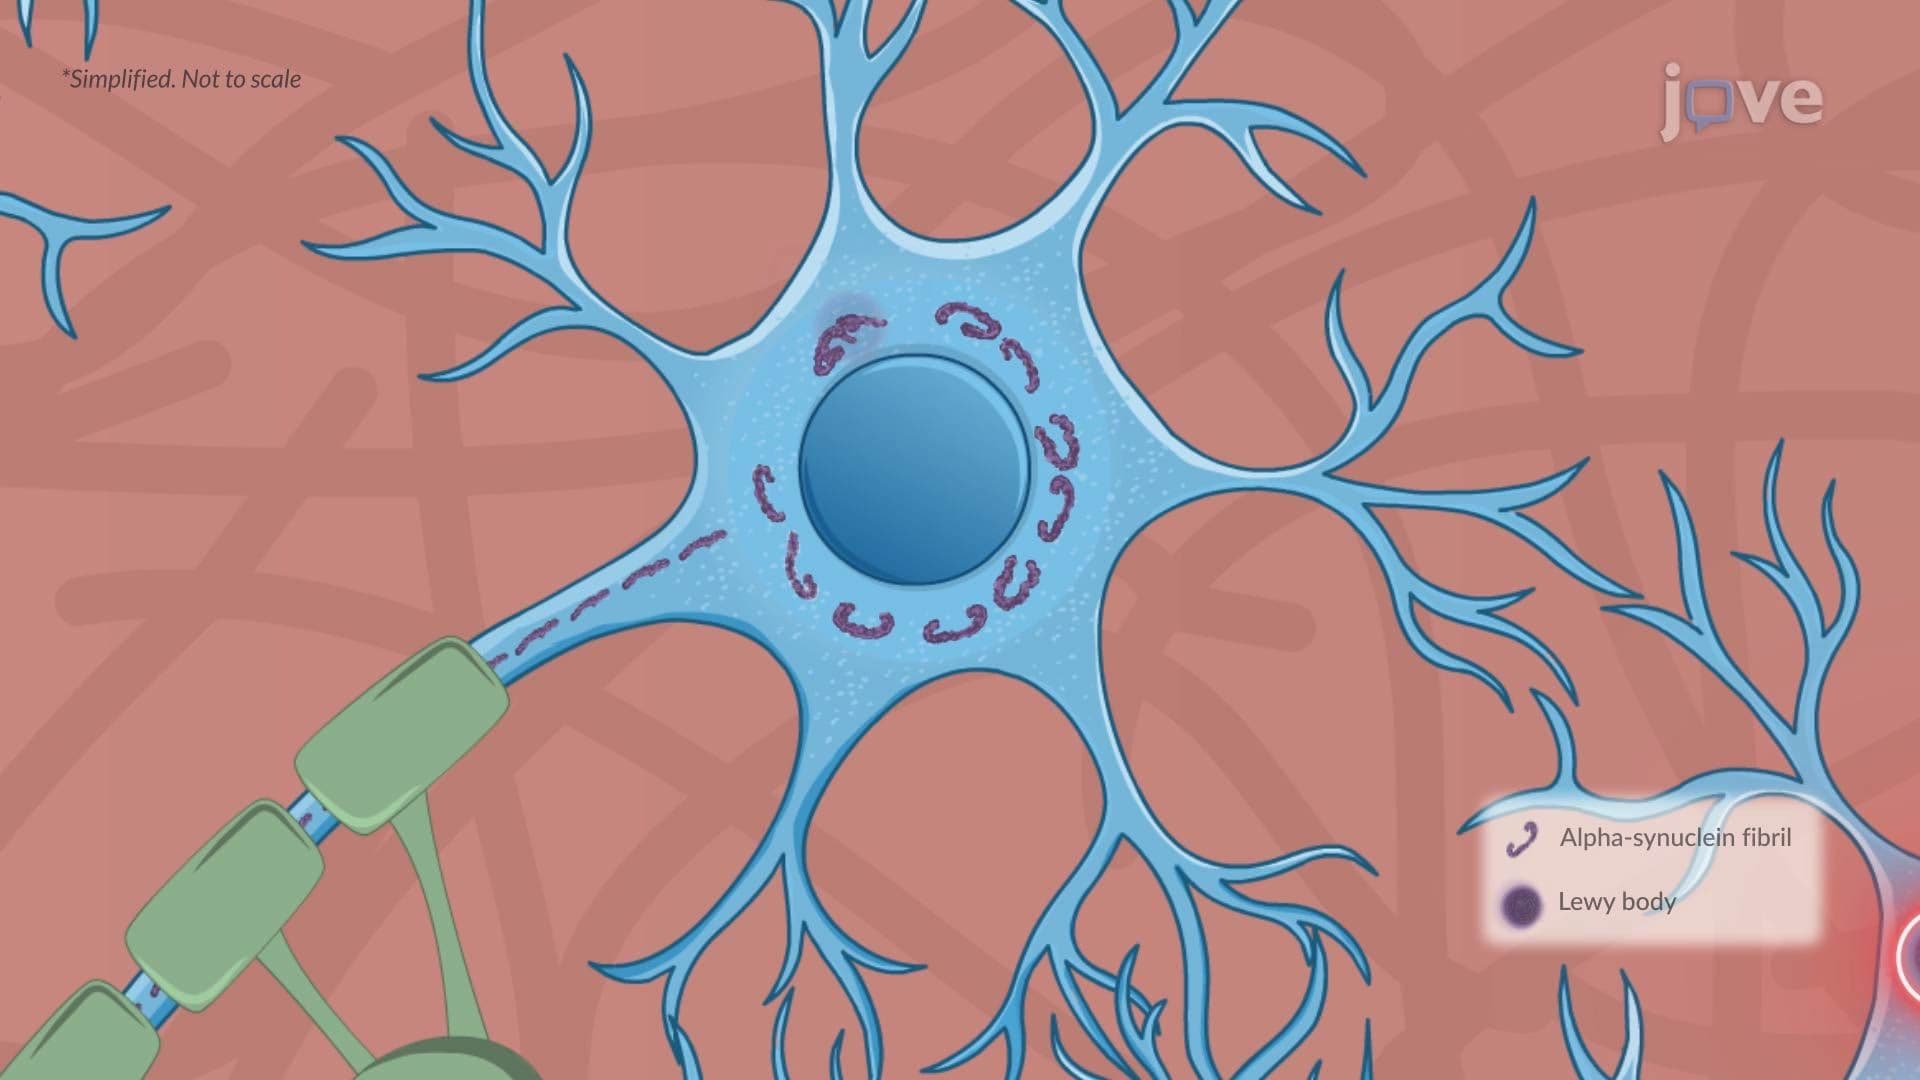
Parkinson Disease ll: Pathophysiology
 01:24
Oligosaccharide Assembly
Multiple sugar molecules that may or may...
 01:23
Alzheimer Disease ll: Pathophysiology
 01:20
Inborn Errors of Metabolism