Related Experiment Video
Updated: Jun 5, 2026
Fractionation for Resolution of Soluble and Insoluble Huntingtin Species
Published on: February 27, 2018
Huntington's disease: from molecular pathogenesis to clinical treatment
Christopher A Ross1, Sarah J Tabrizi
1Departments of Psychiatry, Neurology, Pharmacology, and Neuroscience, and Program in Cellular and Molecular Medicine, Johns Hopkins University School of Medicine, Baltimore, MD, USA. ca.ross@jhu.edu

Insights
Huntington's disease, a neurodegenerative disorder, is caused by a genetic CAG repeat expansion. Research reveals neurodegeneration begins years before symptoms, offering new therapeutic targets.
Area of Science:
- Neuroscience
- Genetics
- Molecular Biology
Background:
- Huntington's disease (HD) is a fatal, progressive neurodegenerative disorder.
- It stems from an expanded CAG repeat in the huntingtin gene, leading to an abnormal polyglutamine repeat in the huntingtin protein.
- HD serves as a crucial model for understanding other neurodegenerative diseases like Alzheimer's and Parkinson's.
Observation:
- Shared features among neurodegenerative disorders include delayed onset, selective neuronal vulnerability, and abnormal protein aggregation.
- Cellular toxicity involves both intrinsic (cell autonomous) and intercellular mechanisms.
- Neurodegeneration in HD initiates years before clinical symptoms, accompanied by subtle prodromal changes.
Findings:
- Pathogenic pathways underlying Huntington's disease are increasingly understood.
- Predictive genetic testing and neuroimaging confirm early-onset neurodegeneration.
- Subtle cognitive, motor, and psychiatric alterations precede overt disease manifestation.
Implications:
- Understanding HD's pathogenic pathways provides targets for novel therapeutic interventions.
- HD serves as a model for developing strategies to delay or prevent disease onset, not just manage symptoms.
- Early detection and intervention hold promise for altering the course of neurodegenerative diseases.
Abstract:
Huntington's disease is a progressive, fatal, neurodegenerative disorder caused by an expanded CAG repeat in the huntingtin gene, which encodes an abnormally long polyglutamine repeat in the huntingtin protein. Huntington's disease has served as a model for the study of other more common neurodegenerative disorders, such as Alzheimer's disease and Parkinson's disease. These disorders all share features including: delayed onset; selective neuronal vulnerability, despite widespread expression of disease-related proteins during the whole lifetime; abnormal protein processing and aggregation; and cellular toxic effects involving both cell autonomous and cell-cell interaction mechanisms. Pathogenic pathways of Huntington's disease are beginning to be unravelled, offering targets for treatments. Additionally, predictive genetic testing and findings of neuroimaging studies show that, as in some other neurodegenerative disorders, neurodegeneration in affected individuals begins many years before onset of diagnosable signs and symptoms of Huntington's disease, and it is accompanied by subtle cognitive, motor, and psychiatric changes (so-called prodromal disease). Thus, Huntington's disease is also emerging as a model for strategies to develop therapeutic interventions, not only to slow progression of manifest disease but also to delay, or ideally prevent, its onset.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:24
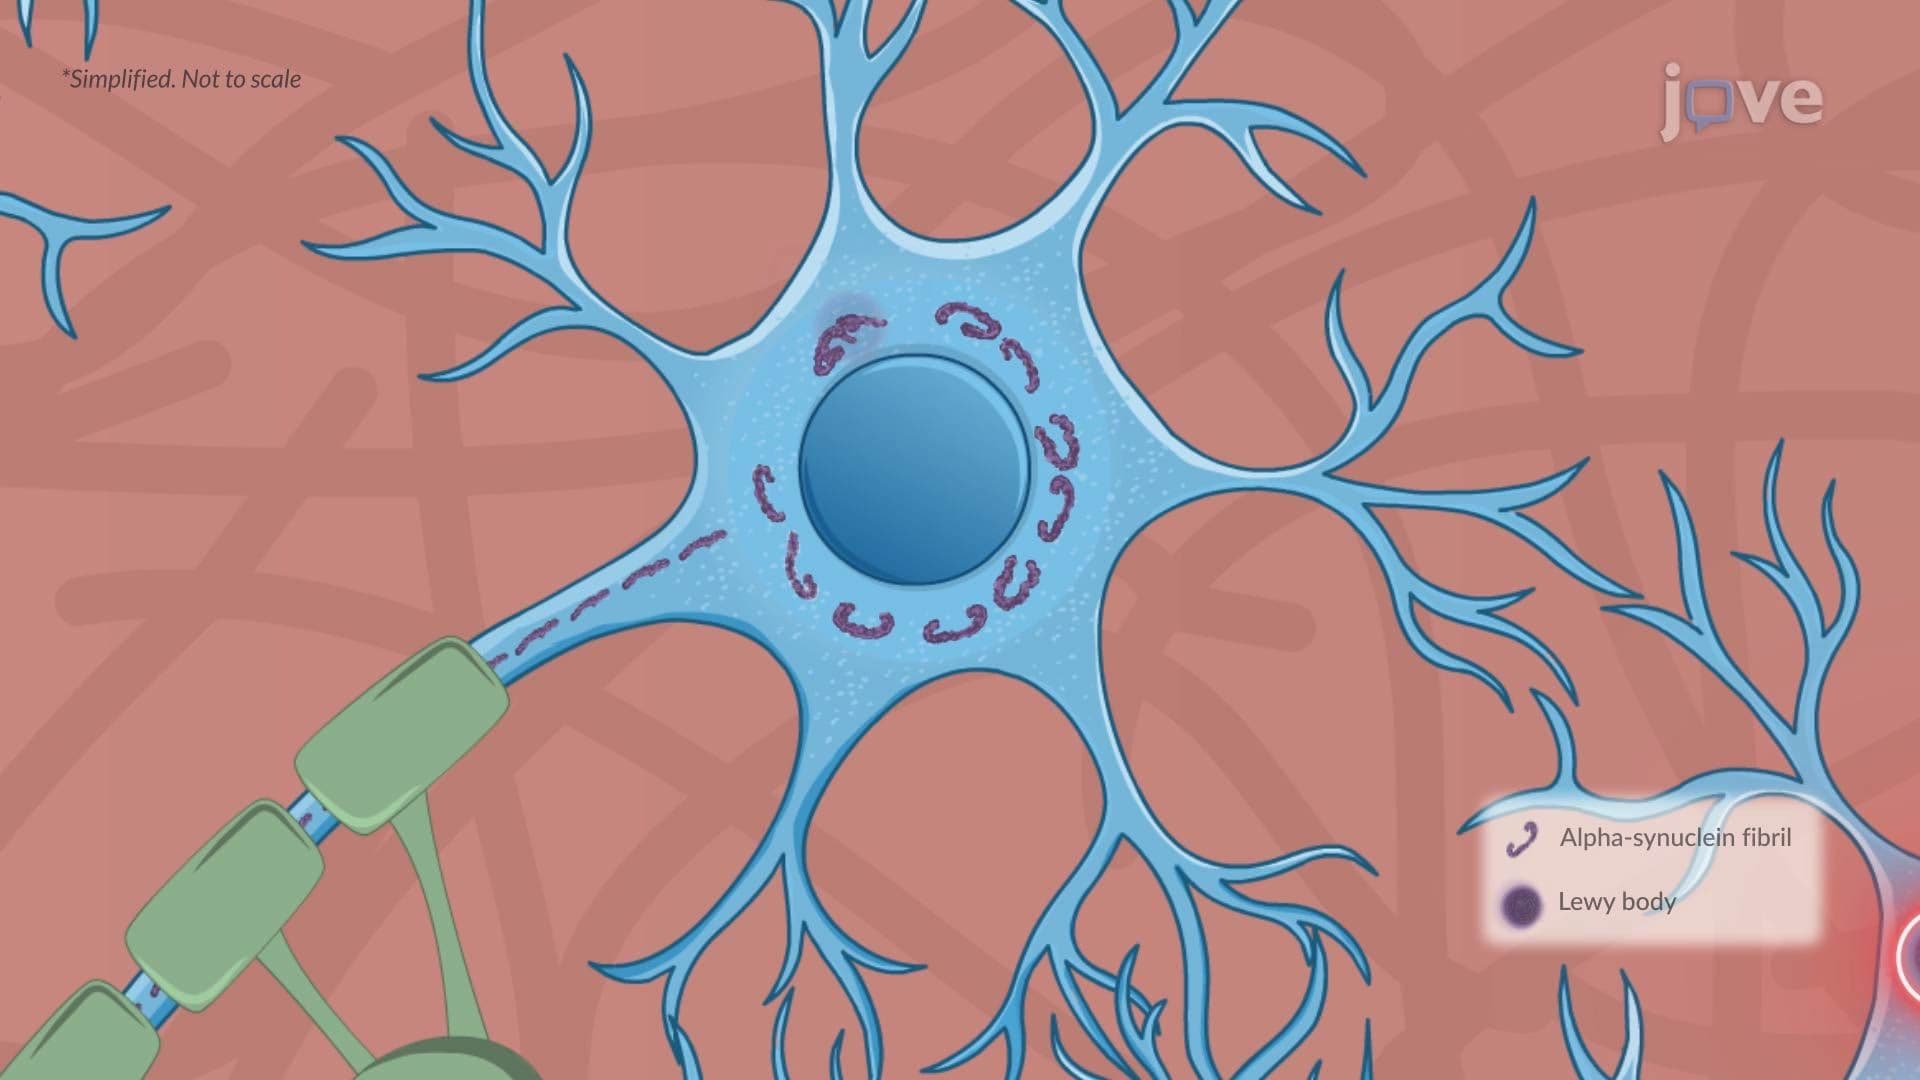
Parkinson Disease ll: Pathophysiology
 01:15
Parkinson's Disease: Overview
 01:24
Parkinson's Disease: Treatment
Parkinson's Disease is primarily a result of the loss of dopaminergic neurons in the substantia nigra pars compacta. The cornerstone of its...
 01:23
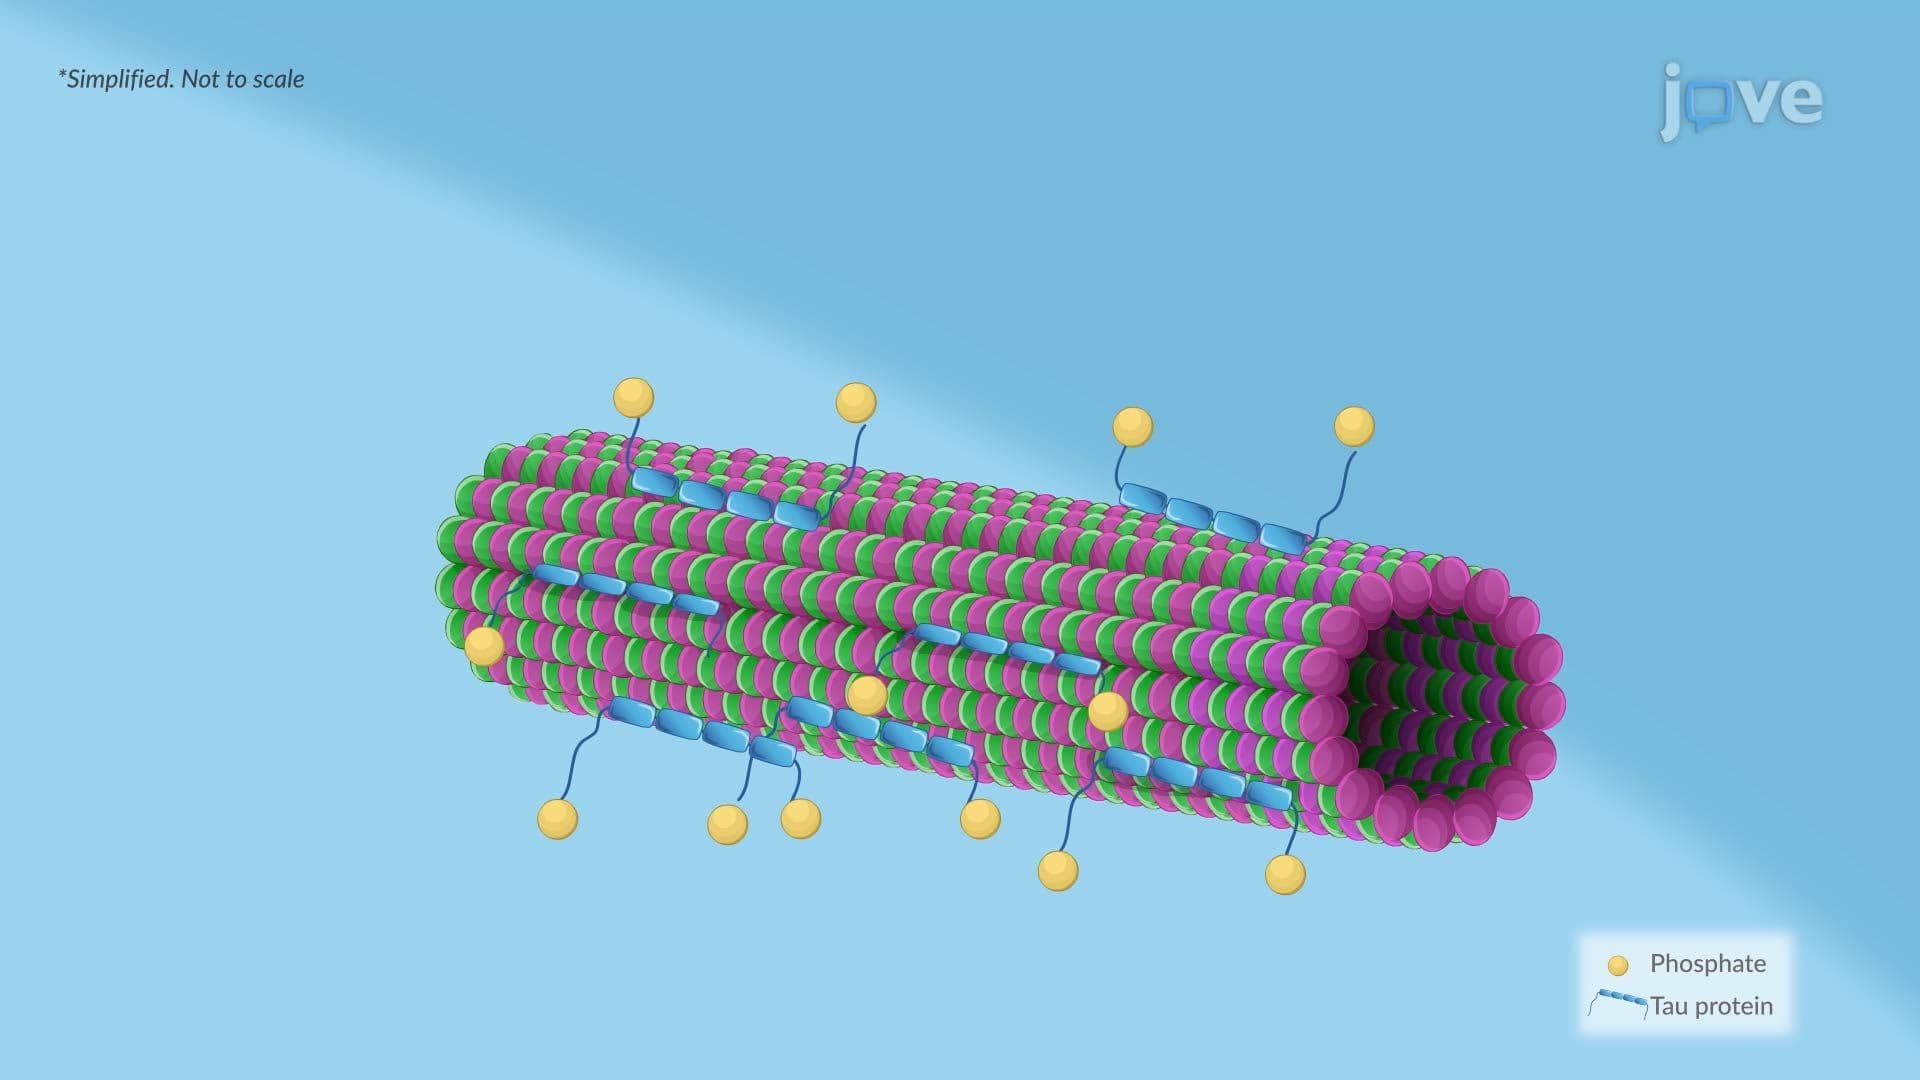
Alzheimer Disease ll: Pathophysiology
 01:22
Alzheimer's Disease: Treatment