Related Experiment Video
Updated: Jun 5, 2026
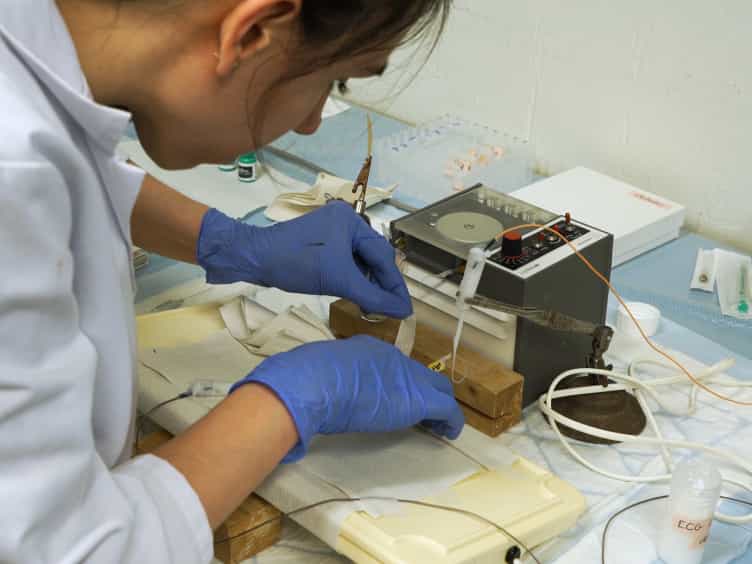

Nasal Potential Difference to Quantify Trans-epithelial Ion Transport in Mice
Published on: July 4, 2018
Chronic ataluren (PTC124) treatment of nonsense mutation cystic fibrosis
M Wilschanski1, L L Miller, D Shoseyov
1Paediatric Gastroenterology, Hadassah University Hospital, Mount Scopus POB 24035, Jerusalem, 91240, Israel. michaelwil@hadassah.org.il

Abstract:
In a subset of patients with cystic fibrosis (CF), nonsense mutations (premature stop codons) disrupt production of full-length, functional CF transmembrane conductance regulator (CFTR). Ataluren (PTC124) allows ribosomal readthrough of premature stop codons in mRNA. We evaluated drug activity and safety in patients with nonsense mutation CF who took ataluren three times daily (morning, midday and evening) for 12 weeks at either a lower dose (4, 4 and 8 mg·kg(-1)) or higher dose (10, 10 and 20 mg·kg(-1)). The study enrolled 19 patients (10 males and nine females aged 19-57 yrs; dose: lower 12, higher seven) with a classic CF phenotype, at least one CFTR nonsense mutation allele, and an abnormal nasal total chloride transport. Both ataluren doses were similarly active, improving total chloride transport with a combined mean change of -5.4 mV (p<0.001), and on-treatment responses (at least -5 mV improvement) and hyperpolarisations (values more electrically negative than -5 mV) in 61% (p<0.001) and 56% (p = 0.002) of patients. CFTR function was greater with time and was accompanied by trends toward improvements in pulmonary function and CF-related coughing. Adverse clinical and laboratory findings were uncommon and usually mild. Chronic ataluren administration produced time-dependent improvements in CFTR activity and clinical parameters with generally good tolerability.
Insights
Ataluren treatment helped patients with cystic fibrosis (CF) who have nonsense mutations. The drug improved CF transmembrane conductance regulator (CFTR) function and showed promising trends in lung function and cough relief.
Area of Science:
- Medical Research
- Pharmacology
- Genetics
Background:
- Nonsense mutations in cystic fibrosis (CF) patients prevent functional CF transmembrane conductance regulator (CFTR) protein production.
- Ataluren is a drug designed to enable ribosomal readthrough of these premature stop codons.
Purpose of the Study:
- To evaluate the efficacy and safety of ataluren in CF patients with nonsense mutations.
- To assess the drug's effect on CFTR function and clinical parameters over 12 weeks.
Main Methods:
- A 12-week study involving 19 CF patients with nonsense mutations.
- Patients received either a lower or higher dose of ataluren three times daily.
- Nasal total chloride transport was measured to assess CFTR function.
Main Results:
- Both ataluren doses significantly improved nasal total chloride transport (mean change -5.4 mV, p<0.001).
- 61% of patients showed treatment response and 56% achieved hyperpolarization, indicating improved CFTR activity.
- Trends towards improved pulmonary function and reduced CF-related coughing were observed.
Conclusions:
- Chronic ataluren administration demonstrated time-dependent improvements in CFTR activity.
- The drug was generally well-tolerated with uncommon and mild adverse events.
- Ataluren shows potential as a therapeutic option for CF patients with specific genetic mutations.
More Related Videos
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 00:59
Gene Therapy
 01:26
Treatment for Pulmonary Arterial Hypertension: Receptor Tyrosine Kinase Inhibitors and Calcium Channel Blockers
TKIs, such as imatinib (Gleevec), are particularly effective in tackling the growth and mitogenic factors that become upregulated in PAH patients. These factors contribute to the...