Related Experiment Video
Updated: Jun 5, 2026
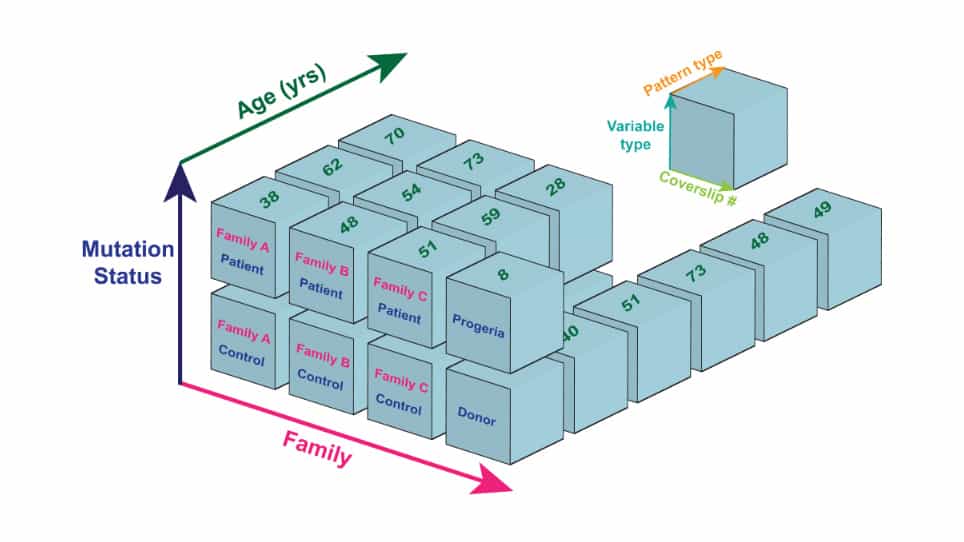

Databases to Efficiently Manage Medium Sized, Low Velocity, Multidimensional Data in Tissue Engineering
Published on: November 22, 2019
[Three cases of Hutchinson-Gilford progeria syndrome]
Y Doubaj1, A Lamzouri, S-C Elalaoui
1Département de génétique médicale, Institut national d'hygiène, 27, avenue Ibn-Batouta, BP 769, Rabat, Maroc. y.doubaj@gmail.com

Insights
Hutchinson-Gilford progeria syndrome (HGPS) is a rare genetic disorder causing accelerated aging. Genetic testing confirmed the mutation in only one of three suspected pediatric cases, highlighting diagnostic challenges.
Area of Science:
- Genetics
- Pediatrics
- Rare Diseases
Background:
- Hutchinson-Gilford progeria syndrome (HGPS) is a rare genetic disorder characterized by accelerated aging in children.
- HGPS presents with distinct clinical features including failure to thrive, alopecia, and severe atherosclerosis, leading to early mortality from cardiovascular complications.
Observation:
- Three pediatric patients aged 5, 11, and 12 years presented with dysmorphic facies and failure to thrive, prompting genetic consultation.
- Clinical assessment raised suspicion for progeria syndrome in all three patients.
Findings:
- Molecular analysis identified the recurrent LMNA gene mutation (c.1824C>T; p.Gly608Gly) in only one of the three patients.
- This case series highlights variability in clinical presentation and the diagnostic utility of genetic testing for HGPS.
Implications:
- Geneticists play a crucial role in diagnosing rare genetic syndromes like HGPS and providing essential genetic counseling.
- Understanding the clinical spectrum and diagnostic criteria for HGPS is vital for accurate and timely diagnosis in affected children.
Abstract:
Progeria, or Hutchinson-Gilford syndrome, is a rare genetic disease, characterized by several clinical features that develop in childhood, in particular, an accelerated aging aspect. Its incidence is 1-4 per 8 million newborns. Children with progeria syndrome usually appear normal at birth and in early infancy. Profound failure to thrive occurs during the 1st year. Characteristic facies, partial alopecia progressing to total alopecia, loss of subcutaneous fat, stiffness of joints, bone changes, and abnormal tightness of the skin over the abdomen and upper thighs usually become apparent during the 2nd to 3rd years. Motor and mental development is normal. Patients develop severe atherosclerosis. Death occurs as a result of complications of cardiac or cerebrovascular disease (heart attack or stroke) generally between ages 6 and 20 years. The diagnosis of Hutchinson-Gilford progeria syndrome (HGPS) is based on recognition of common clinical features and the detection of the recurrent p.Gly608Gly mutation in exon 11 of the LMNA gene, which is present in almost all individuals with HGPS. We present here 3 patients aged 5, 11, and 12 years referred to genetic consultation for dysmorphic facies and failure to thrive. After careful clinical examination and paraclinical assessment, the diagnosis of progeria syndrome was raised. We performed molecular analysis for the 3 patients by searching for the recurrent mutation c.1824C>T (p.Gly608Gly) of the LMNA gene, which was found only in 1 patient. We discuss the geneticist's role in the diagnosis of rare dysmorphic syndromes and their genetic counseling. We also analyze the clinical spectrum of HGPS by comparing the 3 patients.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 02:18
Alternative RNA Splicing
There are five types of alternative RNA splicing that vary in the ways the pre-mRNA segments are removed or retained in the mature mRNA. The first...
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:43
Sex-linked Disorders
 01:33
Pleiotropy
 01:20
The Retinoblastoma Gene
The first-ever tumor suppressor gene called Rb was identified in retinoblastoma - a rare eye tumor in children. In inherited forms of the disease, a child inherits one defective copy of the Rb gene, which predisposes them to retinoblastoma. However,...