Related Experiment Video
Updated: Jun 4, 2026

Expression and Purification of Mammalian Bestrophin Ion Channels
Published on: August 2, 2018
Functional characterization of bestrophin-1 missense mutations associated with autosomal recessive bestrophinopathy
Alice E Davidson1, Ian D Millar, Rosemary Burgess-Mullan
1School of Biomedicine, The University of Manchester, Manchester Academic Health Science Centre, Central Manchester University Hospitals, NHS Foundation Trust, Manchester, United Kingdom.

Purpose:
Autosomal recessive bestrophinopathy (ARB) is a retinal dystrophy affecting macular and retinal pigmented epithelium function resulting from homozygous or compound heterozygous mutations in BEST1. In this study we characterize the functional implications of missense bestrophin-1 mutations that cause ARB by investigating their effect on bestrophin-1's chloride conductance, cellular localization, and stability.
Methods:
The chloride conductance of wild-type bestropin-1 and a series of ARB mutants were determined by whole-cell patch-clamping of transiently transfected HEK cells. The effect of ARB mutations on the cellular localization of bestrophin-1 was determined by confocal immunofluorescence on transiently transfected MDCK II cells that had been polarized on Transwell filters. Protein stability of wild-type and ARB mutant forms of bestrophin-l was determined by the addition of proteasomal or lysosomal inhibitors to transiently transfected MDCK II cells. Lysates were then analyzed by Western blot analysis.
Results:
All ARB mutants investigated produced significantly smaller chloride currents compared to wild-type bestrophin-1. Additionally, co-transfection of compound heterozygous mutants abolished chloride conductance in contrast to co-transfections of a single mutant with wild-type bestrophin-l, reflecting the recessive nature of the condition. In control experiments, expression of two dominant vitelliform macular dystrophy mutants was shown to inhibit wild-type currents. Cellular localization of ARB mutants demonstrated that the majority did not traffic correctly to the plasma membrane and that five of these seven mutants were rapidly degraded by the proteasome. Two ARB-associated mutants (p.D312N and p.V317M) that were not trafficked correctly nor targeted to the proteasome had a distinctive appearance, possibly indicative of aggresome or aggresome-like inclusion bodies.
Conclusions:
Differences in cellular processing mechanisms for different ARB associated mutants lead to the same disease phenotype. The existence of distinct pathogenic disease mechanisms has important ramifications for potential gene replacement therapies since we show that missense mutations associated with an autosomal recessive disease have a pathogenic influence beyond simple loss of function.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:48
ATP Synthase: Mechanism
 01:18
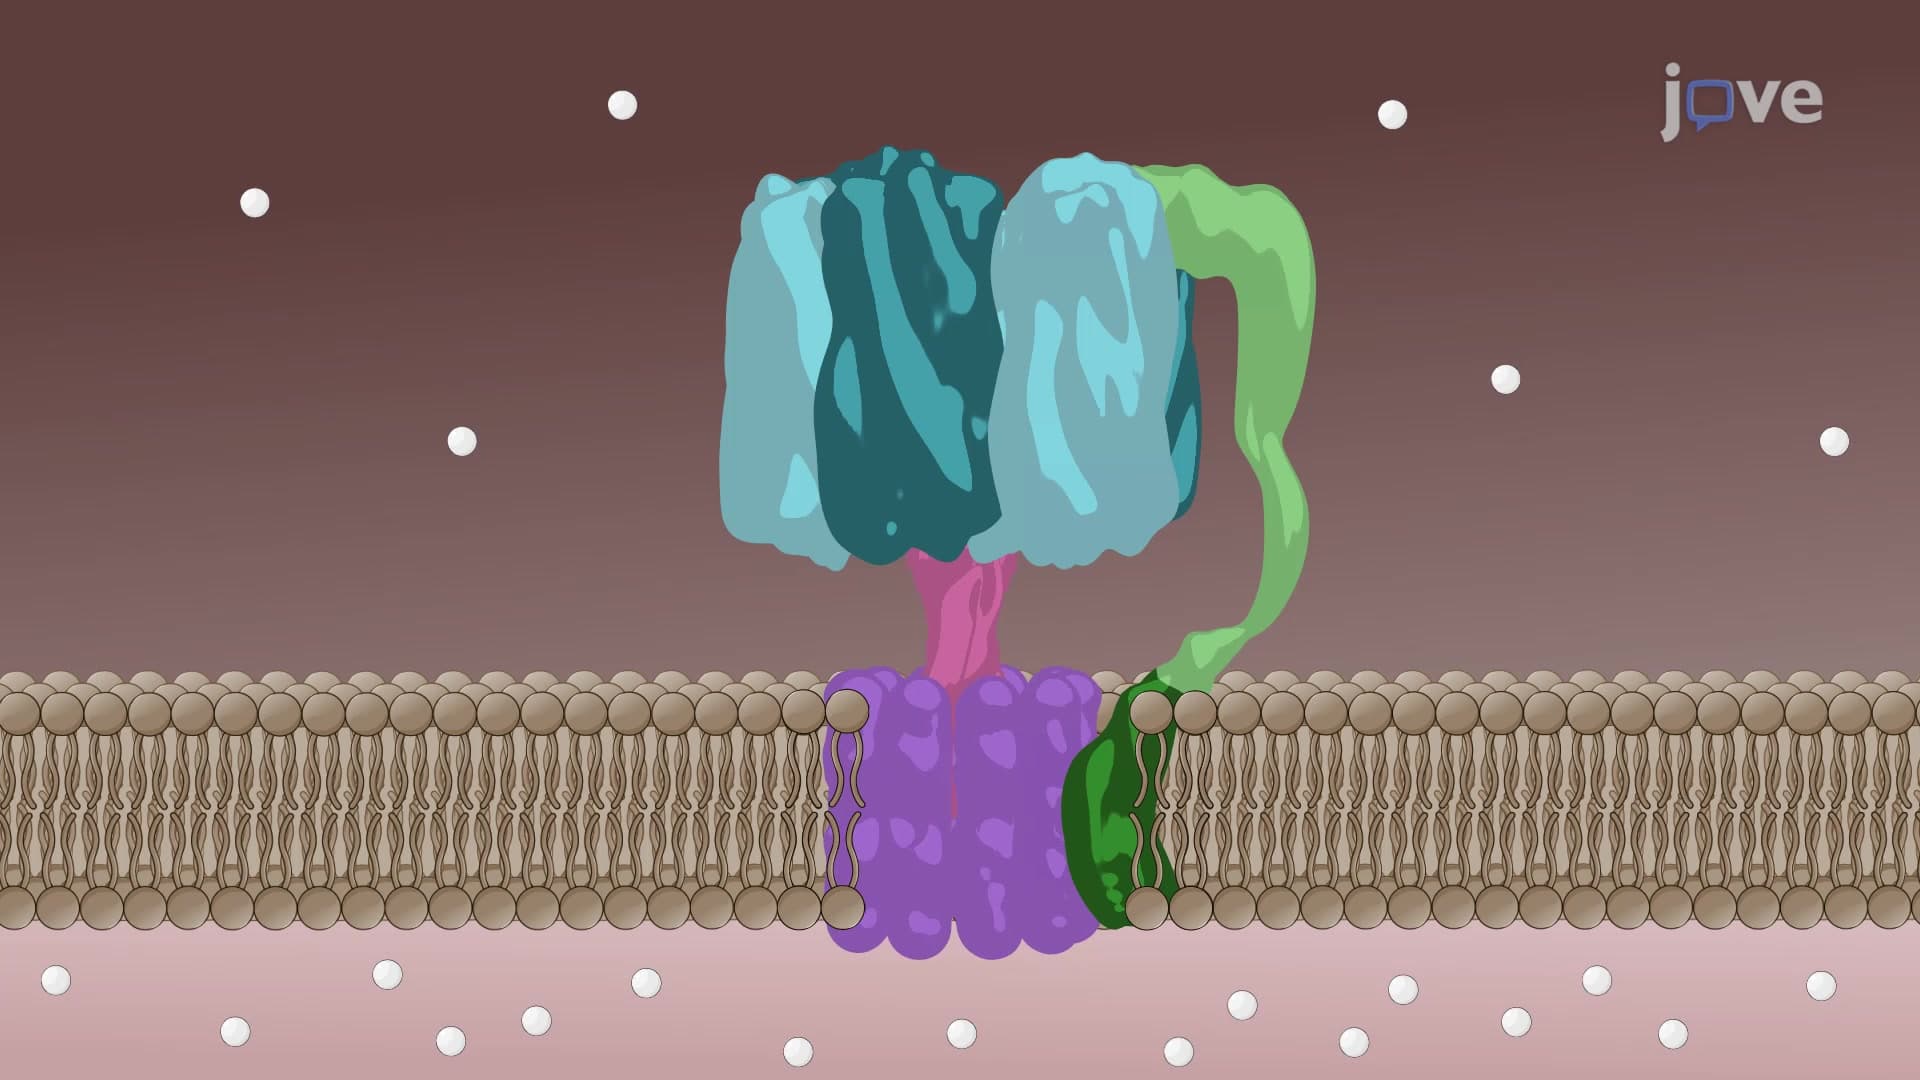
ATP Synthase: Structure
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life
01:31Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life
 01:39
Mutations