Related Experiment Video
Updated: Jun 4, 2026


Identification and Classification of Position-specific GABAA Receptor Subunit Missense Variants for Their Role In Hippocampal Pyramidal Neurons
Published on: June 6, 2025
Structural insights into differences in drug-binding selectivity between two forms of human alpha1-acid glycoprotein
Koji Nishi1, Tomomi Ono, Teruya Nakamura
1Department of Biopharmaceutics, Graduate School of Pharmaceutical Sciences, Kumamoto University, Kumamoto 862-0973, Japan.

Abstract:
Human α(1)-acid glycoprotein (hAGP) in serum functions as a carrier of basic drugs. In most individuals, hAGP exists as a mixture of two genetic variants, the F1*S and A variants, which bind drugs with different selectivities. We prepared a mutant of the A variant, C149R, and showed that its drug-binding properties were indistinguishable from those of the wild type. In this study, we determined the crystal structures of this mutant hAGP alone and complexed with disopyramide (DSP), amitriptyline (AMT), and the nonspecific drug chlorpromazine (CPZ). The crystal structures revealed that the drug-binding pocket on the A variant is located within an eight-stranded β-barrel, similar to that found in the F1*S variant and other lipocalin family proteins. However, the binding region of the A variant is narrower than that of the F1*S variant. In the crystal structures of complexes with DSP and AMT, the two aromatic rings of each drug interact with Phe-49 and Phe-112 at the bottom of the binding pocket. Although the structure of CPZ is similar to those of DSP and AMT, its fused aromatic ring system, which is extended in length by the addition of a chlorine atom, appears to dictate an alternative mode of binding, which explains its nonselective binding to the F1*S and A variant hAGPs. Modeling experiments based on the co-crystal structures suggest that, in complexes of DSP, AMT, or CPZ with the F1*S variant, Phe-114 sterically hinders interactions with DSP and AMT, but not CPZ.
Related Concept Videos
 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In contrast,...
 01:20
Factors Affecting Protein-Drug Binding: Protein-Related Factors
The physicochemical properties of a drug play a significant role in its ability to bind to proteins. Lipophilic drugs, which dissolve in fats, oils, and lipids, can be bound by...
 01:25
Dose-Response Relationship: Selectivity and Specificity
 01:31
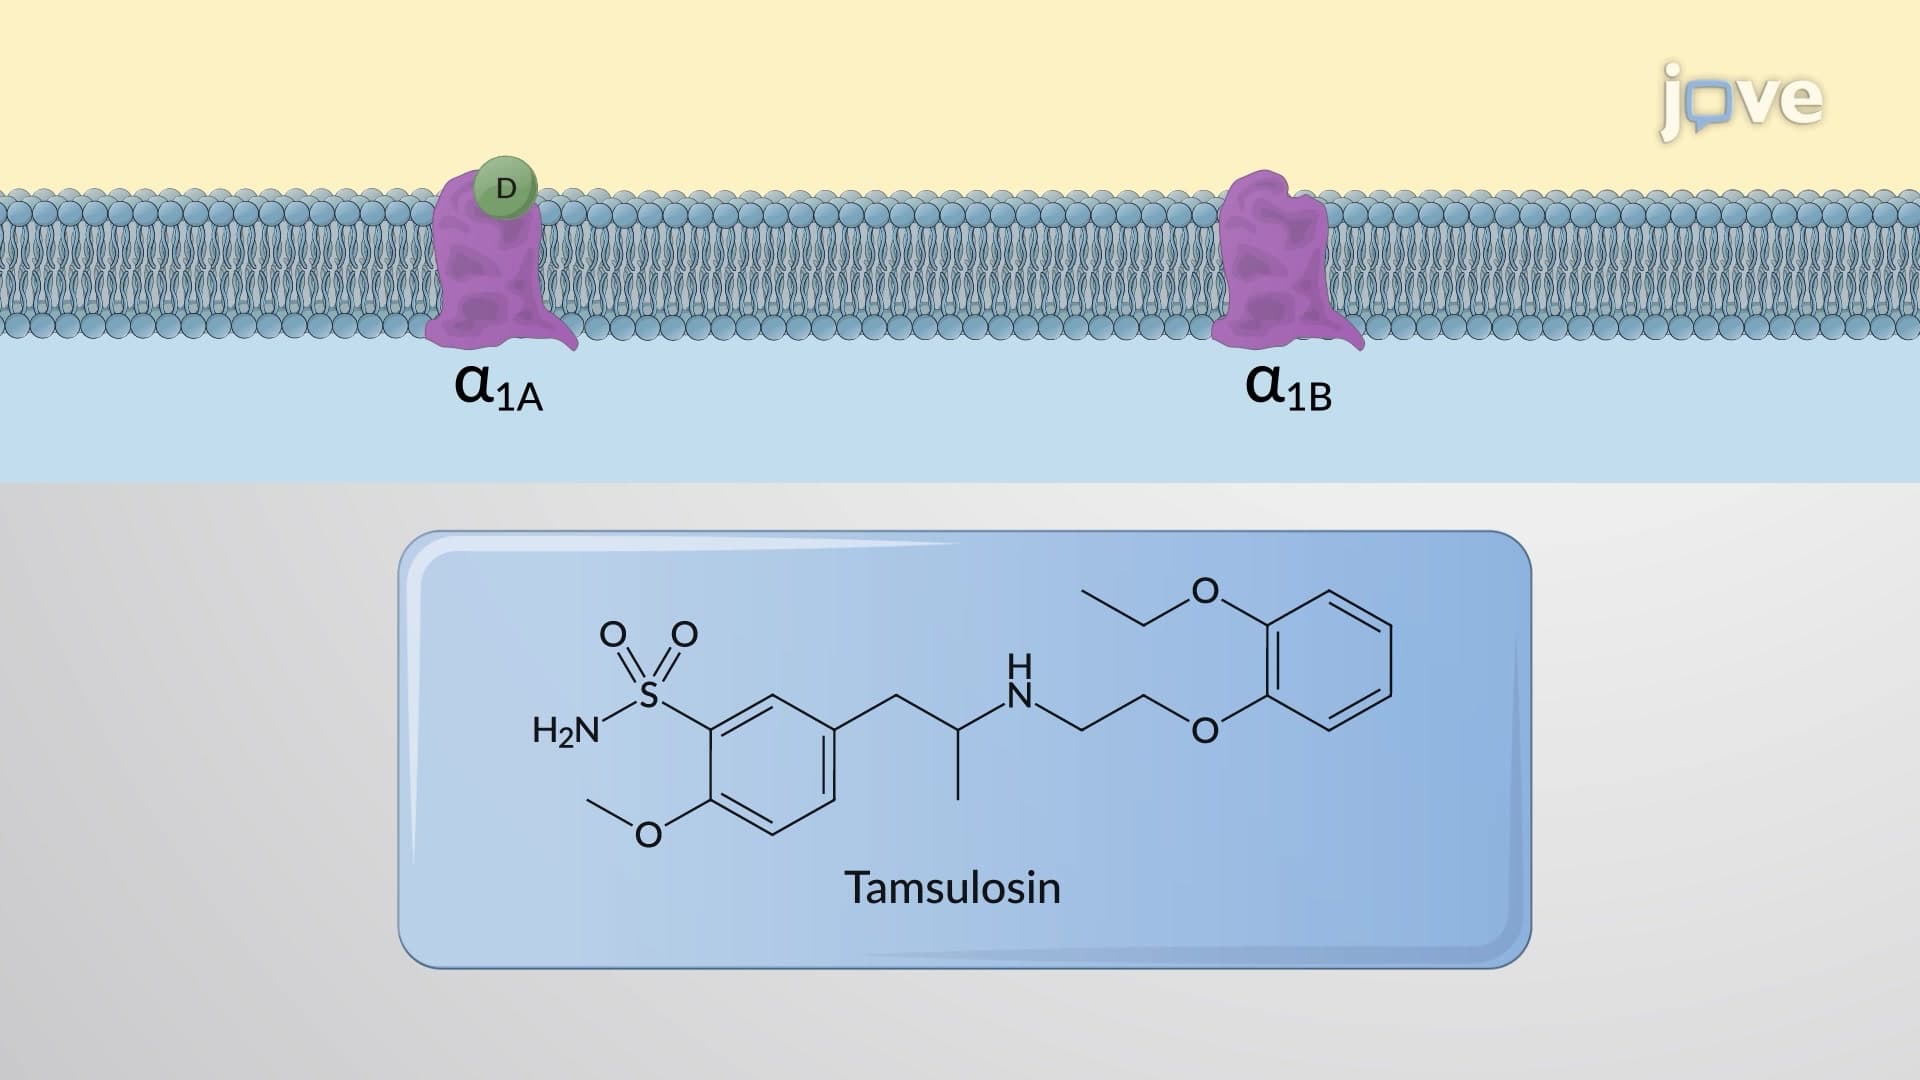
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:29
Factors Affecting Protein-Drug Binding: Patient-Related Factors
Age stands as a key determinant in protein-drug binding. Neonates, characterized by low albumin content, experience heightened concentrations of unbound drugs such as phenytoin and...
 01:30
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...