Related Experiment Video
Updated: Jun 4, 2026
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
P2(2-) and P(3-) units in the [Rh8P9]δ- polyanion of La4Rh8P9
Ulrike Pfannenschmidt1, Dirk Johrendt, Frederik Behrends
1Institut für Anorganische und Analytische Chemie, Universität Münster, Münster, Germany.

Abstract:
The phosphide La(4)Rh(8)P(9) was synthesized from the elements in a bismuth flux. The structure was refined from single crystal diffractometer data: space group Cmcm, a = 1303.1(2), b = 1893.2(2), c = 576.70(6) pm, wR2 = 0.0277, 1380 F(2) values, 65 variables. The rhodium and phosphorus atoms build up a three-dimensional [Rh(8)P(9)] polyanion which leaves larger cages for the three crystallographically independent lanthanum sites. The rhodium atoms have between four and six phosphorus neighbors at Rh-P distance ranging from 229 to 254 pm. Three of the four crystallographically independent phosphorus atoms are isolated (P(3-) units), while the P4 atoms form dimers with double bond character (208 pm P-P). The P(2)(2-) diphosphenide units bond side-on to a Rh3 and end-on to four Rh5 atoms. (31)P magic angle spinning (MAS) NMR spectroscopy is able to resolve three of the four crystallographically distinct phosphorus sites. The doubly bonded phosphorus site P4 is characterized by an axially symmetric shielding tensor of moderate anisotropy Δσ = σ(33) - σ(iso) = 257 ppm. Electronic band structure calculations prove the metallic character and reveal the significant difference between the isolated P(3-) and the phosphorus atoms of the P(2)(2-) units. Magnetic susceptibility measurement reveals Pauli paramagnetism.
Related Concept Videos
 01:18
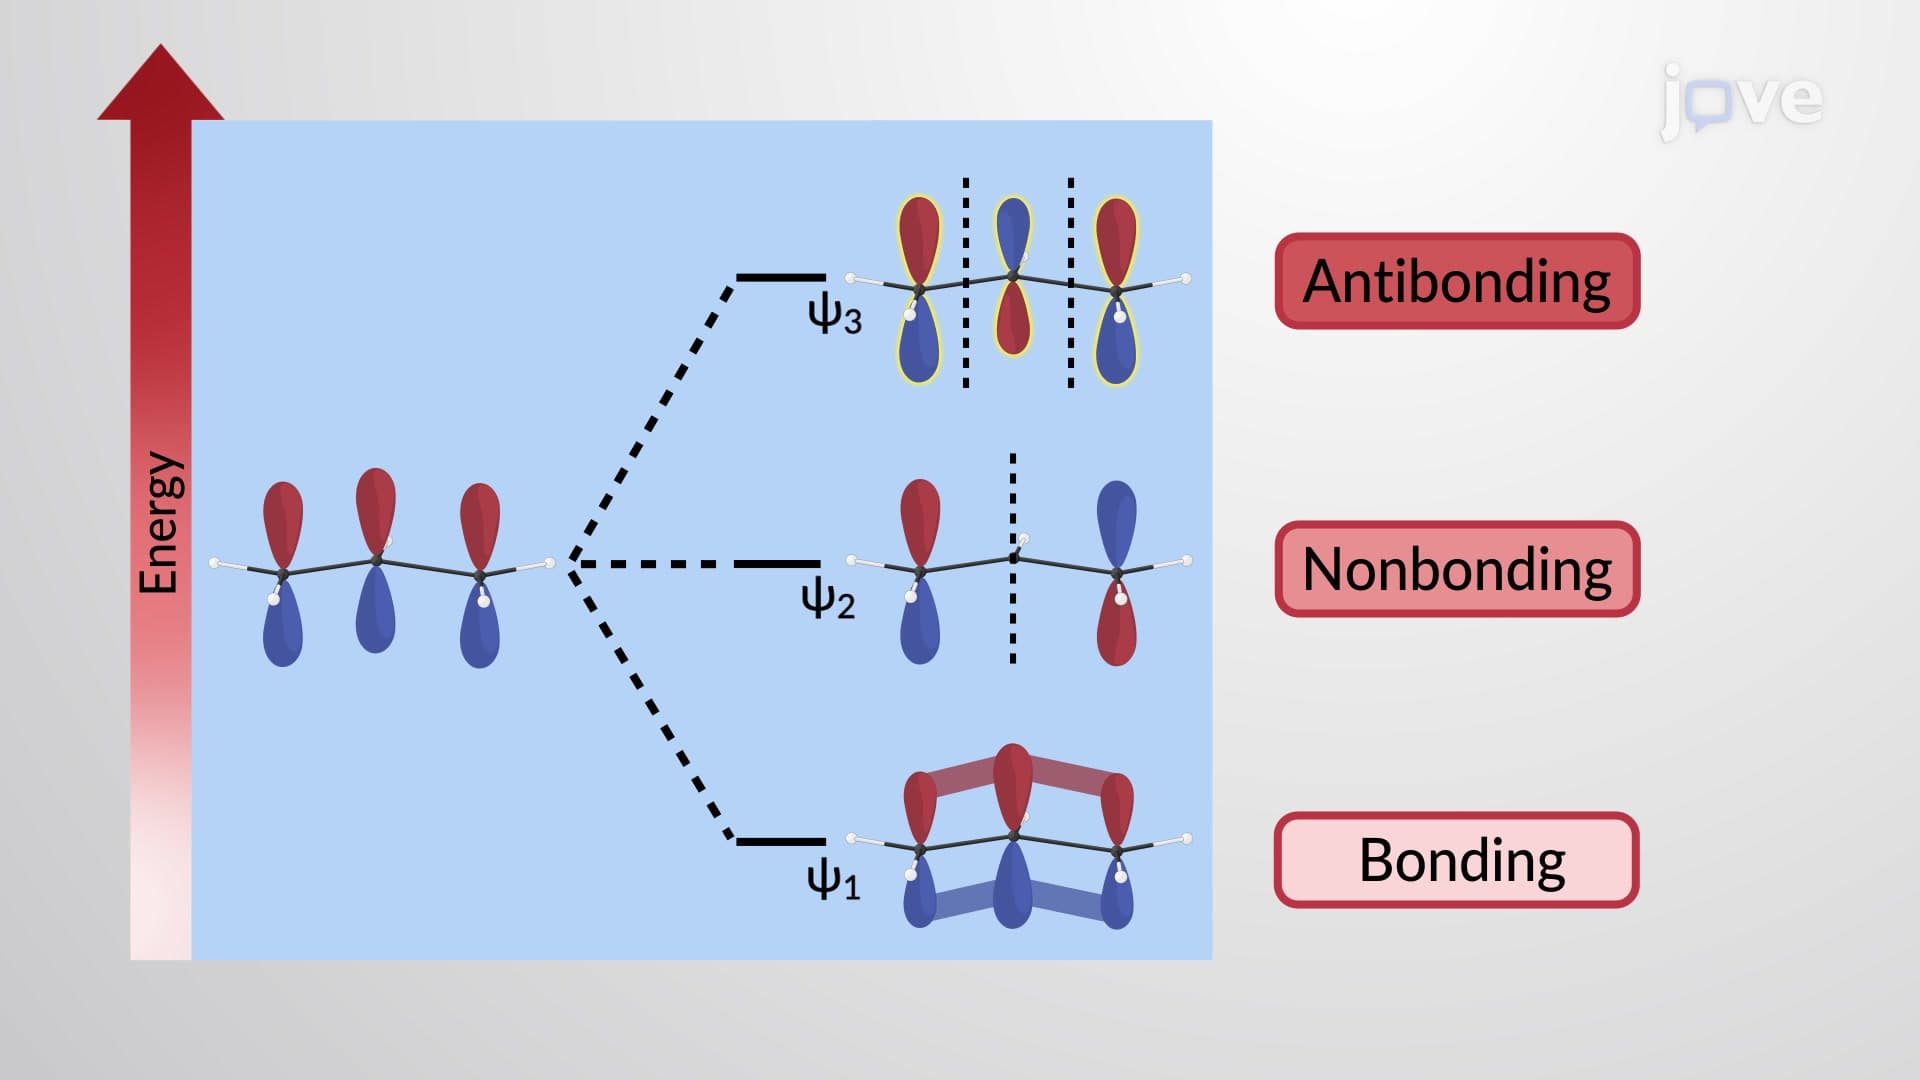
π Molecular Orbitals of the Allyl Cation and Anion
 03:02
The Aufbau Principle and Hund's Rule
 04:01
VSEPR Theory and the Effect of Lone Pairs
 03:35
Hybridization of Atomic Orbitals II
 02:27
Predicting Molecular Geometry
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...