Related Experiment Video
Updated: May 27, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Density-functional approaches to noncovalent interactions: a comparison of dispersion corrections (DFT-D),
Lori A Burns1, Alvaro Vázquez-Mayagoitia, Bobby G Sumpter
1Center for Computational Molecular Science and Technology, School of Chemistry and Biochemistry, Georgia Institute of Technology, Atlanta, Georgia 30332-0400, USA.

Abstract:
A systematic study of techniques for treating noncovalent interactions within the computationally efficient density functional theory (DFT) framework is presented through comparison to benchmark-quality evaluations of binding strength compiled for molecular complexes of diverse size and nature. In particular, the efficacy of functionals deliberately crafted to encompass long-range forces, a posteriori DFT+dispersion corrections (DFT-D2 and DFT-D3), and exchange-hole dipole moment (XDM) theory is assessed against a large collection (469 energy points) of reference interaction energies at the CCSD(T) level of theory extrapolated to the estimated complete basis set limit. The established S22 [revised in J. Chem. Phys. 132, 144104 (2010)] and JSCH test sets of minimum-energy structures, as well as collections of dispersion-bound (NBC10) and hydrogen-bonded (HBC6) dissociation curves and a pairwise decomposition of a protein-ligand reaction site (HSG), comprise the chemical systems for this work. From evaluations of accuracy, consistency, and efficiency for PBE-D, BP86-D, B97-D, PBE0-D, B3LYP-D, B970-D, M05-2X, M06-2X, ωB97X-D, B2PLYP-D, XYG3, and B3LYP-XDM methodologies, it is concluded that distinct, often contrasting, groups of these elicit the best performance within the accessible double-ζ or robust triple-ζ basis set regimes and among hydrogen-bonded or dispersion-dominated complexes. For overall results, M05-2X, B97-D3, and B970-D2 yield superior values in conjunction with aug-cc-pVDZ, for a mean absolute deviation of 0.41 - 0.49 kcal/mol, and B3LYP-D3, B97-D3, ωB97X-D, and B2PLYP-D3 dominate with aug-cc-pVTZ, affording, together with XYG3/6-311+G(3df,2p), a mean absolute deviation of 0.33 - 0.38 kcal/mol.


Related Concept Videos
 01:24
Van der Waals Interactions
 03:13
Intermolecular Forces
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:36
Molecular Geometry and Dipole Moments
 01:14
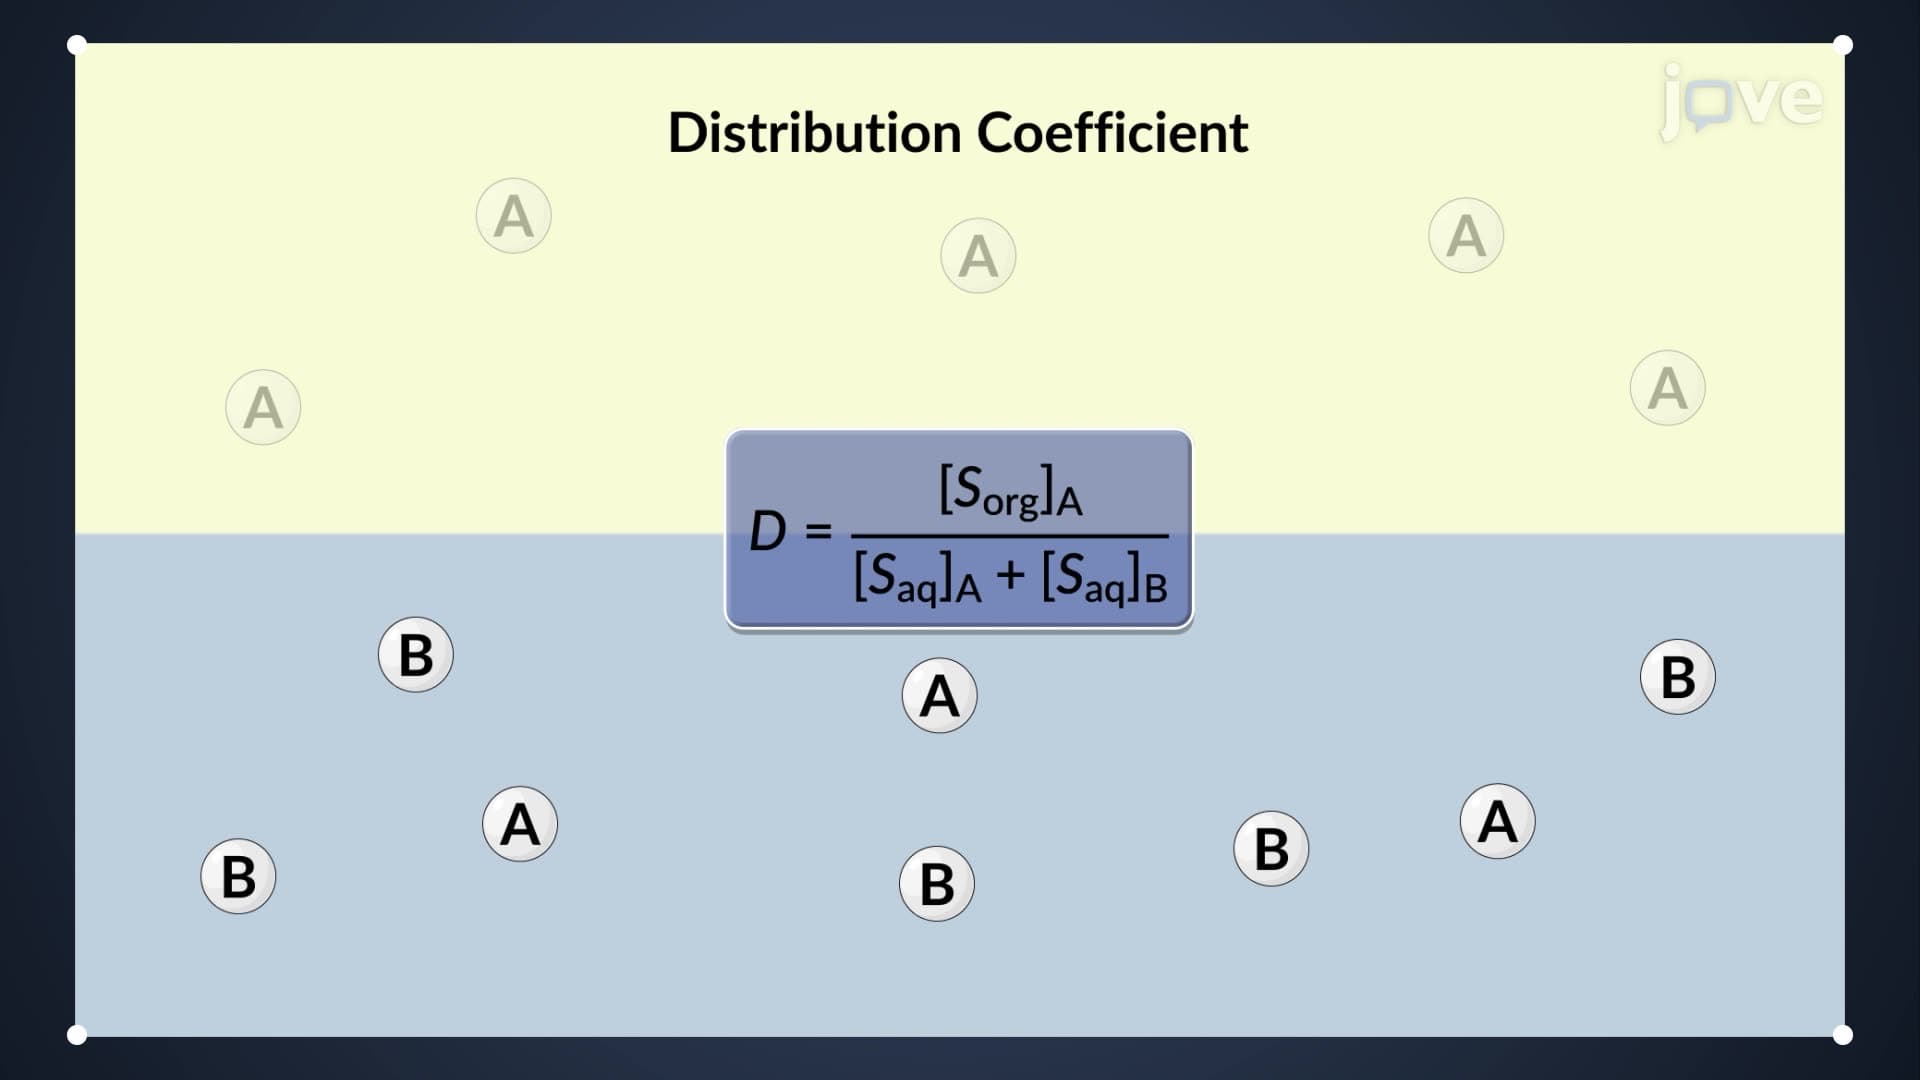
Extraction: Partition and Distribution Coefficients
For extracting a solute from an aqueous phase into an...