Related Experiment Video
Updated: Jun 4, 2026

Inducing Meningococcal Meningitis Serogroup C in Mice via Intracisternal Delivery
Published on: November 5, 2019
Neisseria meningitidis is structured in clades associated with restriction modification systems that modulate
Sonia Budroni1, Emilio Siena, Julie C Dunning Hotopp
1Research Center, Novartis Vaccines and Diagnostics, 53100 Siena, Italy.

Abstract:
Molecular data on a limited number of chromosomal loci have shown that the population of Neisseria meningitidis (Nm), a deadly human pathogen, is structured in distinct lineages. Given that the Nm population undergoes substantial recombination, the mechanisms resulting in the evolution of these lineages, their persistence in time, and the implications for the pathogenicity of the bacterium are not yet completely understood. Based on whole-genome sequencing, we show that Nm is structured in phylogenetic clades. Through acquisition of specific genes and through insertions and rearrangements, each clade has acquired and remodeled specific genomic tracts, with the potential to impact on the commensal and virulence behavior of Nm. Despite this clear evidence of a structured population, we confirm high rates of detectable recombination throughout the whole Nm chromosome. However, gene conversion events were found to be longer within clades than between clades, suggesting a DNA cleavage mechanism associated with the phylogeny of the species. We identify 22 restriction modification systems, probably acquired by horizontal gene transfer from outside of the species/genus, whose distribution in the different strains coincides with the phylogenetic clade structure. We provide evidence that these clade-associated restriction modification systems generate a differential barrier to DNA exchange consistent with the observed population structure. These findings have general implications for the emergence of lineage structure and virulence in recombining bacterial populations, and they could provide an evolutionary framework for the population biology of a number of other bacterial species that show contradictory population structure and dynamics.
Insights
Neisseria meningitidis (Nm) exhibits distinct phylogenetic clades shaped by gene acquisition and genomic remodeling. Clade-specific restriction modification systems likely drive this lineage structure and influence bacterial virulence.
Area of Science:
- Microbiology
- Genomics
- Evolutionary Biology
Background:
- Neisseria meningitidis (Nm) is a significant human pathogen with a complex population structure.
- Previous studies using limited molecular data suggested distinct Nm lineages, but the evolutionary mechanisms and impact on pathogenicity remained unclear.
- The high rate of recombination in Nm populations complicates understanding lineage evolution and persistence.
Purpose of the Study:
- To investigate the population structure of Neisseria meningitidis using whole-genome sequencing.
- To identify genomic features contributing to lineage diversification and their impact on virulence.
- To elucidate the role of restriction modification systems in shaping Nm population structure and DNA exchange.
Main Methods:
- Whole-genome sequencing of Neisseria meningitidis strains.
- Phylogenetic analysis to determine clade structure.
- Identification and distribution analysis of genomic tracts, including restriction modification systems.
- Assessment of gene conversion lengths within and between clades.
Main Results:
- Neisseria meningitidis is structured into distinct phylogenetic clades, each with unique genomic tracts acquired and remodeled.
- High rates of recombination persist across the Nm chromosome, but gene conversion is longer within clades.
- Twenty-two restriction modification systems were identified, distributed according to clade structure, suggesting horizontal gene transfer.
- These systems create differential barriers to DNA exchange, consistent with the observed population structure.
Conclusions:
- Genomic remodeling and clade-specific restriction modification systems are key drivers of lineage structure in Neisseria meningitidis.
- These findings provide insights into the evolution of virulence in recombining bacterial populations.
- The study offers an evolutionary framework applicable to other bacterial species with complex population dynamics.
Related Concept Videos
 02:53
CRISPR and crRNAs
The CRISPR-Cas system stores a copy of foreign DNA in the host genome and uses it to identify the foreign DNA upon reinfection. CRISPR-Cas has three different...
 01:23
The Antiviral System of Bacteria and Archaea: CRISPR
 01:26
Bacterial Meningitis II: Pathophysiology
 01:11
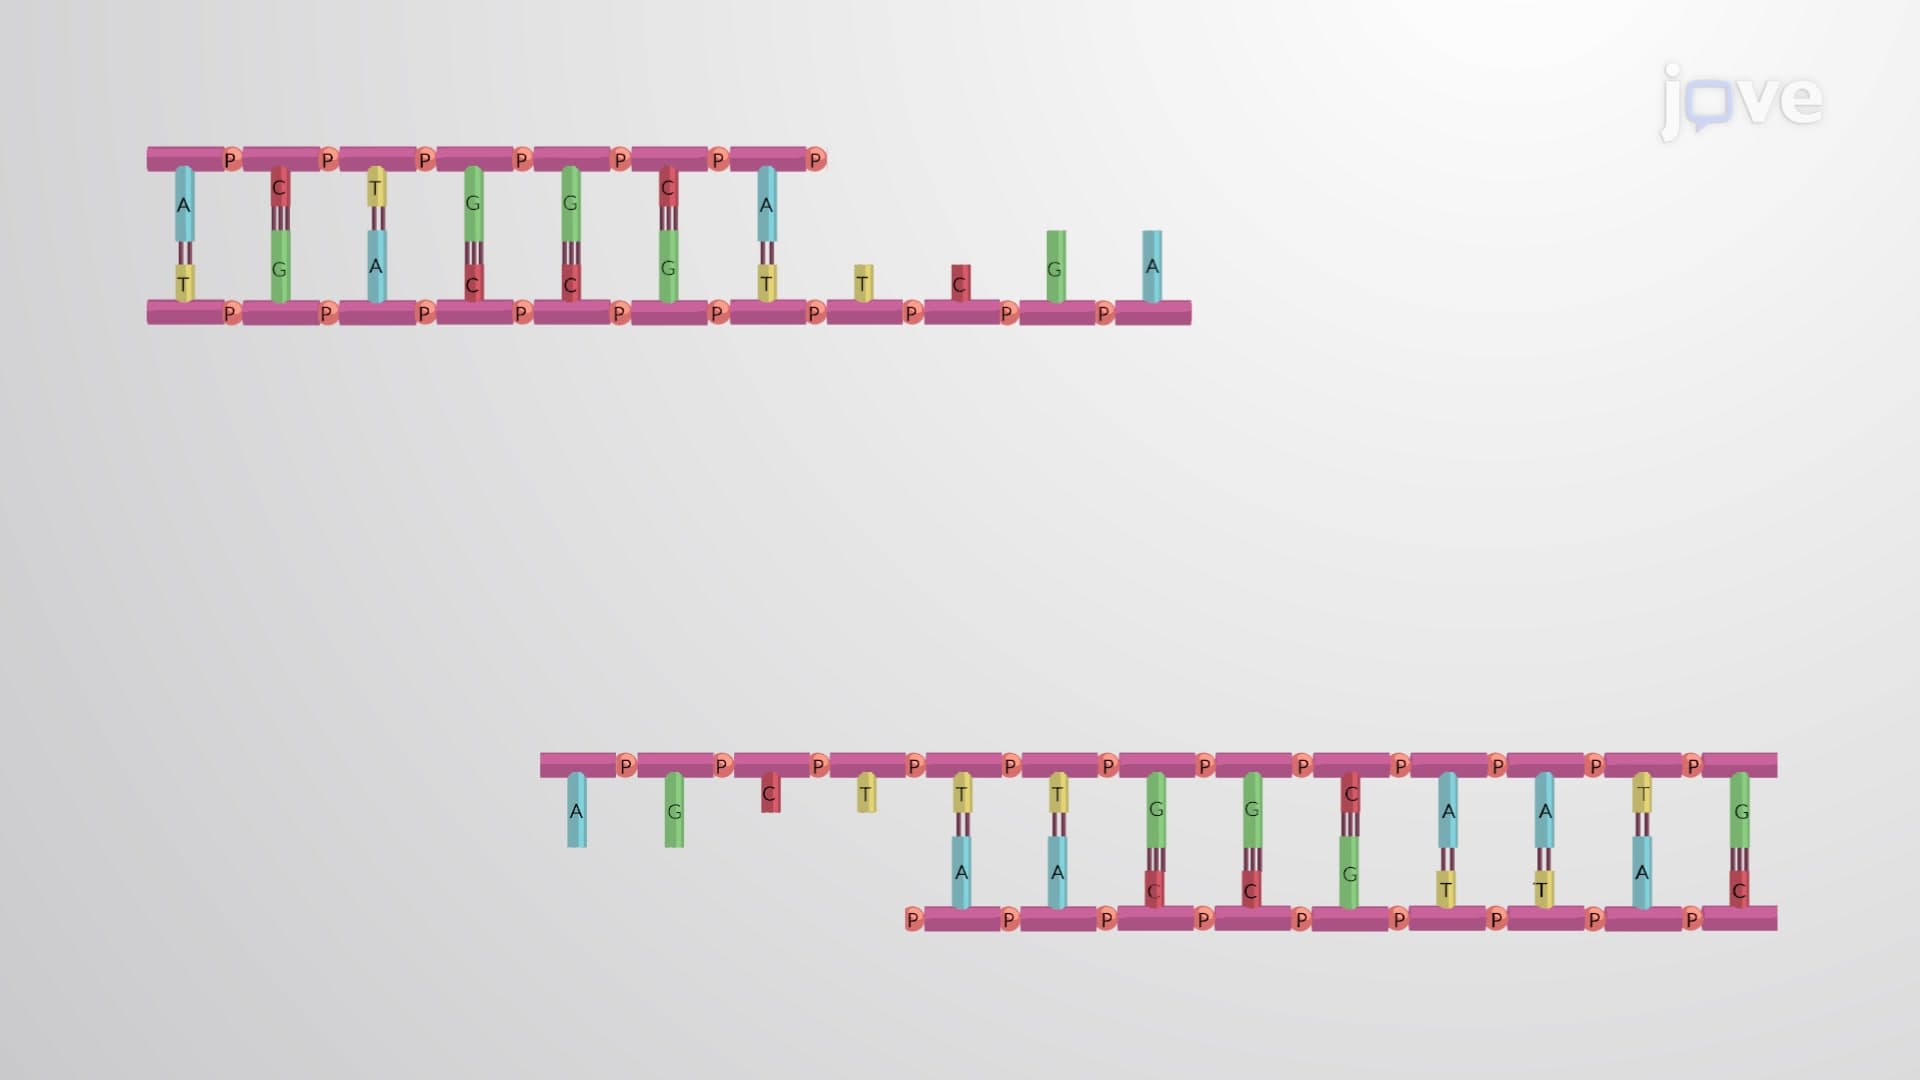
Restriction Enzymes
The host bacteria protect their own genomic DNA from these enzymes by methylating these sites. Some...
 01:24
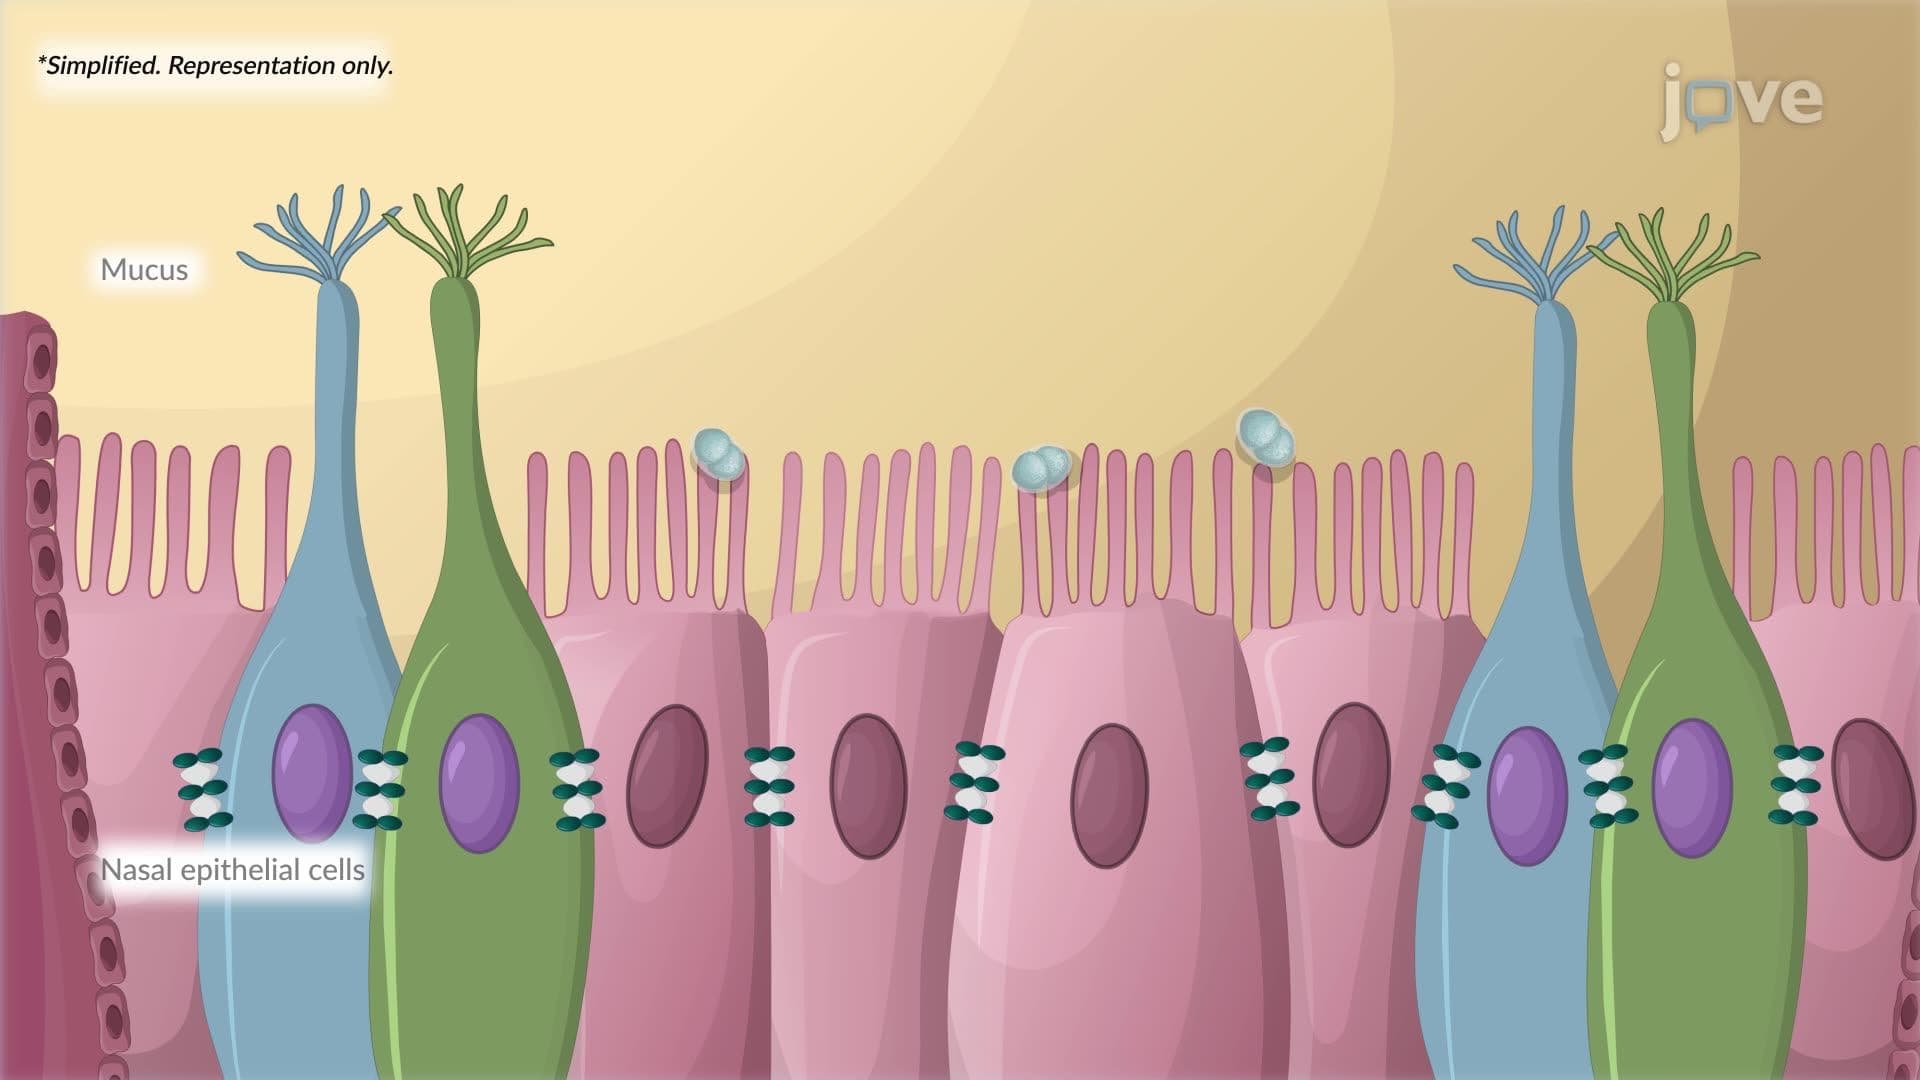
Bacterial Meningitis
 02:53
Conservative Site-specific Recombination and Phase Variation
The recognition sites for Cre recombinase called LoxP...