Related Experiment Video
Updated: Jun 3, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Atomic and molecular adsorption on transition-metal carbide (111) surfaces from density-functional theory: a trend
A Vojvodic1, C Ruberto, B I Lundqvist
1Department of Applied Physics, Chalmers University of Technology, Göteborg, Sweden. alevoj@chalmers.se

Abstract:
This study explores atomic and molecular adsorption on a number of early transition-metal carbides (TMCs) in NaCl structure by means of density-functional theory calculations. The investigated substrates are the TM-terminated TMC(111) surfaces, of interest because of the presence of different types of surface resonances (SRs) on them and because of their technological importance in growth processes. Also, TM compounds have shown potential in catalysis applications. Trend studies are conducted with respect to both period and group in the periodic table, choosing the substrates ScC, TiC, VC, ZrC, NbC, δ-MoC, TaC, and WC (in NaCl structure) and the adsorbates H, B, C, N, O, F, NH, NH(2), and NH(3). Trends in adsorption strength are explained in terms of surface electronic factors, by correlating the calculated adsorption-energy values with the calculated surface electronic structures. The results are rationalized by use of a concerted-coupling model (CCM), which has previously been applied successfully to the description of adsorption on TiC(111) and TiN(111) surfaces (Ruberto et al 2007 Solid State Commun. 141 48). First, the clean TMC(111) surfaces are characterized by calculating surface energies, surface relaxations, Bader charges, and surface-localized densities of states (DOSs). Detailed comparisons between surface and bulk DOSs reveal the existence of transition-metal localized SRs (TMSRs) in the pseudogap and of several C-localized SRs (CSRs) in the upper valence band on all considered TMC(111) surfaces. The spatial extent and the dangling bond nature of these SRs are supported by real-space analyses of the calculated Kohn-Sham wavefunctions. Then, atomic and molecular adsorption energies, geometries, and charge transfers are presented. An analysis of the adsorbate-induced changes in surface DOSs reveals a presence of both adsorbate-TMSR and adsorbate-CSRs interactions, of varying strengths depending on the surface and the adsorbate. These variations are correlated to the variations in adsorption energies. The results are used to generalize the content and applications of the previously proposed CCM to this larger class of substrates and adsorbates. Implications for other classes of materials, for catalysis, and for other surface processes are discussed.
More Related Videos
 07:47
Reverse Microemulsion-mediated Synthesis of Monometallic and Bimetallic Early Transition Metal Carbide and Nitride Nanoparticles
Published on: November 27, 2015
 10:57
Synthesis and Performance Characterizations of Transition Metal Single Atom Catalyst for Electrochemical CO2 Reduction
Published on: April 10, 2018
Related Concept Videos
 01:25
Adsorption Isotherms II
 01:28
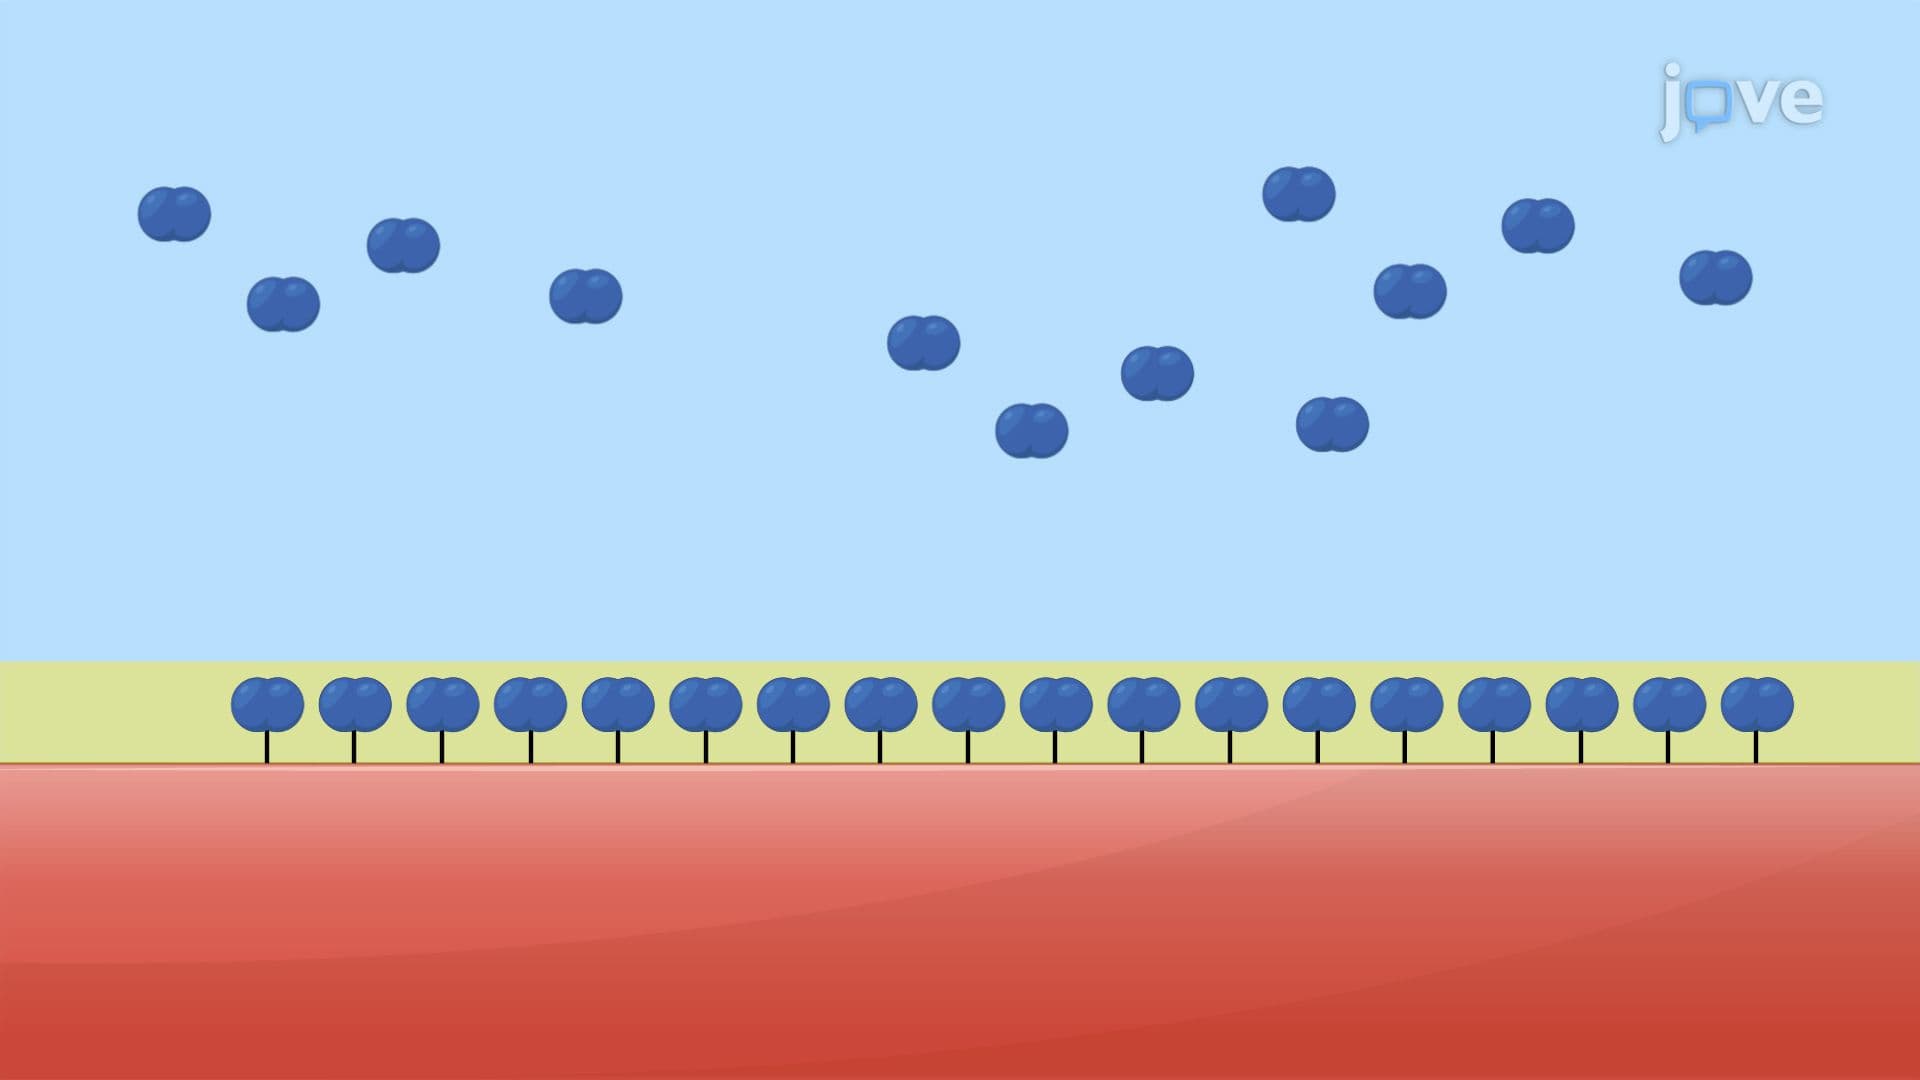
Adsorption of Gases on Solids
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:29
Adsorption Isotherms I
 02:42
Valence Bond Theory
 01:08
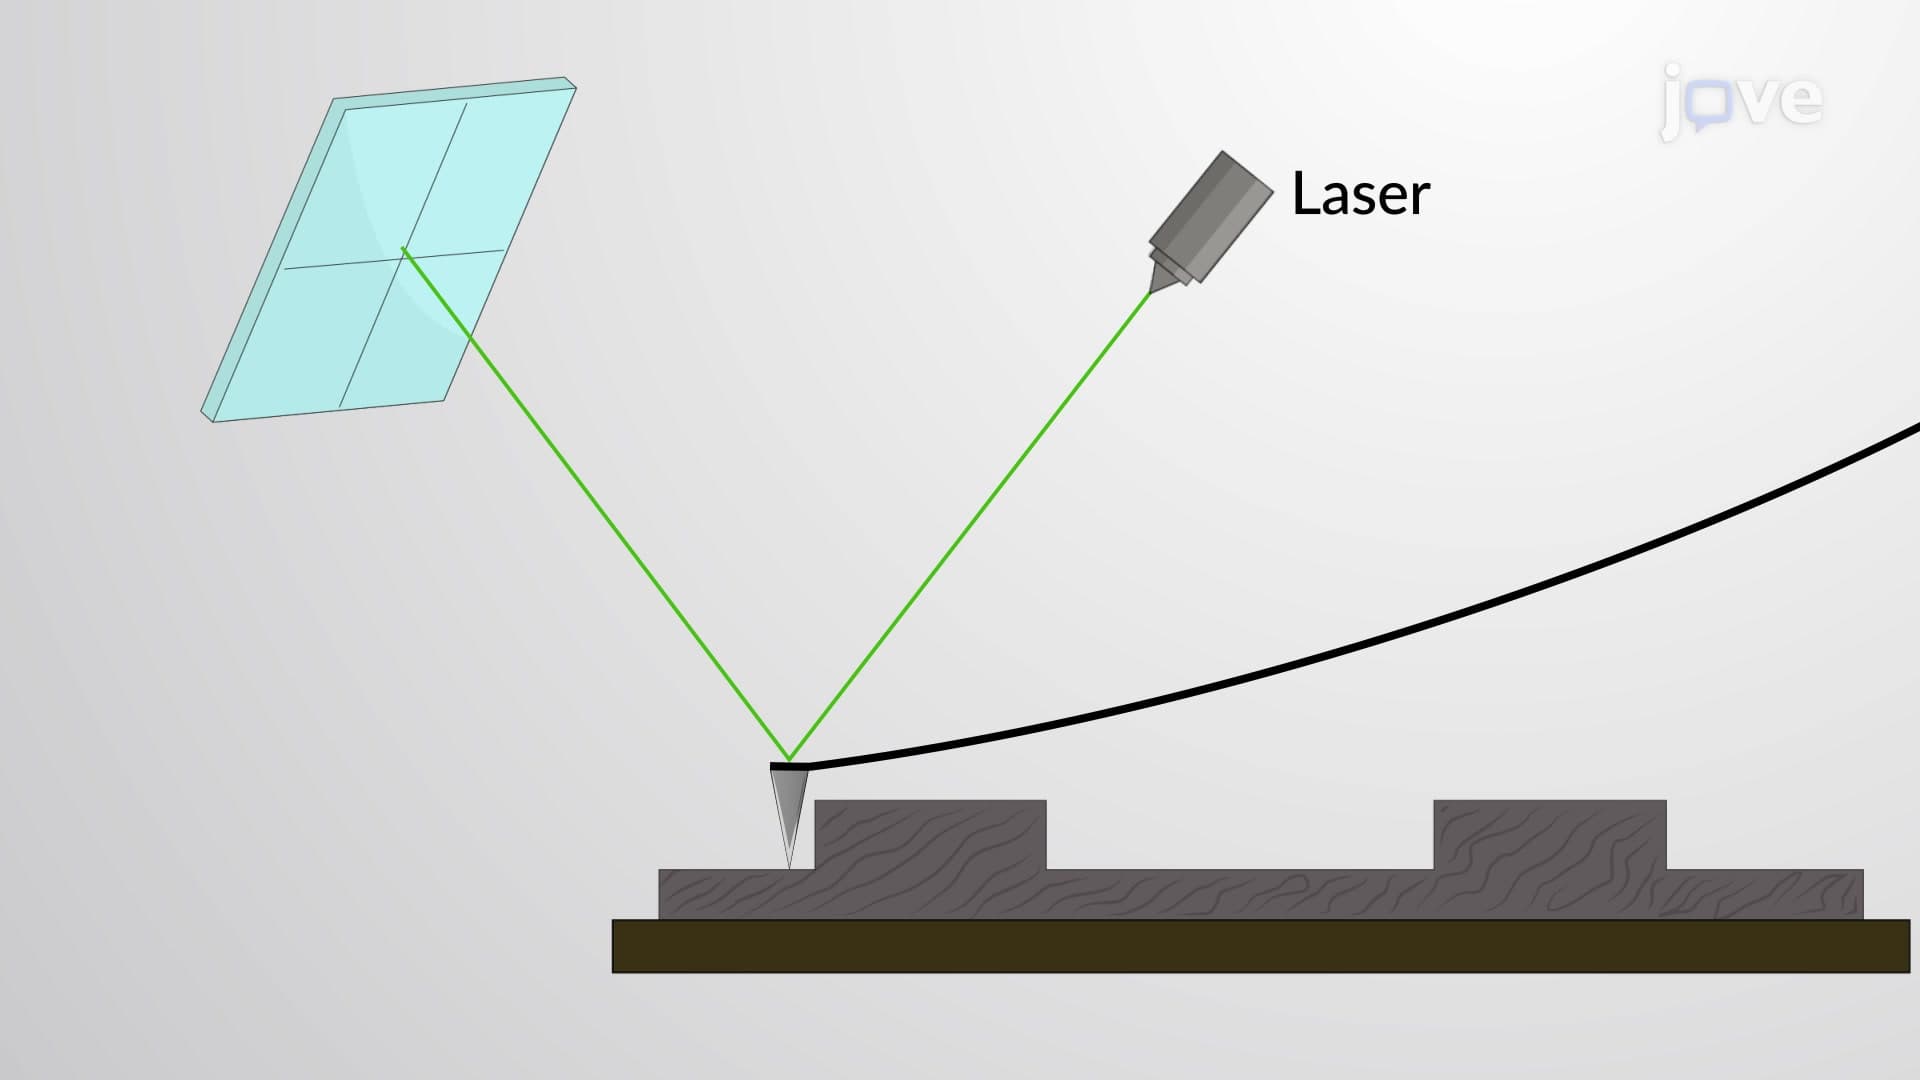
Atomic Force Microscopy
The AFM Probe
The probe is regarded as the heart of any AFM setup and comprises the...