Related Experiment Video
Updated: Jun 3, 2026

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Efficacy of surface error corrections to density functional theory calculations of vacancy formation energy in
Prithwish Kumar Nandi1, M C Valsakumar, Sharat Chandra
1Materials Science Group, Indira Gandhi Centre for Atomic Research, Kalpakkam, Tamil Nadu, India.

Abstract:
We calculate properties like equilibrium lattice parameter, bulk modulus and monovacancy formation energy for nickel (Ni), iron (Fe) and chromium (Cr) using Kohn-Sham density functional theory (DFT). We compare the relative performance of local density approximation (LDA) and generalized gradient approximation (GGA) for predicting such physical properties for these metals. We also make a relative study between two different flavors of GGA exchange correlation functional, namely PW91 and PBE. These calculations show that there is a discrepancy between DFT calculations and experimental data. In order to understand this discrepancy in the calculation of vacancy formation energy, we introduce a correction for the surface intrinsic error corresponding to an exchange correlation functional using the scheme implemented by Mattsson et al (2006 Phys. Rev. B 73 195123) and compare the effectiveness of the correction scheme for Al and the 3d transition metals.
More Related Videos
 06:57
Theoretical Calculation and Experimental Verification for Dislocation Reduction in Germanium Epitaxial Layers with Semicylindrical Voids on Silicon
Published on: July 17, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 01:12
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:29
Imperfections in Crystal Structure: Non-Stoichiometric Defects
 01:10
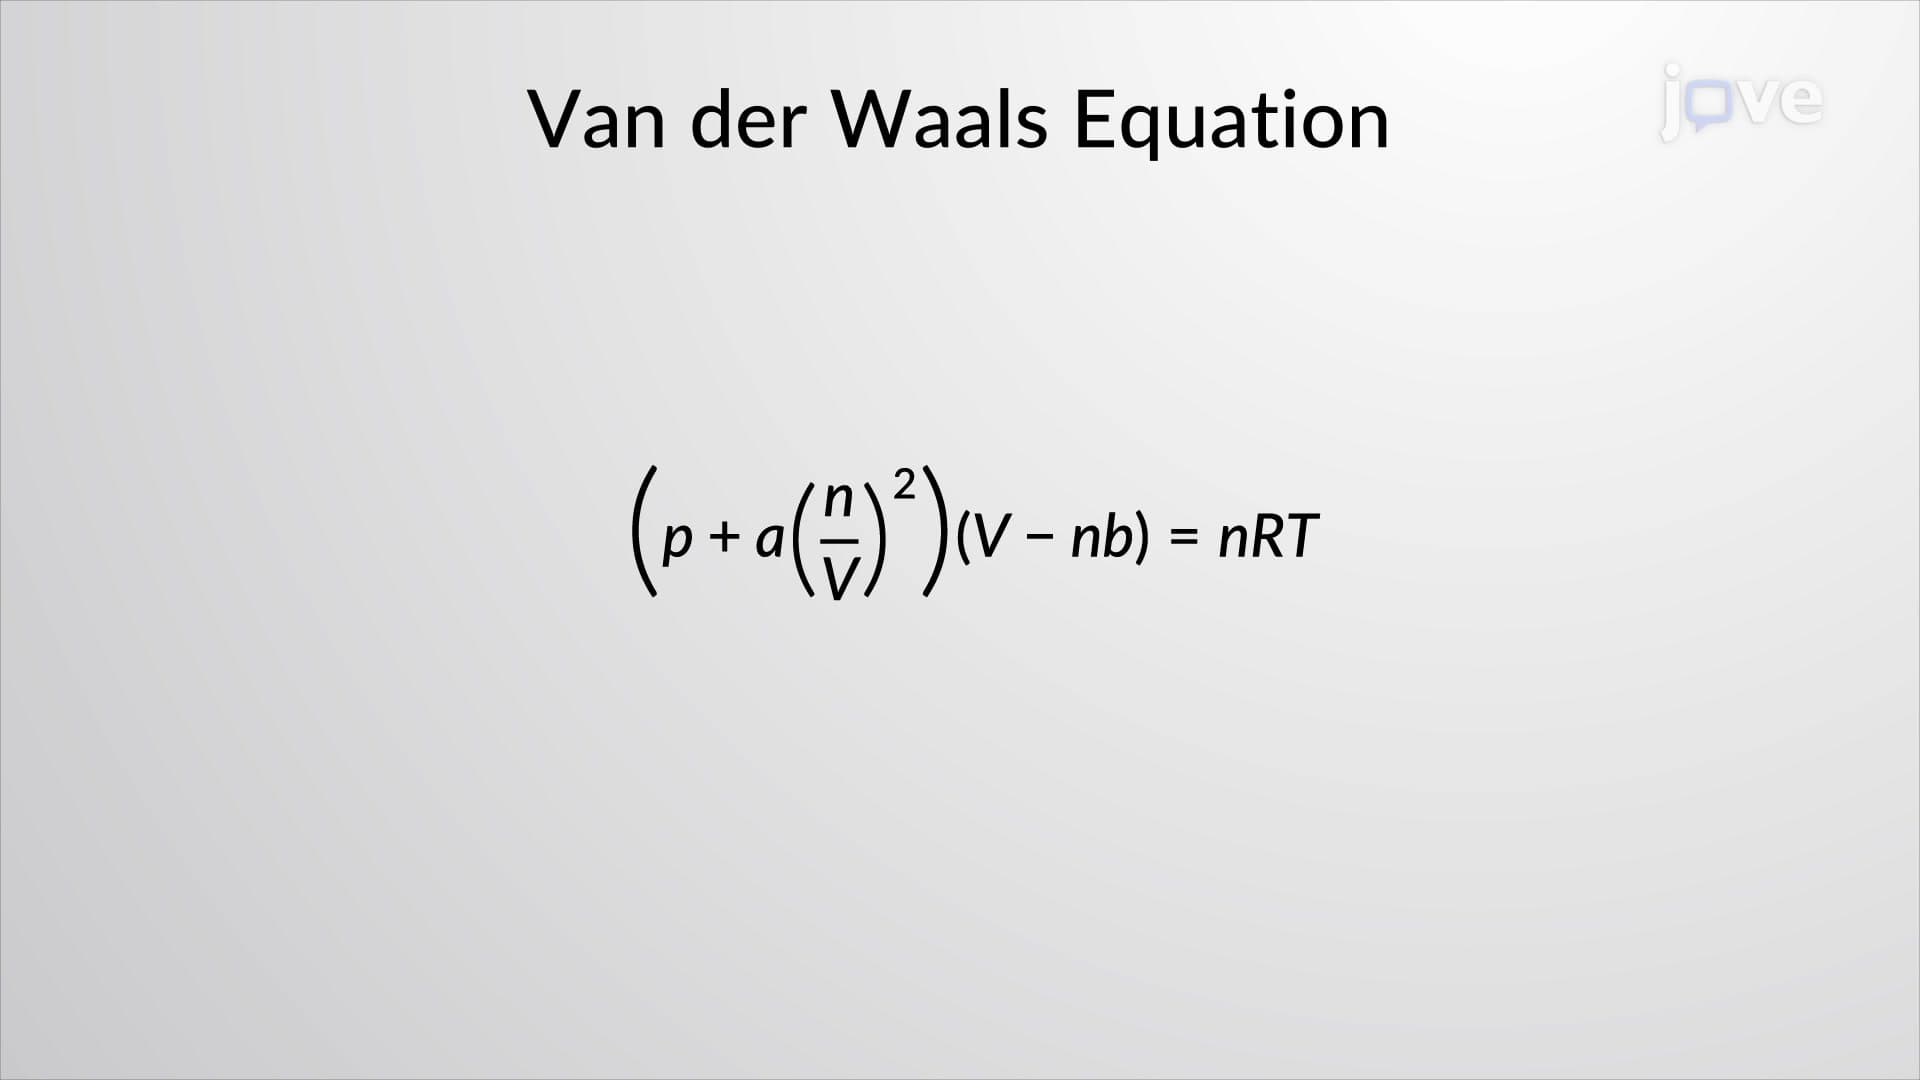
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...