Related Experiment Video
Updated: Jun 2, 2026


Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
Functional group dependence of solute partitioning to various locations within a DOPC bilayer: a comparison of
Ravindra W Tejwani1, Malcolm E Davis, Bradley D Anderson
1Department of Pharmaceutical Sciences, College of Pharmacy, University of Kentucky, Lexington, Kentucky 40506, USA.

Abstract:
Atomic-level molecular dynamics simulations of 1,2-dioleoyl-sn-glycero-3-phosphocholine (DOPC) bilayers containing small, amphiphilic, drug-like molecules were carried out to examine the influence of polar functionality on membrane partitioning and transport. Three related molecules (tyramine, phenethylamine, and 4-ethylphenol) were chosen to allow a detailed study of the isolated effects of the amine and hydroxyl functionalities on the preferred solute location, free energies of transfer, and the effect of combining both functional groups in a same molecule. Transfer free energy profiles (from water) generated from molecular dynamics (MD) simulations as a function of bilayer depth compared favorably to comparable experimental results. The simulations allowed the determination of the location of the barrier domain for permeability where the transfer free energy is highest and the preferred binding region at which the free energy is a minimum for each of the three solutes. Comparisons of the free energy profiles reveal that the hydrocarbon chain interior is the region most selective to chemical structure of different solutes because the free energies of transfer in that region vary to a significantly greater extent than in other regions of the bilayer. The contributions of the hydroxyl and amino groups to the free energies of solute transfer from water to the interfacial region were close to zero in both the MD simulations and experimental measurements. This suggests that the free energy decrease observed for solute transfer into the head group region occurs with minimal loss in solvation by hydrogen bonding to polar functional groups on the solute and is largely driven by hydrophobicity. Overall, the joint experimental and simulation studies suggest that the assumption of additivity of free energy contributions from multiple polar functional groups on the same molecule may hold for predictions of passive bilayer permeability coefficients providing that the groups are well isolated. However, this assumption does not hold for predictions of relative liposome-binding affinities.
More Related Videos
 12:18
Assembly of Cell Mimicking Supported and Suspended Lipid Bilayer Models for the Study of Molecular Interactions
Published on: August 3, 2021
 10:08
Phase Behavior of Charged Vesicles Under Symmetric and Asymmetric Solution Conditions Monitored with Fluorescence Microscopy
Published on: October 24, 2017
Related Concept Videos
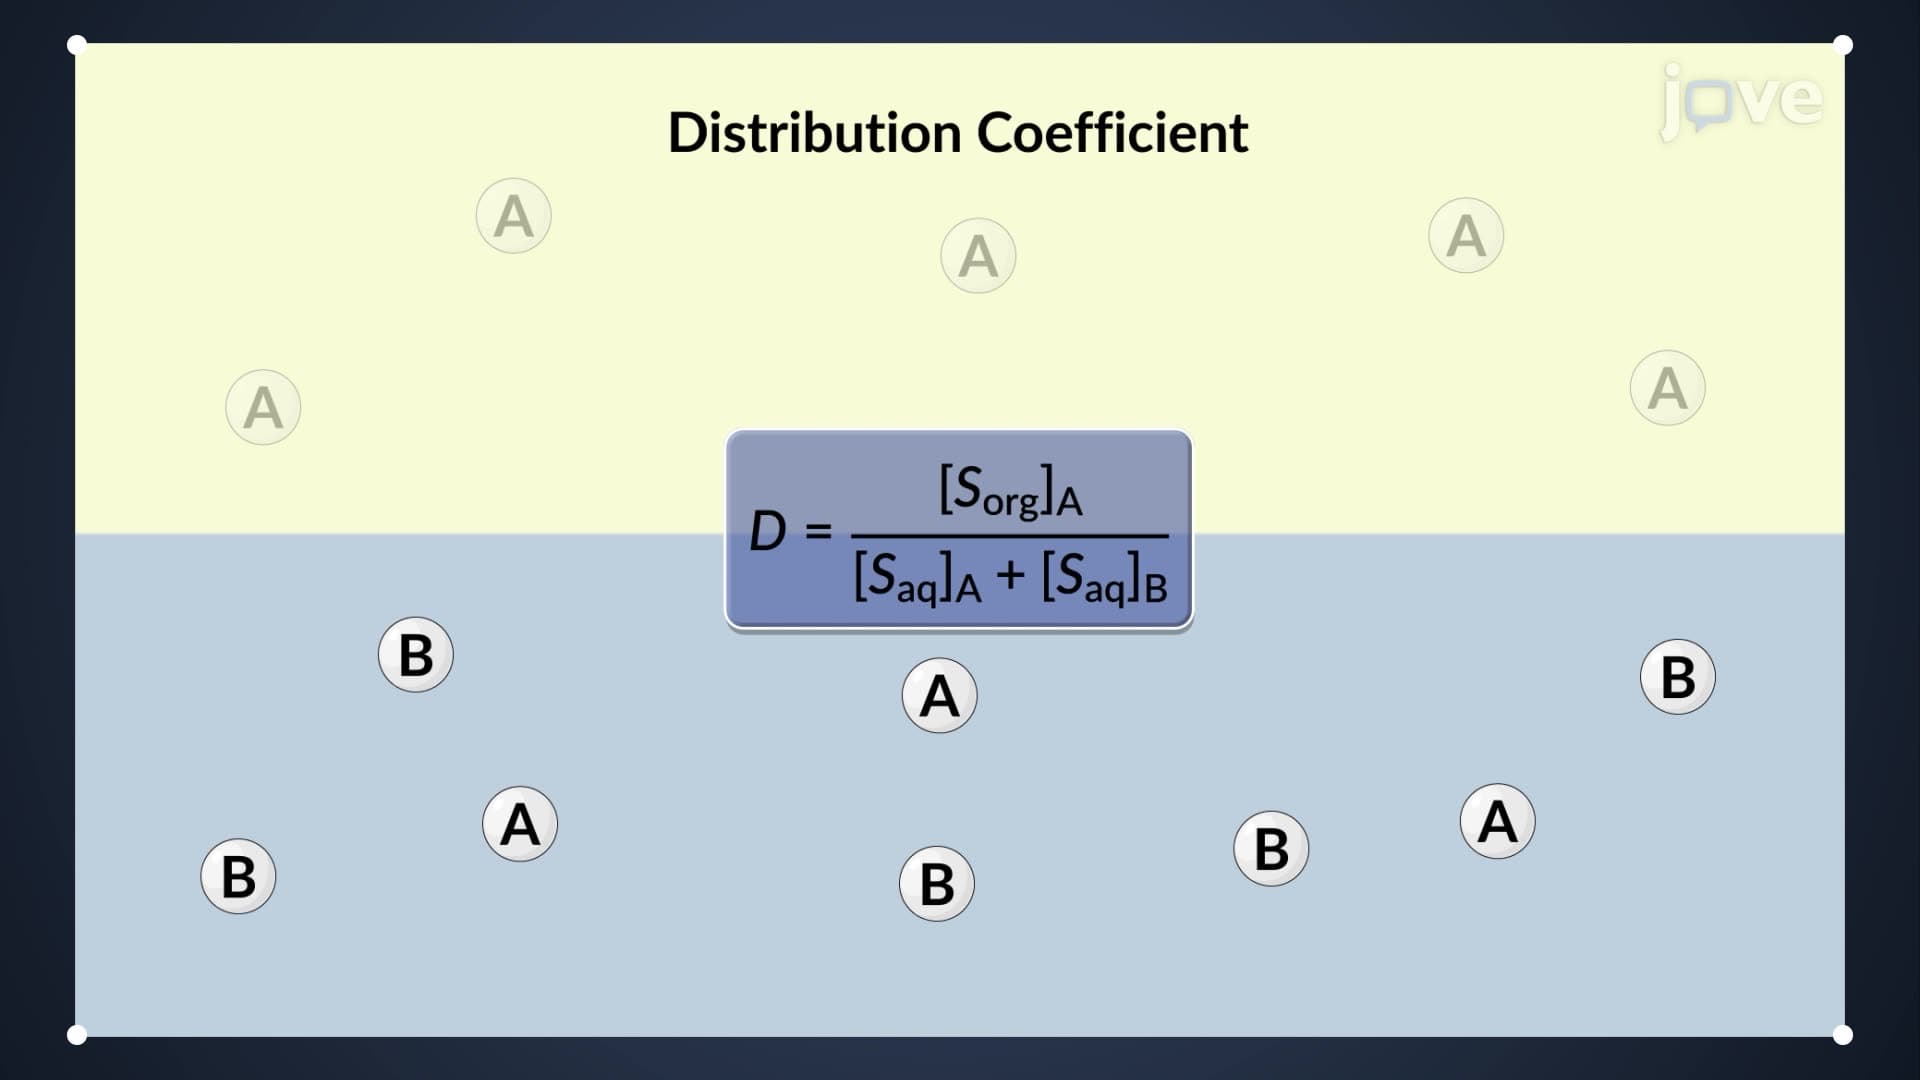 01:14
Extraction: Partition and Distribution Coefficients
For extracting a solute from an aqueous phase into an organic...
 01:21
Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH
A drug's pKa and the pH of the gastrointestinal (GI) tract play crucial roles in drug...
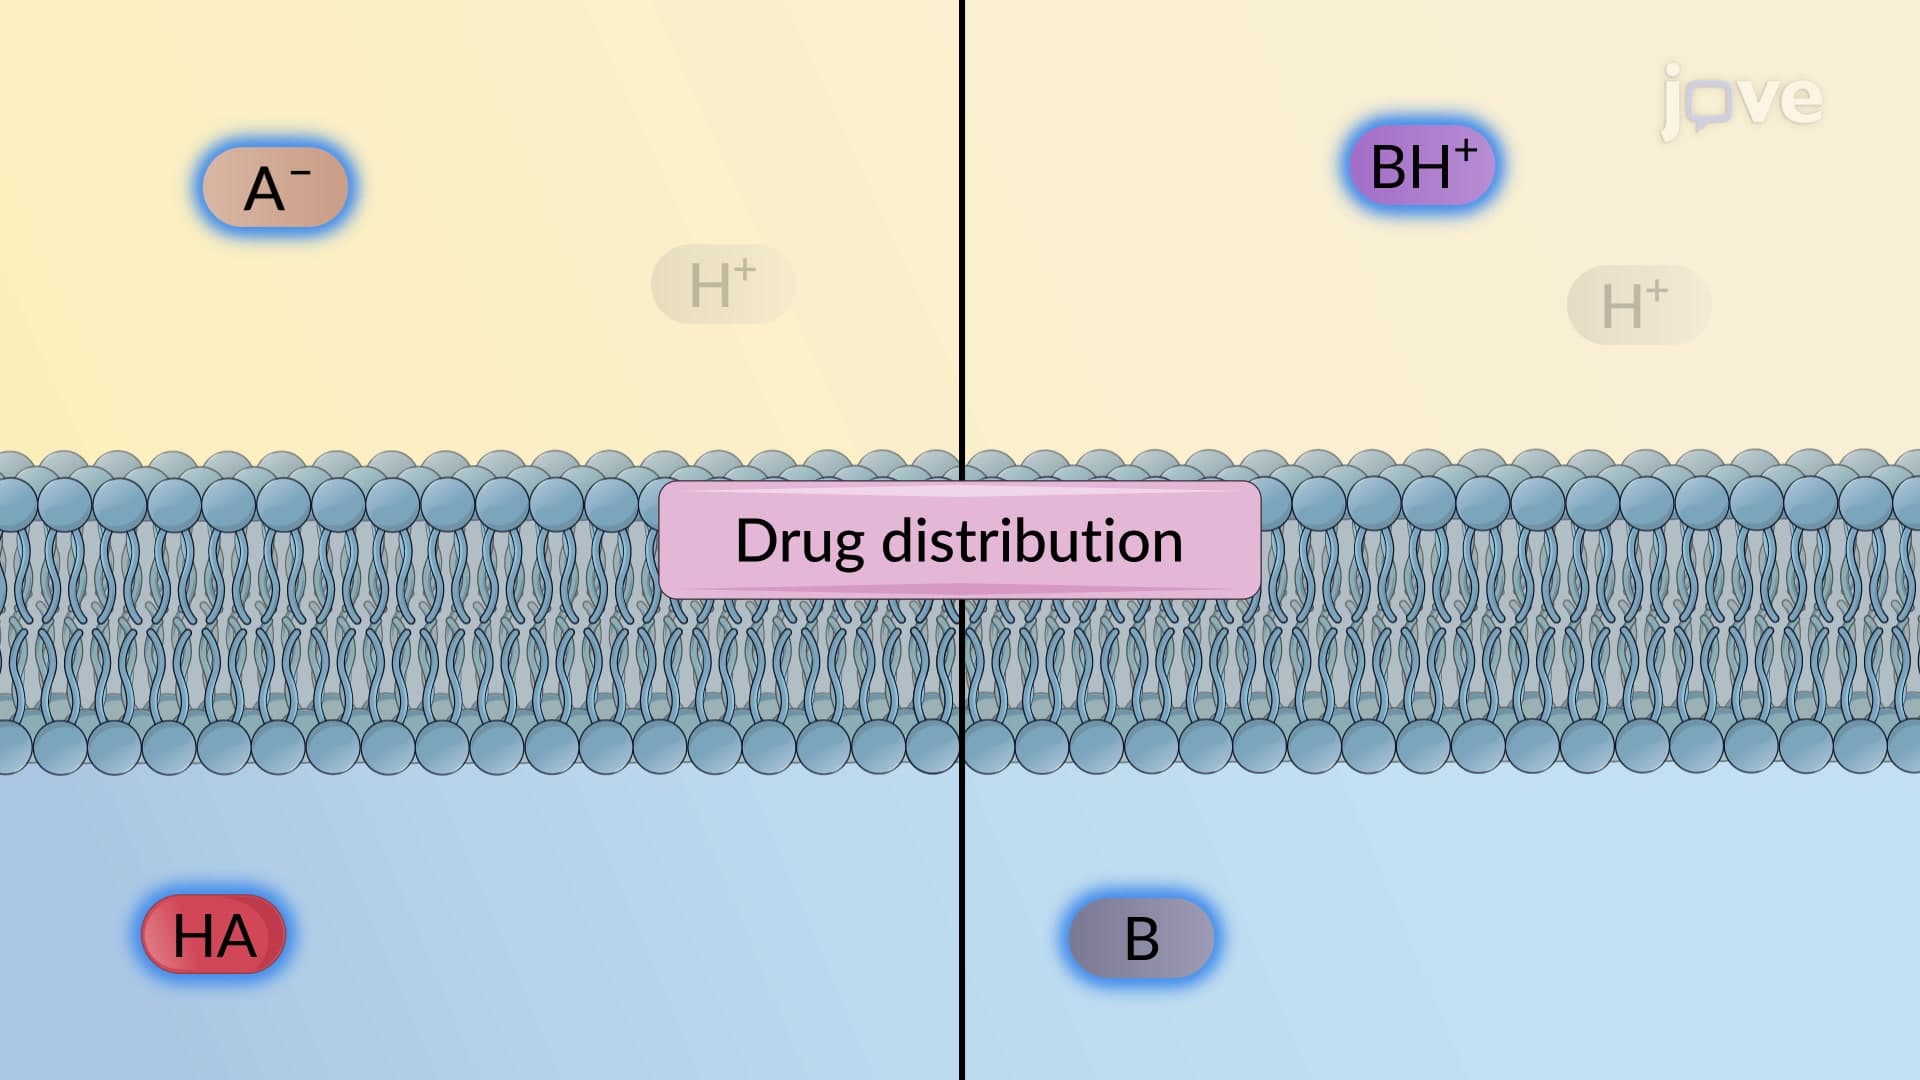 01:23
Drug Absorption Mechanism: Passive Membrane Transport
 03:13
Intermolecular Forces
 03:00
Solubility
A solution is a homogeneous mixture composed of a solvent, the major component, and a solute, the minor component. The physical state of a solution—solid, liquid, or gas—is typically the same as that of the solvent. Solute concentrations are often described with qualitative terms such as dilute (of relatively low concentration) and concentrated (of relatively high concentration).
In a solution, the solute particles (molecules, atoms, and/or ions)...
 01:09
Theories of Dissolution: The Danckwerts' Model and Interfacial Barrier Model