Related Experiment Video
Updated: Jun 2, 2026

Modification and Functionalization of the Guanidine Group by Tailor-made Precursors
Published on: April 27, 2017
Sequential catalytic role of bifunctional bicyclic guanidine in asymmetric phospha-Michael reaction
Bokun Cho1, Choon-Hong Tan, Ming Wah Wong
1Department of Chemistry, National University of Singapore, 3 Science Drive 3, Singapore, 117543.

Abstract:
The catalytic mechanism and origin of enantioselectivity of bicyclic guanidine-catalyzed phospha-Michael reaction between diphenyl phosphine oxide and β-nitrostyrene were investigated by DFT calculations at M06-2X/cc-pVTZ//M06-2X/cc-pVDZ level in conjunction with the implicit SMD solvation method. The catalyst is found to be involved in all 3 steps of the proposed catalytic cycle, namely (1) tautomerization of phosphine oxide, (2) C-P bond formation and (3) concerted hydrogen transfer. The bifunctional role of the guanidine catalyst is clearly demonstrated in all 3 key steps. Due to the geometry of the bicyclic guanidine catalyst, the preferred orientation of the reactants in the transition state of enantioselective C-P bond forming step favours the R enantiomer, in excellent accord with the observed enantioselectivity. Analysis of various transition states suggests that the asymmetric C-P bond formation is controlled by the hydrogen bonding interaction and steric effect between the catalyst and substrate. Various weaker C-H···X (X = N, O and π) interactions also play a role in stabilizing the key transition states.
More Related Videos
 09:04
A Direct, Early Stage Guanidinylation Protocol for the Synthesis of Complex Aminoguanidine-containing Natural Products
Published on: September 9, 2016
 10:17
Efficient Construction of Drug-like Bispirocyclic Scaffolds Via Organocatalytic Cycloadditions of α-Imino γ-Lactones and Alkylidene Pyrazolones
Published on: February 7, 2019
Related Concept Videos
 01:11
Nucleophilic Aromatic Substitution: Elimination–Addition
 01:15
α-Bromination of Carboxylic Acids: Hell–Volhard–Zelinski Reaction
 02:26
Preparation and Reactions of Sulfides
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:08
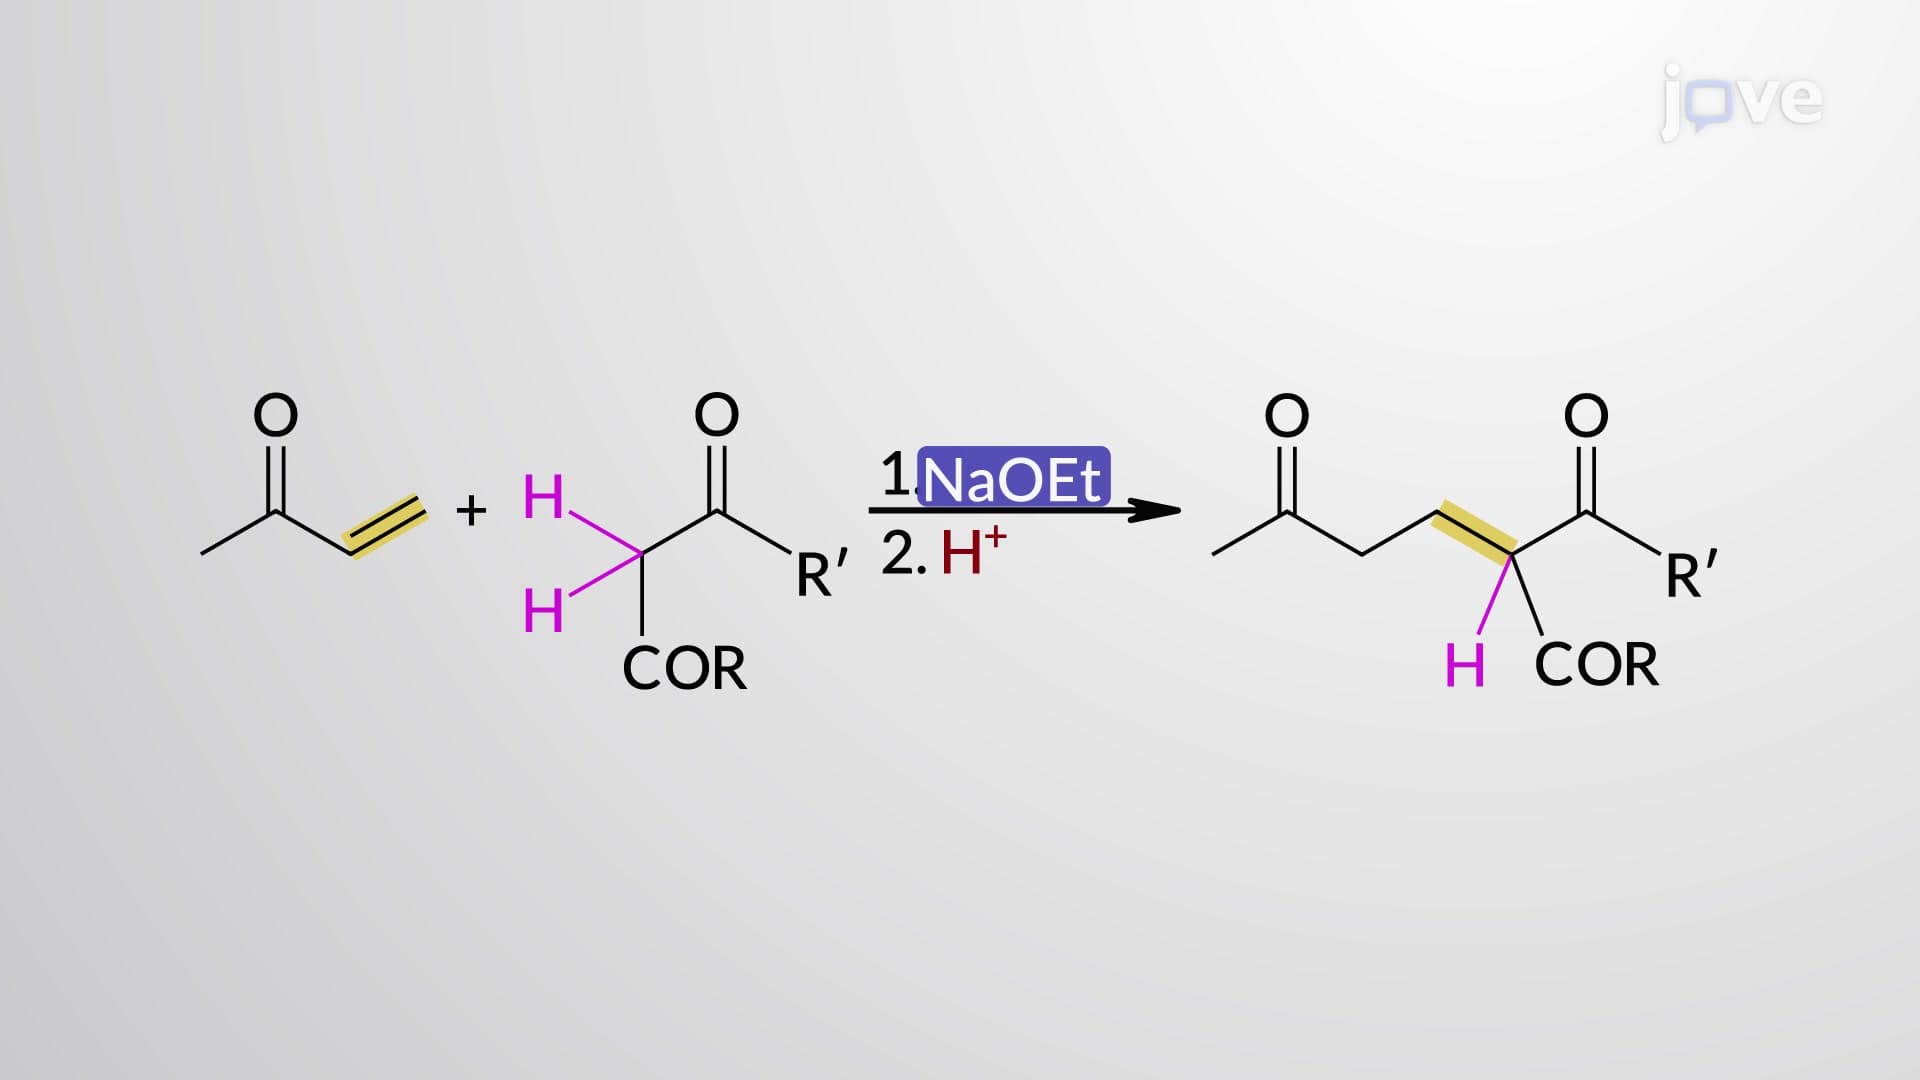
Conjugate Addition of Enolates: Michael Addition
 01:13
Electrophilic Aromatic Substitution: Fluorination and Iodination of Benzene