Related Experiment Video
Updated: Jun 2, 2026
Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
Three-state trajectory surface hopping studies of the photodissociation dynamics of formaldehyde on ab initio
Bina Fu1, Benjamin C Shepler, Joel M Bowman
1Department of Chemistry and Cherry L. Emerson Center for Scientific Computation, Emory University, Atlanta, Georgia 30322, USA.

Abstract:
Full-dimensional, three-state, surface hopping calculations of the photodissociation dynamics of formaldehyde are reported on ab initio potential energy surfaces (PESs) for electronic states S(1), T(1), and S(0). This is the first such study initiated on S(1) with ab initio-calculated spin-orbit couplings among the three states. We employ previous PESs for S(0) and T(1), and a new PES for S(1), which we describe here, as well as new spin-orbit couplings. The time-dependent electronic state populations and the branching ratio of radical products produced from S(0) and T(1) states and that of total radical products and molecular products at three total energies are calculated. Details of the surface hopping dynamics are described, and a novel pathway for isomerization on T(1) via S(0) is reported. Final translational energy distributions of H + HCO products from S(0) and T(1) are also reported as well as the translational energy distribution and final rovibrational distributions of H(2) products from the molecular channel. The present results are compared to previous trajectory calculations initiated from the global minimum of S(0). The roaming pathway leading to low rotational distribution of CO and high vibrational population of H(2) is observed in the present calculations.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:16
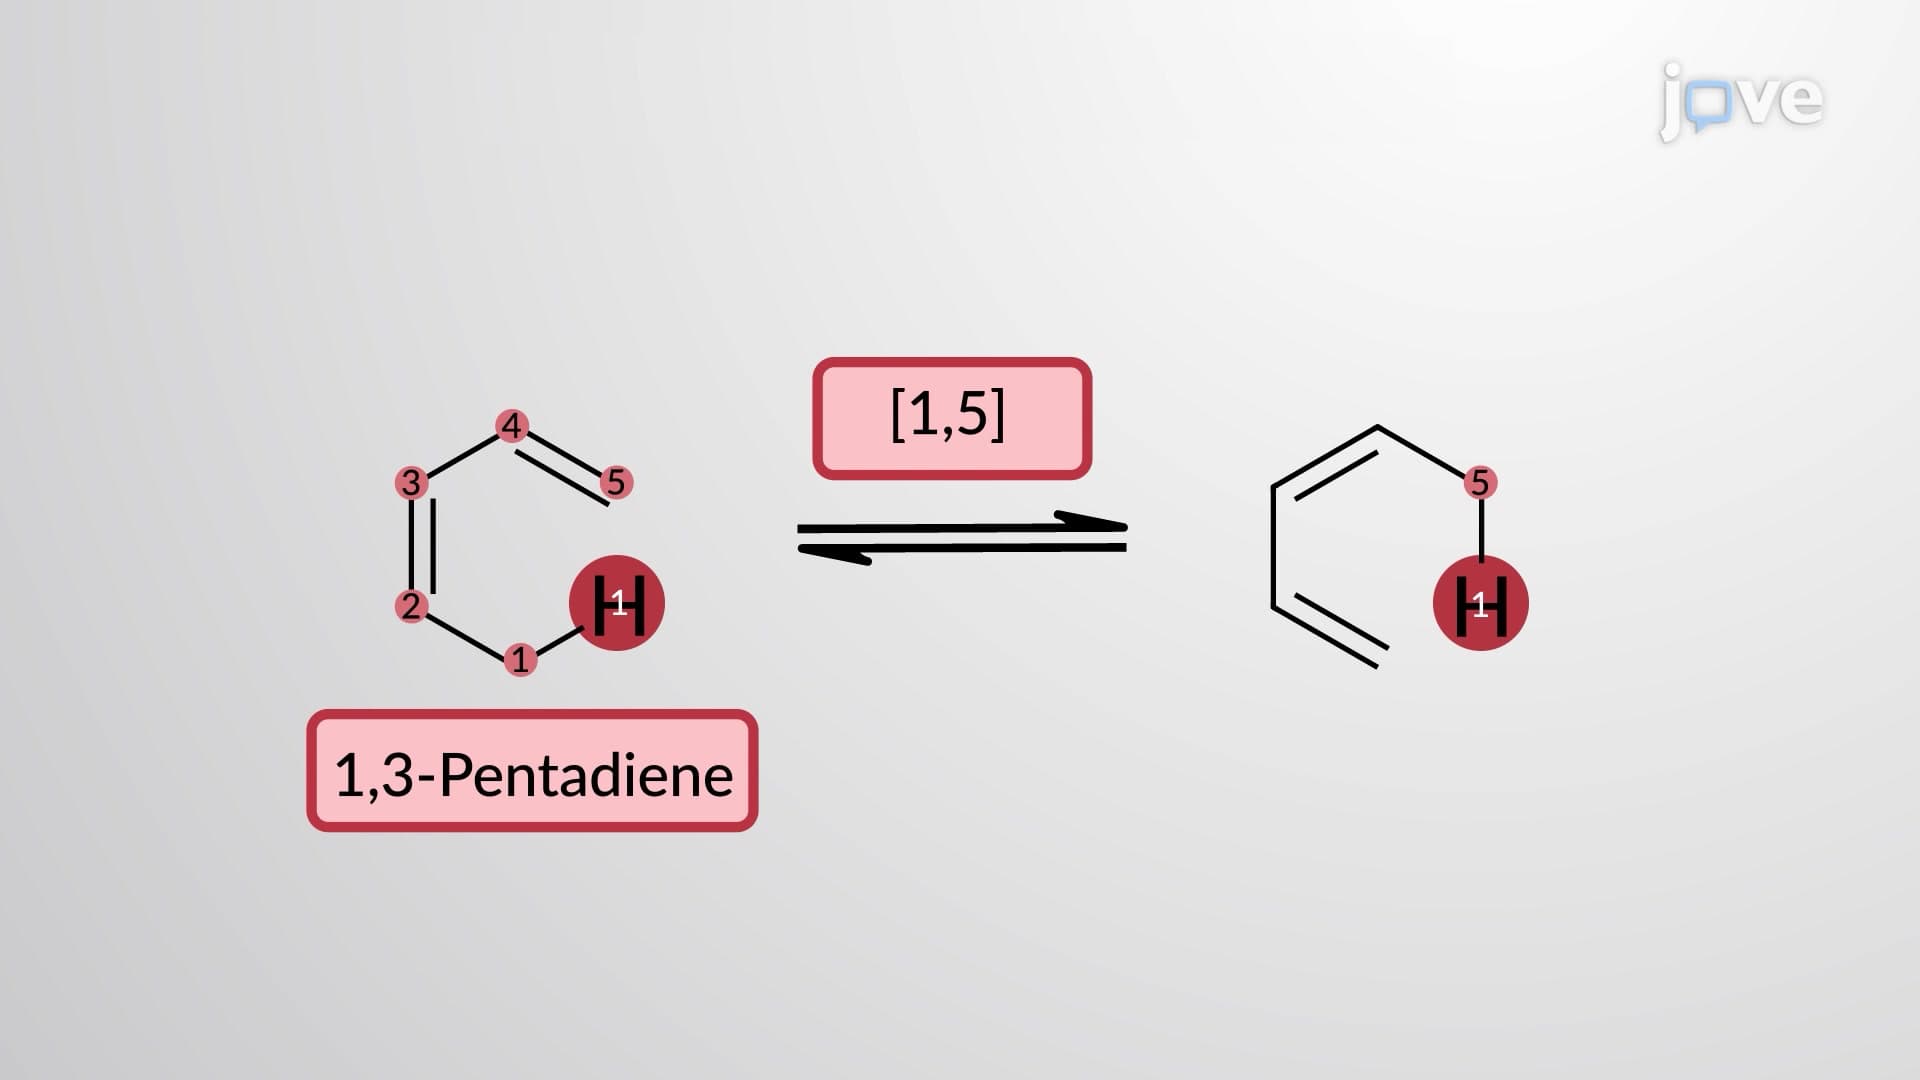
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred to as...
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:25
Deactivation Processes: Jablonski Diagram
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:25
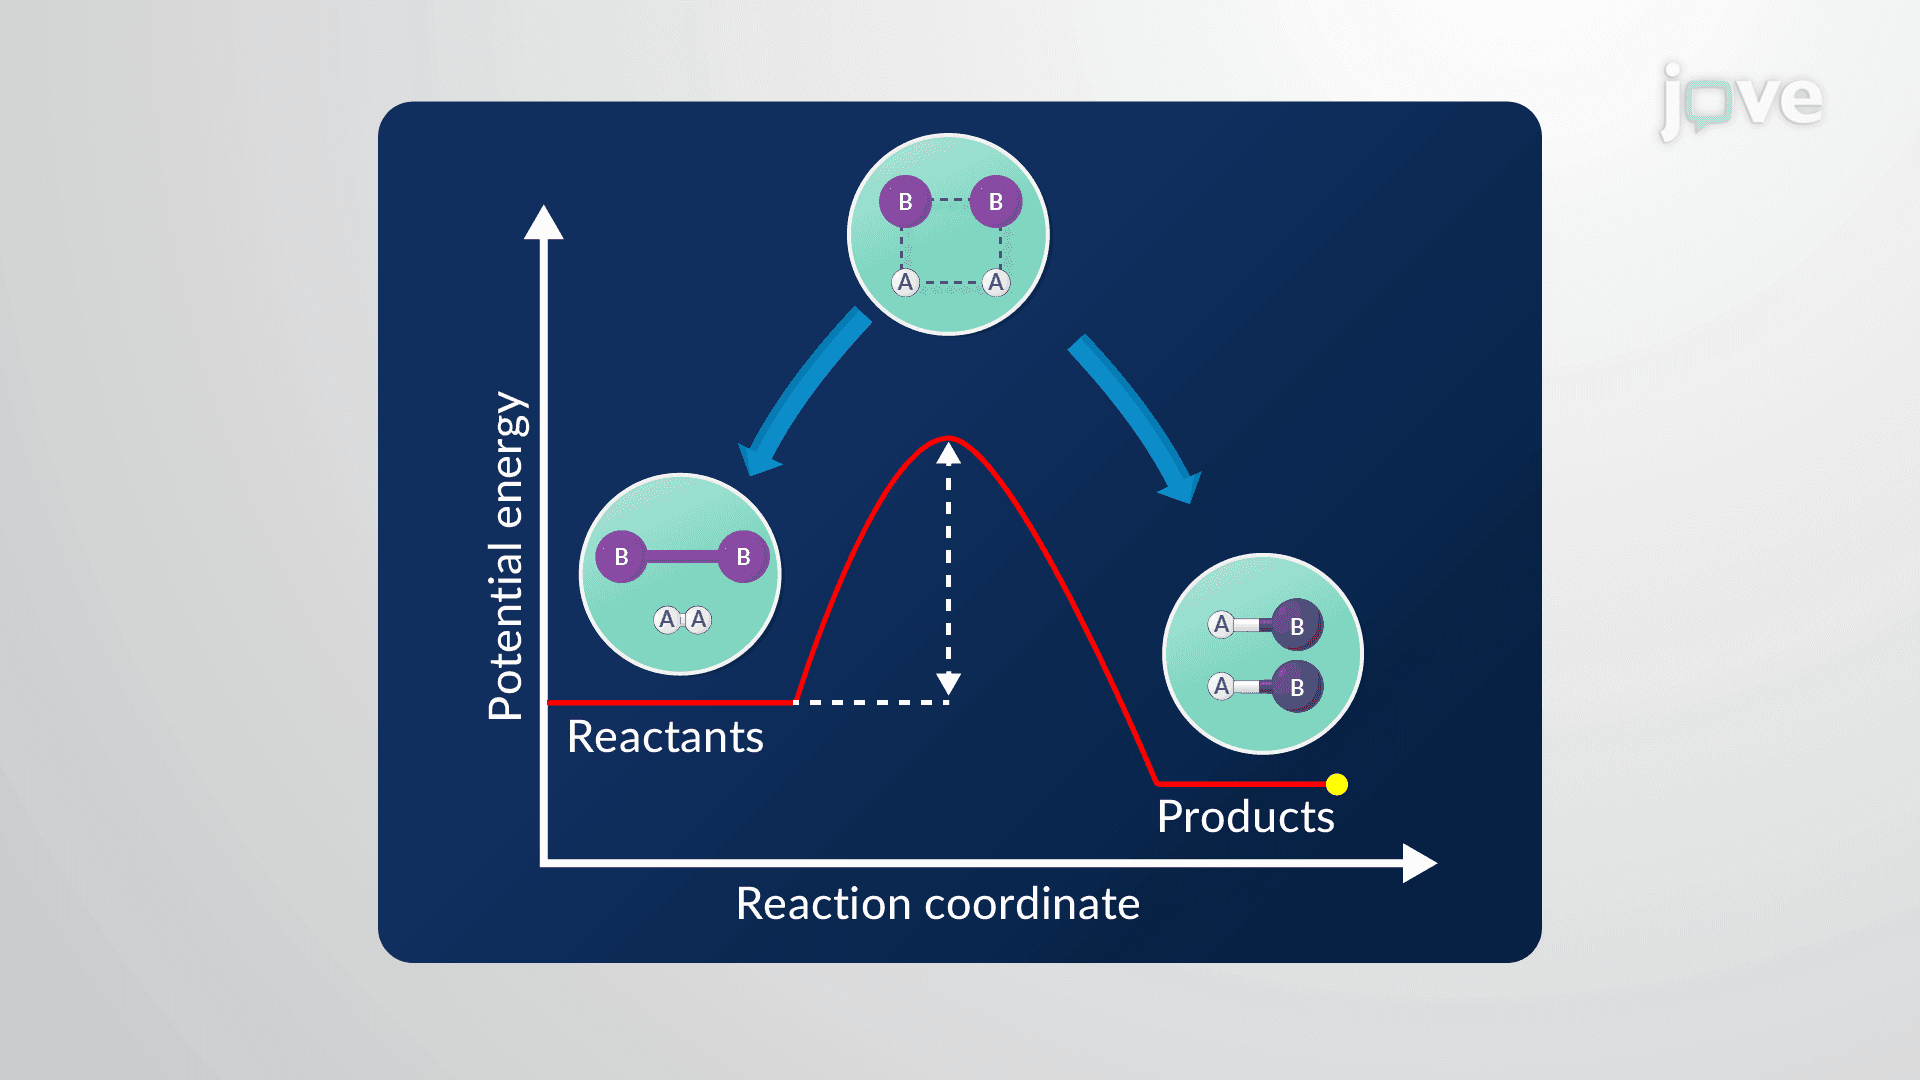
Transition State Theory
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions