Related Experiment Video
Updated: Jun 2, 2026

Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
On the electronic properties of two-dimensional honeycomb GaInN and GaAlN alloys: a molecular analysis
Ernesto Chigo Anota1, Heriberto Hernández Cocoletzi
1Cuerpo Académico Ingeniería en Materiales, Facultad de Ingeniería Química, Benemérita Universidad Autónoma de Puebla, CU San Manuel, CP 72570, Puebla, Mexico. echigoa@yahoo.es

Abstract:
We have performed first principles total energy calculations to investigate the structural and the electronic properties of two-dimensional honeycomb GaAlN and GaInN alloys. Calculations were done using a coronene-like (C(24)H(12)) cluster and for different numbers of Ga, Al, and In atoms. The exchange and correlation potential energies were treated within the generalized gradient approximation (GGA). The bond length, dipole moment, binding energy, and gap between the HOMO and the LUMO are reported as a function of x. The stability of the structures depends on the site of the substituted atom; for example, when three Ga atoms are substituted, the GaInN alloy becomes unstable. The gap in the GaAlN increases from 3.76 eV (GaN) to 4.51 eV (AlN), and in the GaInN decreases to 2.11 eV. The biggest polarity occurs when eight and four Ga atoms are substituted, for GaAlN and GaInN, respectively.
Related Concept Videos
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability. Many...
 03:51
Molecular Orbital Theory II
 02:42
Valence Bond Theory
 02:40
MO Theory and Covalent Bonding
 01:18
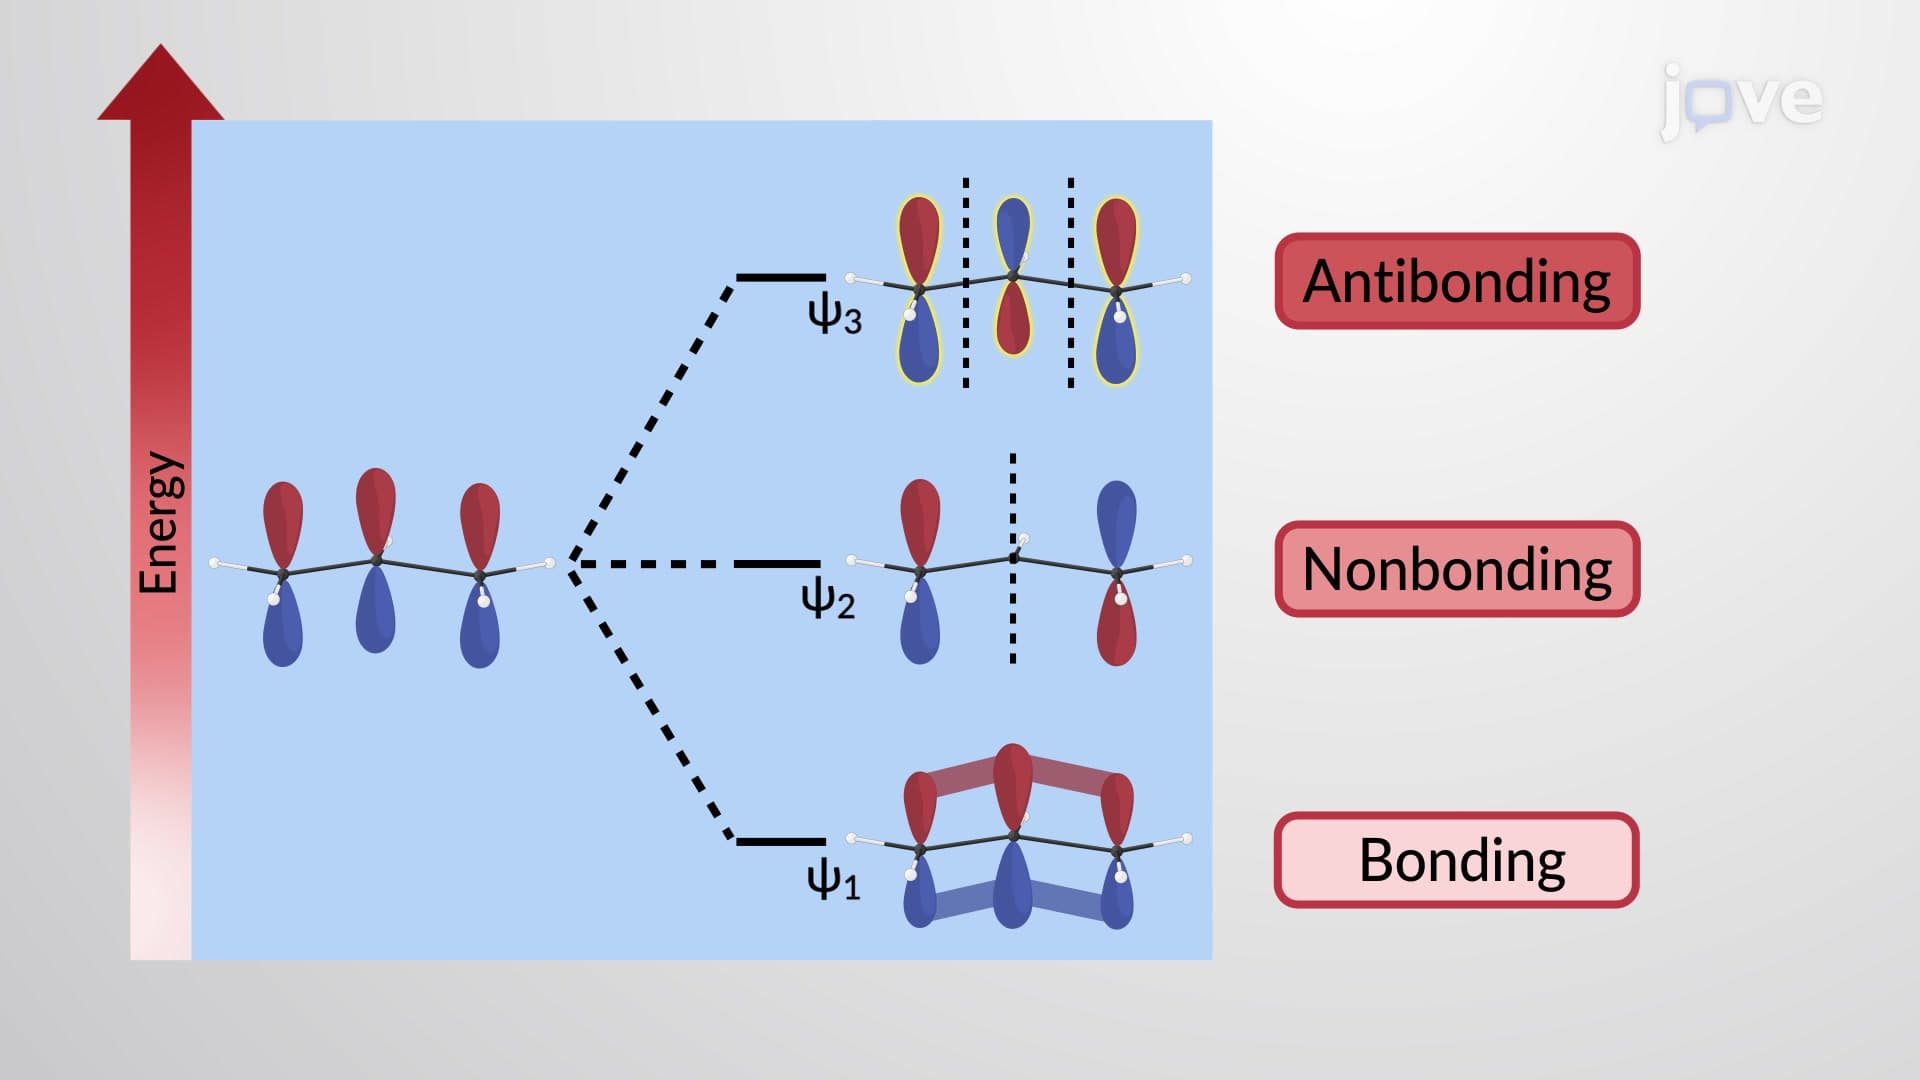
π Molecular Orbitals of the Allyl Cation and Anion
 02:35
Molecular Orbital Theory I