

Calculation of the structure, potential energy surface, vibrational dynamics, and electric dipole properties for the
M Preller1, J Grunenberg, V P Bulychev
1Institut für Organische Chemie, Technische Universität Braunschweig, Braunschweig, Deutschland. Preller.matthias@mh-hannover.de

Abstract:
We report the structure and spectroscopic characteristics for the Xe:HI van der Waals binary isomers determined from variational solutions of two-dimensional and three-dimensional (3D) vibrational Schrödinger equations. The solutions are based on a potential energy surface computed at the coupled-cluster level of theory including single and double excitations and a non-iterative perturbation treatment of triple excitations [CCSD(T)]. The dipole moment surface was calculated using quadratic configuration interaction (QCISD). The global potential minimum is shown to be located at the anti-hydrogen-bonded Xe-IH isomer, 21 cm(-1) below the secondary local minimum associated with the hydrogen-bonded Xe-HI isomeric form. The dissociation energy from the global minimum is 245.9 cm(-1). 3D Schrödinger equations are solved for the rotational quantum numbers J = k = 0, 1, and 2, without invoking an adiabatic separation of high- and low-frequency degrees of freedom. The vibrational ground state resides in the Xe-HI potential well, while the first excited state, 8.59 cm(-1) above the ground, occupies the Xe-IH well. We find that intra-complex dynamics exhibits a sudden transformation upon increase of the r(HI) bond length, accompanied by abrupt changes in the geometric and dipole parameters. A similar chaotic behavior is predicted to occur for Xe:DI at a shorter r(DI) bond length, which implies stronger coupling between low- and high-frequency motions in the heavier complex. Our calculations confirm a strong enhancement for the r(HI) stretch fundamental and a significant weakening for the first overtone vibrational transitions in Xe:HI, as compared to those in the free HI molecule. A qualitative explanation of this, earlier experimentally detected effect is suggested.


Related Concept Videos
 01:10
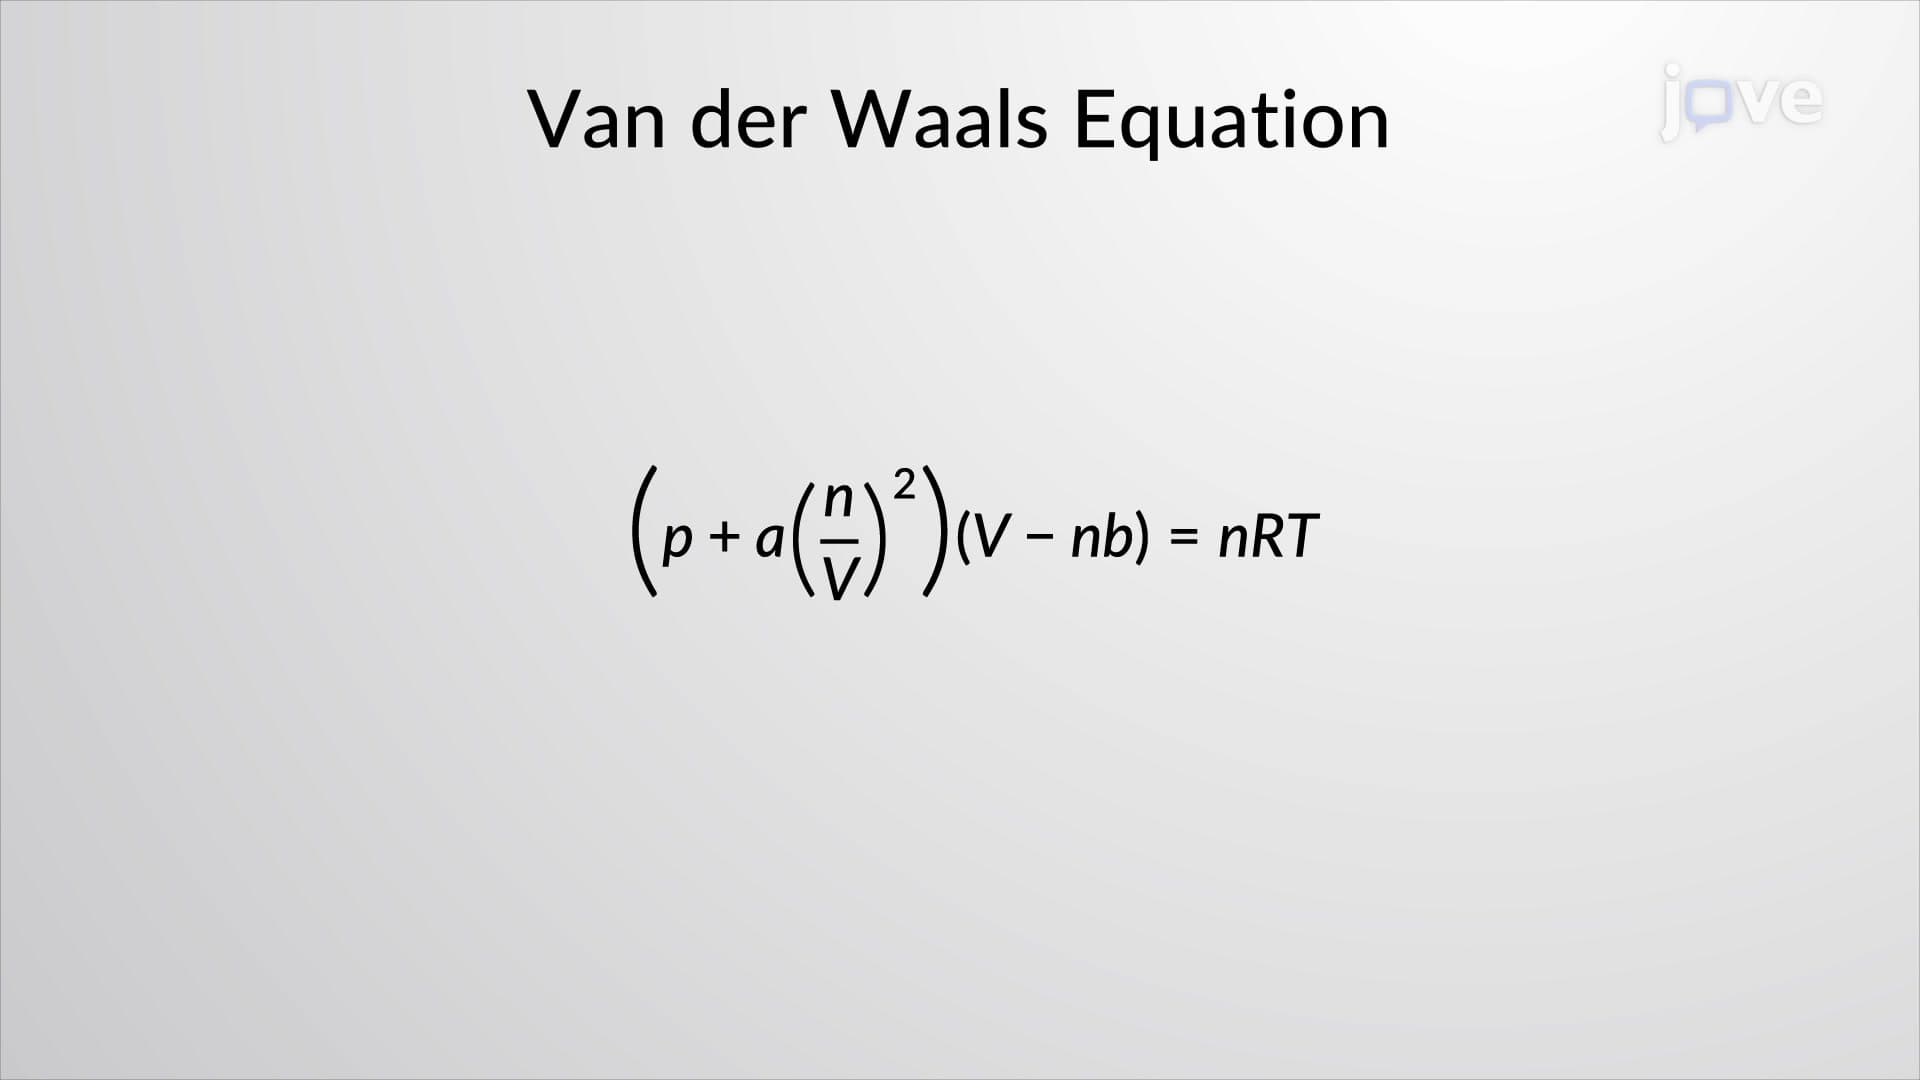
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:27
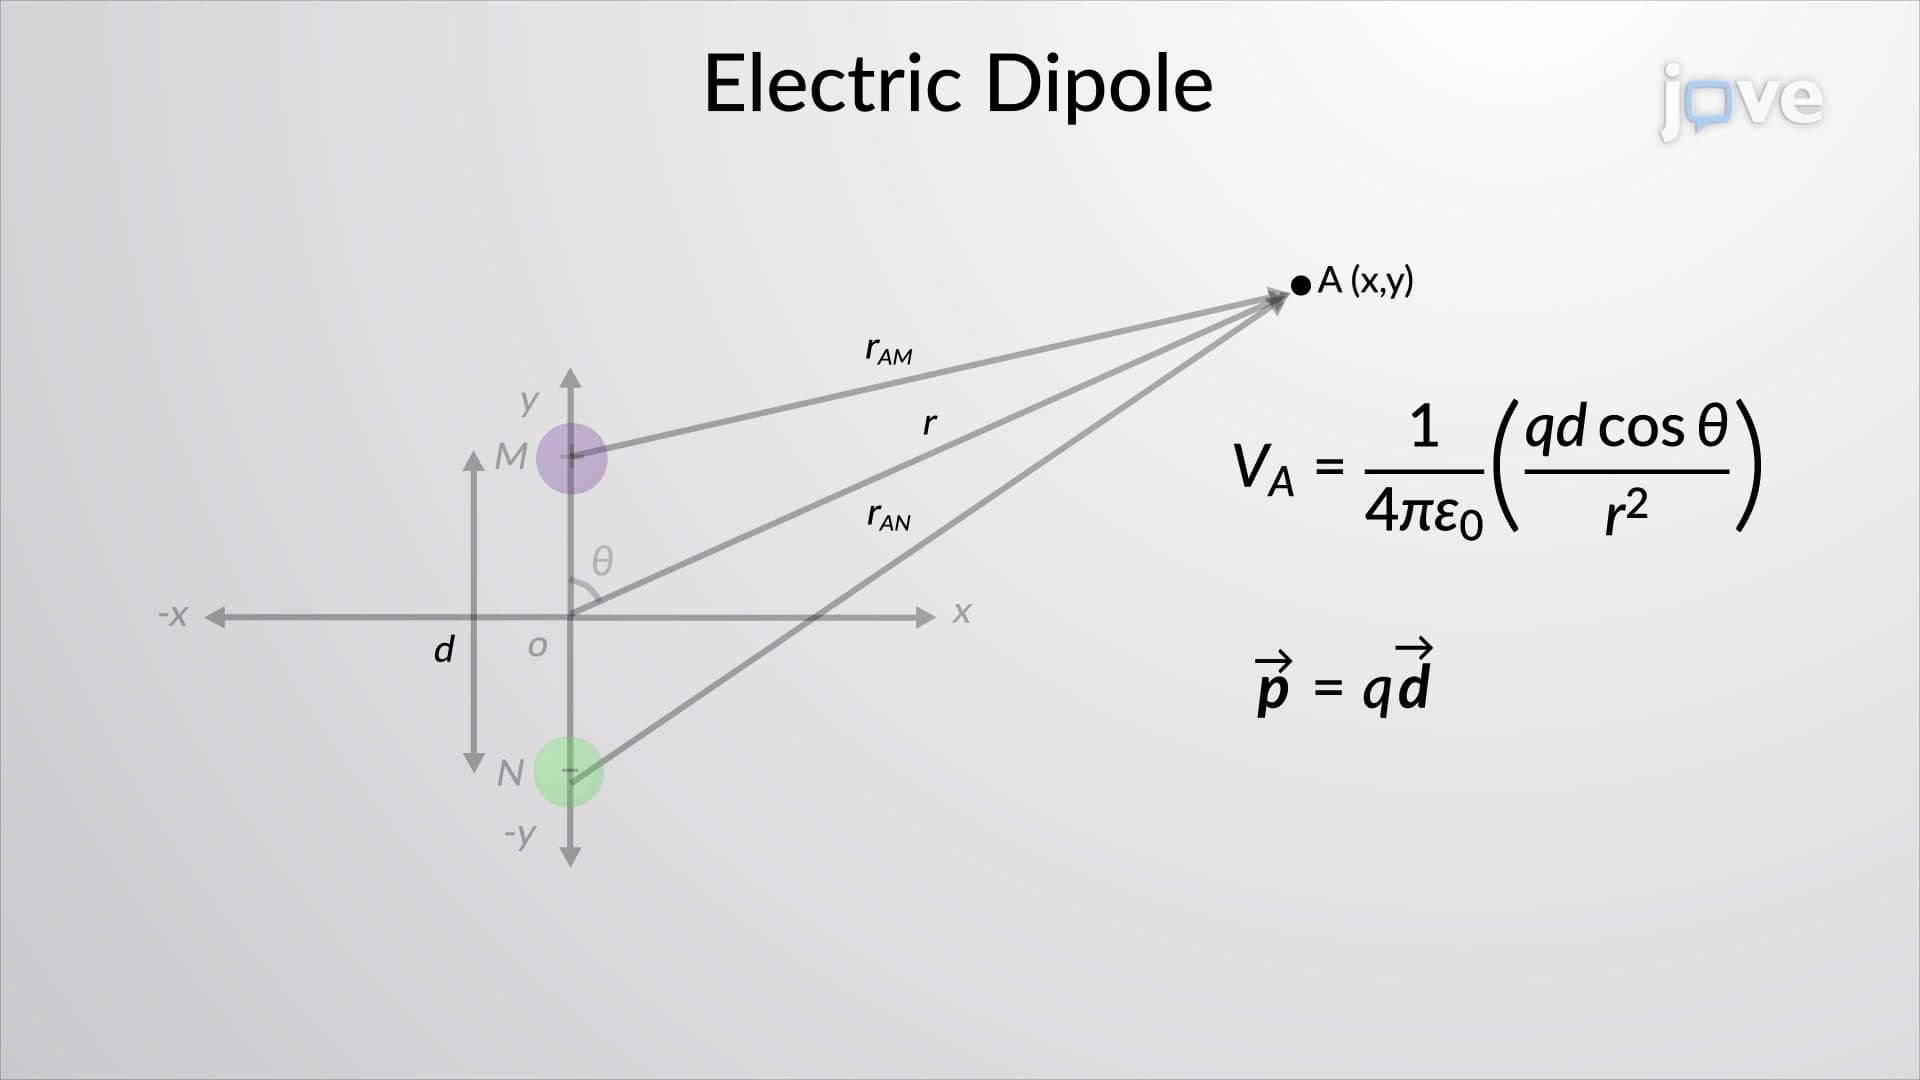
Calculations of Electric Potential II
Consider a...
 01:24
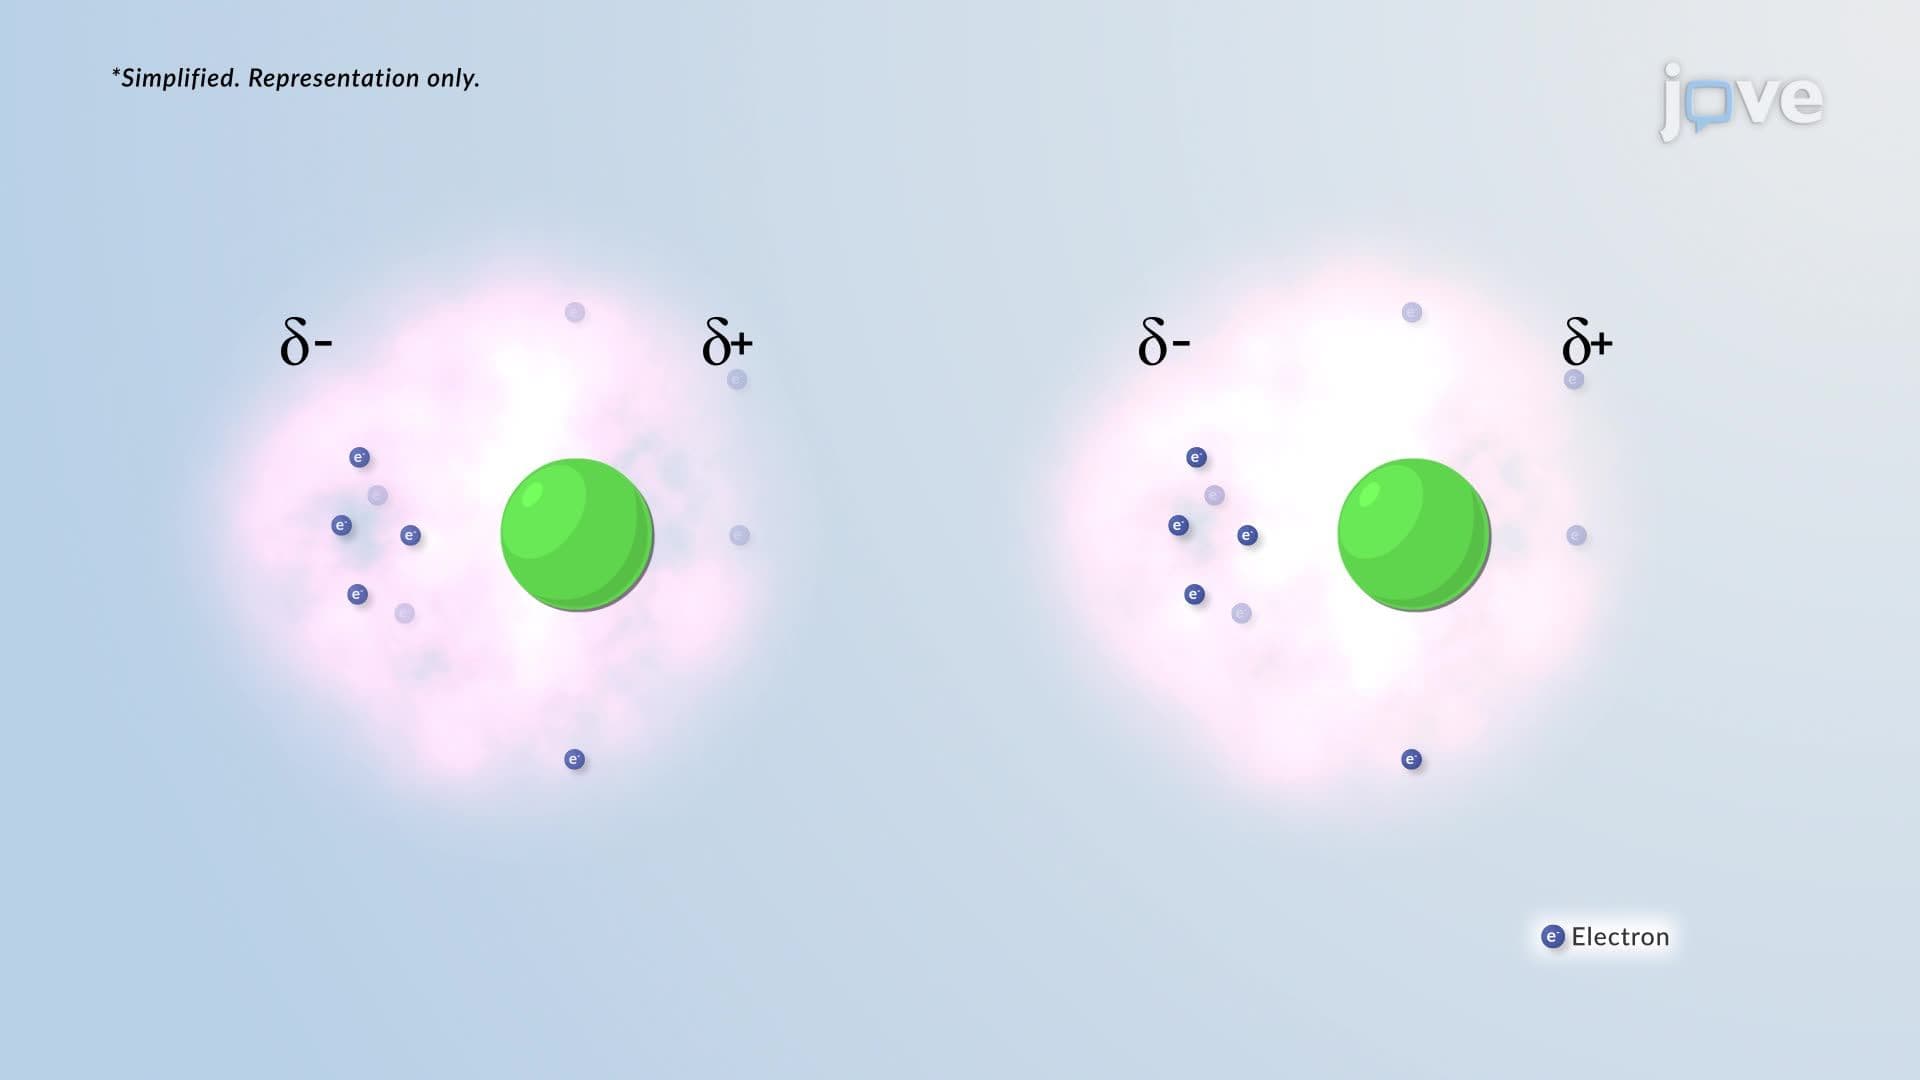
Van der Waals Interactions
 01:13
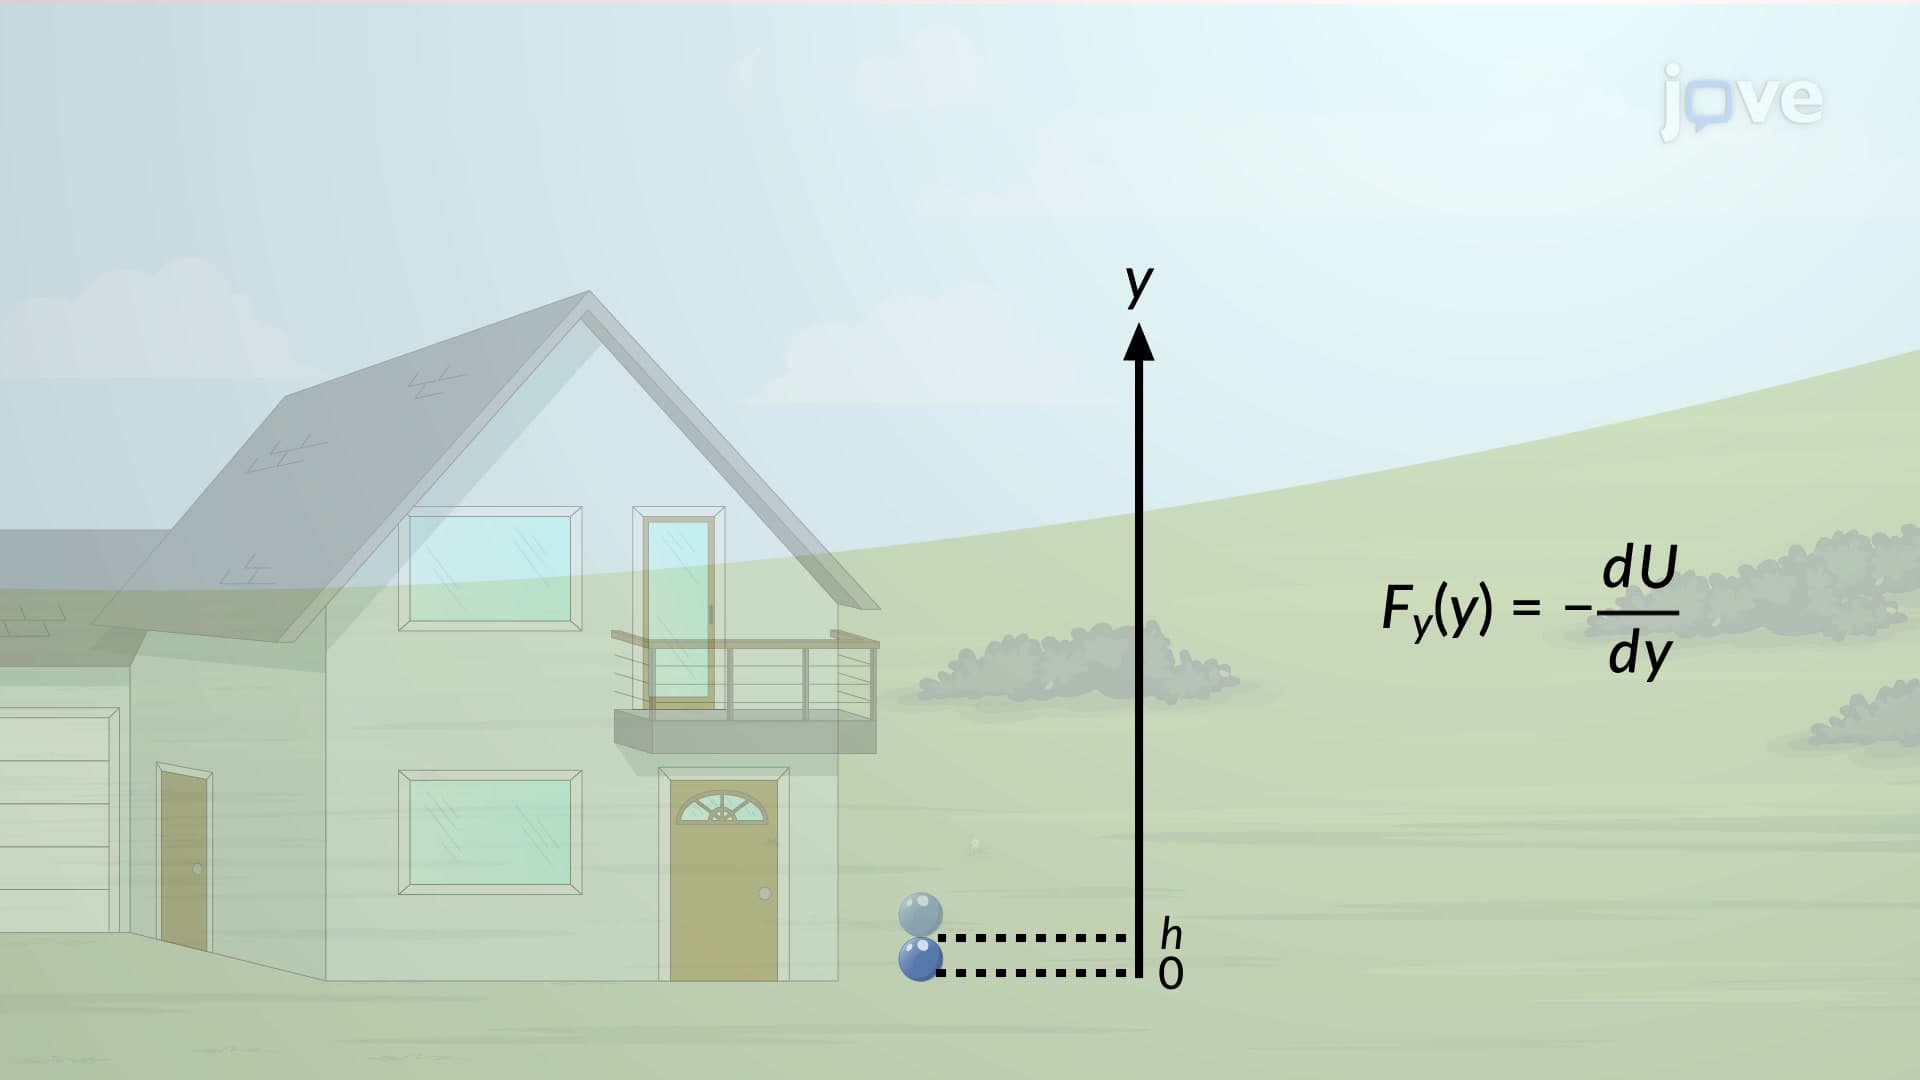
Force and Potential Energy in One Dimension
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...