Related Experiment Video
Updated: Jun 2, 2026

Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
Exome sequencing identifies mitochondrial alanyl-tRNA synthetase mutations in infantile mitochondrial cardiomyopathy
Alexandra Götz1, Henna Tyynismaa, Liliya Euro
1Research Programs Unit, Molecular Neurology, Biomedicum-Helsinki, University of Helsinki, Helsinki, Finland.

Abstract:
Infantile cardiomyopathies are devastating fatal disorders of the neonatal period or the first year of life. Mitochondrial dysfunction is a common cause of this group of diseases, but the underlying gene defects have been characterized in only a minority of cases, because tissue specificity of the manifestation hampers functional cloning and the heterogeneity of causative factors hinders collection of informative family materials. We sequenced the exome of a patient who died at the age of 10 months of hypertrophic mitochondrial cardiomyopathy with combined cardiac respiratory chain complex I and IV deficiency. Rigorous data analysis allowed us to identify a homozygous missense mutation in AARS2, which we showed to encode the mitochondrial alanyl-tRNA synthetase (mtAlaRS). Two siblings from another family, both of whom died perinatally of hypertrophic cardiomyopathy, had the same mutation, compound heterozygous with another missense mutation. Protein structure modeling of mtAlaRS suggested that one of the mutations affected a unique tRNA recognition site in the editing domain, leading to incorrect tRNA aminoacylation, whereas the second mutation severely disturbed the catalytic function, preventing tRNA aminoacylation. We show here that mutations in AARS2 cause perinatal or infantile cardiomyopathy with near-total combined mitochondrial respiratory chain deficiency in the heart. Our results indicate that exome sequencing is a powerful tool for identifying mutations in single patients and allows recognition of the genetic background in single-gene disorders of variable clinical manifestation and tissue-specific disease. Furthermore, we show that mitochondrial disorders extend to prenatal life and are an important cause of early infantile cardiac failure.
More Related Videos
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase (COX/SDH) Double-labeling Histochemistry
Published on: November 23, 2011
 07:24
Genotyping Single Nucleotide Polymorphisms in the Mitochondrial Genome by Pyrosequencing
Published on: February 10, 2023
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 02:59
Animal Mitochondrial Genetics
 01:39
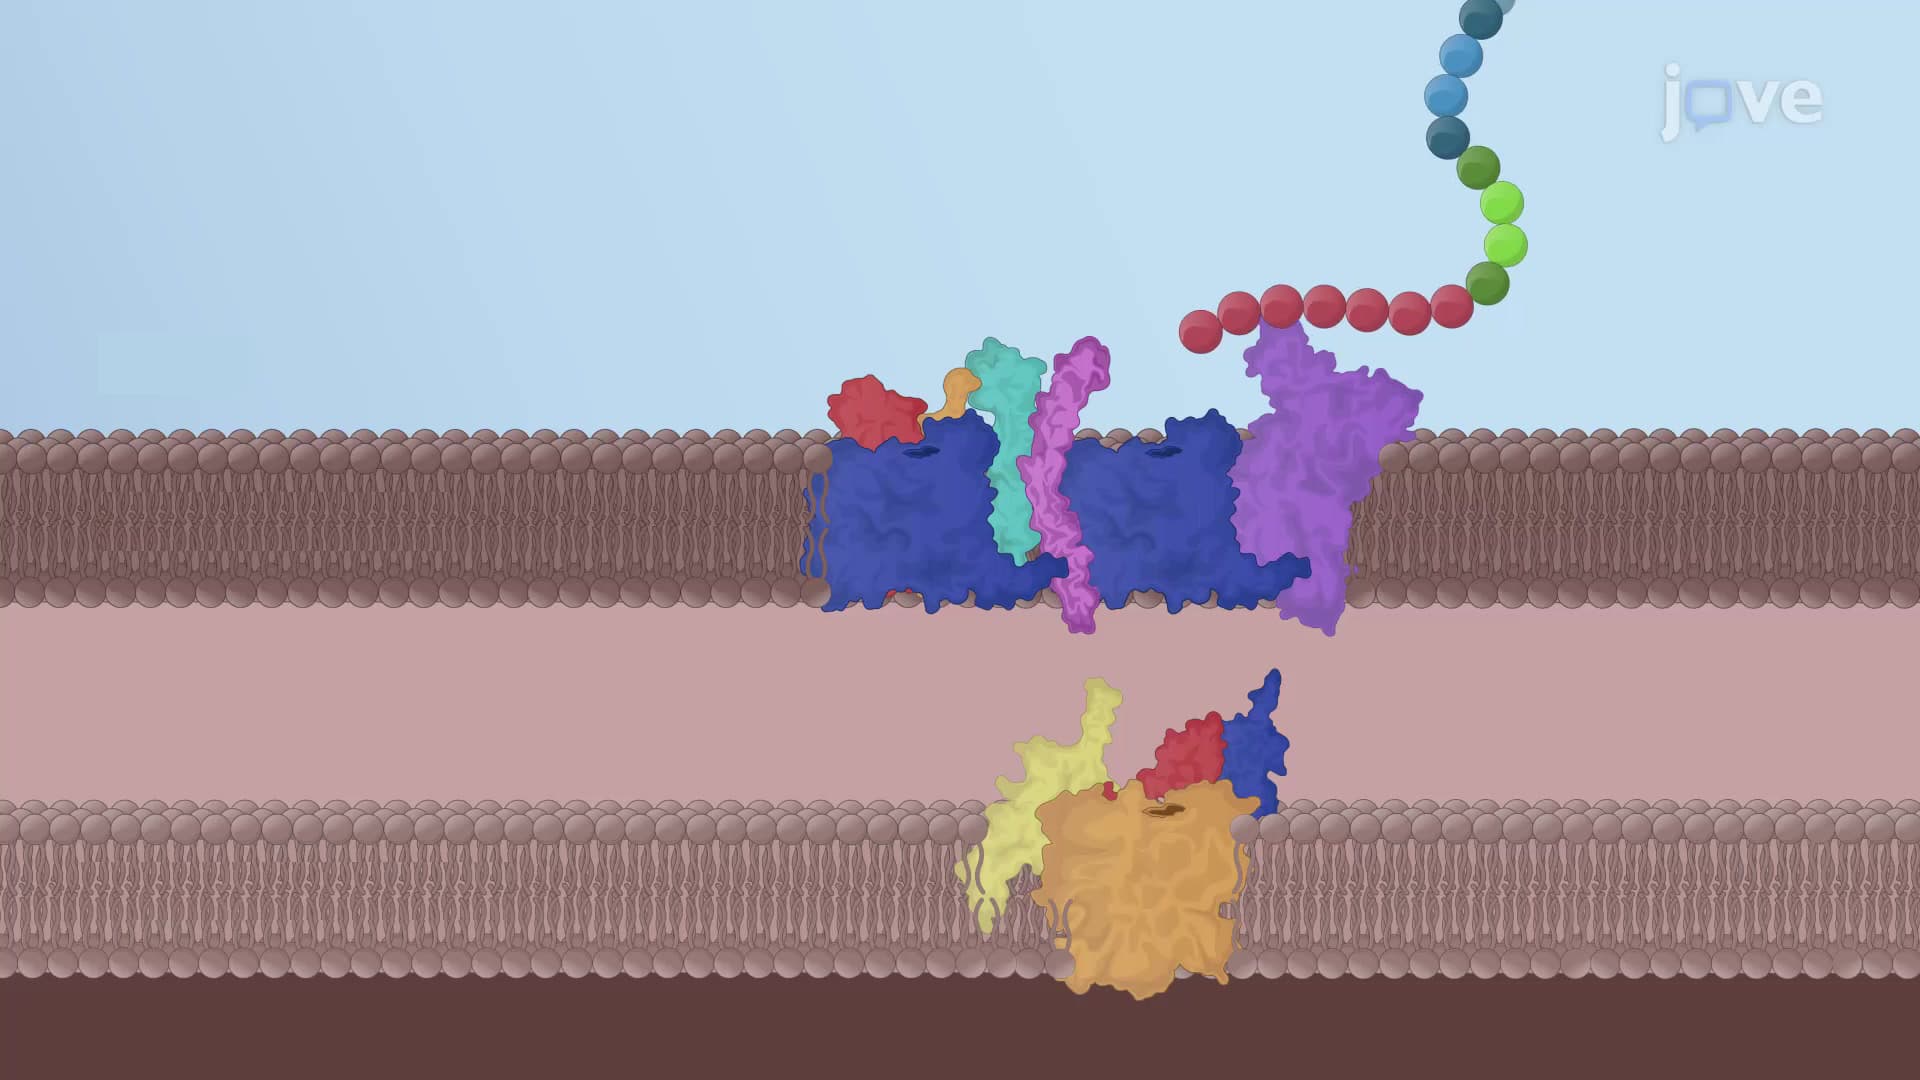
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...
 01:39
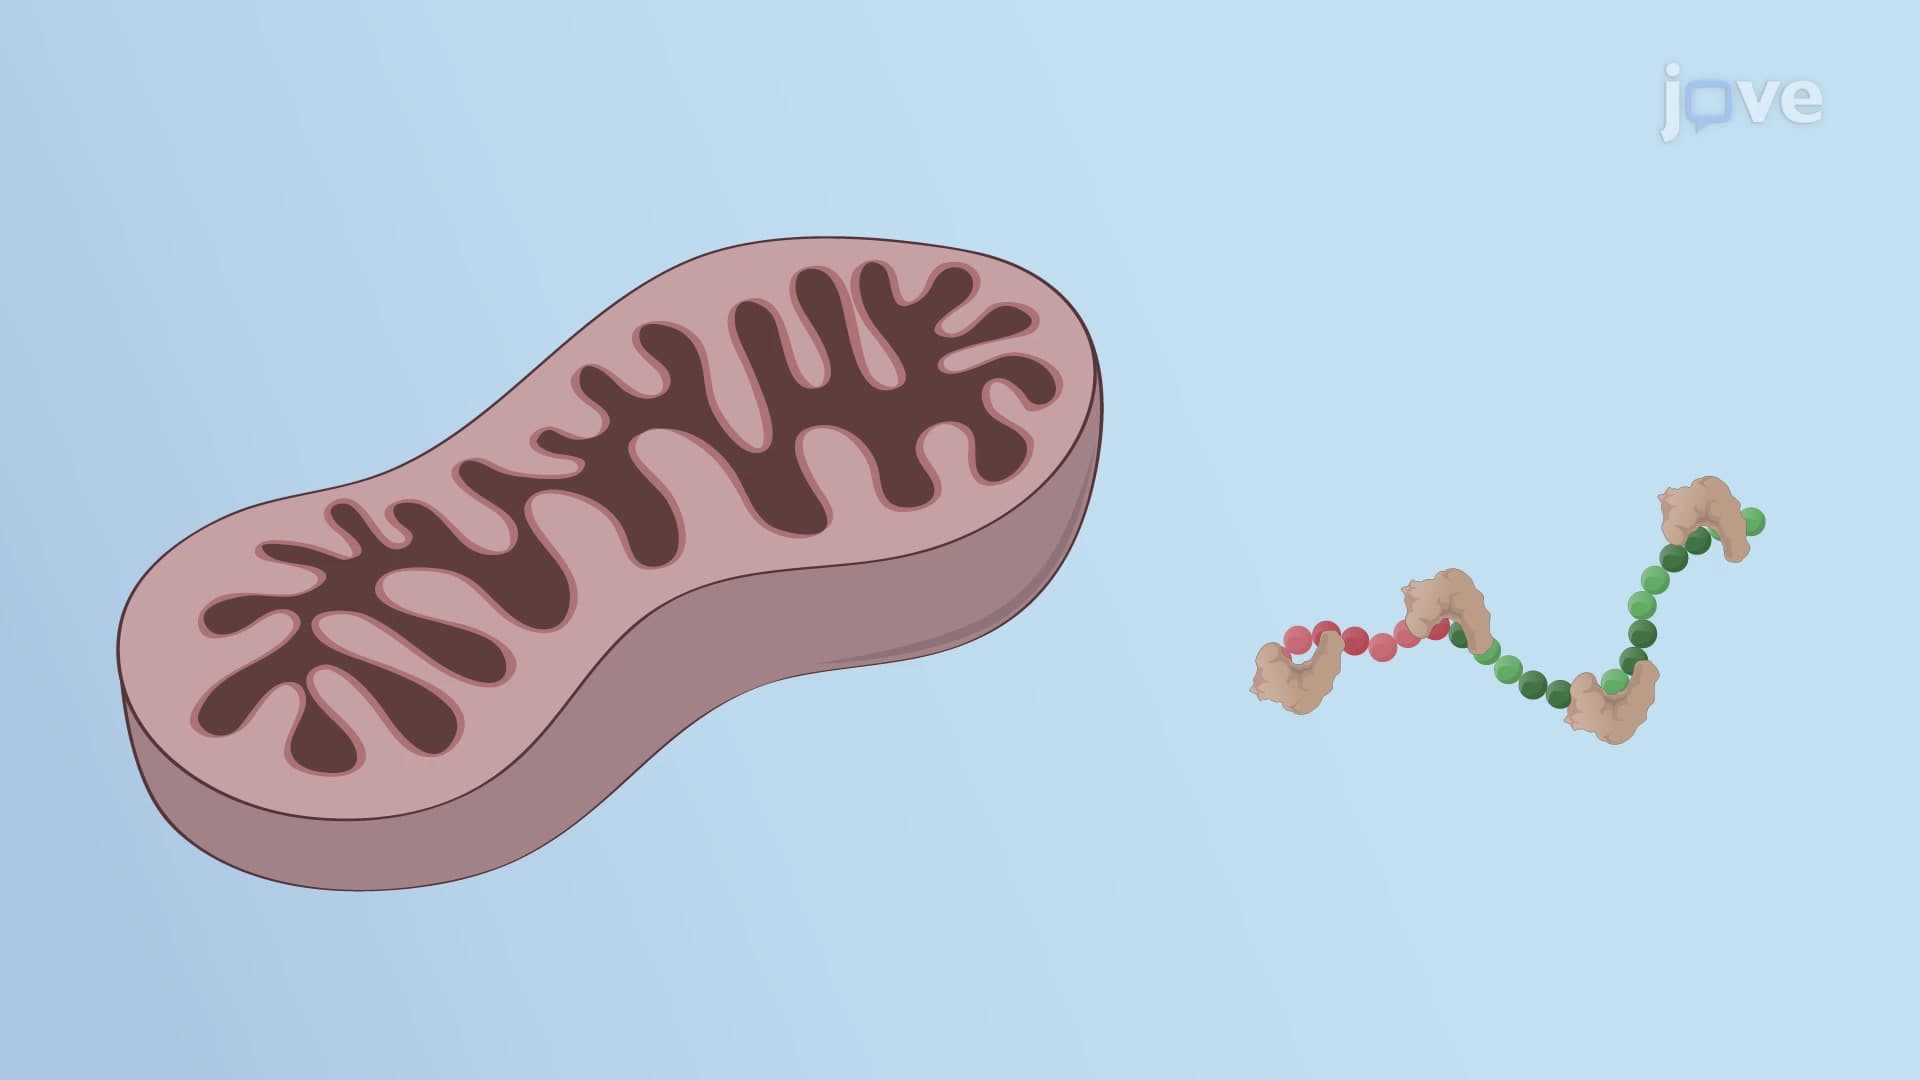
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:18
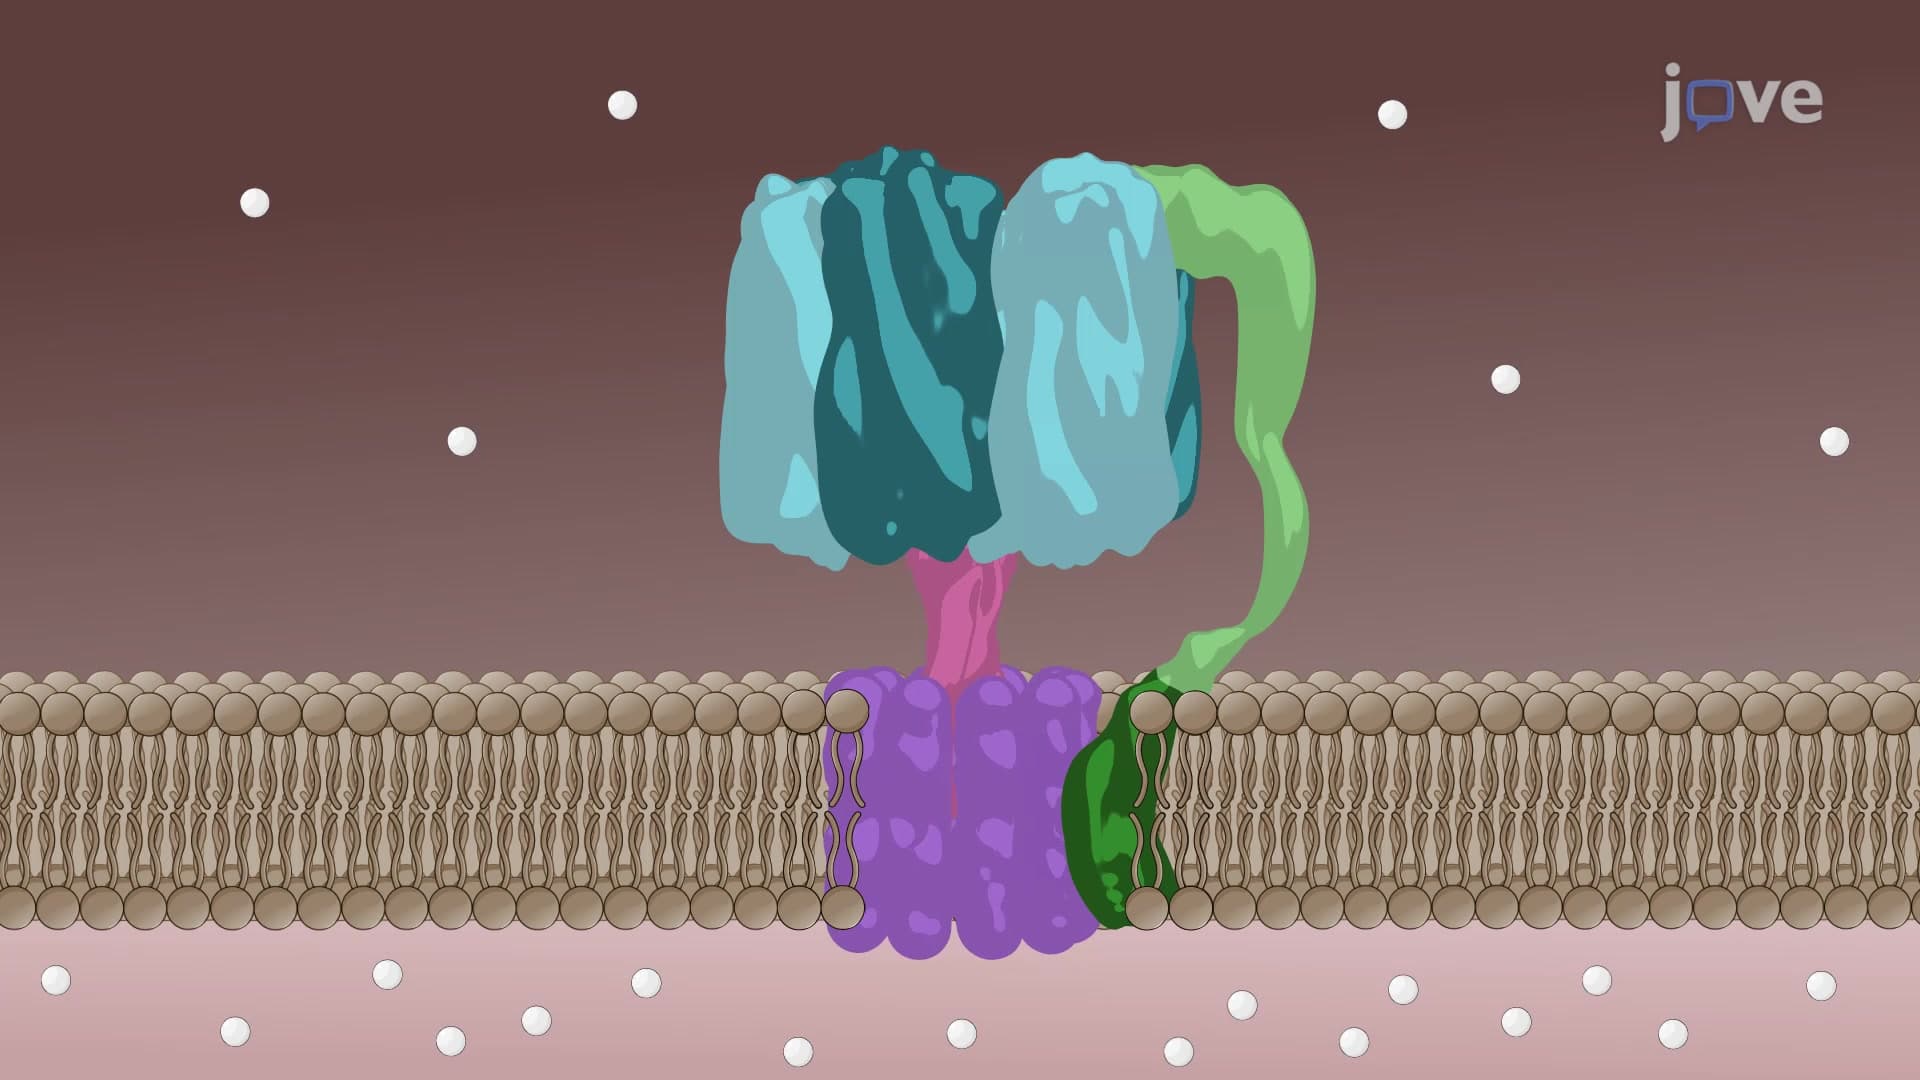
ATP Synthase: Structure
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life