Related Experiment Video
Updated: Jun 1, 2026
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Exhaustive sampling of docking poses reveals binding hypotheses for propafenone type inhibitors of P-glycoprotein
Freya Klepsch1, Peter Chiba, Gerhard F Ecker
1Department of Medicinal Chemistry, University of Vienna, Vienna, Austria.

Abstract:
Overexpression of the xenotoxin transporter P-glycoprotein (P-gp) represents one major reason for the development of multidrug resistance (MDR), leading to the failure of antibiotic and cancer therapies. Inhibitors of P-gp have thus been advocated as promising candidates for overcoming the problem of MDR. However, due to lack of a high-resolution structure the concrete mode of interaction of both substrates and inhibitors is still not known. Therefore, structure-based design studies have to rely on protein homology models. In order to identify binding hypotheses for propafenone-type P-gp inhibitors, five different propafenone derivatives with known structure-activity relationship (SAR) pattern were docked into homology models of the apo and the nucleotide-bound conformation of the transporter. To circumvent the uncertainty of scoring functions, we exhaustively sampled the pose space and analyzed the poses by combining information retrieved from SAR studies with common scaffold clustering. The results suggest propafenone binding at the transmembrane helices 5, 6, 7 and 8 in both models, with the amino acid residue Y307 playing a crucial role. The identified binding site in the non-energized state is overlapping with, but not identical to, known binding areas of cyclic P-gp inhibitors and verapamil. These findings support the idea of several small binding sites forming one large binding cavity. Furthermore, the binding hypotheses for both catalytic states were analyzed and showed only small differences in their protein-ligand interaction fingerprints, which indicates only small movements of the ligand during the catalytic cycle.

Related Concept Videos
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:16
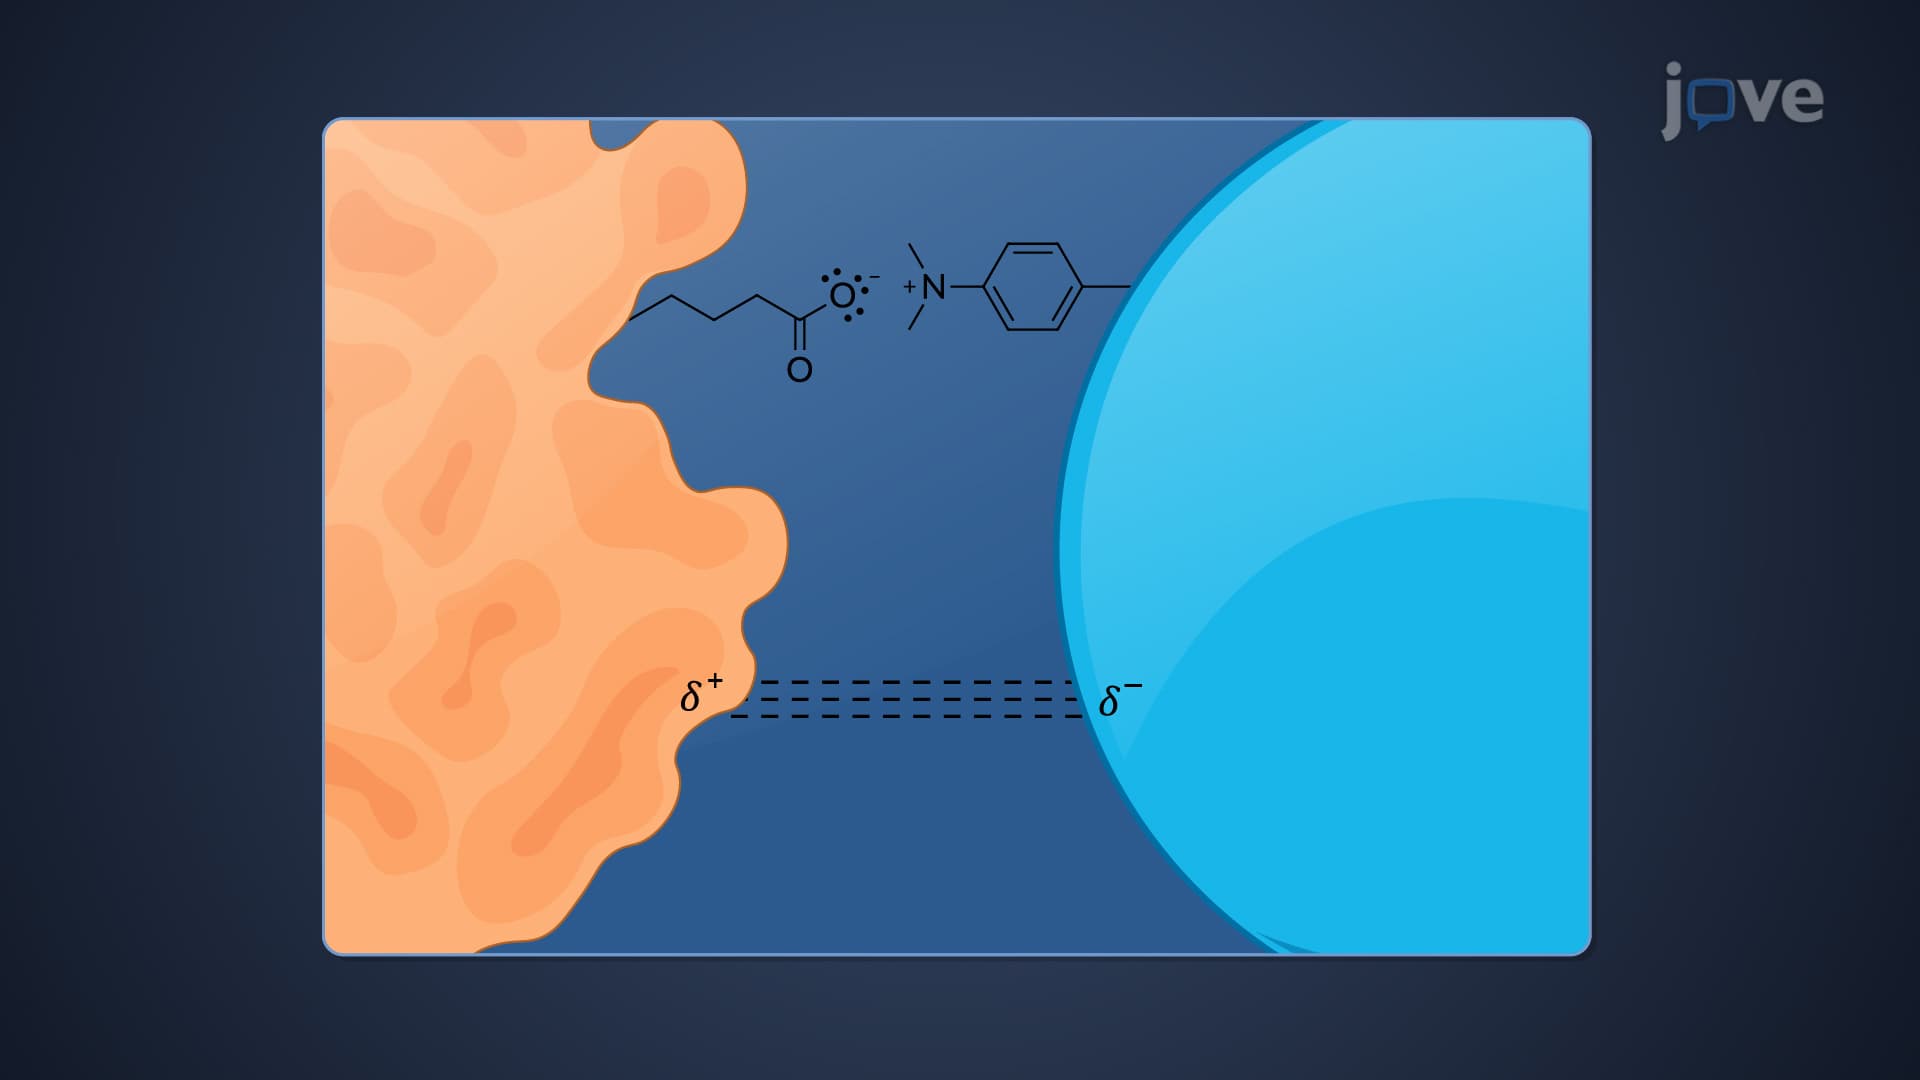
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:09
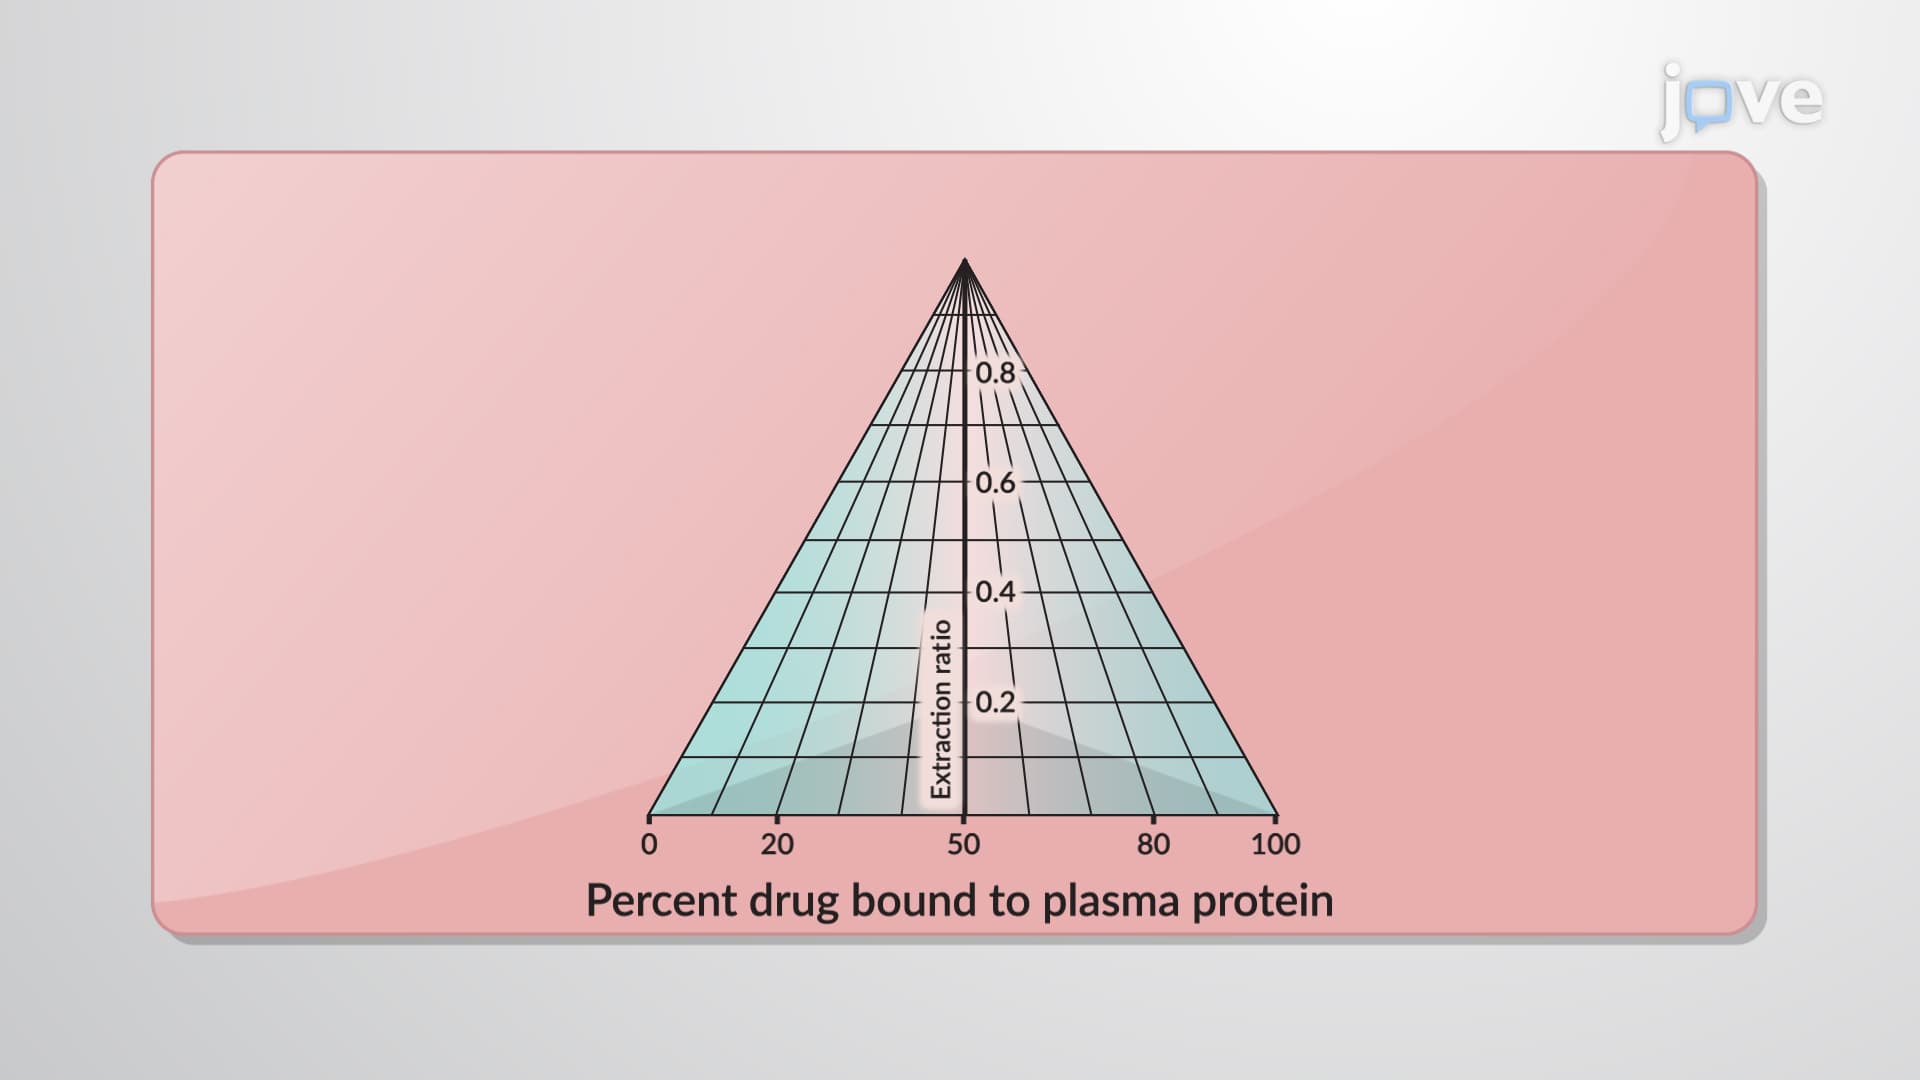
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:30
Drug Binding to Blood Components
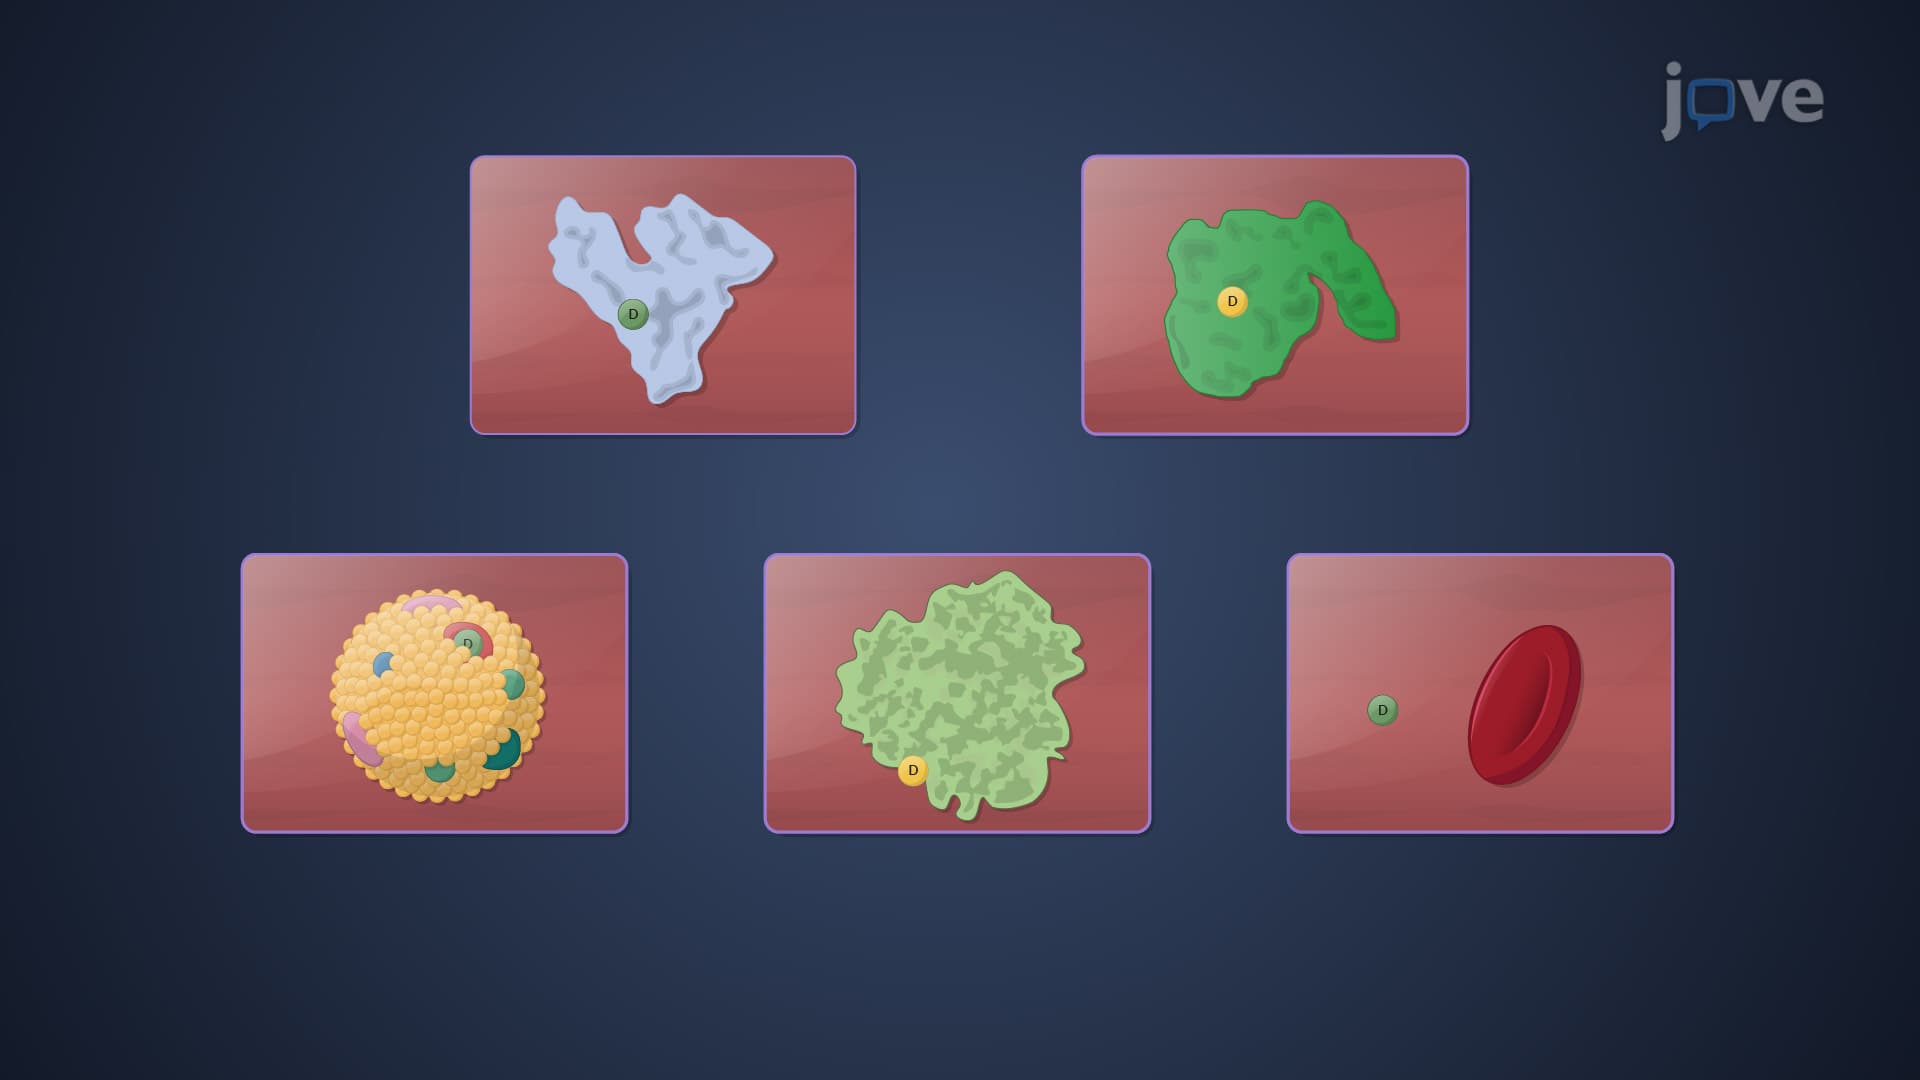
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...
 02:04
Protein-protein Interfaces