Related Experiment Video
Updated: Jun 1, 2026

Adapting Taylor Dispersion to Measure the Dispersion Coefficient of Electrolyte Solutions via an Accessible Microfluidic Setup
Published on: October 7, 2025
Contributions of chemical exchange to T1ρ dispersion in a tissue model
Jared G Cobb1, Jingping Xie, John C Gore
1Institute of Imaging Science, Vanderbilt University, Nashville, Tennessee 37232-2310, USA. Jared.G.Cobb@Vanderbilt.Edu

Abstract:
Variations in T(1ρ) with locking-field strength (T(1ρ) dispersion) may be used to estimate proton exchange rates. We developed a novel approach utilizing the second derivative of the dispersion curve to measure exchange in a model system of cross-linked polyacrylamide gels. These gels were varied in relative composition of comonomers, increasing stiffness, and in pH, modifying exchange rates. Magnetic resonance images were recorded with a spin-locking sequence as described by Sepponen et al. These measurements were fit to a mono-exponential decay function yielding values for T(1ρ) at each locking-field measured. These values were then fit to a model by Chopra et al. for estimating exchange rates. For low stiffness gels, the calculated exchange values increased by a factor of 4 as pH increased, consistent with chemical exchange being the dominant contributor to T(1ρ) dispersion. Interestingly, calculated chemical exchange rates also increased with stiffness, likely due to modified side-chain exchange kinetics as the composition varied. This article demonstrates a new method to assess the structural and chemical effects on T(1ρ) relaxation dispersion with a suitable model. These phenomena may be exploited in an imaging context to emphasize the presence of nuclei of specific exchange rates, rather than chemical shifts.
Related Concept Videos
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:06
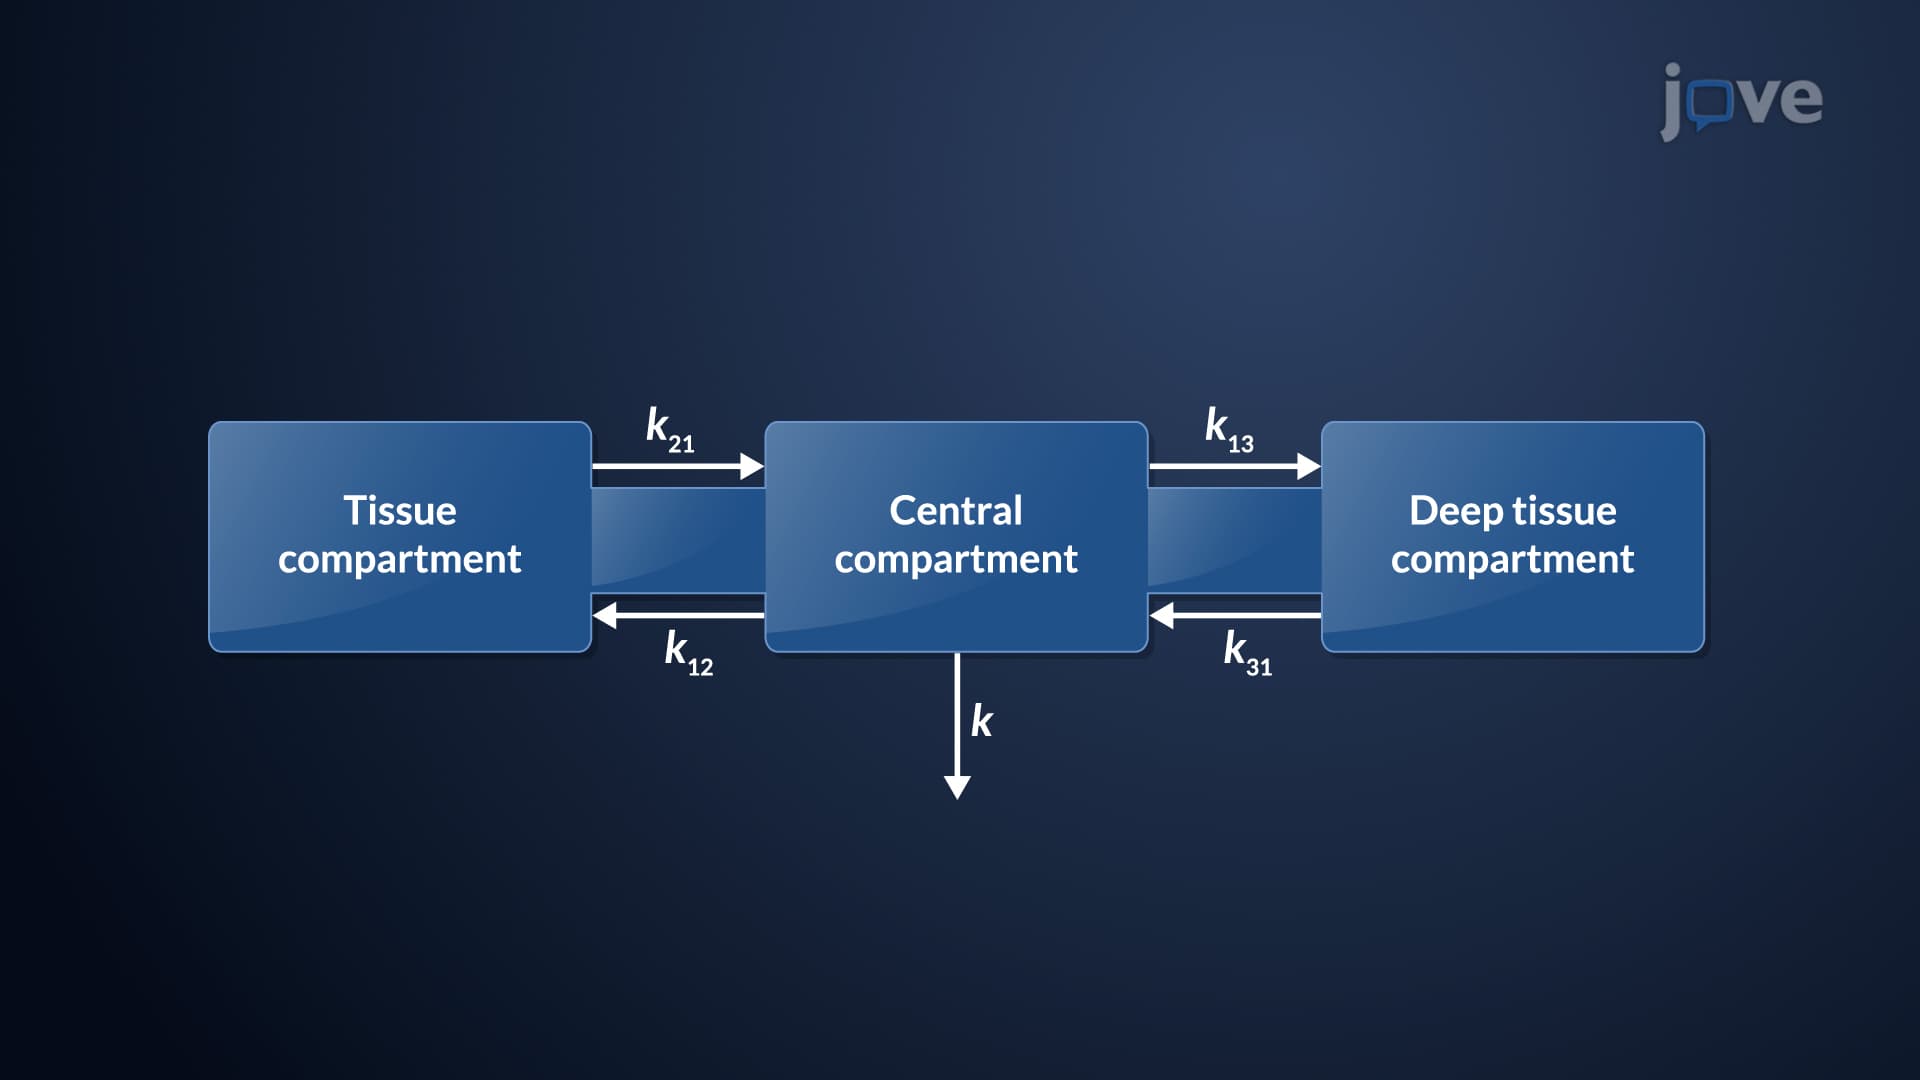
Three-Compartment Open Model
 00:57
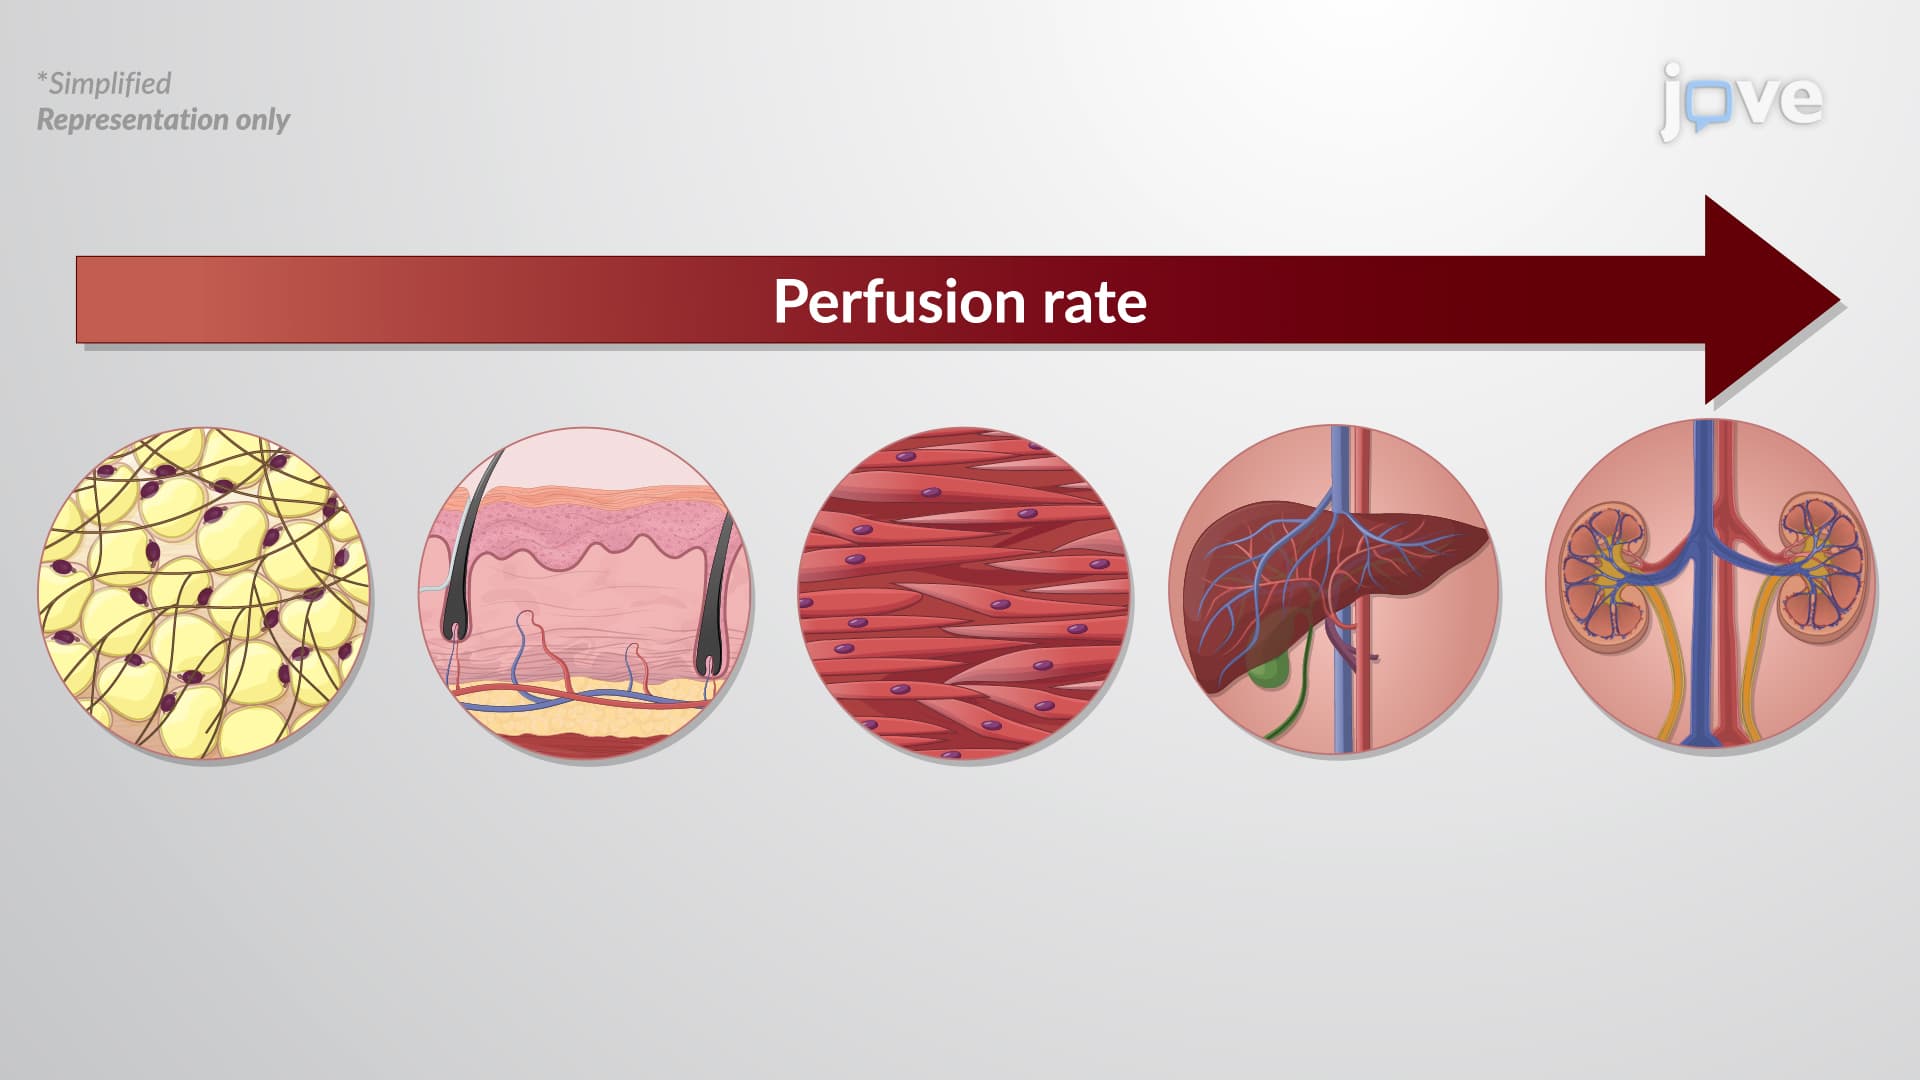
Physiological Pharmacokinetic Models: Blood Flow-Limited Versus Diffusion-Limited Models
 01:15
Model Approaches for Pharmacokinetic Data: Physiological Models
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:05
Two-Compartment Open Model: Overview
The...