Computer simulation of trifluoromethane properties with ab initio force field
Related Experiment Video
Updated: Jun 1, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Yi-Hsing Chung1, Arvin Huang-Te Li, Sheng D Chao
1Institute of Applied Mechanics, National Taiwan University, Taipei, Taiwan, Republic of China.

Abstract:
Intermolecular interaction potentials of the trifluoromethane dimer in 15 orientations have been calculated using the Hartree-Fock (HF) self-consistent theory and the second-order Møller-Plesset (MP2) perturbation theory. Single point energies at important geometries were also calibrated by the coupled cluster with single and double and perturbative triple excitation [CCSD(T)] calculations. We have employed Pople's medium size basis sets [up to 6-311++G(3df,3pd)] and Dunning's correlation consistent basis sets (up to aug-cc-pVQZ). Basis set limit potential values were obtained through well-studied extrapolation methods. The calculated MP2 potential data were employed to parameterize a 5-site force field for molecular simulations. We performed molecular dynamics simulations using the constructed ab initio force field and compared the simulation results with experiments. Quantitative agreements for the atom-wise radial distribution functions and the self-diffusion coefficients over a wide range of experimental conditions can be obtained, thus validating the ab initio force field without using experimental data a priori.
More Related Videos
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:34
Comparing Intermolecular Forces: Melting Point, Boiling Point, and Miscibility
Temporary attractive forces like dispersion are present in all molecules, whether they are polar or nonpolar. They...
 03:37
Molecular Shape and Polarity
 02:52
VSEPR Theory and the Basic Shapes
 02:18
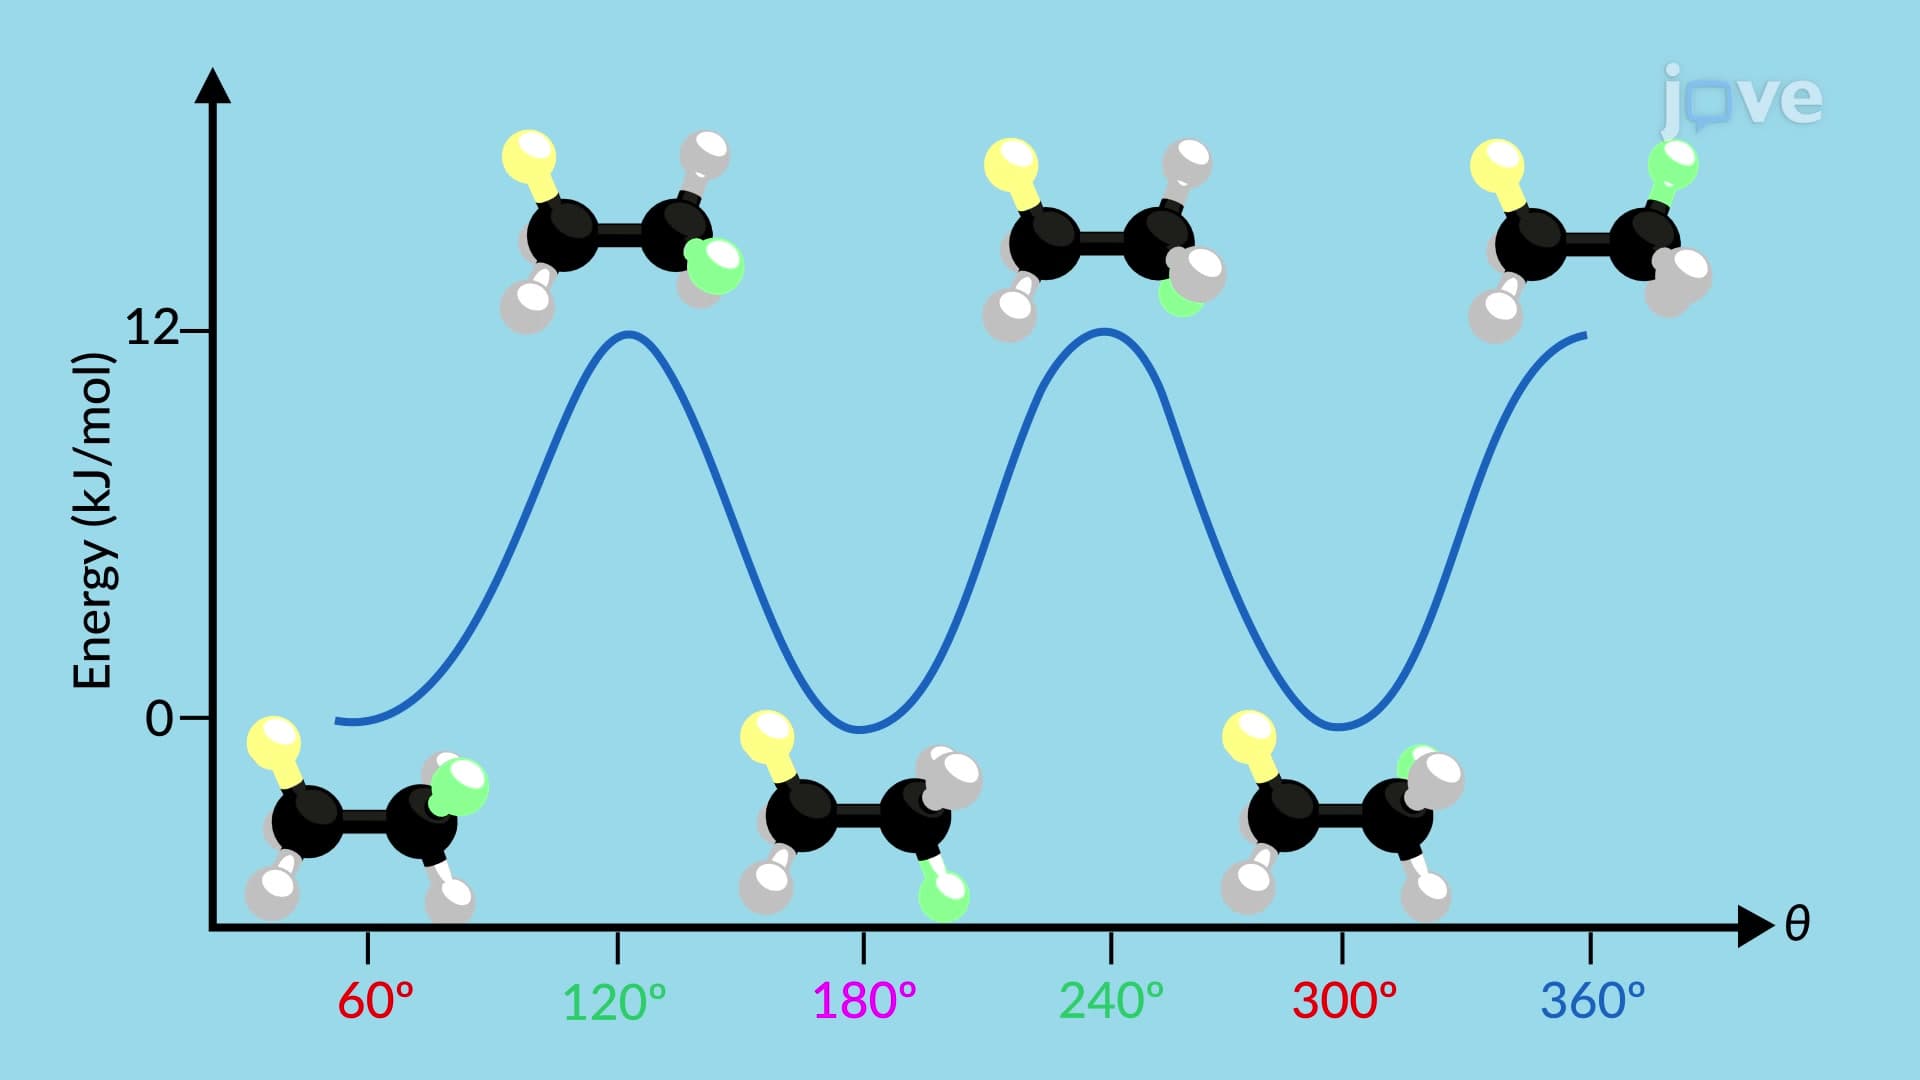
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...