Related Experiment Video
Updated: Jun 1, 2026

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
A comparison of different QSAR approaches to modeling CYP450 1A2 inhibition
Sergii Novotarskyi1, Iurii Sushko, Robert Körner
1eADMET GmbH, Ingolstädter Landstrasse 1, D-85764 Neuherberg, Germany.

Abstract:
Prediction of CYP450 inhibition activity of small molecules poses an important task due to high risk of drug-drug interactions. CYP1A2 is an important member of CYP450 superfamily and accounts for 15% of total CYP450 presence in human liver. This article compares 80 in-silico QSAR models that were created by following the same procedure with different combinations of descriptors and machine learning methods. The training and test sets consist of 3745 and 3741 inhibitors and noninhibitors from PubChem BioAssay database. A heterogeneous external test set of 160 inhibitors was collected from literature. The studied descriptor sets involve E-state, Dragon and ISIDA SMF descriptors. Machine learning methods involve Associative Neural Networks (ASNN), K Nearest Neighbors (kNN), Random Tree (RT), C4.5 Tree (J48), and Support Vector Machines (SVM). The influence of descriptor selection on model accuracy was studied. The benefits of "bagging" modeling approach were shown. Applicability domain approach was successfully applied in this study and ways of increasing model accuracy through use of applicability domain measures were demonstrated as well as fragment-based model interpretation was performed. The most accurate models in this study achieved values of 83% and 68% correctly classified instances on the internal and external test sets, respectively. The applicability domain approach allowed increasing the prediction accuracy to 90% for 78% of the internal and 17% of the external test sets, respectively. The most accurate models are available online at http://ochem.eu/models/Q5747 .
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 05:47
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
Related Concept Videos
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:24
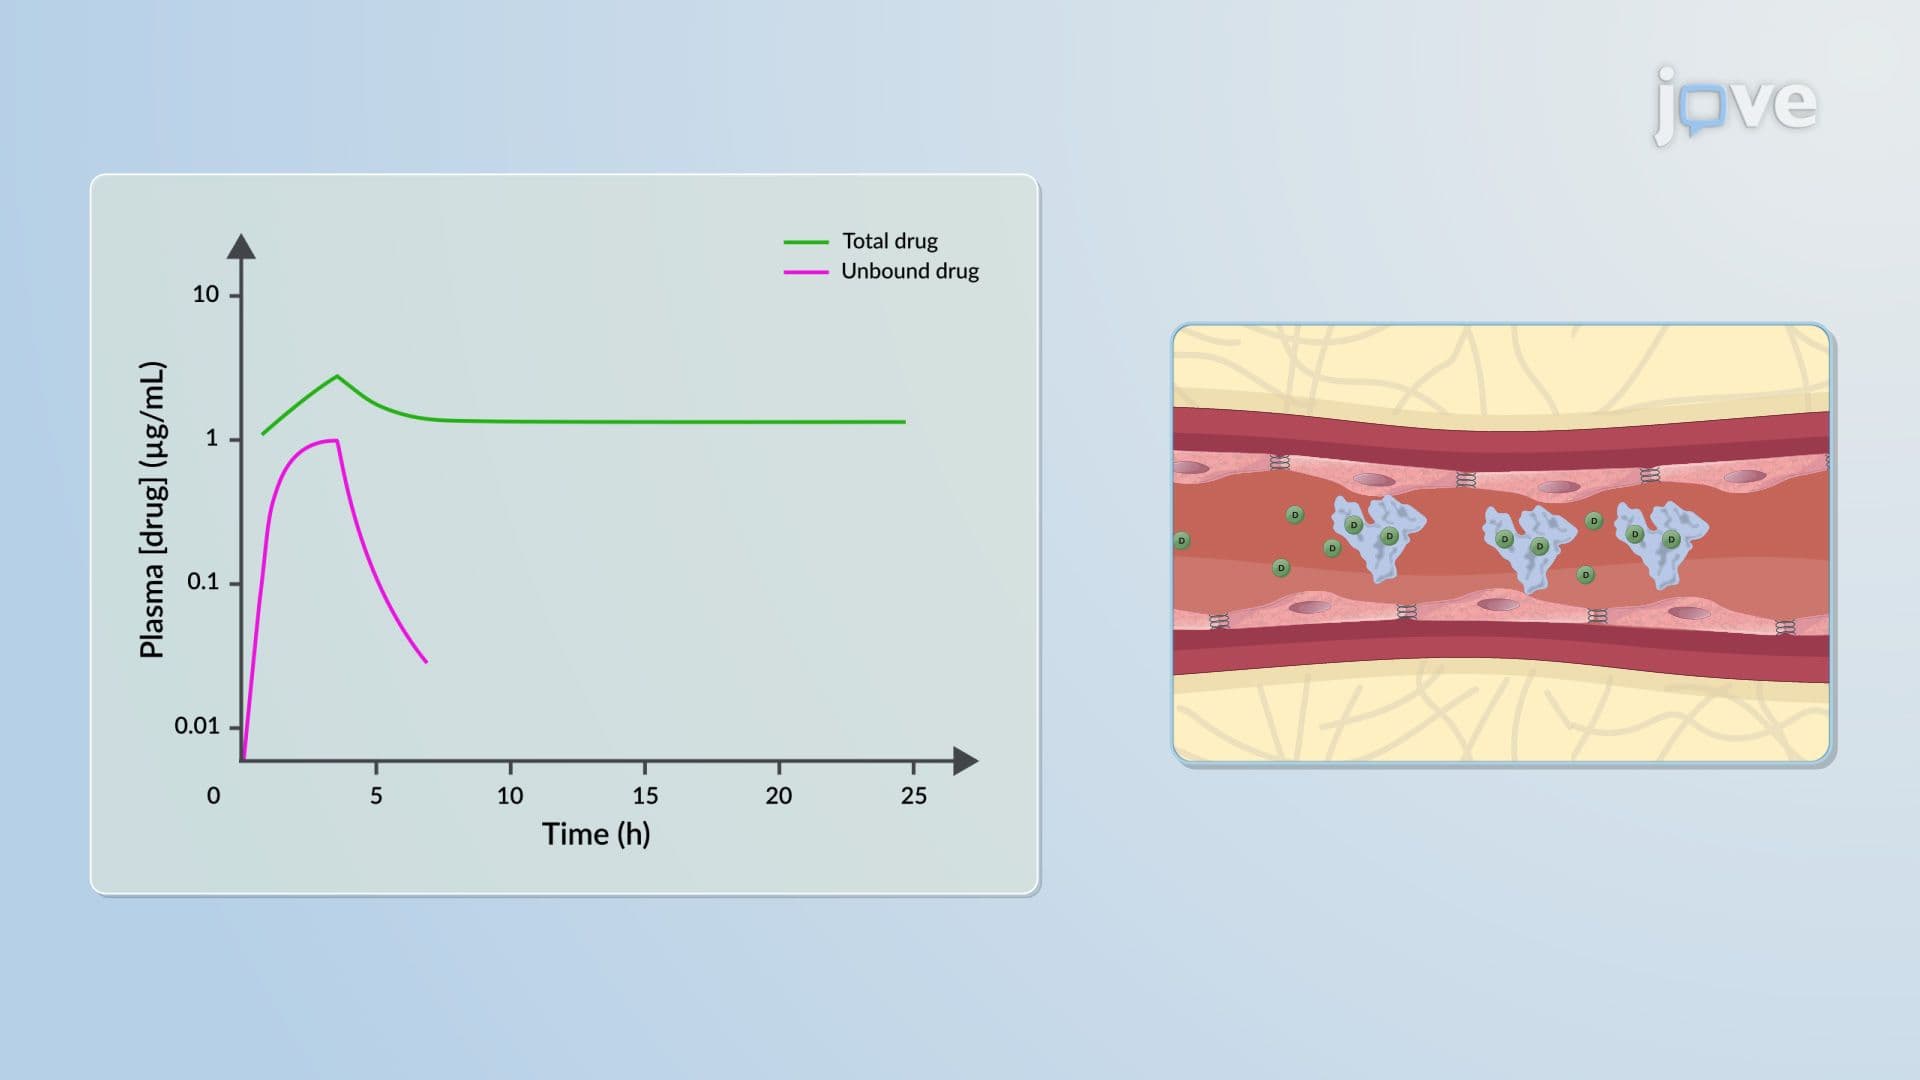
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model