Theoretical unimolecular kinetics for CH4 + M ⇄ CH3 + H + M in eight baths, M = He, Ne, Ar, Kr, H2, N2, CO, and CH4
Related Experiment Video
Updated: Jun 1, 2026
Conducting Miller-Urey Experiments
Published on: January 21, 2014
Ahren W Jasper1, James A Miller
1Combustion Research Facility, Sandia National Laboratories, P.O. Box 969, Livermore, California 94551-0969, United States. ajasper@sandia.gov

Abstract:
Ensembles of classical trajectories are used to study collisional energy transfer in highly vibrationally excited CH(4) for eight bath gases. Several simplifying assumptions for the CH(4) + M interaction potential energy surface are tested against full dimensional direct dynamics trajectory calculations for M = He, Ne, and H(2). The calculated energy transfer averages are confirmed to be sensitive to the shape of the repulsive wall of the intermolecular potential, with an exponential repulsive wall required for quantitative predictions. For the diatomic baths, the usual "separable pairwise" approximation for the interaction potential is unable to describe the orientation dependence of the interaction potential accurately, and the ambiguity in the resulting parametrizations contributes an additional uncertainty to the predicted energy transfer averages of 20-40%. On the other hand, the energy transfer averages are shown to be insensitive to the level of theory used to describe the intramolecular CH(4) potential, with a computationally efficient semiempirical tight binding potential for hydrocarbons performing equally well as an MP2 potential. The relative collisional energy transfer efficiencies of the eight bath gases are discussed and shown to be a function of temperature. The ensemble-averaged energy transferred in deactivating collisions <ΔE(d)> for each bath is used to parametrize a single-exponential-down model for collisional energy transfer in master equation calculations. The predicted decomposition rate coefficients for CH(4) agree well with available experimental rate coefficients for M = He, Ar, Kr, and CH(4). The effect of vibrational anharmonicity on the predicted rate coefficients is considered briefly.
More Related Videos
 07:24
Combustion Chemistry of Fuels: Quantitative Speciation Data Obtained from an Atmospheric High-temperature Flow Reactor with Coupled Molecular-beam Mass Spectrometer
Published on: February 19, 2018
 09:46
Protocol for Measuring the Thermal Properties of a Supercooled Synthetic Sand-water-gas-methane Hydrate Sample
Published on: March 21, 2016
Related Concept Videos
 02:14
SN2 Reaction: Kinetics
In a chemical reaction, a relationship exists between the concentration of reactants and the rate at which the reaction proceeds. The study to measure this relationship is known as the kinetics of a chemical reaction. Kinetic studies are used to deduce the rate law of a chemical reaction, which provides information about the species involved during the transition state of the rate-determining step. Thus, kinetic studies help to derive the mechanism of a reaction.
 01:29
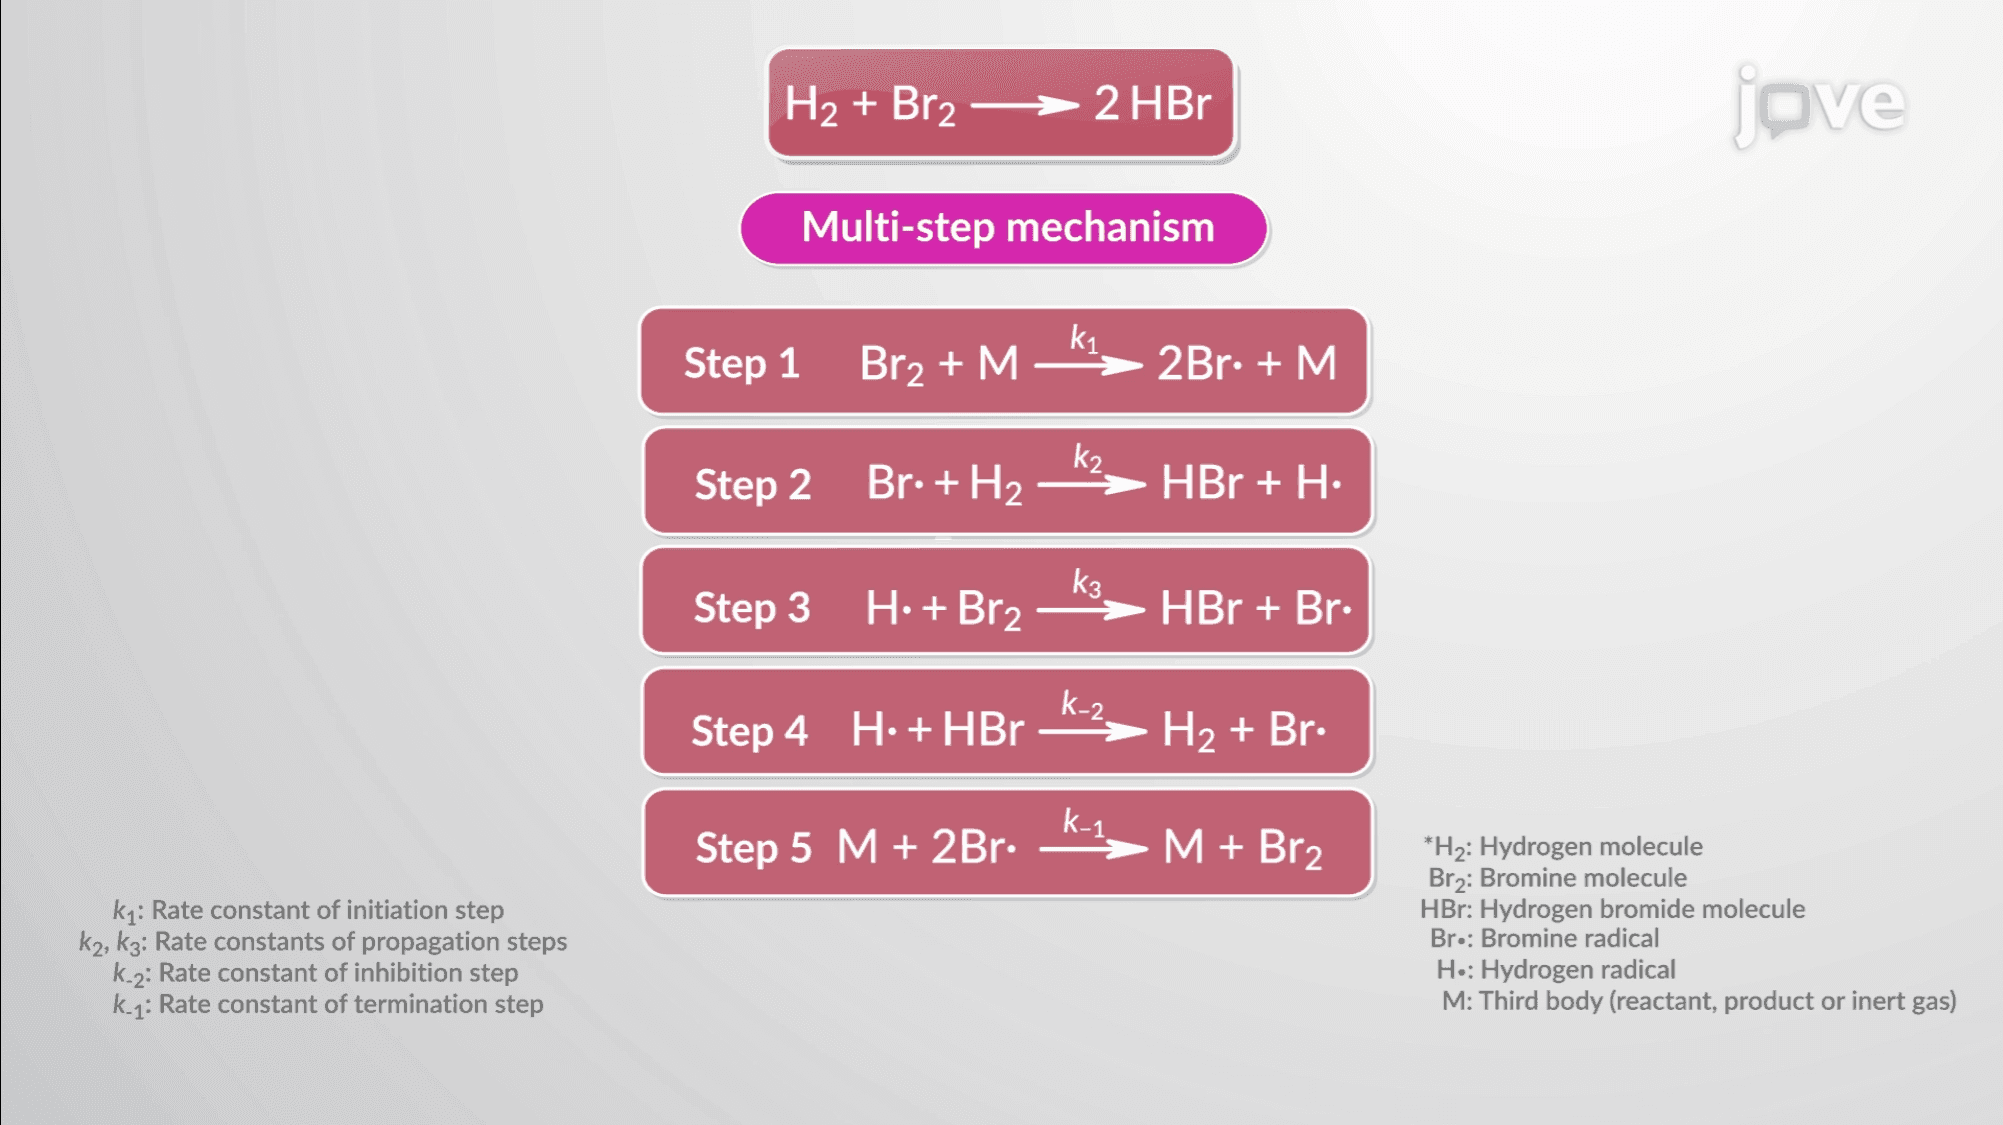
Chain Reactions
 01:32
Radical Anti-Markovnikov Addition to Alkenes: Thermodynamics
 01:34
Radical Halogenation: Thermodynamics
 03:40
Hess's Law
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration