Related Experiment Video
Updated: Jun 1, 2026

Manufacturing and Using Piggy-back Multibarrel Electrodes for In vivo Pharmacological Manipulations of Neural Responses
Published on: January 18, 2013
Inhibition of human Na(v)1.5 sodium channels by strychnine and its analogs
Chunhua Yuan1, Lirong Sun, Meng Zhang
1Department of Anatomy and Neurobiology, Southern Medical University, Guangzhou 510515, China.

Abstract:
Strychnine and brucine from the seeds of the plant Strychnos nux vomica have been shown to have interesting pharmacological effects on several neurotransmitter receptors. In this study, we have characterized the pharmacological properties of strychnine and its analogs on human Na(v)1.5 channels to assess their potential therapeutic advantage in certain arrhythmias. Among the eight alkaloids, only strychnine and icajine exhibited inhibition potency on the Na(v)1.5 channel with the half-maximum inhibition (IC(50)) values of 83.1μM and 104.6μM, respectively. Structure-function analysis indicated that the increased bulky methoxy groups on the phenyl ring or the negatively charged oxygen atom may account for this lack of inhibition on the Na(v)1.5 channel. Strychnine and icajine may bind to the channel by cation-π interactions. The substitution with a large side chain on the phenyl ring or the increased molecular volume may alter the optimized position for the compound close to the binding sites of the channel. Strychnine and icajine bind to the Na(v)1.5 channel with a new mechanism that is different from TTX and local anesthetics. They bind to the outer vestibule of the channel pore with fast association and dissociation rates at resting state. Strychnine and icajine had little effect on steady-state fast inactivation but markedly shifted the slow inactivation of Na(v)1.5 currents toward more hyperpolarized potentials. The property of icajine influencing slow-inactivated state of Na(v)1.5 channel would be potential therapeutic advantages in certain arrhythmias.
More Related Videos
 15:05
Functional Evaluation of Biological Neurotoxins in Networked Cultures of Stem Cell-derived Central Nervous System Neurons
Published on: February 5, 2015
 08:30
Preparation of Acute Spinal Cord Slices for Whole-cell Patch-clamp Recording in Substantia Gelatinosa Neurons
Published on: January 18, 2019
Related Concept Videos
 01:21
Drugs Affecting Neurotransmitter Release or Uptake
 01:28
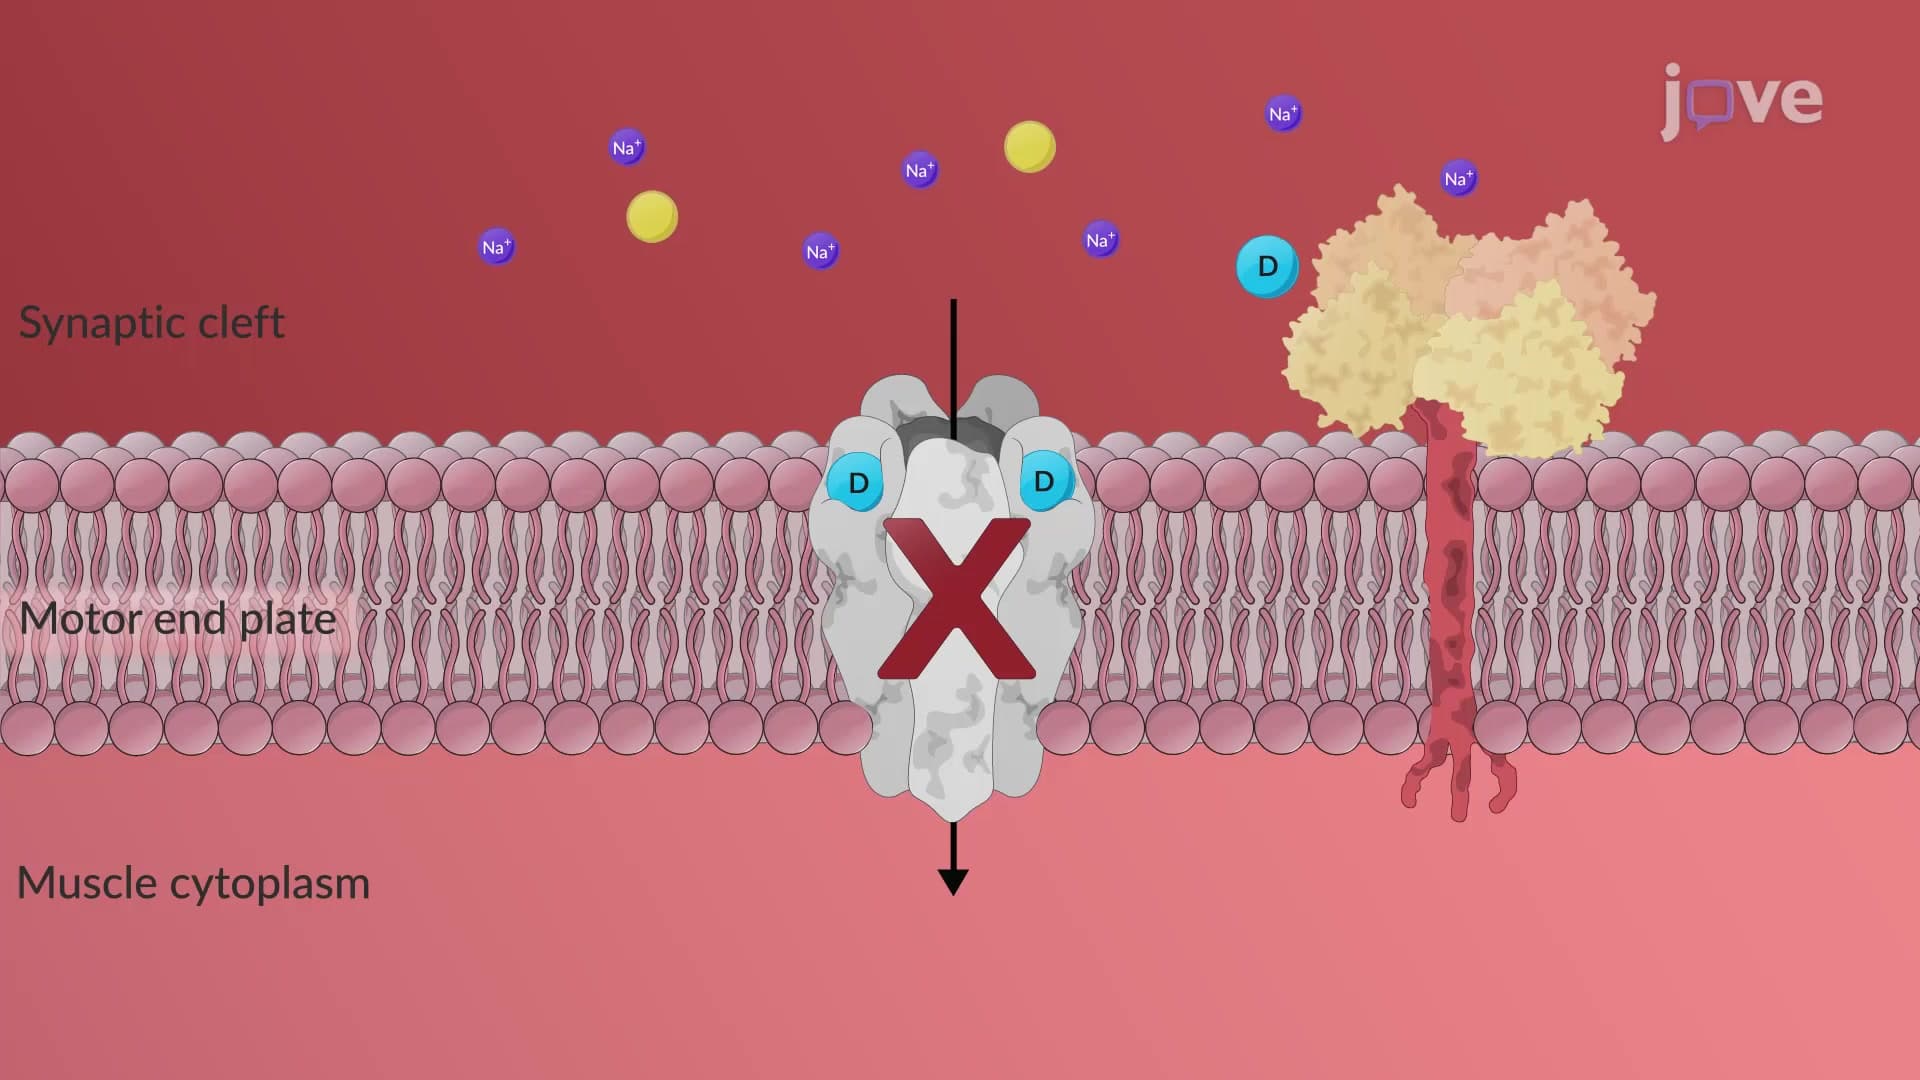
Depolarizing Blockers: Mechanism of Action
Succinylcholine is the most commonly used depolarizing blocker. Chemically, it constitutes two molecules of acetylcholine joined together by an acetate methyl group. They act on the receptors in the same way as acetylcholine. Because succinylcholine...
 01:17
Nondepolarizing (Competitive) Neuromuscular Blockers: Mechanism of Action
Competitive antagonists prevent acetylcholine from binding to its receptor, inhibiting membrane depolarization. Without conformational changes or intrinsic...
 01:22
Antiarrhythmic Drugs: Class I Agents as Sodium Channel Blockers
Class 1A Antiarrhythmic Drugs: These drugs work by moderately blocking sodium channels,...
 01:19
Depolarizing Blockers: Pharmocokinetics
 01:27
Nondepolarizing (Competitive) Neuromuscular Blockers: Pharmacological Actions
Although all competitive neuromuscular blockers are designed...