Related Experiment Video
Updated: Jun 1, 2026
Fluorescence-Activated Cell Sorting-Radioligand Treated Tissue (FACS-RTT) to Determine the Cellular Origin of Radioactive Signal
Published on: September 10, 2021
Pharmacokinetics, distribution, and excretion of 125I-labeled human plasma-derived-FVIIa and -FX with MC710 (FVIIa/FX
Yasushi Nakatomi1, Manami Tsuji, Teruhisa Nakashima
1Therapeutic Protein Product Research Department, The Chemo-Sero-Therapeutic Research Institute, KAKETSUKEN, Kumamoto, Japan.

Introduction:
MC710 is a mixture agent consisting of plasma-derived activated factor VII (FVIIa) and factor X (FX) at a weight ratio of 1:10 developed as a novel bypassing agent for the management of the bleeding of hemophilia patients with inhibitors. The pharmacokinetics, distribution, and excretion of (125)I-labeled-FVIIa ((125)I-FVIIa) and -FX ((125)I-FX) were studied in male rats after a single intravenous administration of (125)I-FVIIa or (125)I-FX combined with MC710.
Methods:
(125)I-FVIIa or (125)I-FX was administered intravenously with MC710 to male rats in a single dosage (FVIIa 0.4 mg and FX 4 mg/kg body weight) and radioactivity and antigen levels in plasma were quantified for 24h. Urine and feces were sampled to study the excretion of radioactivity during 168 h after dosing. Whole-body autoradiography was performed to evaluate the qualitative distribution of radioactivity 168 h after dosing.
Results And Conclusions:
The half-life (t(1/2)α and t(1/2)β) of radioactivity and FVIIa antigen were 0.704 and 6.27 h, and 0.496 and 1.66 h, respectively and the area under the plasma concentration-time curve (AUC(0-∞)) of radioactivity and FVIIa antigen were 17,932 and 8671 ng·h/mL, respectively. The t(1/2) of radioactivity and FX antigen were 4.06 and 3.05 h, respectively, and the AUC(0-∞) of radioactivity and FX antigen were 320,143 and 395,794 ng·h/mL, respectively. About 80% of the administered dose of radioactivity was excreted in urine and feces by 168 h after administration. Tissue distribution experiments showed that FVIIa- and FX-related (125)I accumulated in bone and bone marrow, and disappeared slowly.
More Related Videos
 11:58
Initial Evaluation of Antibody-conjugates Modified with Viral-derived Peptides for Increasing Cellular Accumulation and Improving Tumor Targeting
Published on: March 8, 2018
 11:03
Automated Radiochemical Synthesis of [18F]3F4AP: A Novel PET Tracer for Imaging Demyelinating Diseases
Published on: May 29, 2017
Related Concept Videos
 01:18
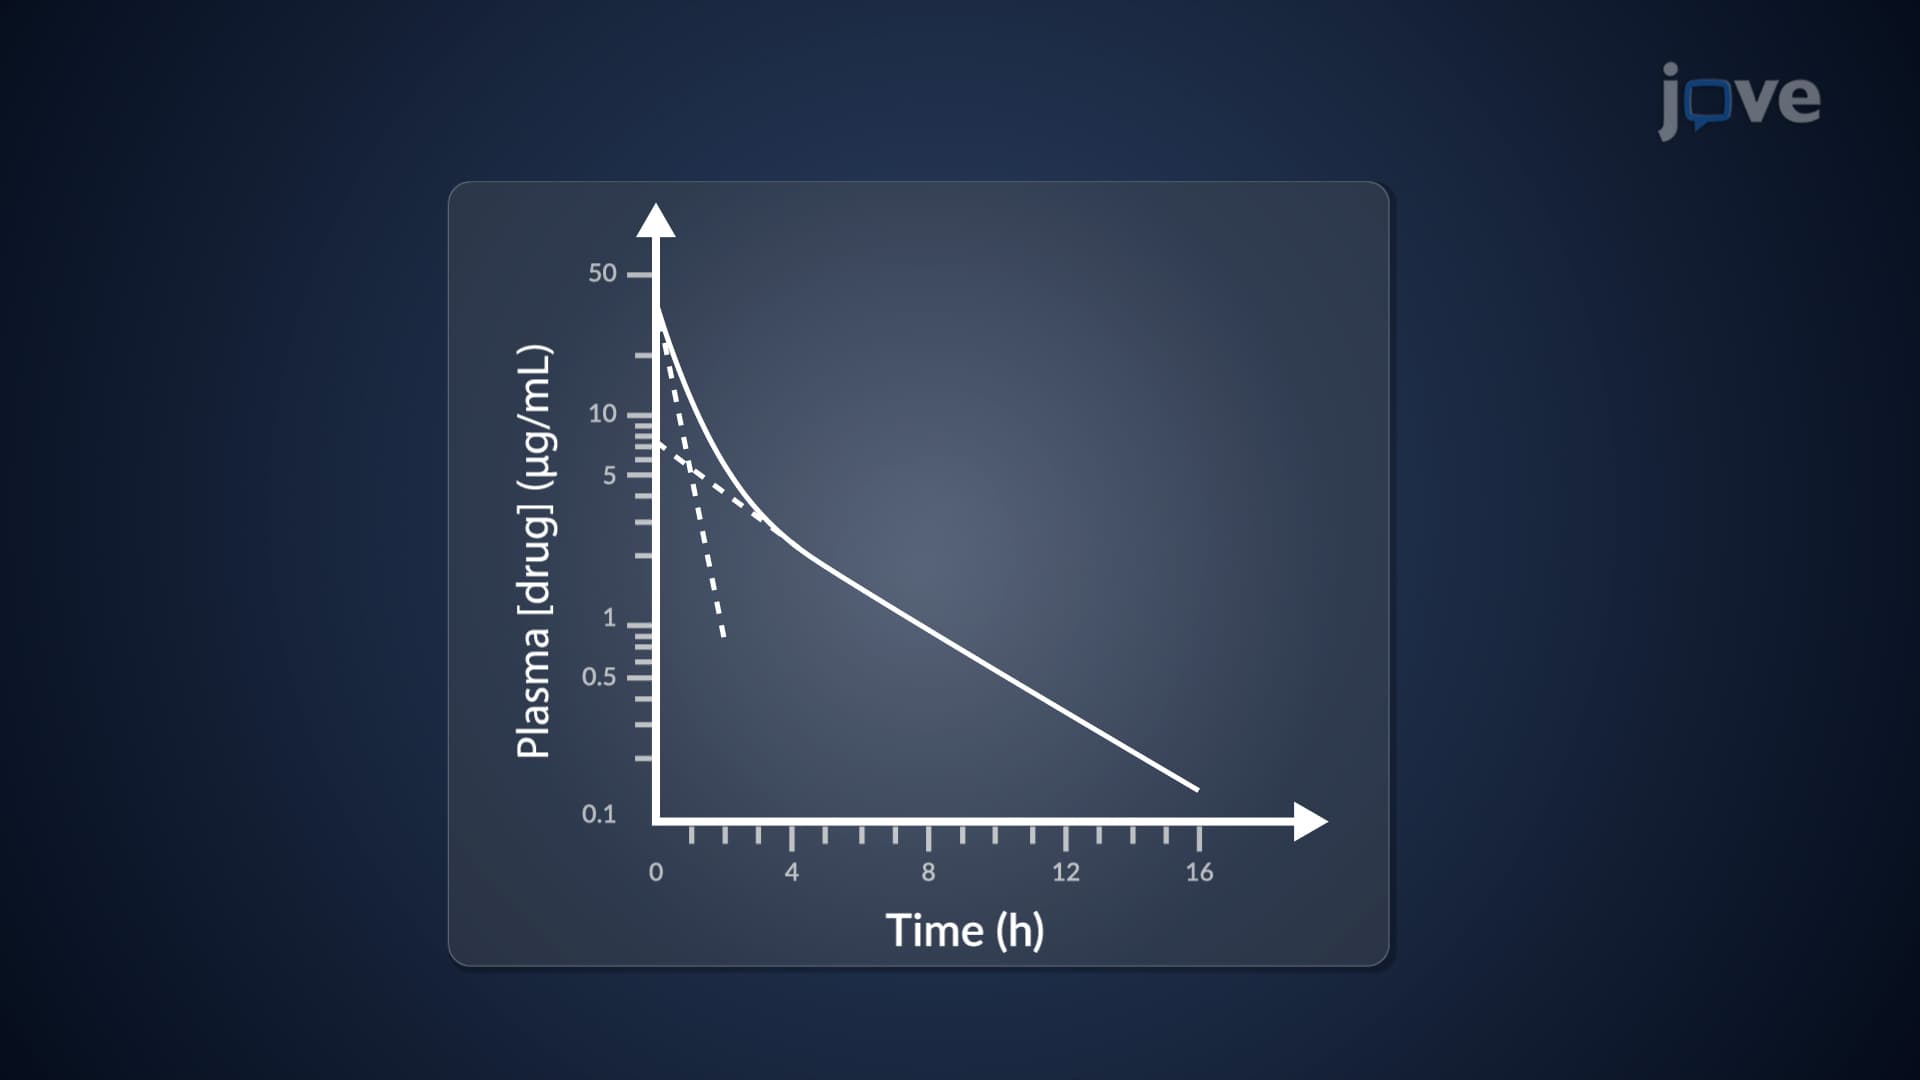
Two-Compartment Open Model: IV Bolus Administration
The disparity between drug input and the sum of drug transfer rates between...
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
 01:20
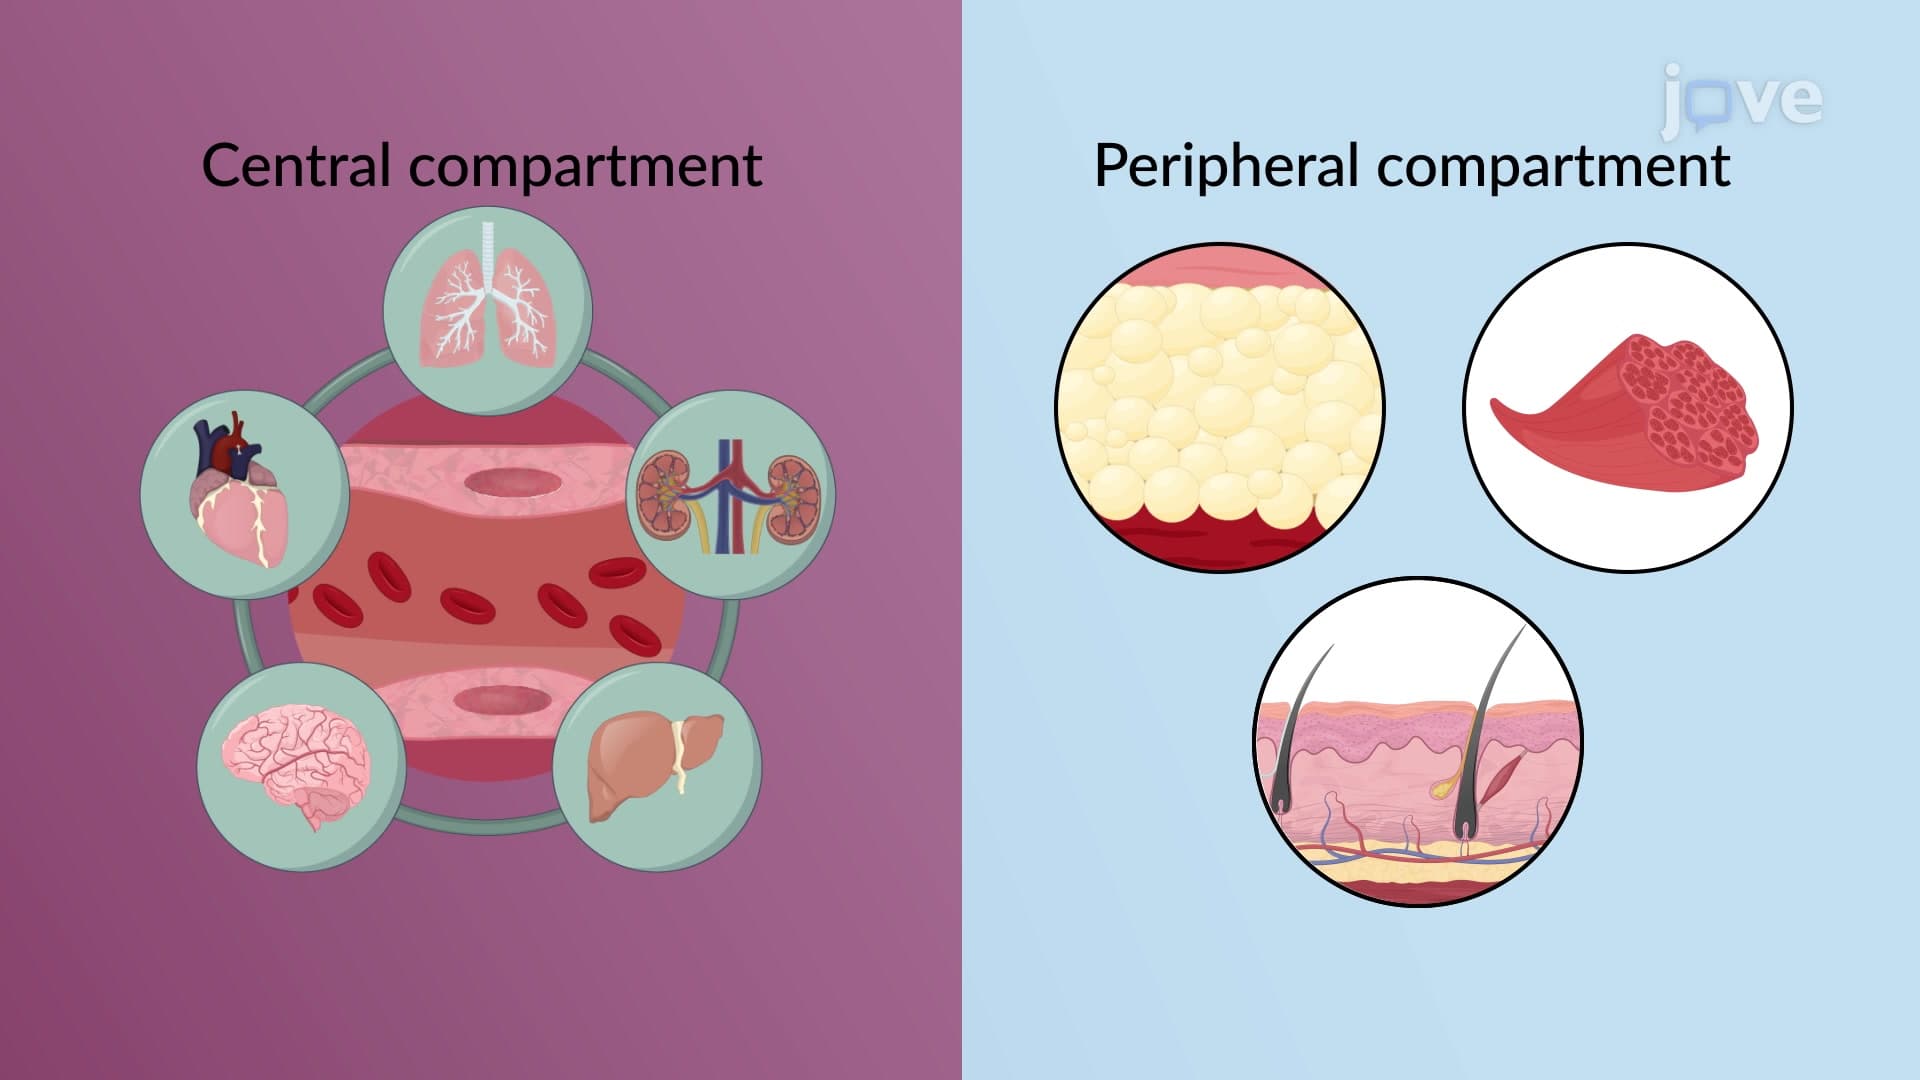
Compartment Models: Two-Compartment Model
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Duration of Dose-Effect Relationship
 01:09
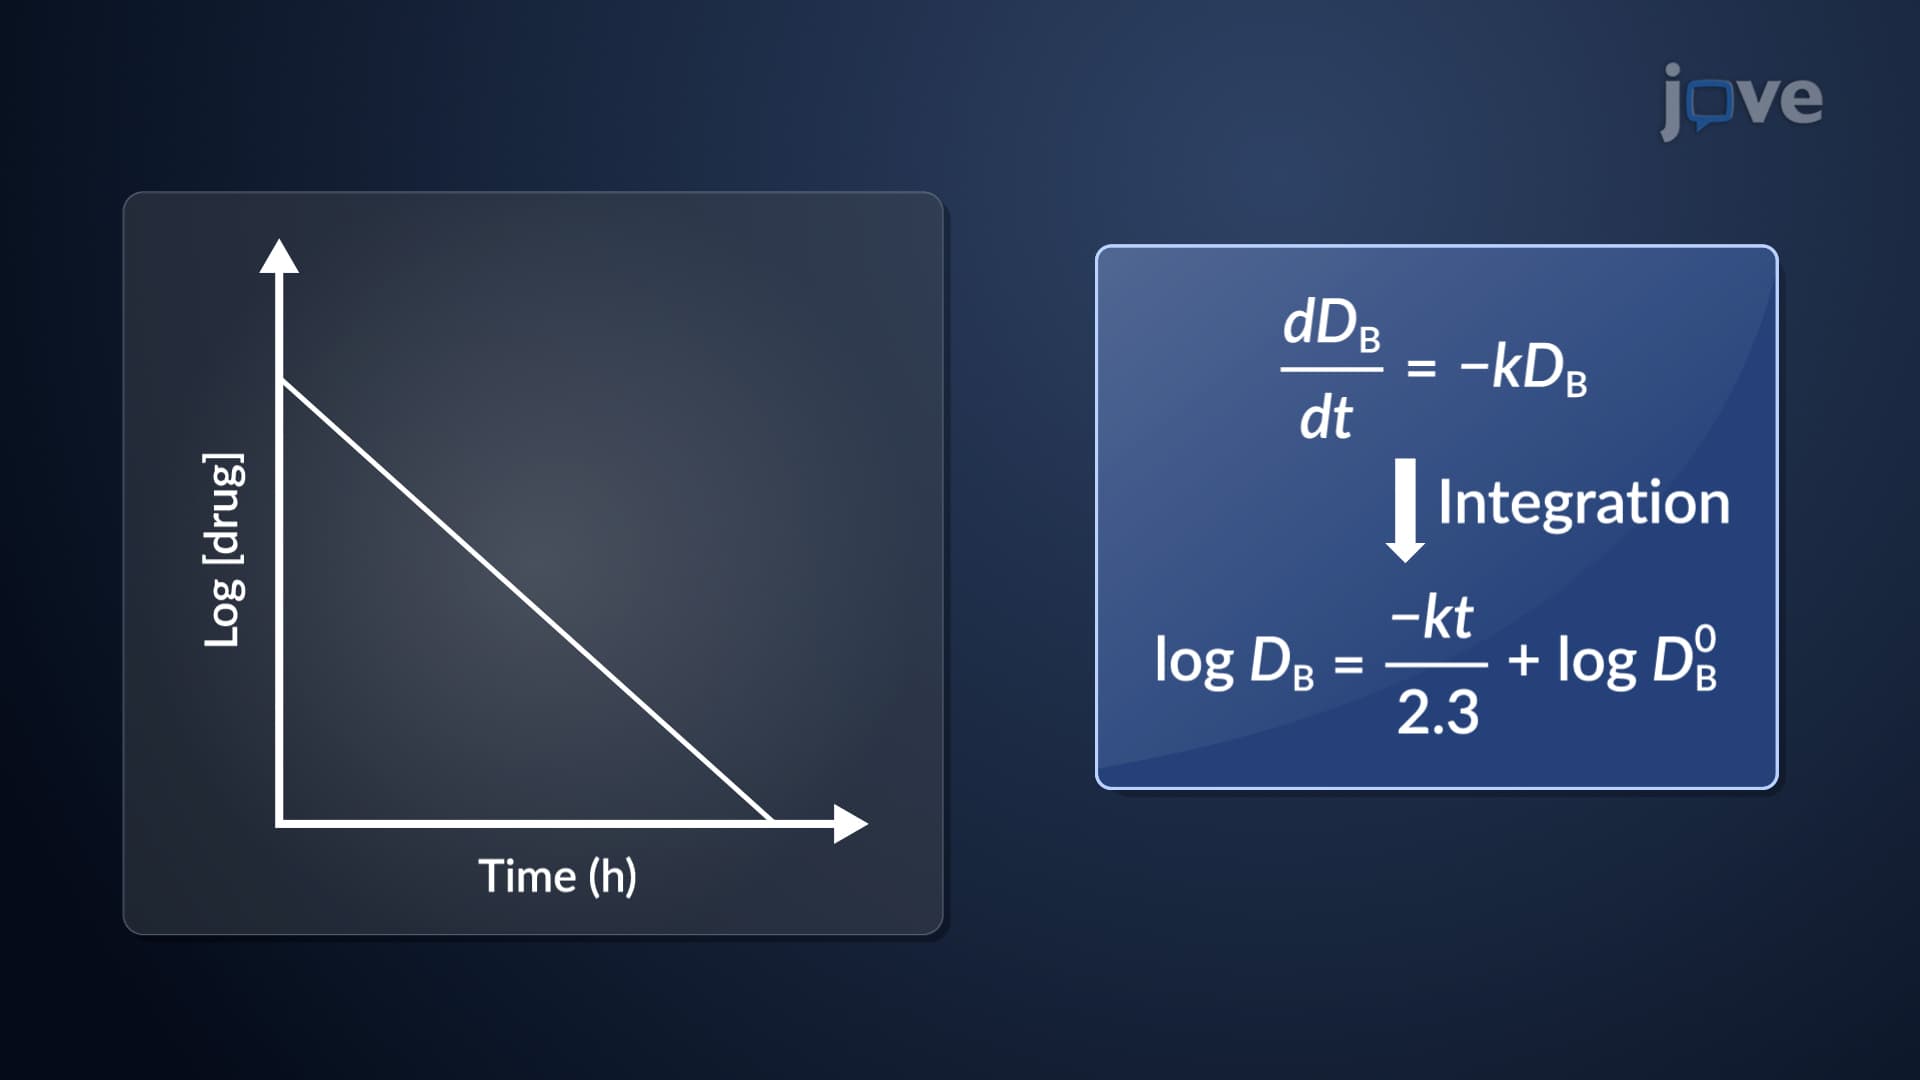
One-Compartment Open Model for IV Bolus Administration: Estimation of Elimination Rate Constant, Half-Life and Volume of Distribution
 00:54
Noncompartmental Analysis: Miscellaneous Pharmacokinetic Parameters
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's overall...