Related Experiment Video
Updated: May 31, 2026

Evaluating Cell Death Signaling by Immunofluorescence in a Rat Model of Ischemic Stroke
Published on: January 3, 2025
CaMKII in cerebral ischemia
Steven J Coultrap1, Rebekah S Vest, Nicole M Ashpole
1Department of Pharmacology, University of Colorado Denver-School of Medicine, Aurora, CO 80045, USA.

Abstract:
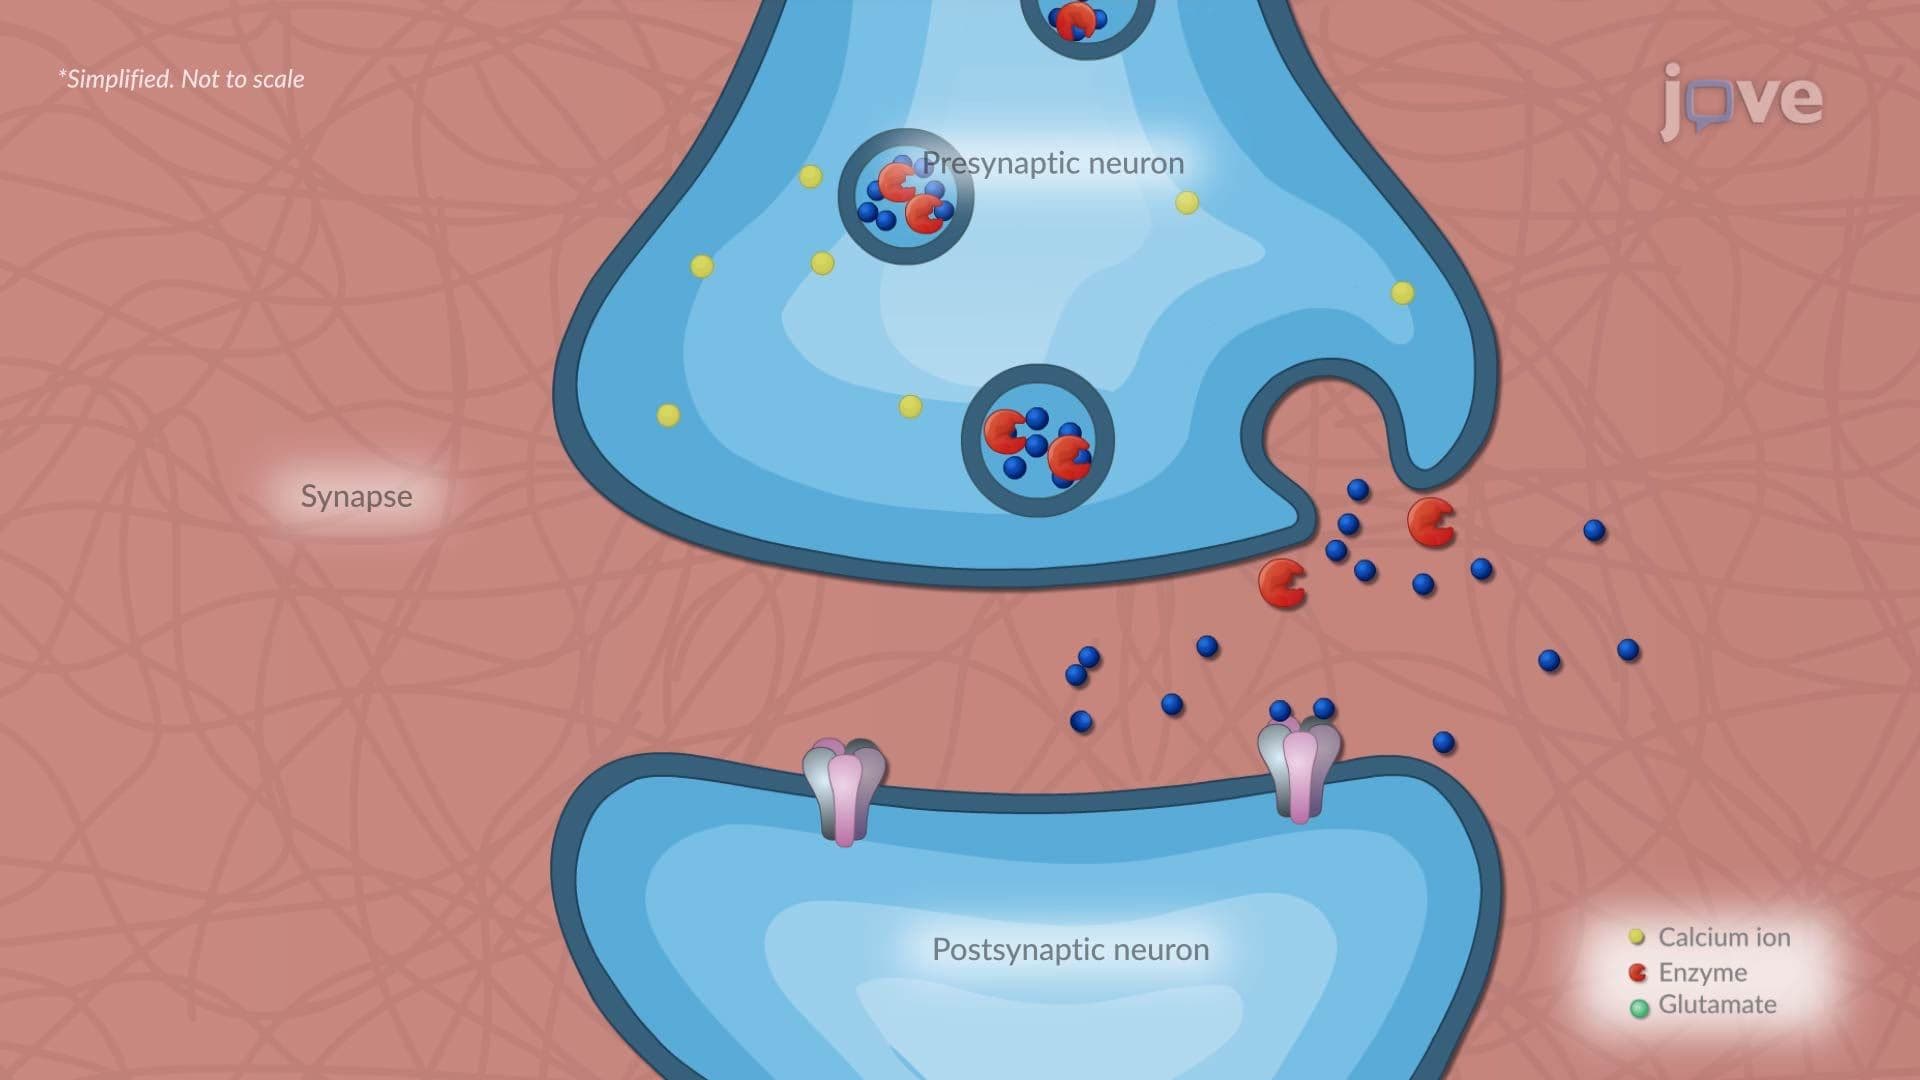
Ischemic insults on neurons trigger excessive, pathological glutamate release that causes Ca²⁺ overload resulting in neuronal cell death (excitotoxicity). The Ca²⁺/calmodulin (CaM)-dependent protein kinase II (CaMKII) is a major mediator of physiological excitatory glutamate signals underlying neuronal plasticity and learning. Glutamate stimuli trigger autophosphorylation of CaMKII at T286, a process that makes the kinase "autonomous" (partially active independent from Ca²⁺ stimulation) and that is required for forms of synaptic plasticity. Recent studies suggested autonomous CaMKII activity also as potential drug target for post-insult neuroprotection, both after glutamate insults in neuronal cultures and after focal cerebral ischemia in vivo. However, CaMKII and other members of the CaM kinase family have been implicated in regulation of both neuronal death and survival. Here, we discuss past findings and possible mechanisms of CaM kinase functions in excitotoxicity and cerebral ischemia, with a focus on CaMKII and its regulation.
Related Concept Videos
 01:15
Ischemic Stroke ll: Pathophysiology
 01:15
Ischemic Stroke l: Introduction