Related Experiment Video
Updated: May 31, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
[An extension strategy of Discovery Studio 2.0 for non-bonded interaction energy automatic calculation at the residue
1State Key Laboratory of Genetic Resources and Evolution, Kunming Institute of Zoology, the Chinese Academy of Sciences, Kunming 650223, China.

Abstract:
Non-bonded interaction forces play crucial roles in molecular recognition and binding in biological systems. However, it is difficult for traditional methods to automatically calculate and batch the non-bonded energy at the residue level. In recent years, many studies have focused on non-bonded interactions and developed tools to calculate and analyze such interactions. In this study, we present a highly automated approach for the calculation of non-bonded energy. Our strategy invoked protocols relevant to non-bonded interactions within Discovery Studio 2.0 (DS2.0, Accelrys Inc.) bottom module using Perl script, and determined the direct command line operation of calculating non-bonded interaction energy batches without accessing the graphical interface of DS. This approach extended the DS2.0 module and was applied to a recent study of complex structure analysis.


Related Concept Videos
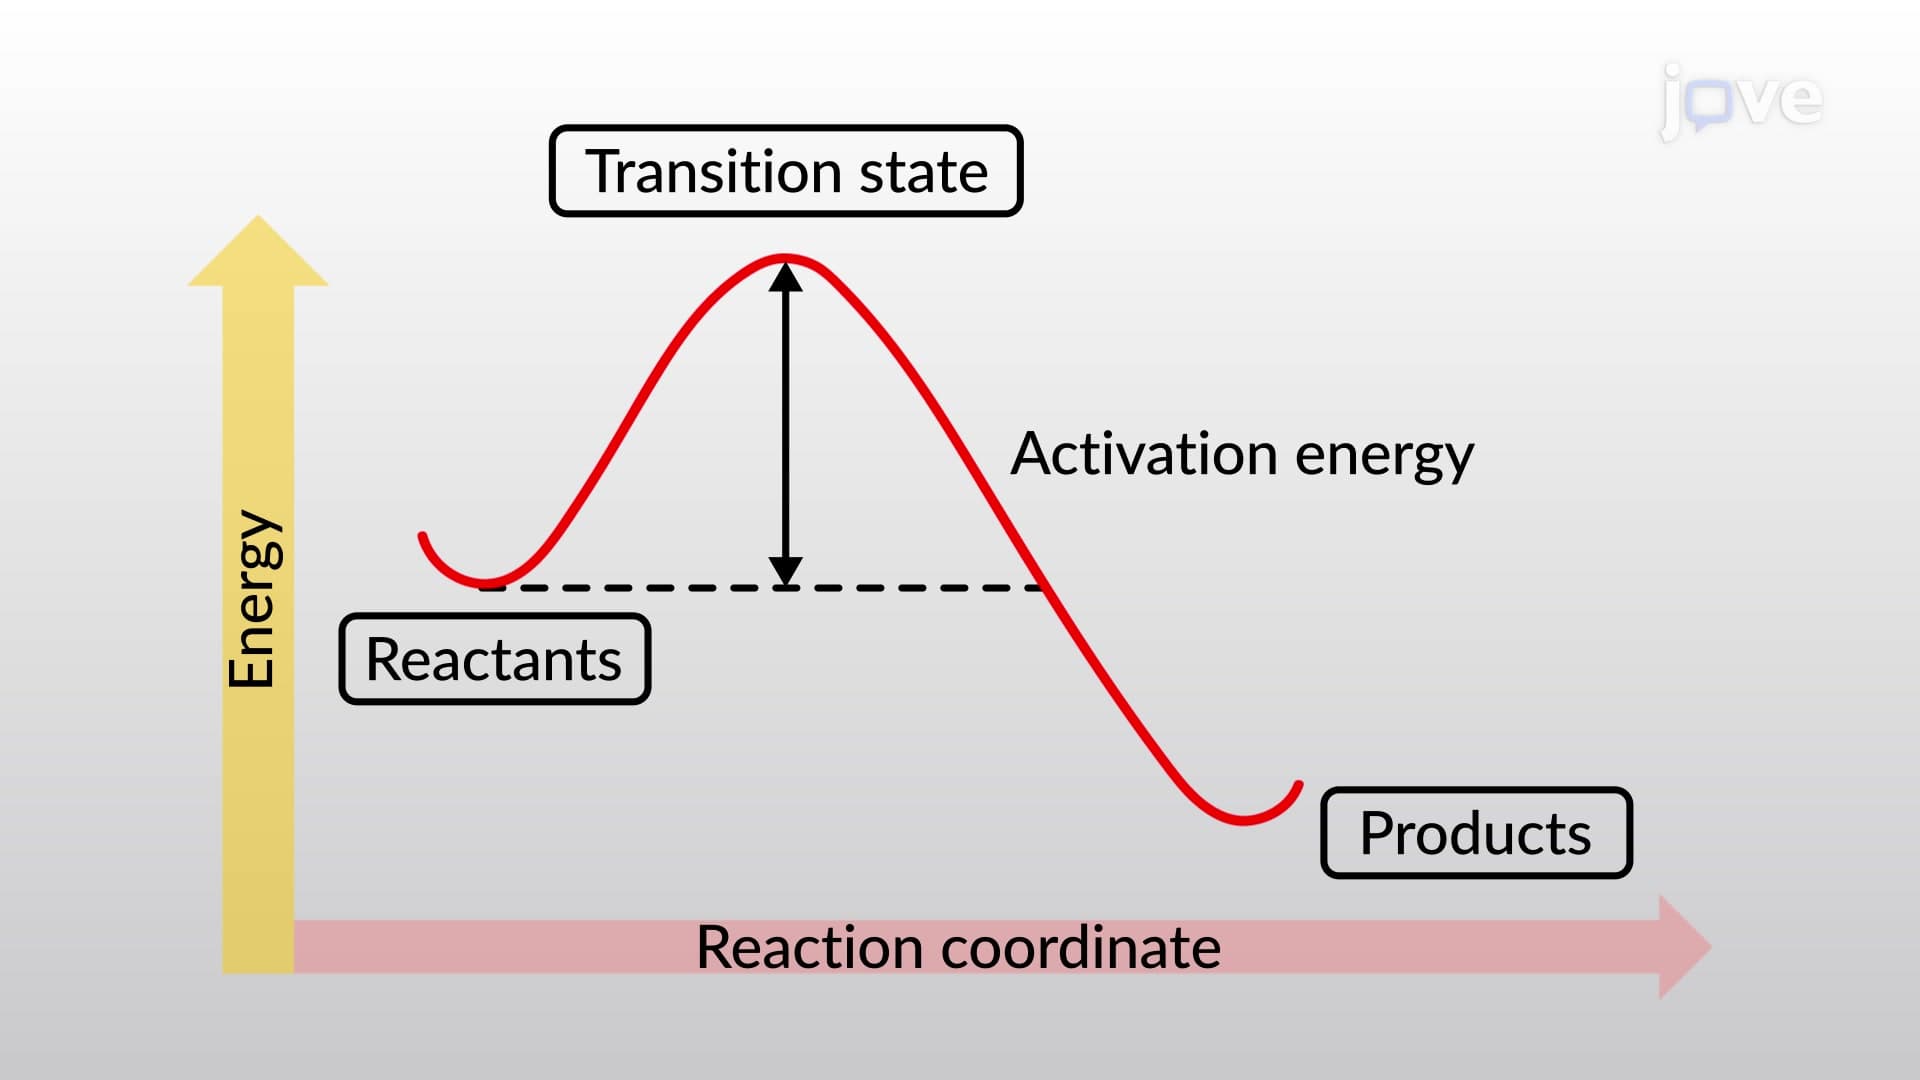 02:13
Bond Dissociation Energy and Activation Energy
 01:29
Determination of Crystal Structures
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 03:35
Hybridization of Atomic Orbitals II