Related Experiment Video
Updated: May 31, 2026

Accelerated Type 1 Diabetes Induction in Mice by Adoptive Transfer of Diabetogenic CD4+ T Cells
Published on: May 6, 2013
The microRNA-21-PDCD4 axis prevents type 1 diabetes by blocking pancreatic beta cell death
Qingguo Ruan1, Ting Wang, Vasumathi Kameswaran
1Department of Pathology and Laboratory of Medicine, University of Pennsylvania School of Medicine, Philadelphia, PA 19104, USA.

Abstract:
Death of pancreatic β cells is a pathological hallmark of type 1 diabetes (T1D). However, the molecular mechanisms of β cell death and its regulation are poorly understood. Here we describe a unique regulatory pathway of β cell death that comprises microRNA-21, its target tumor suppressor PDCD4, and its upstream transcriptional activator nuclear factor-κB (NF-κB). In pancreatic β cells, c-Rel and p65 of the NF-κB family activated the mir21 gene promoter and increased miR-21 RNA levels; miR-21 in turn decreased the level of PDCD4, which is able to induce cell death through the Bax family of apoptotic proteins. Consequently, PDCD4 deficiency in pancreatic β cells renders them resistant to death, and PDCD4 deficiency in NOD or C57BL/6 mice conferred resistance to spontaneous diabetes and diabetes induced by autoimmune T cells or the β cell toxin streptozotocin (STZ). Thus, the NF-κB-microRNA-21-PDCD4 axis plays a crucial role in T1D and represents a unique therapeutic target for treating the disease.
Insights
A novel pathway involving nuclear factor-κB (NF-κB), microRNA-21, and PDCD4 regulates pancreatic β cell death in type 1 diabetes (T1D). Targeting this axis offers a new therapeutic strategy for T1D.
Area of Science:
- Immunology
- Molecular Biology
- Endocrinology
Background:
- Pancreatic β cell death is central to type 1 diabetes (T1D) pathogenesis.
- The precise molecular regulators of β cell apoptosis remain incompletely understood.
- Identifying novel pathways is crucial for developing effective T1D therapies.
Purpose of the Study:
- To elucidate a unique molecular pathway regulating pancreatic β cell death.
- To investigate the roles of microRNA-21, PDCD4, and nuclear factor-κB (NF-κB) in β cell apoptosis.
- To assess the therapeutic potential of targeting the NF-κB-microRNA-21-PDCD4 axis in T1D.
Main Methods:
- Analysis of the NF-κB activation of the microRNA-21 gene promoter in pancreatic β cells.
- Quantification of miR-21 and PDCD4 RNA levels.
- Assessment of PDCD4's role in apoptosis induction via Bax proteins.
- Evaluation of PDCD4 deficiency effects on diabetes development in mouse models (NOD, C57BL/6) under various diabetogenic conditions (autoimmune, streptozotocin-induced).
Main Results:
- NF-κB (c-Rel and p65) activates the mir21 gene promoter, increasing miR-21 levels in pancreatic β cells.
- miR-21 downregulates the tumor suppressor PDCD4, which promotes cell death through Bax proteins.
- PDCD4 deficiency confers resistance to β cell death and protects against spontaneous and induced diabetes in mice.
- The NF-κB-microRNA-21-PDCD4 signaling pathway is critical for T1D pathogenesis.
Conclusions:
- A novel regulatory axis (NF-κB-microRNA-21-PDCD4) governs pancreatic β cell death.
- This pathway is integral to the development of type 1 diabetes.
- Targeting this axis presents a promising therapeutic avenue for T1D treatment.
Related Concept Videos
 01:26
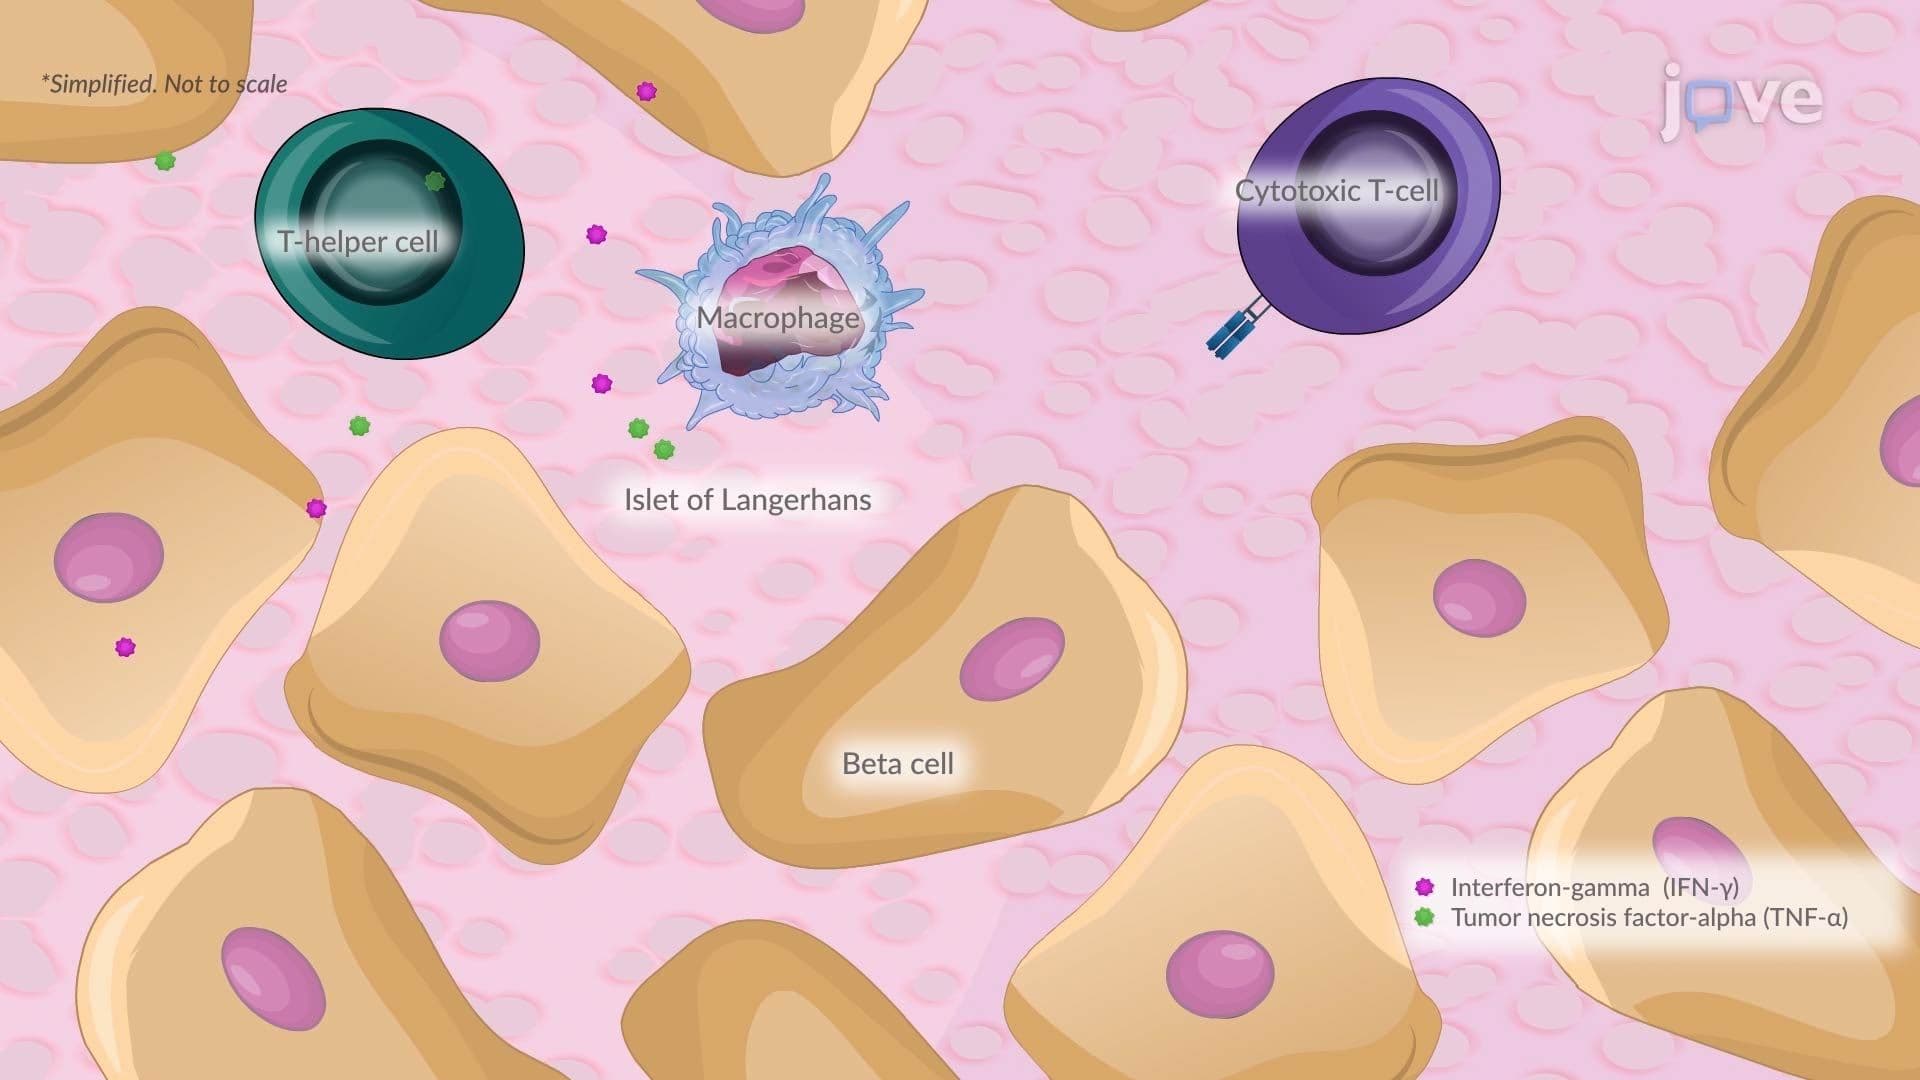
Type I Diabetes II: Pathophysiology
 01:12
Type I Diabetes I: Introduction
 01:24
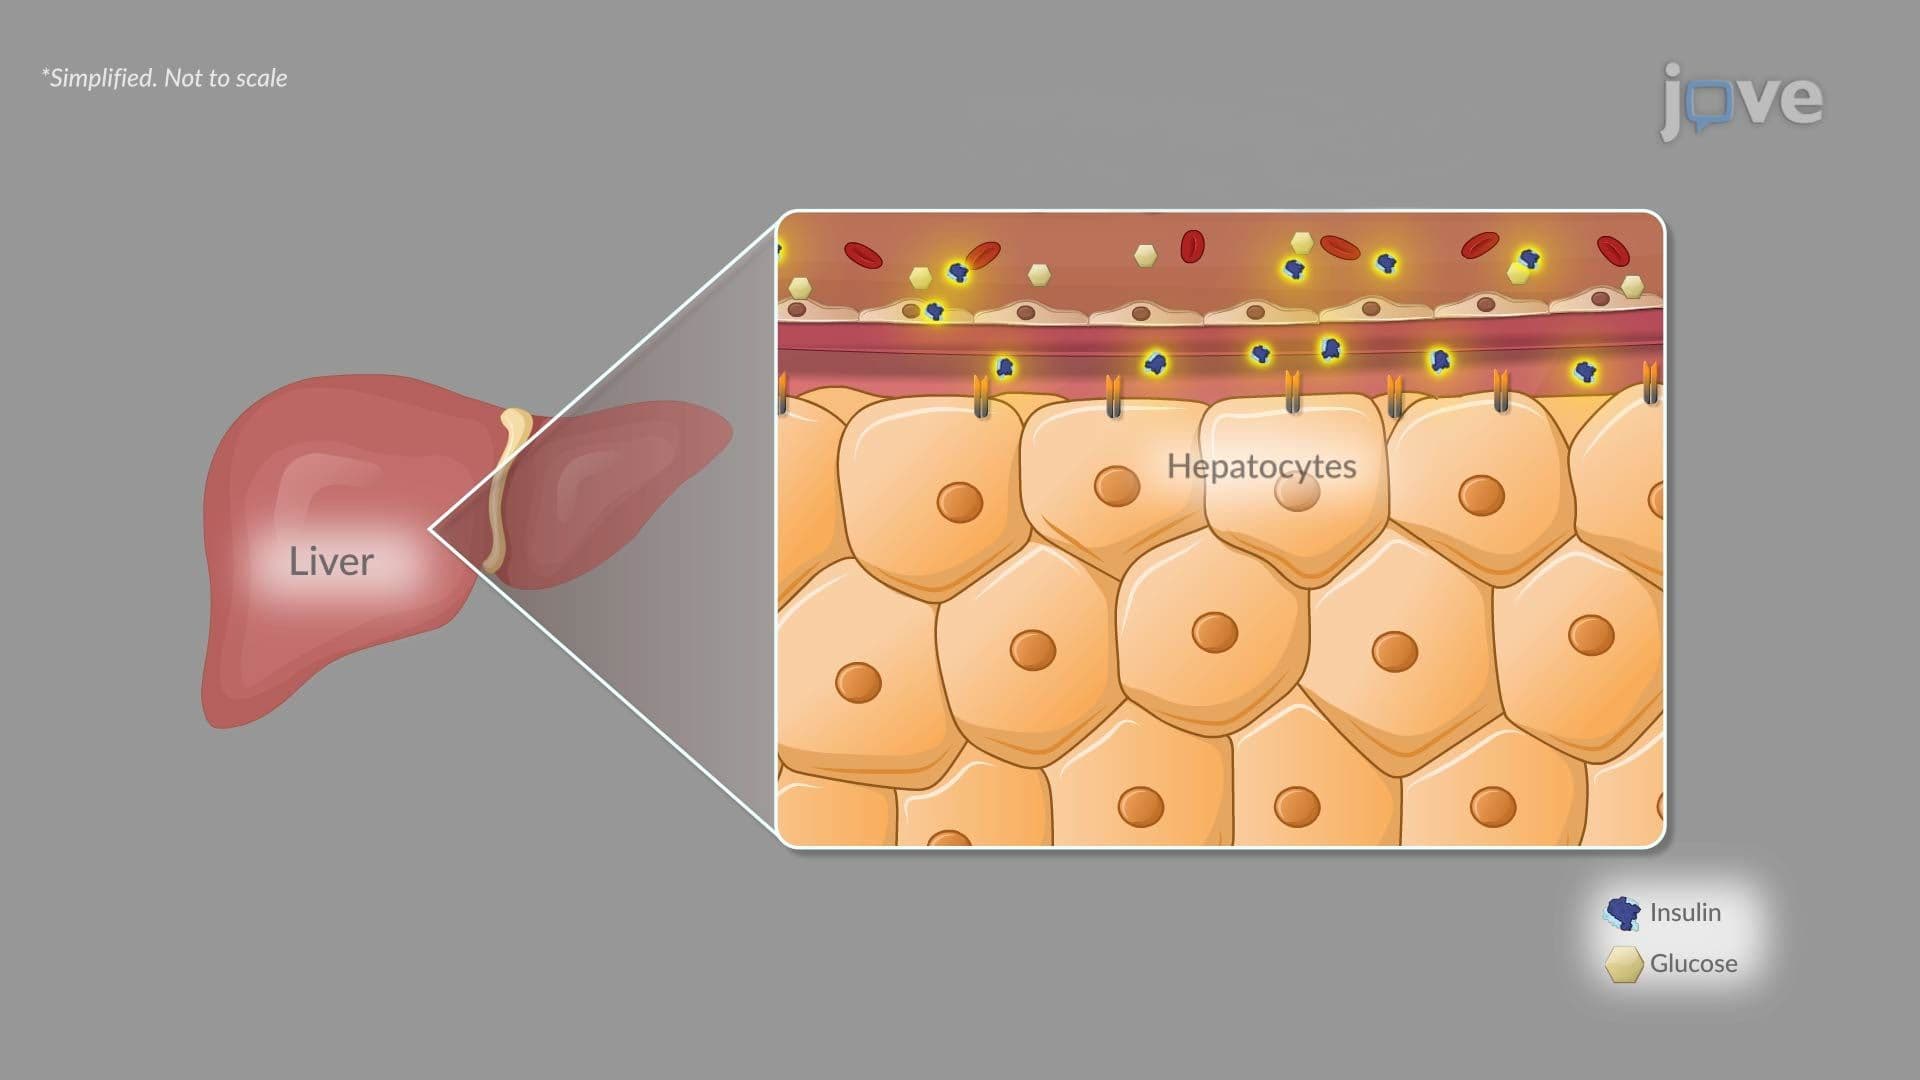
Type II Diabetes II: Pathophysiology
 01:26
Type II Diabetes I: Introduction
 01:22
Diabetes Mellitus: Overview and Type I Subtype
Type 1 diabetes is an autoimmune disease in which the immune system mistakenly attacks and destroys the insulin-producing beta cells in the pancreas. As a result, the body is unable to produce sufficient insulin, and individuals with...
 01:20
Pathophysiology of Diabetes
Type 1 diabetes is characterized by autoimmune-mediated destruction of pancreatic β cells, with environmental factors potentially triggering this process in genetically susceptible individuals. Despite many not having a family history, certain genes increase susceptibility, suggesting a...