Related Experiment Video
Updated: May 31, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
'Lassoing' a phylogenetic tree I: basic properties, shellings, and covers
Andreas W M Dress1, Katharina T Huber, Mike Steel
1CAS-MPG Partner Institute and Key Lab for Computational Biology, Shanghai, China. andreas.dress@infinity-3.de

Abstract:
A classical result, fundamental to evolutionary biology, states that an edge-weighted tree T with leaf set X, positive edge weights, and no vertices of degree 2 can be uniquely reconstructed from the leaf-to-leaf distances between any two elements of X. In biology, X corresponds to a set of taxa (e.g. extant species), the tree T describes their phylogenetic relationships, the edges correspond to earlier species evolving for a time until splitting in two or more species by some speciation/bifurcation event, and their length corresponds to the genetic change accumulating over that time in such a species. In this paper, we investigate which subsets of [Formula: see text] suffice to determine ('lasso') the tree T from the leaf-to-leaf distances induced by that tree. The question is particularly topical since reliable estimates of genetic distance-even (if not in particular) by modern mass-sequencing methods-are, in general, available only for certain combinations of taxa.
Related Concept Videos
 03:21
Phylogenetic Trees
03:21Phylogenetic Trees
 01:23
Phylogeny
 01:28
Microbial Phylogeny
 02:54
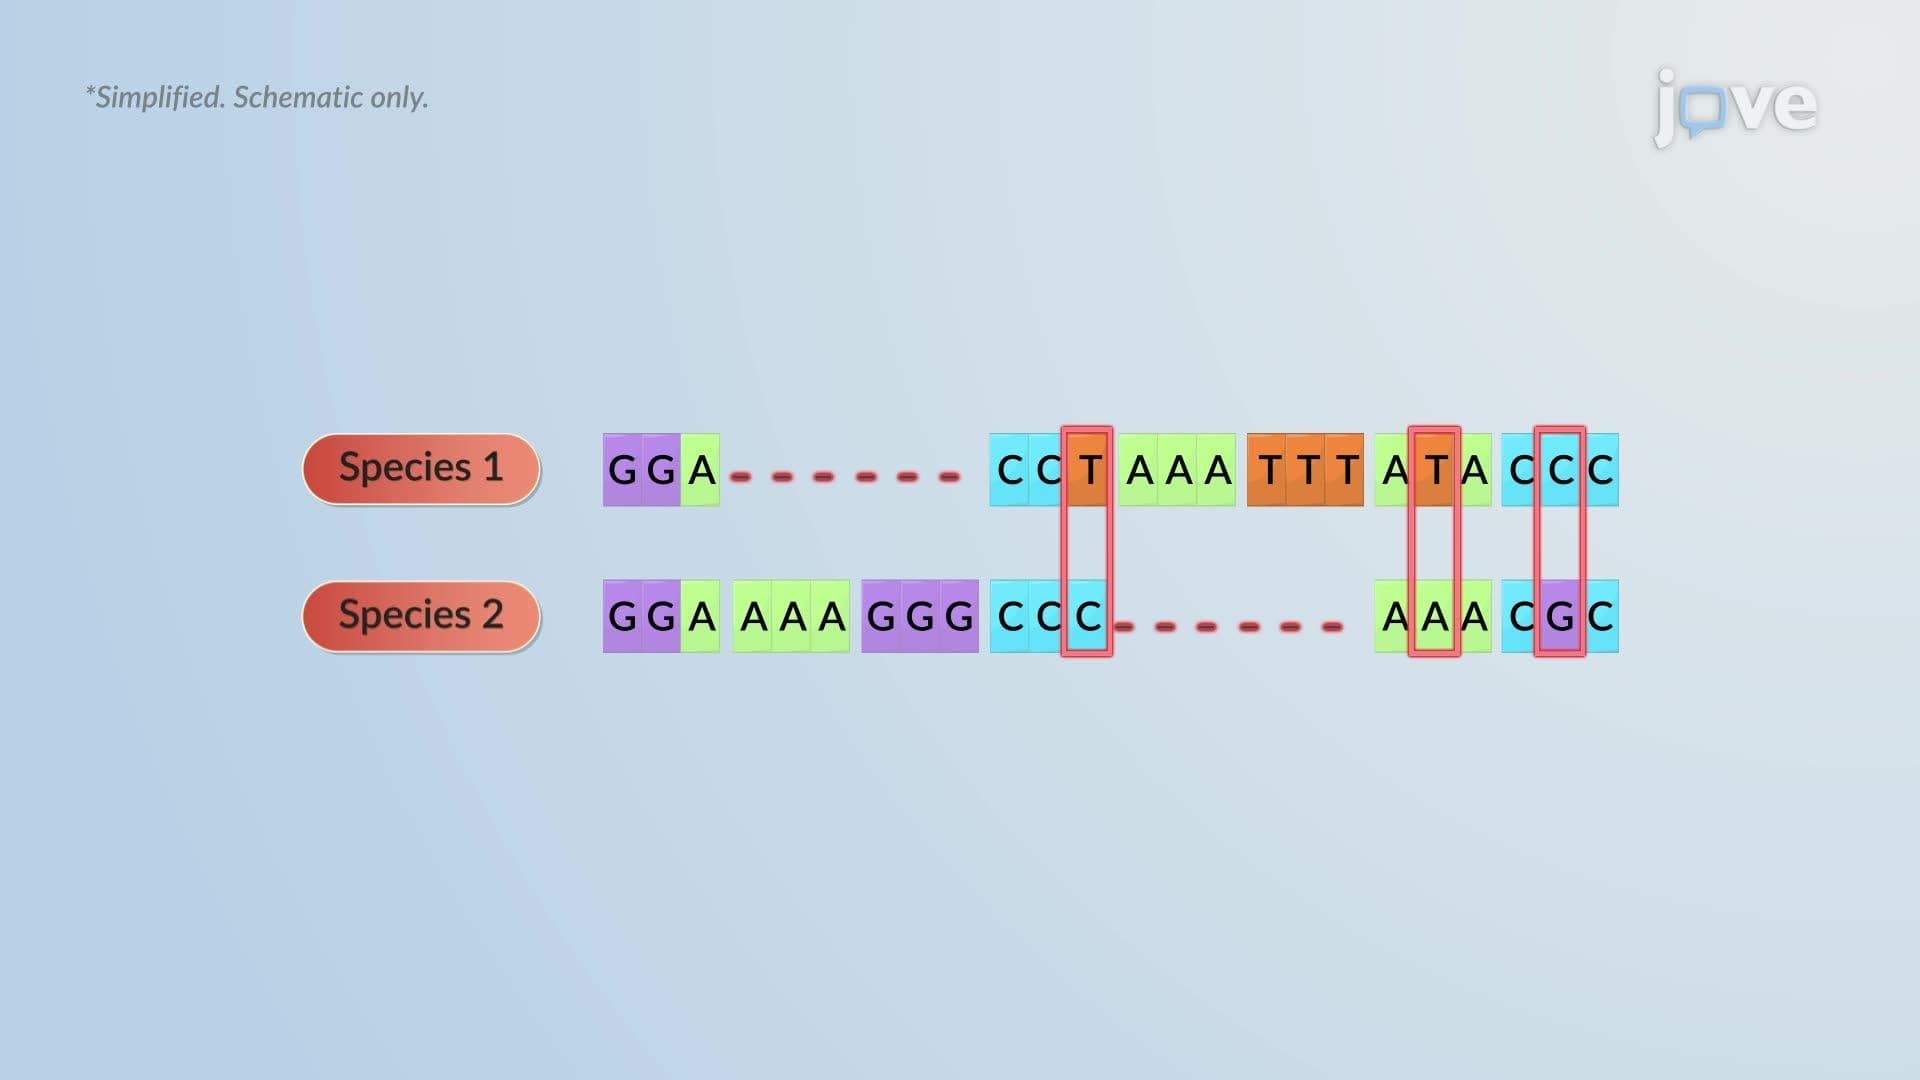
Evolutionary Relationships through Genome Comparisons
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...