Related Experiment Video
Updated: May 31, 2026

An Integrated System to Remotely Trigger Intracellular Signal Transduction by Upconversion Nanoparticle-mediated Kinase Photoactivation
Published on: August 30, 2017
Calculation of pK(a) in proteins with the microenvironment modulated-screened coulomb potential
1Department of Physiology and Biophysics, Weill Cornell Medical College of Cornell University, New York, New York 10065, USA.

Abstract:
The MM-SCP has been applied to predict pK(a) values of titratable residues in wild type and mutants of staphylococcal nuclease (SNase). The calculations were based on crystal structures made available by the Garcia-Moreno Laboratory. In the mutants, mostly deeply buried hydrophobic residues were replaced with ionizable residues, and thus their pK(a) values could be measured and calculated using various methods. The data set used here consisted of a set of WT SNase for which His pK(a) at several ionic strengths had been measured, a set of mutants for which measured pK(a) were available and a set of 11 mutants for which the measured pK(a) were not known at the time of calculation. For this latter set, blind predictions were submitted to the protein pK(a) cooperative, 2009 workshop at Telluride, where the results of the blind predictions were discussed (the RMSD of the submitted set was 1.10 pH units). The calculations on the structures with known pK(a) indicated that in addition to weaknesses of the method, structural issues were observed that led to larger errors (>1) in pK(a) predictions. For example, different crystallographic conditions or steric clashes can lead to differences in the local environment around the titratable residue, which can produce large differences in the calculated pK(a) . To gain further insight into the reliability of the MM-SCP, pK(a) of an extended set of 54 proteins belonging to several structural classes were carried out. Here some initial results from this study are reported to help place the SNase results in the appropriate context.
Related Concept Videos
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Model Components
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:22
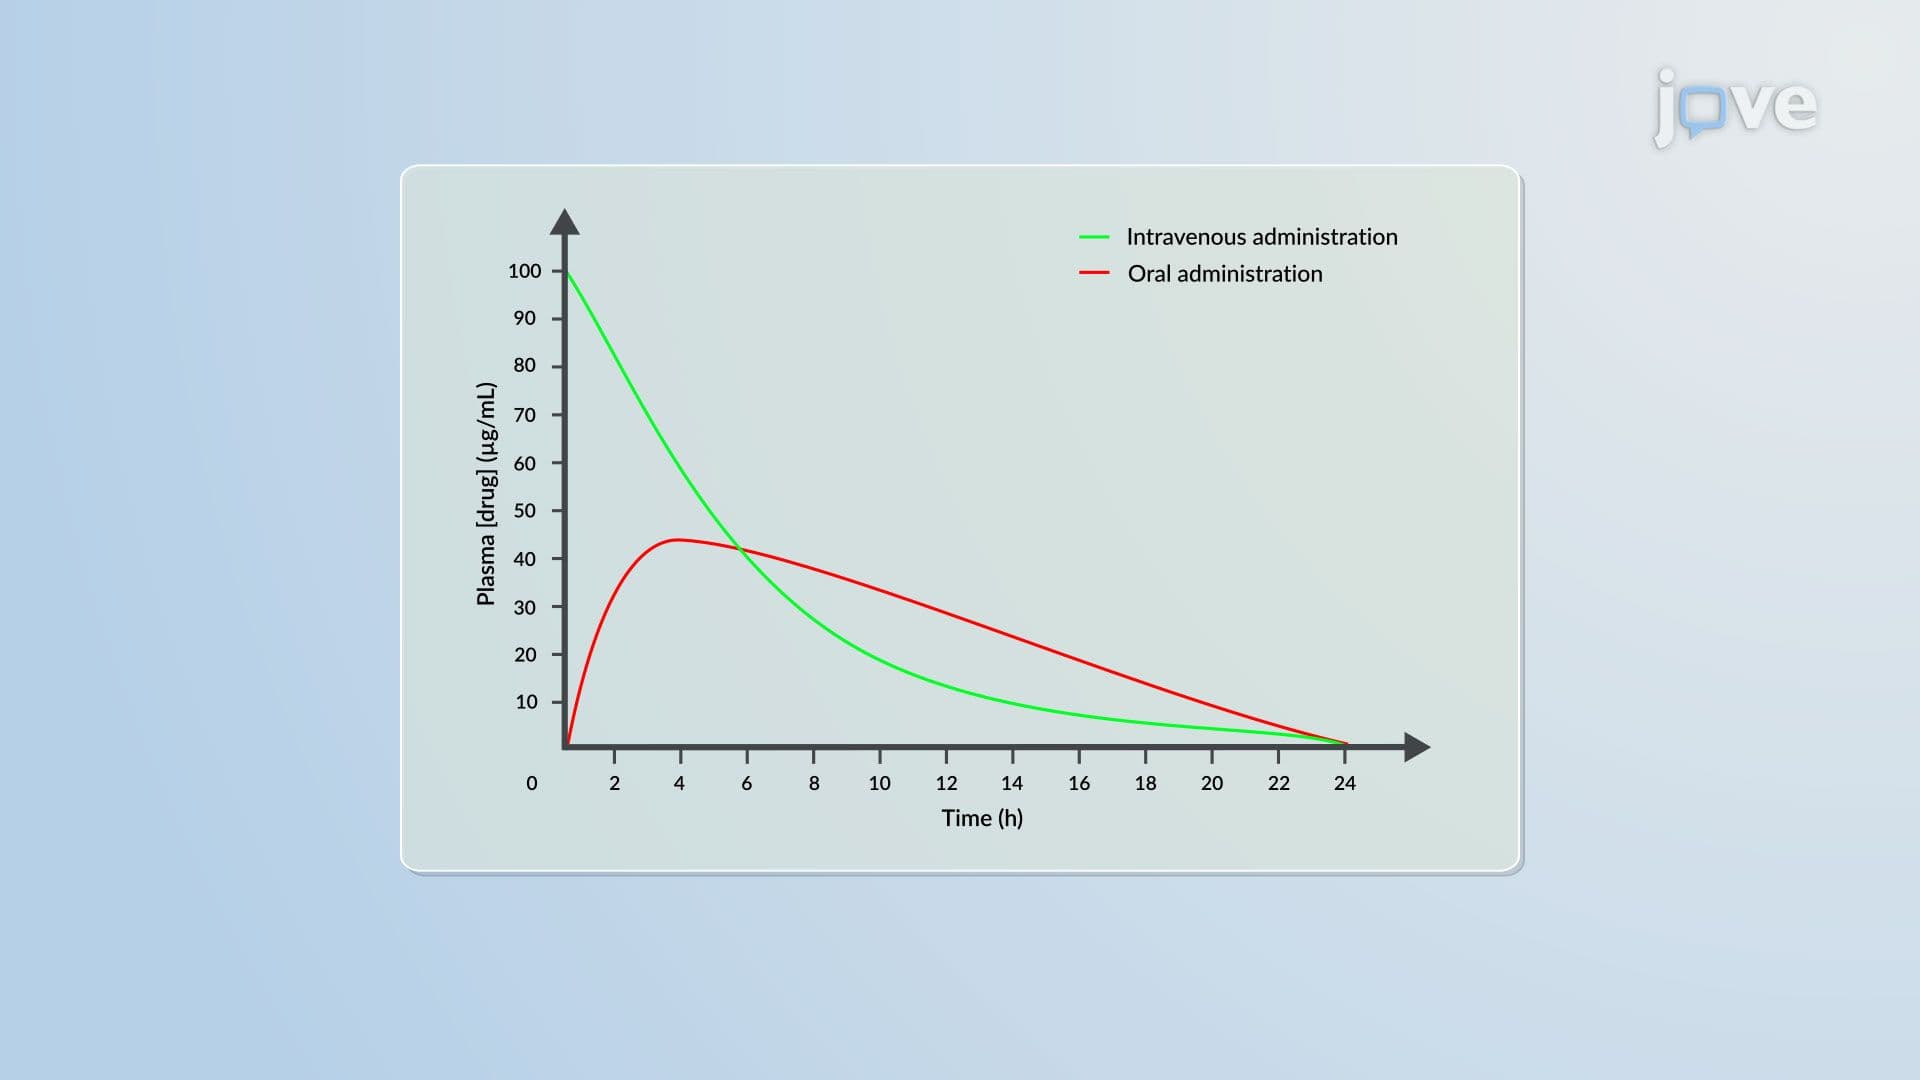
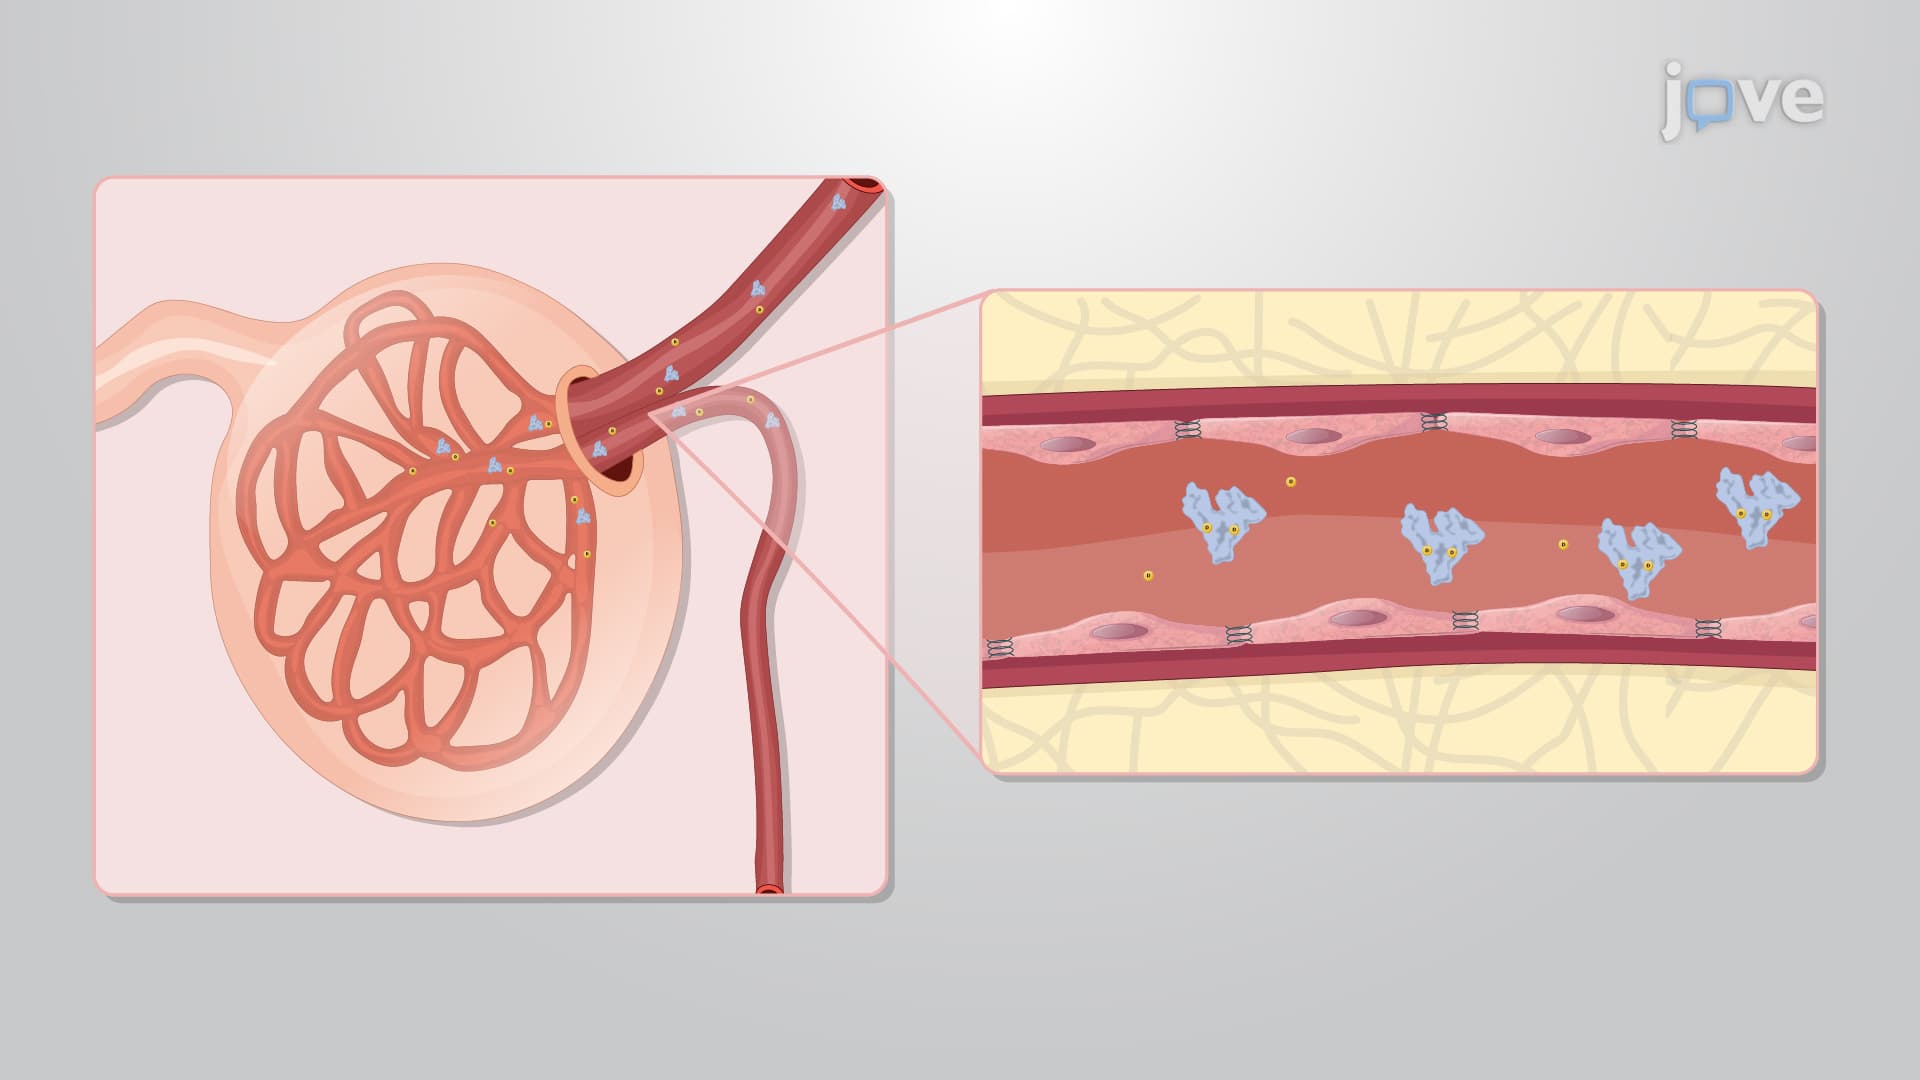
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:24
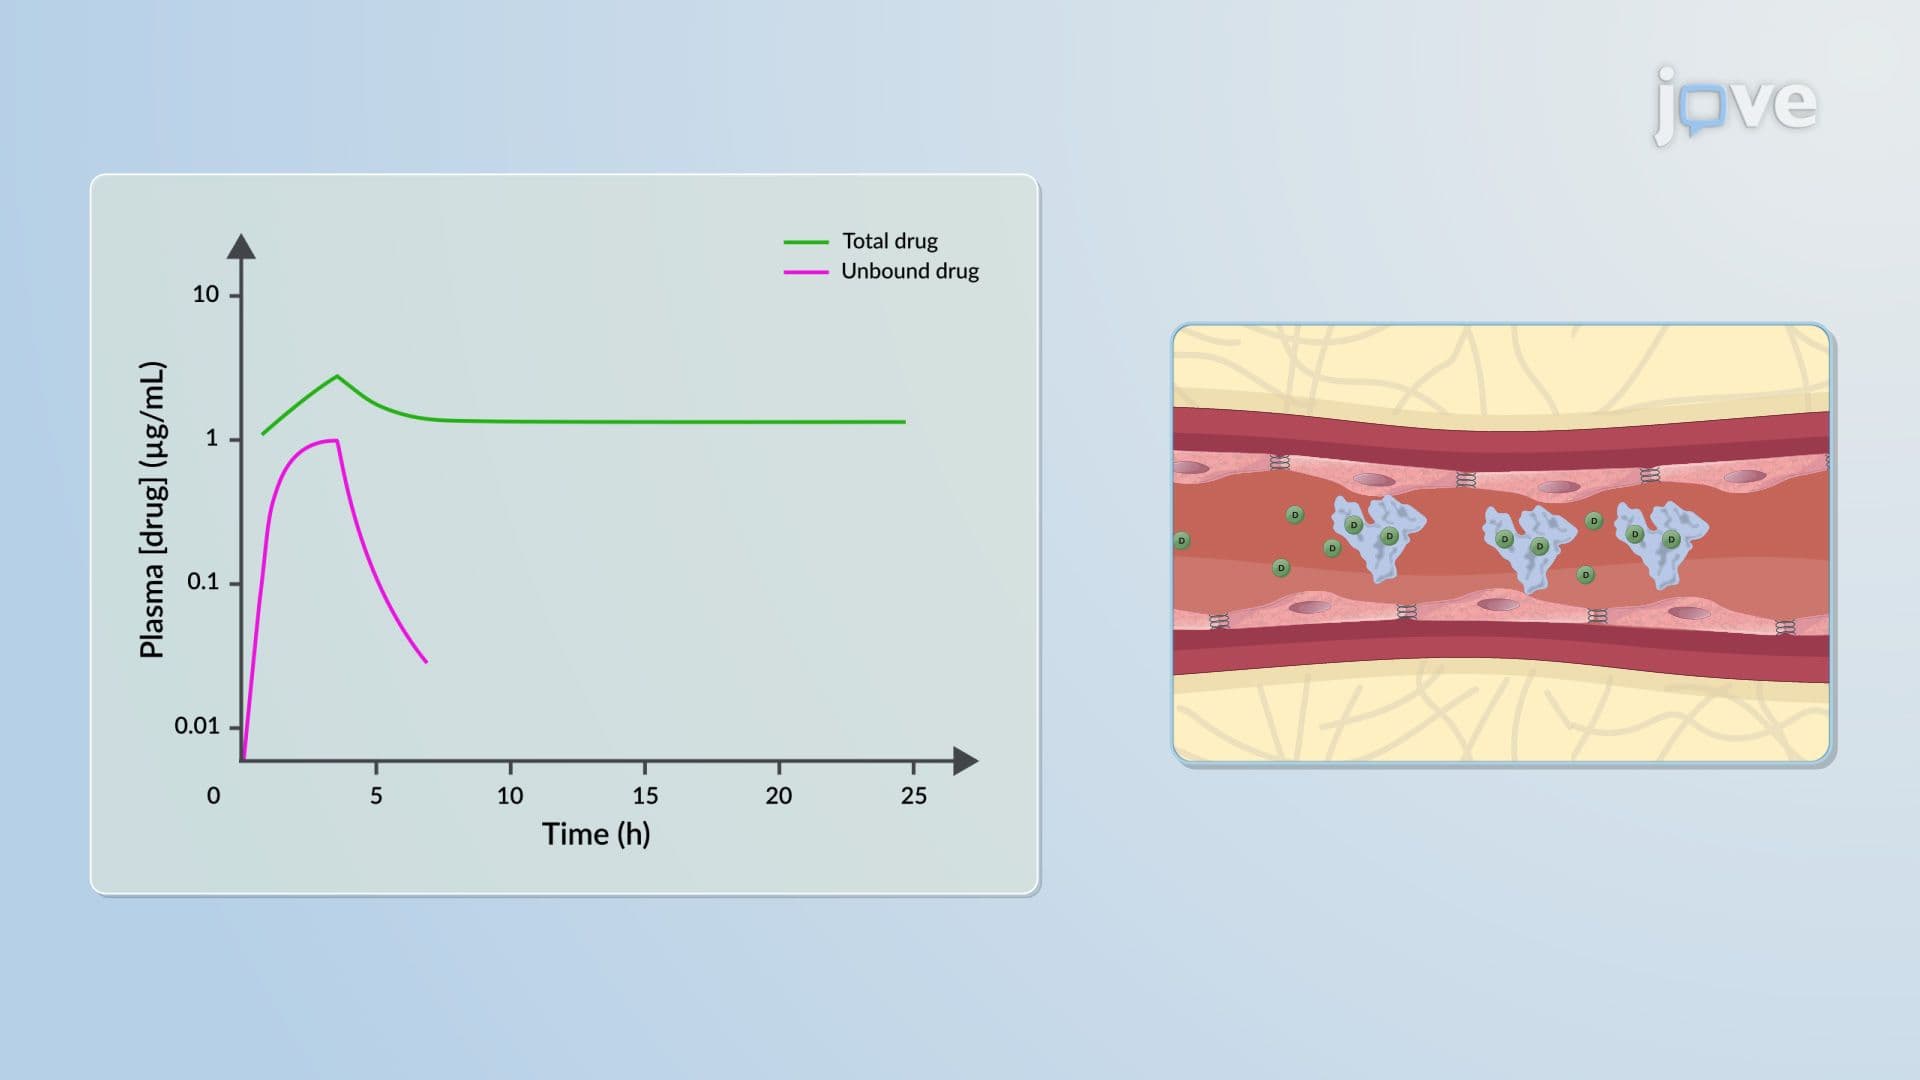
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:24
One-Compartment Open Model: Wagner-Nelson and Loo Riegelman Method for ka Estimation
On...