Related Experiment Video
Updated: May 30, 2026
Determining Surface Areas and Pore Volumes of Metal-Organic Frameworks
Published on: March 8, 2024
Adsorption of benzene on noble metal surfaces studied by density functional theory with Van der Waals correction
Kenji Toyoda1, Ikutaro Hamada, Susumu Yanagisawa
1The Institute of Scientific and Industrial Research (ISIR), Osaka University, 8-1 Mihogaoka, Ibaraki, Osaka 567-0047, Japan.

Abstract:
We have studied the atomic geometries and the electronic properties of benzene/metal interfaces by using density functional theoretical (DFT) calculations with van der Waals corrections. Adsorption energies of benzene on Cu(111), Ag(111), and Au(111) surfaces calculated by van der Waals density functional proposed by Dion and co-workers agree reasonably well with experimentally reported values, while those calculated by a semi-empirical van der Waals correction proposed by Grimme are overestimated slightly. The work function change induced by benzene adsorption on the three surfaces are quite well reproduced by the semi-empirical correction, suggesting that weak adsorption geometries can be quite well reproduced by DFT with a semi-empirical dispersion correction scheme.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:25
Adsorption Isotherms II
 01:07
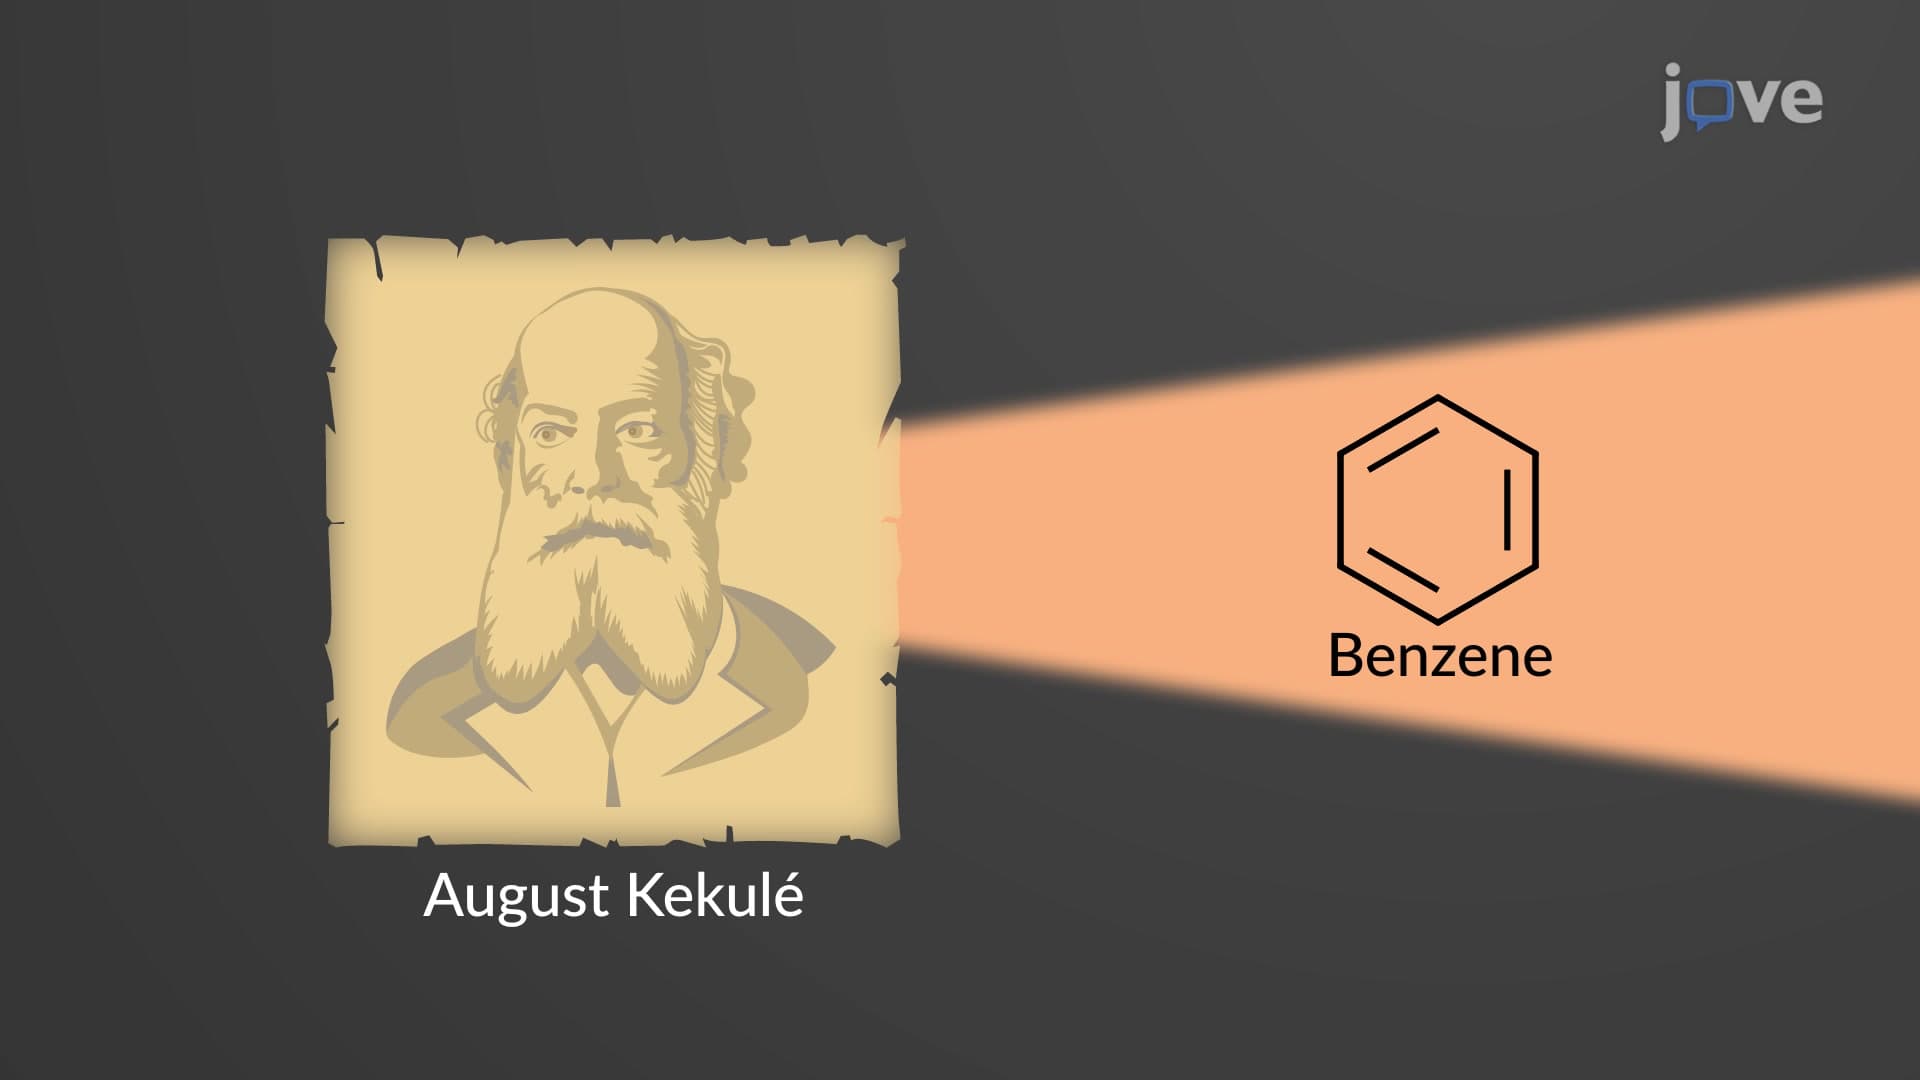
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 03:51
Molecular Orbital Theory II
 01:28
Adsorption of Gases on Solids
 01:29
Adsorption Isotherms I