Related Experiment Video
Updated: Aug 12, 2026
An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
Clinical significance of pharmacokinetic models of hepatic elimination
1Department of Pharmaceutics, Victorian College of Pharmacy, Melbourne, Australia.

Abstract:
Various pharmacokinetic models, both simple and complex, have been developed to describe the way in which the rate of hepatic elimination of drugs depends on hepatic blood flow, hepatic intrinsic clearance and unbound fraction of drug in blood. A model is necessary because it is not possible to measure the average blood concentration of drug within the liver, i.e. the concentration at the site of drug elimination. However, the predictions of these models can differ markedly for drugs of high hepatic clearance, especially with the oral route of administration. Investigations of the models have mostly involved studies with in vitro experimental preparations, such as isolated perfused livers. While such studies have advanced our understanding of the mechanism of hepatic uptake and elimination processes, the implications for clinical drug usage have been somewhat neglected. Use of one of the available models is necessary for the assessment of the capacity of in vivo hepatic drug metabolism processes (i.e. hepatic intrinsic clearance) and for predicting the effect of increasing dose on blood concentrations of high clearance drugs exhibiting Michaelis-Menten elimination kinetics, especially those that undergo a nonlinear hepatic first-pass effect. Clinically significant differences between the models can occur under these circumstances. A model is also required for quantitative prediction of the effect on blood drug concentrations of changes in hepatic blood flow, hepatic intrinsic clearance or drug-protein binding in blood. It is in predicting these changes that differences of major clinical significance can occur between the models. The greatest differences are seen in predicting the effect for orally administered drugs of changes of hepatic blood flow on blood concentrations, and changes of protein binding on unbound blood concentrations of drug. These changes can result from disease processes, altered physiology (old age or pregnancy), food intake or concomitant administration of other drugs. A model is also required for determining the mechanism by which such clinical changes occur. When considering these effects on hepatic elimination, it is essential to appreciate that the conclusions may depend markedly on the particular model chosen. Until more data on the applicability of the models are obtained in humans, the undistributed sinusoidal and venous equilibrium models, which represent the opposite extremes of behaviour among the available models, should both be used in assessing hepatic drug elimination.
Related Concept Videos
 01:20
Pharmacokinetic Models: Overview
There are three primary types of models: empirical, compartment, and physiological. Empirical models, with minimal assumptions,...
 01:15
Model Approaches for Pharmacokinetic Data: Physiological Models
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:14
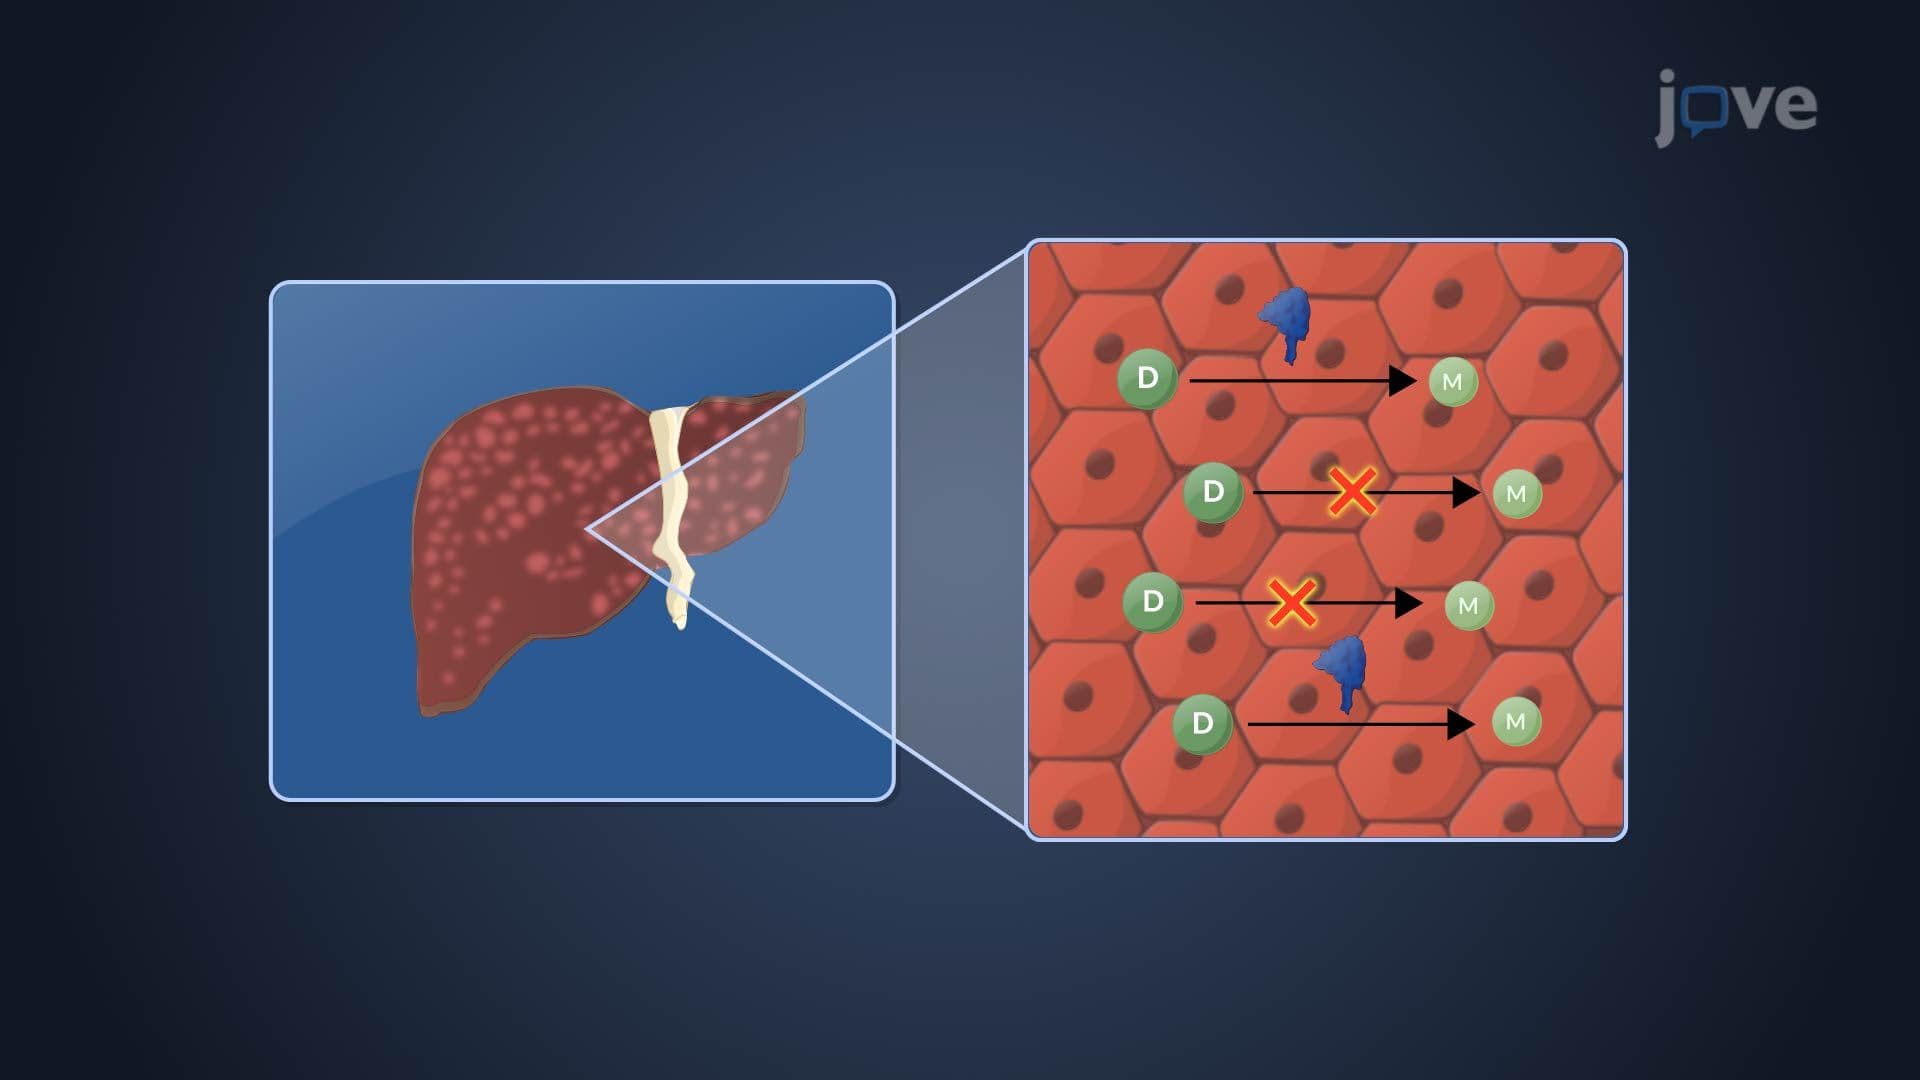
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug