Related Experiment Video
Updated: May 30, 2026

Preparation and Characterization of Individual and Multi-drug Loaded Physically Entrapped Polymeric Micelles
Published on: August 28, 2015
Correlation between rheological properties, in vitro release, and percutaneous permeation of tetrahydropalmatine
Chunmei Li1, Chao Liu, Jie Liu
1Department of Pharmaceutical Sciences, Shenyang Pharmaceutical University, Liaoning, China.

Abstract:
The aim of the present work was to investigate the influence of formulation factors including different grades of Carbopol® matrices and penetration enhancers on the percutaneous permeation of tetrahydropalmatine (THP), rheological properties, and in vitro release; and the correlation behind rheological properties, in vitro release, and percutaneous permeation. Transdermal penetration of THP through excised rabbit skin and in vitro release of THP across transparent Cellophane® were performed by vertical Franz diffusion cell. Rheological analyses were proceeded in terms of "steady flow tests", "oscillation stress sweep", and "creep recovery". The result of percutaneous penetration of THP indicated that, the emulgel prepared with Carbopol® 971P (Cp 971P) as the matrix and N-methyl-2-pyrrolidone (NMP) as the penetration enhancer had the highest cumulative permeation amount (118.19 μg/cm(2)). All the experimental data showed a good fit to the Casson model in viscosimetric studies no matter what the types of matrices or the kinds of penetration enhancers were. The release profile fitted the zero-order release kinetics model with Cp 971P as the matrix without any penetration enhancers. However, when adding penetration enhancers, in vitro release of THP presented anomalous (non-Fickian) release kinetics. Clarifying the relationship behind percutaneous permeation of THP, rheological properties, and in vitro release will provide us with profound insights and facilitate the design of specific emulgel.
More Related Videos
Related Concept Videos
 01:20
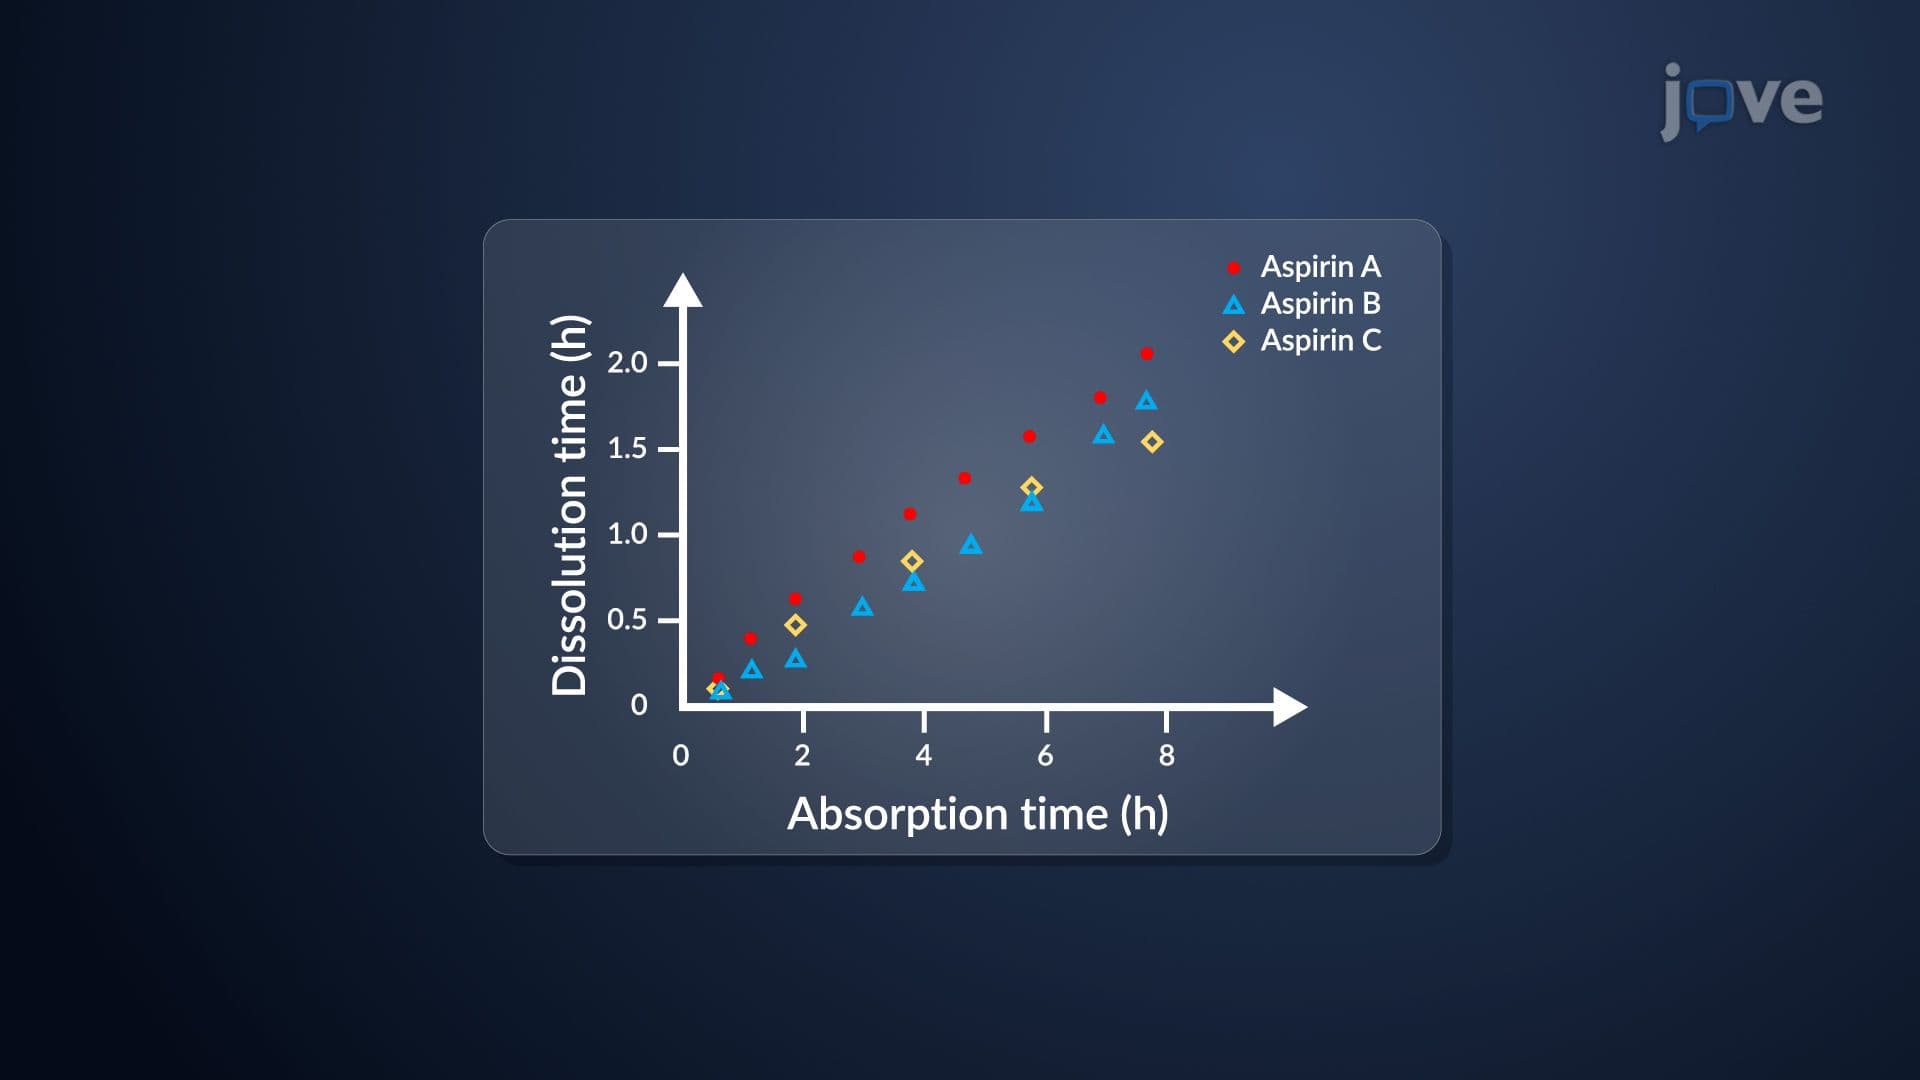
Drug Product Performance: In Vitro–In Vivo Correlation
 01:09
In Vitro Drug Dissolution: Compendial Testing Models II
 01:17
In Vitro Drug Dissolution: Alternative Methods
 01:21
Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH
A drug's pKa and the pH of the gastrointestinal (GI) tract play crucial roles in drug...
 01:27
Factors Influencing Drug Absorption: Drug Dissolution
 01:02
Noncompartmental Analysis: Mean Transit, Absorption and Dissolution Time
One of the key parameters is the mean transit time (MTT), which refers to the total duration required for drug molecules to transit through the body. MTT is determined by calculating the ratio of the area under the moment curve to the area...