Related Experiment Video
Updated: May 30, 2026
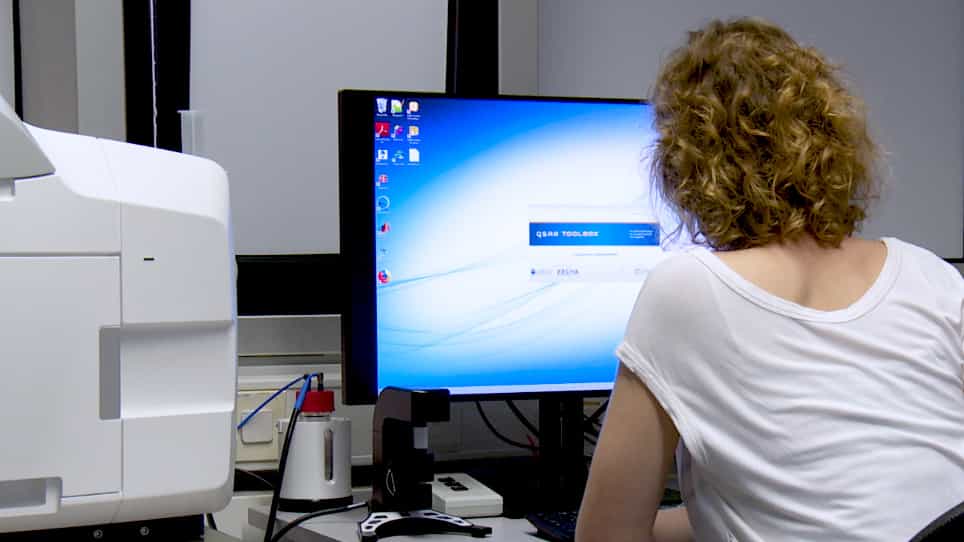

In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
Crystal structure and density functional theory studies of toxic quinone metabolites of polychlorinated biphenyls
Yang Song1, Jyothirmai Ambati, Sean Parkin
1Key Laboratory on Luminescence and Real-Time Analysis, Ministry of Education, College of Pharmaceutical Sciences, Southwest University, Chong Qing 400716, PR China.

Abstract:
Lower chlorinated polychlorinated biphenyls (PCBs) are readily metabolized via hydroxylated metabolites to reactive PCB quinones. Although these PCB metabolites elicit biochemical changes by mechanisms involving cellular target molecules, such as the aryl hydrocarbon receptor, and toxicity by interacting with enzymes like topoisomerases, only few PCB quinones have been synthesized and their conformational properties investigated. Similar to the parent compounds, knowledge of the three-dimensional structure of PCB quinones may therefore be important to assess their fate and risk. To address this gap in our knowledge, the gas phase molecular structure of a series of PCB quinones was predicted using HF/3-21G, B3LYP/6-31G∗∗ and UB3LYP/6-311G∗∗ calculations and compared to the respective solid state structure. All three methods overestimated the Cl-C bond length, but otherwise provided a reasonable approximation of the solid state bond angles and bond lengths. Overall, the UB3LYP/6-311G∗∗ level of theory yielded the best approximation of the molecular structure of PCB quinones in the solid state. Chlorine addition at the ortho position of both rings was found to increase the dihedral angle of the resulting quinone compound, which may have important implications for their interaction with cellular targets and, thus, their toxicity.
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory