Dentatorubral-pallidoluysian atrophy
Related Experiment Video
Updated: May 30, 2026
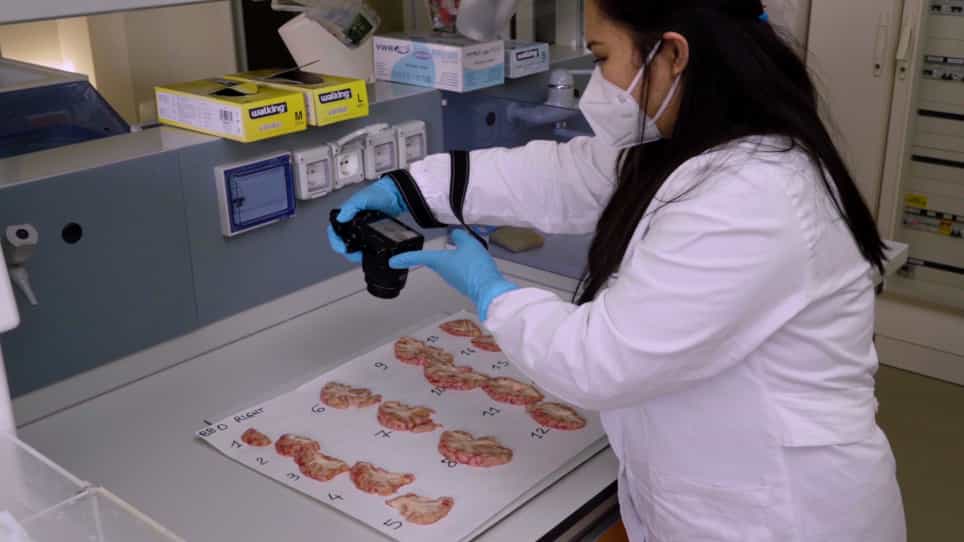
Abbiategrasso Brain Bank Protocol for Collecting, Processing and Characterizing Aging Brains
Published on: June 3, 2020
1Department of Neurology, University of Tokyo, Graduate School of Medicine, Tokyo, Japan. tsuji@m.u-tokyo.ac.jp

Abstract:
Dentatorubral-pallidoluysian atrophy (DRPLA) is a rare autosomal dominant neurodegenerative disorder clinically characterized by various combinations of cerebellar ataxia, choreoathetosis, myoclonus, epilepsy, dementia, and psychiatric symptoms. The most striking clinical features of DRPLA are the considerable heterogeneity in clinical presentation, depending on the age of onset, and the prominent genetic anticipation. DRPLA is caused by unstable expansion of CAG repeats coding for polyglutamine stretches located in exon 5 of the DRPLA gene. DRPLA is characterized by prominent anticipation, with paternal transmission resulting in more prominent anticipation than does maternal transmission, which is now understood based on the intergenerational stability of the CAG repeats. DRPLA protein (also called atrophin-1) is localized in the nucleus and functions as a transcription co-regulator. Recent immunohistochemical studies on autopsied tissues of patients with DRPLA have demonstrated that diffuse accumulation of mutant DRPLA protein (atrophin-1) in the neuronal nuclei, rather than the formation of neuronal intranuclear inclusions (NIIs), is the predominant pathologic condition and involves a wide range of central nervous system regions far beyond the systems previously reported to be affected. Thus, age-dependent and CAG repeat-dependent intranuclear accumulation of mutant DRPLA leading to nuclear dysfunctions are suggested to be the essential pathophysiologic mechanisms in DRPLA.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:23
Alzheimer Disease ll: Pathophysiology
 01:24
Parkinson Disease ll: Pathophysiology
 01:15
Parkinson's Disease: Overview
 01:24
Parkinson Disease l: Introduction
 01:22
Dementia l: Introduction