Related Experiment Video
Updated: May 30, 2026

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Newborn screening for homocystinuria
John H Walter1, Nikki Jahnke, Tracey Remmington
1Willink Biochemical Genetics Unit, Royal Manchester Children's Hospital, Pendlebury, Manchester, UK, M27 4HA.

Insights
Newborn screening for homocystinuria (a rare inherited disorder) lacks controlled studies. Uncontrolled data suggest early diagnosis and treatment benefit patients, but more research is needed.
Area of Science:
- Genetics
- Metabolic Disorders
- Public Health
Background:
- Homocystinuria is a rare inherited metabolic disorder caused by cystathionine beta-synthase deficiency.
- Affected individuals appear normal at birth but develop severe complications in childhood.
- Early diagnosis and intervention can prevent or mitigate these complications.
Purpose of the Study:
- To evaluate the clinical benefits of newborn population screening for homocystinuria.
- To compare outcomes between early diagnosis via screening and later clinical diagnosis.
Main Methods:
- Searched the Cochrane Cystic Fibrosis and Genetic Disorders Group's Inborn Errors of Metabolism Trials Register.
- Included randomized controlled trials and controlled clinical trials comparing screened versus non-screened populations for neonatal homocystinuria diagnosis.
- Most recent search conducted on June 27, 2011.
Main Results:
- No eligible studies were identified for inclusion in the review.
- The review could not identify any randomized controlled trials or controlled clinical trials on this topic.
Conclusions:
- Unable to draw conclusions from controlled studies due to lack of eligible research.
- Uncontrolled case series suggest newborn screening and early treatment for homocystinuria are effective.
- Future multicenter, long-term randomized controlled trials are needed to establish robust evidence and cost-effectiveness.
Background:
Homocystinuria is a rare inherited disorder due to a deficiency in cystathionine beta synthase. Individuals with this condition appear normal at birth but develop serious complications in childhood. Diagnosis and treatment started sufficiently early in life can effectively prevent or reduce the severity of these complications.
Objectives:
To determine if newborn population screening for the diagnosis of homocystinuria due to cystathionine beta synthase deficiency leads to clinical benefit compared to later clinical diagnosis.
Search Strategy:
We searched the Cochrane Cystic Fibrosis and Genetic Disorders Group's Inborn Errors of Metabolism Trials Register.Date of the most recent search of the Inborn Errors of Metabolism Register: 27 June 2011.
Selection Criteria:
Randomised controlled trials and controlled clinical trials assessing the use of any neonatal screening test to diagnose infants with homocystinuria before the condition becomes clinically evident. Eligible studies compare a screened population versus a non-screened population.
Data Collection And Analysis:
No studies were identified for inclusion in the review.
Main Results:
No studies were identified for inclusion in the review.
Authors' Conclusions:
We were unable to identify eligible studies for inclusion in this review and hence it is not possible to draw any conclusions based on controlled studies; however, we are aware of uncontrolled case-series which support the efficacy of newborn screening for homocystinuria and its early treatment. Any future randomised controlled trial would need to be both multicentre and long term in order to provide robust evidence for or against screening and to allow a cost effectiveness analysis to be undertaken.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:23
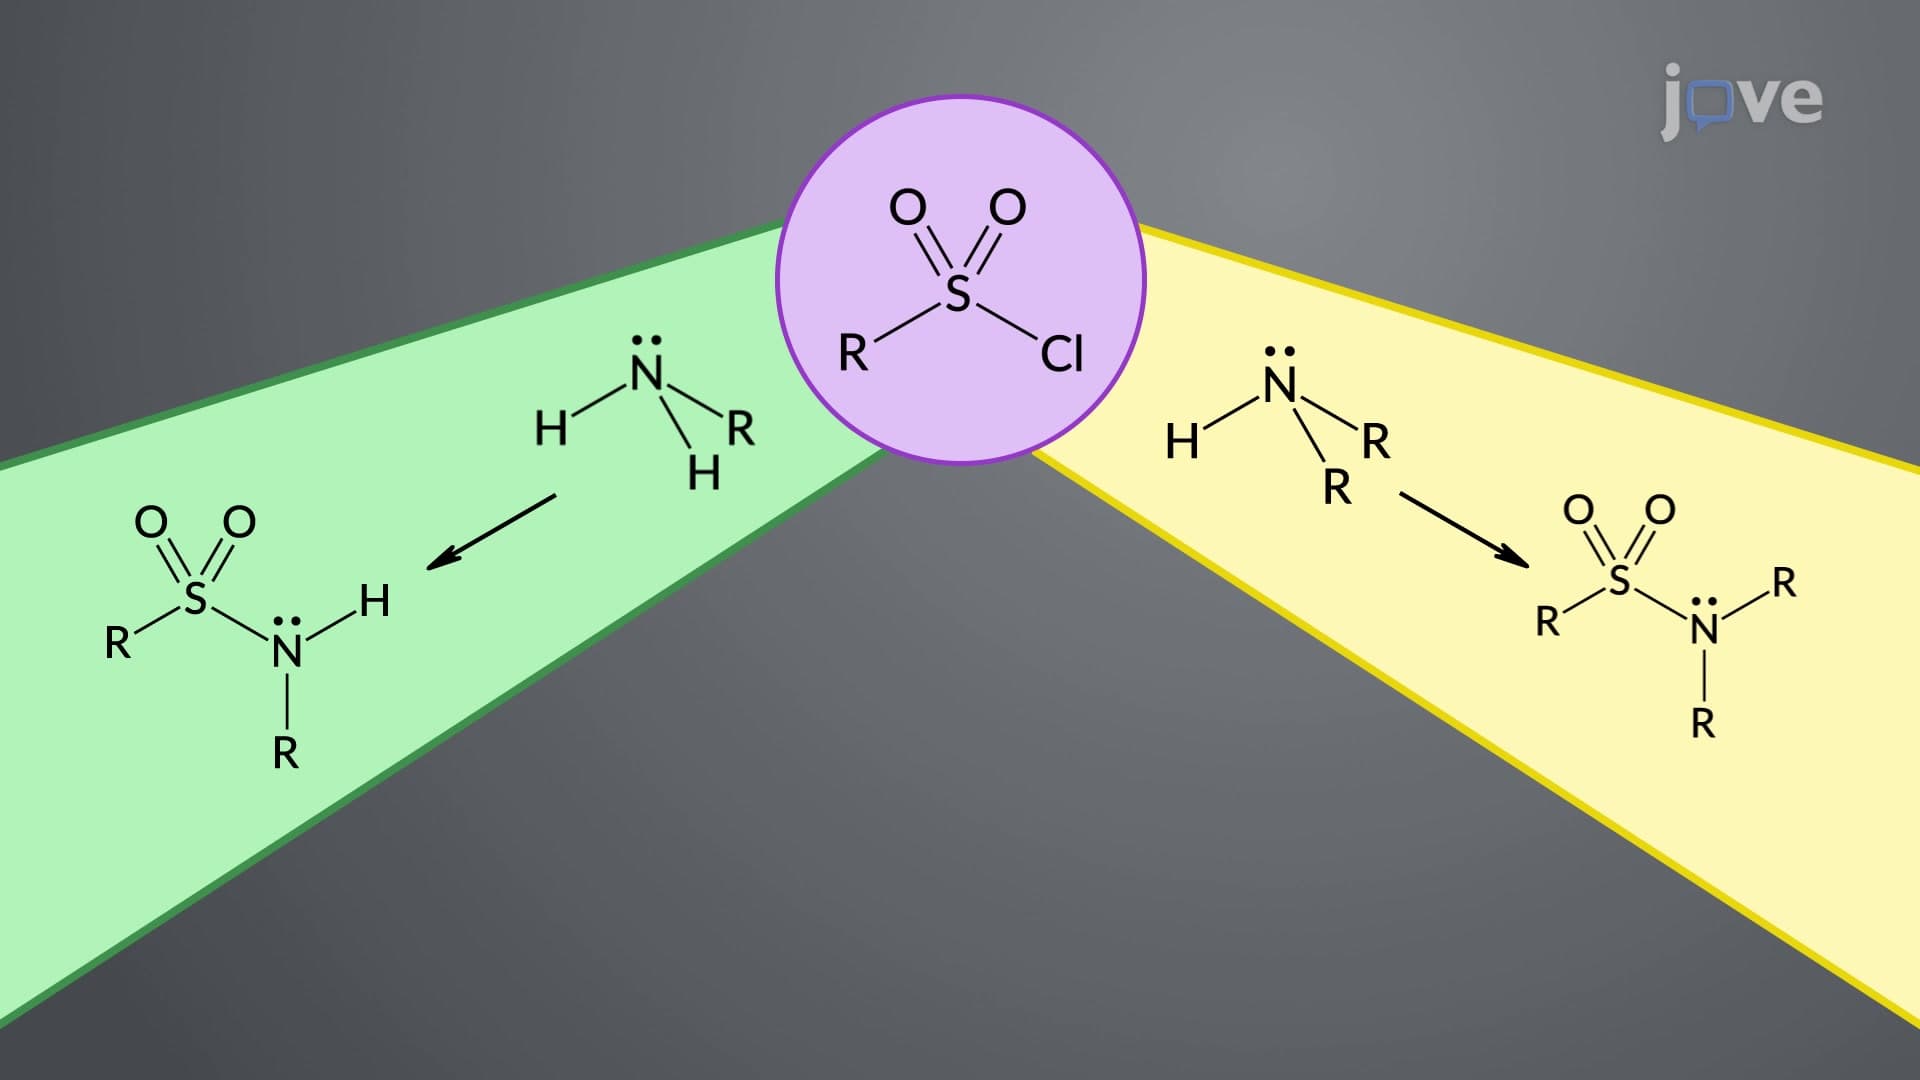
Amines to Sulfonamides: The Hinsberg Test
Generally, a primary amine reacts with the Hinsberg reagent to produce an N-substituted benzenesulfonamide. The electron-withdrawing sulfonyl...