Absolute configuration of podophyllotoxin related lignans from Bursera fagaroides using vibrational circular

René Velázquez-Jiménez1, J Martín Torres-Valencia, Carlos M Cerda-García-Rojas

  • 1Área Académica de Química, Universidad Autónoma del Estado de Hidalgo, Km 4.5 Carretera Pachuca-Tulancingo, Mineral de la Reforma, Hidalgo 42184, Mexico.

Phytochemistry
|August 16, 2011
PubMed

Related Concept Videos

UV–Vis Spectroscopy: Woodward–Fieser Rules01:29

UV–Vis Spectroscopy: Woodward–Fieser Rules

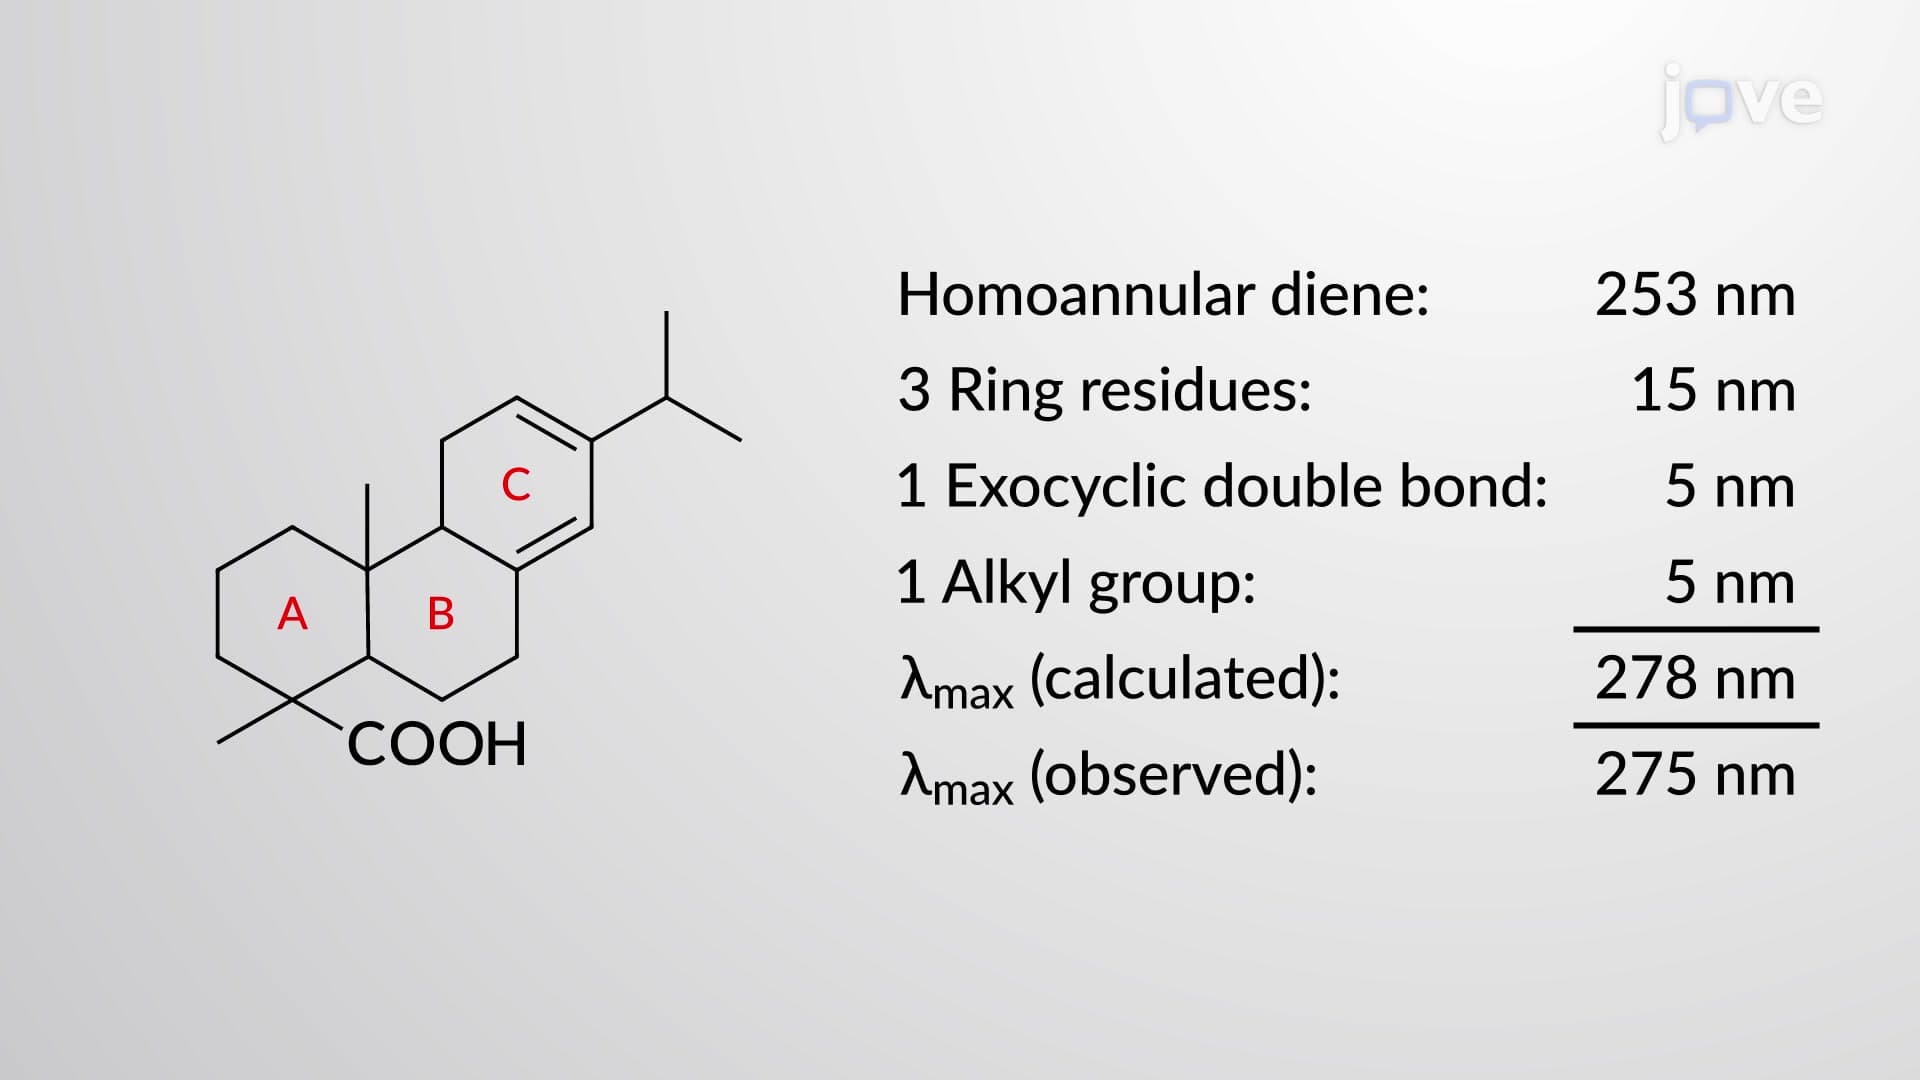
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the contributions...
UV–Vis Spectroscopy of Conjugated Systems01:32

UV–Vis Spectroscopy of Conjugated Systems

Organic compounds with conjugated double bonds show strong absorption features in the UV–visible region of the electromagnetic spectrum attributed to π → π* electronic excitations. Generally, a UV–vis absorption spectrum is recorded as a plot of absorbance vs wavelength. The wavelength of maximum absorbance, which manifests as a peak in the absorption spectrum, is denoted as λmax.
One of the factors influencing λmax is the extent of conjugation in the...
Stereoisomerism of Cyclic Compounds02:33

Stereoisomerism of Cyclic Compounds
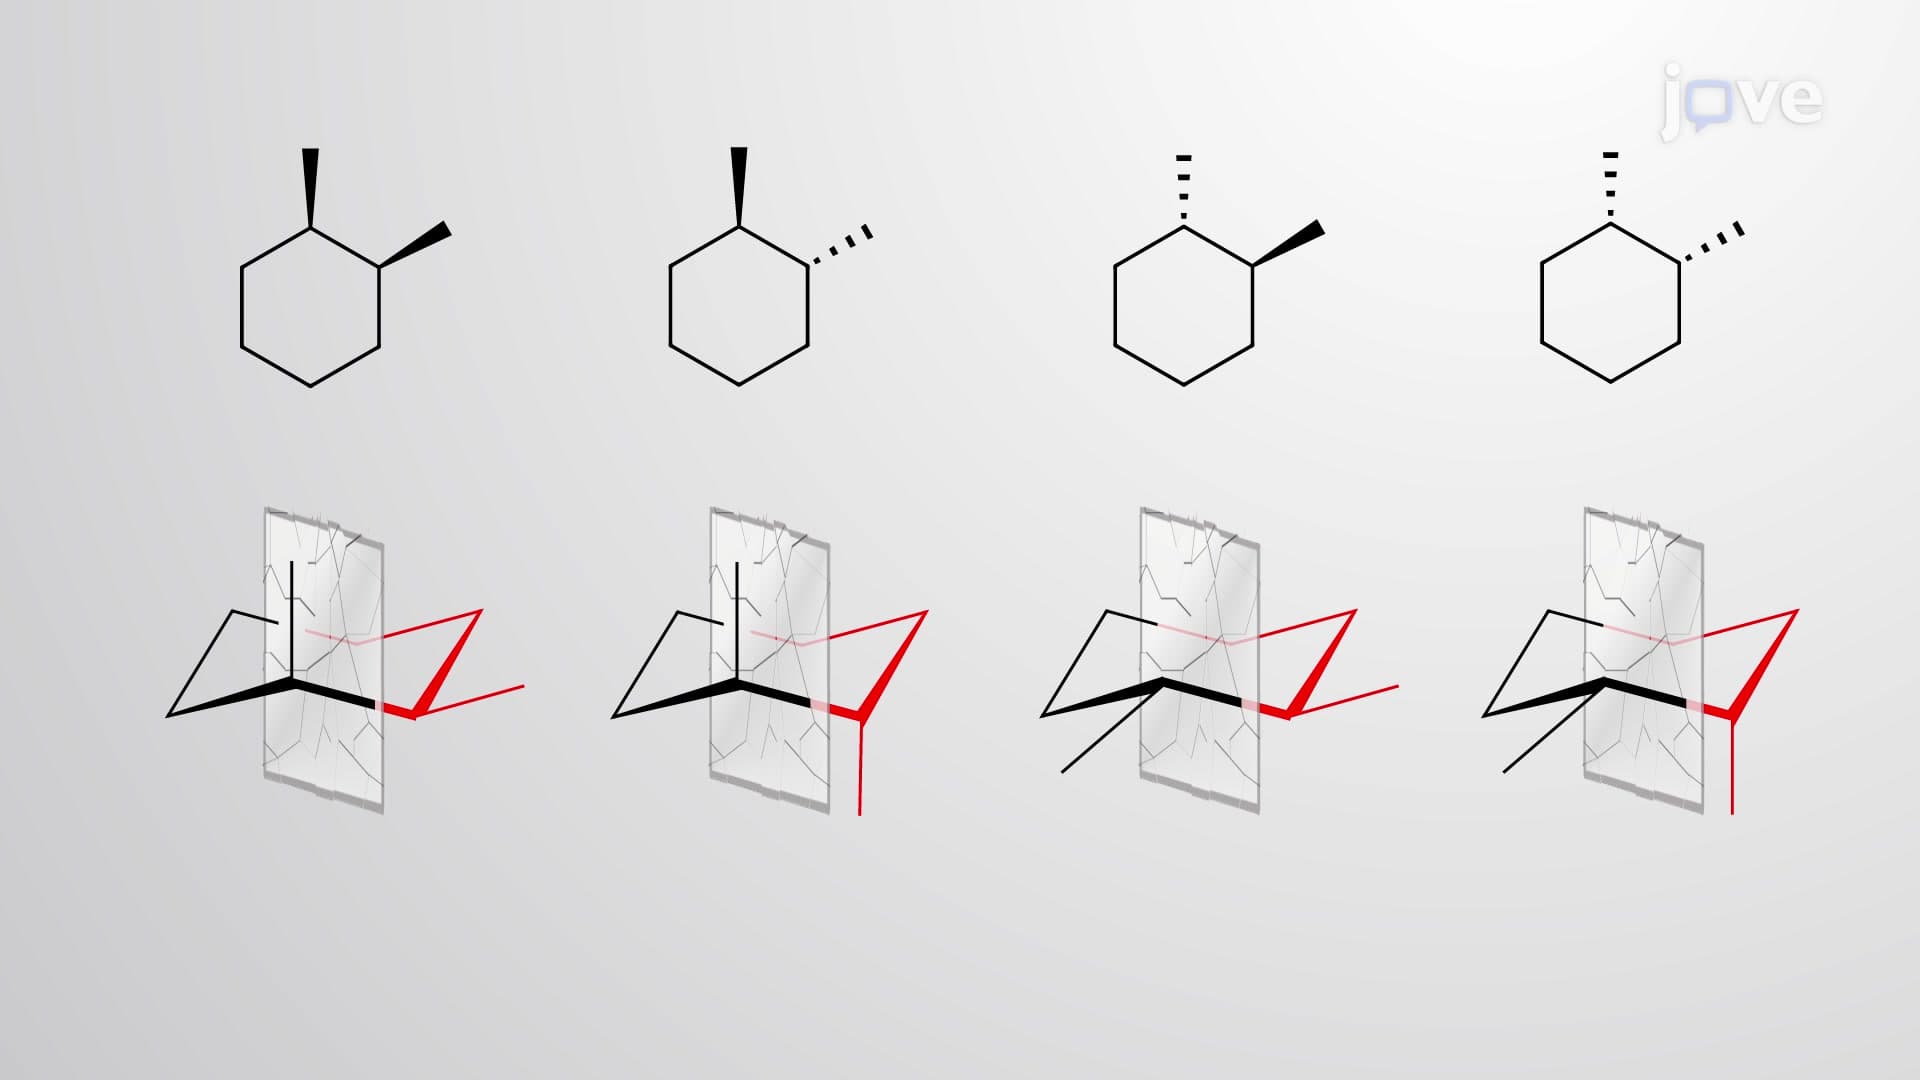
In this lesson, we delve into the role of ring conformation and its stability, which determines the spatial arrangement and, consequently, the molecular symmetry and stereoisomerism of cyclic compounds. 1,2-Dimethylcyclohexane is used as a case study to evaluate the possible number of stereoisomers. Here, given the multiple (n = 2) chiral centers, there are 2n = 4 possible configurations that lack a plane of symmetry, as the ring skeleton exists in a non-planar chair conformation. In addition,...
NMR Spectroscopy of Aromatic Compounds01:14

NMR Spectroscopy of Aromatic Compounds

Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range. Consider...
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds01:14

π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
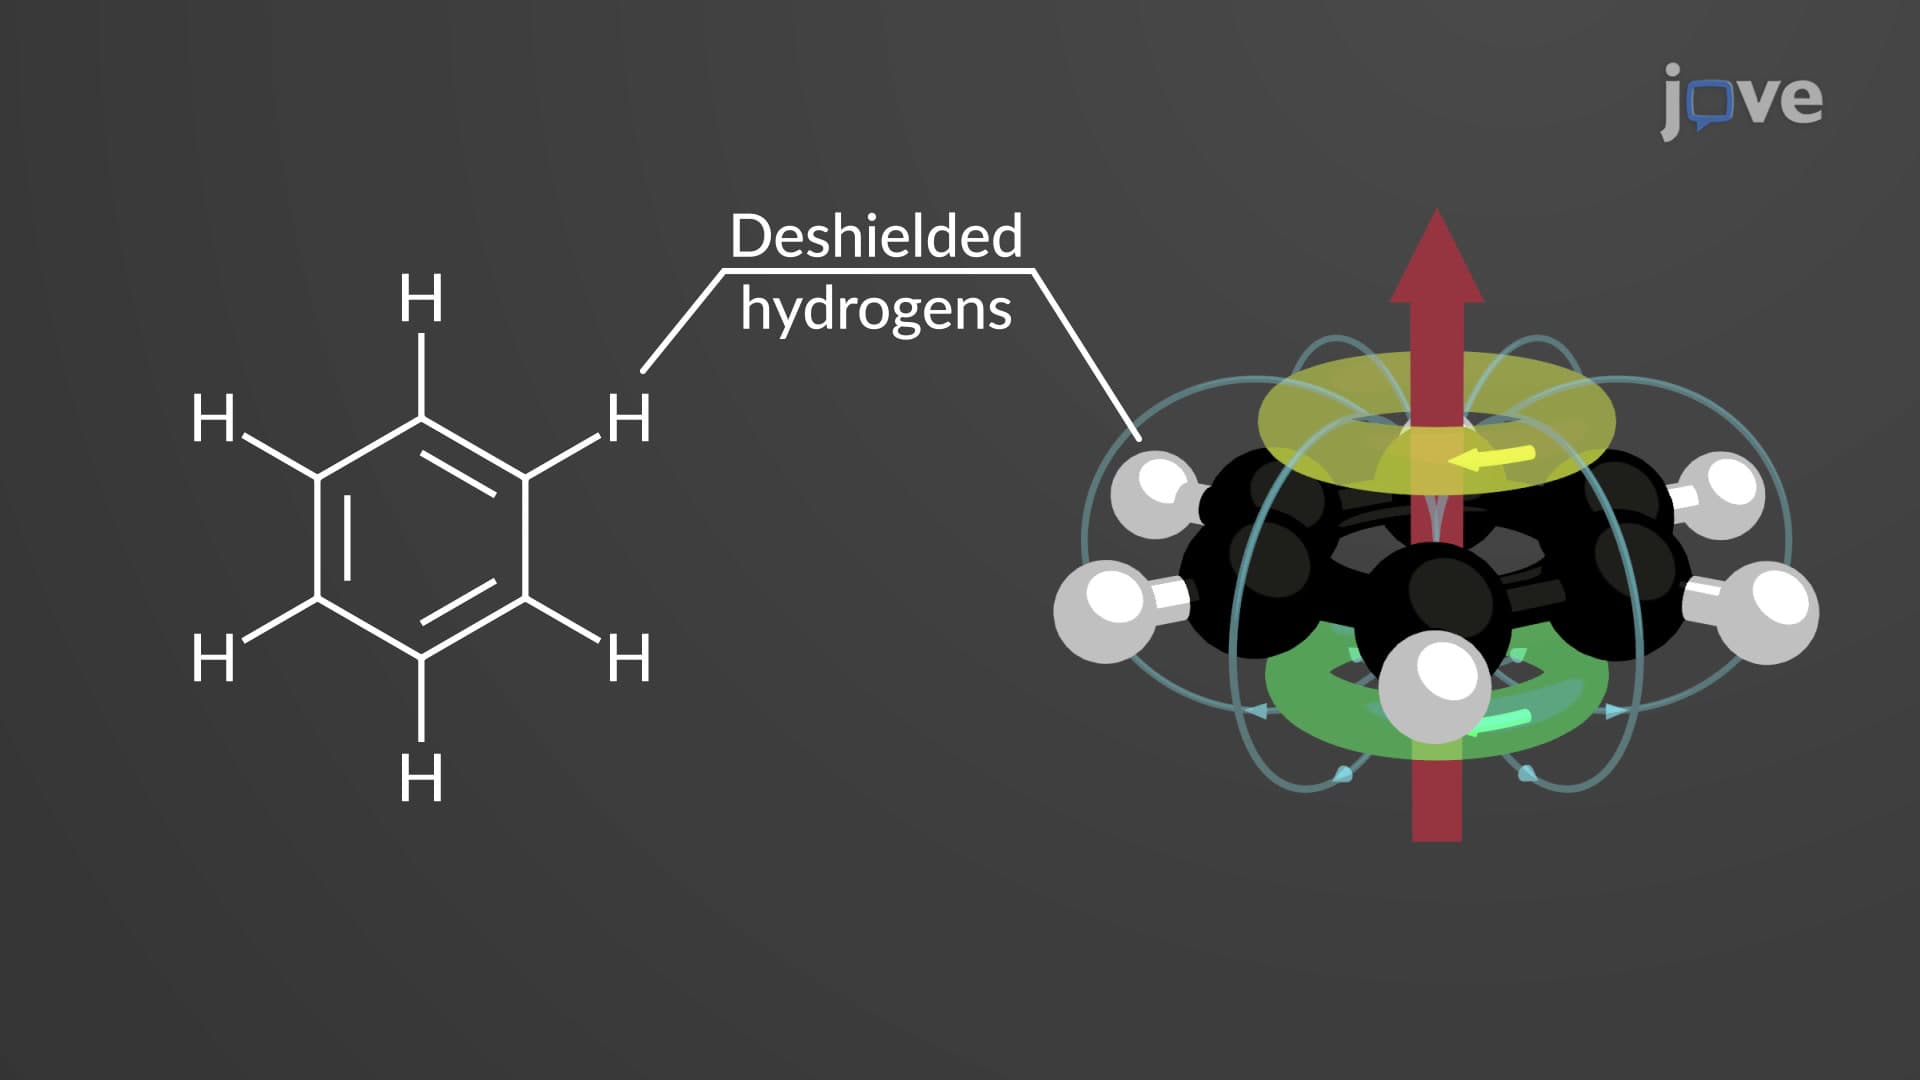
In aromatic compounds, such as benzene, the circulation of (4n + 2) π-electrons sets up a diamagnetic or diatropic ring current around the perimeter of the molecule. This current induces a magnetic field that opposes the external field inside the ring and reinforces it on the outside. The protons in benzene are deshielded and exhibit high chemical shifts in the range 6.5–8.5 ppm. The shielding effect at the center of the ring is evident in complex aromatic molecules, such as annulenes. In...
NMR Spectroscopy of Benzene Derivatives01:37

NMR Spectroscopy of Benzene Derivatives

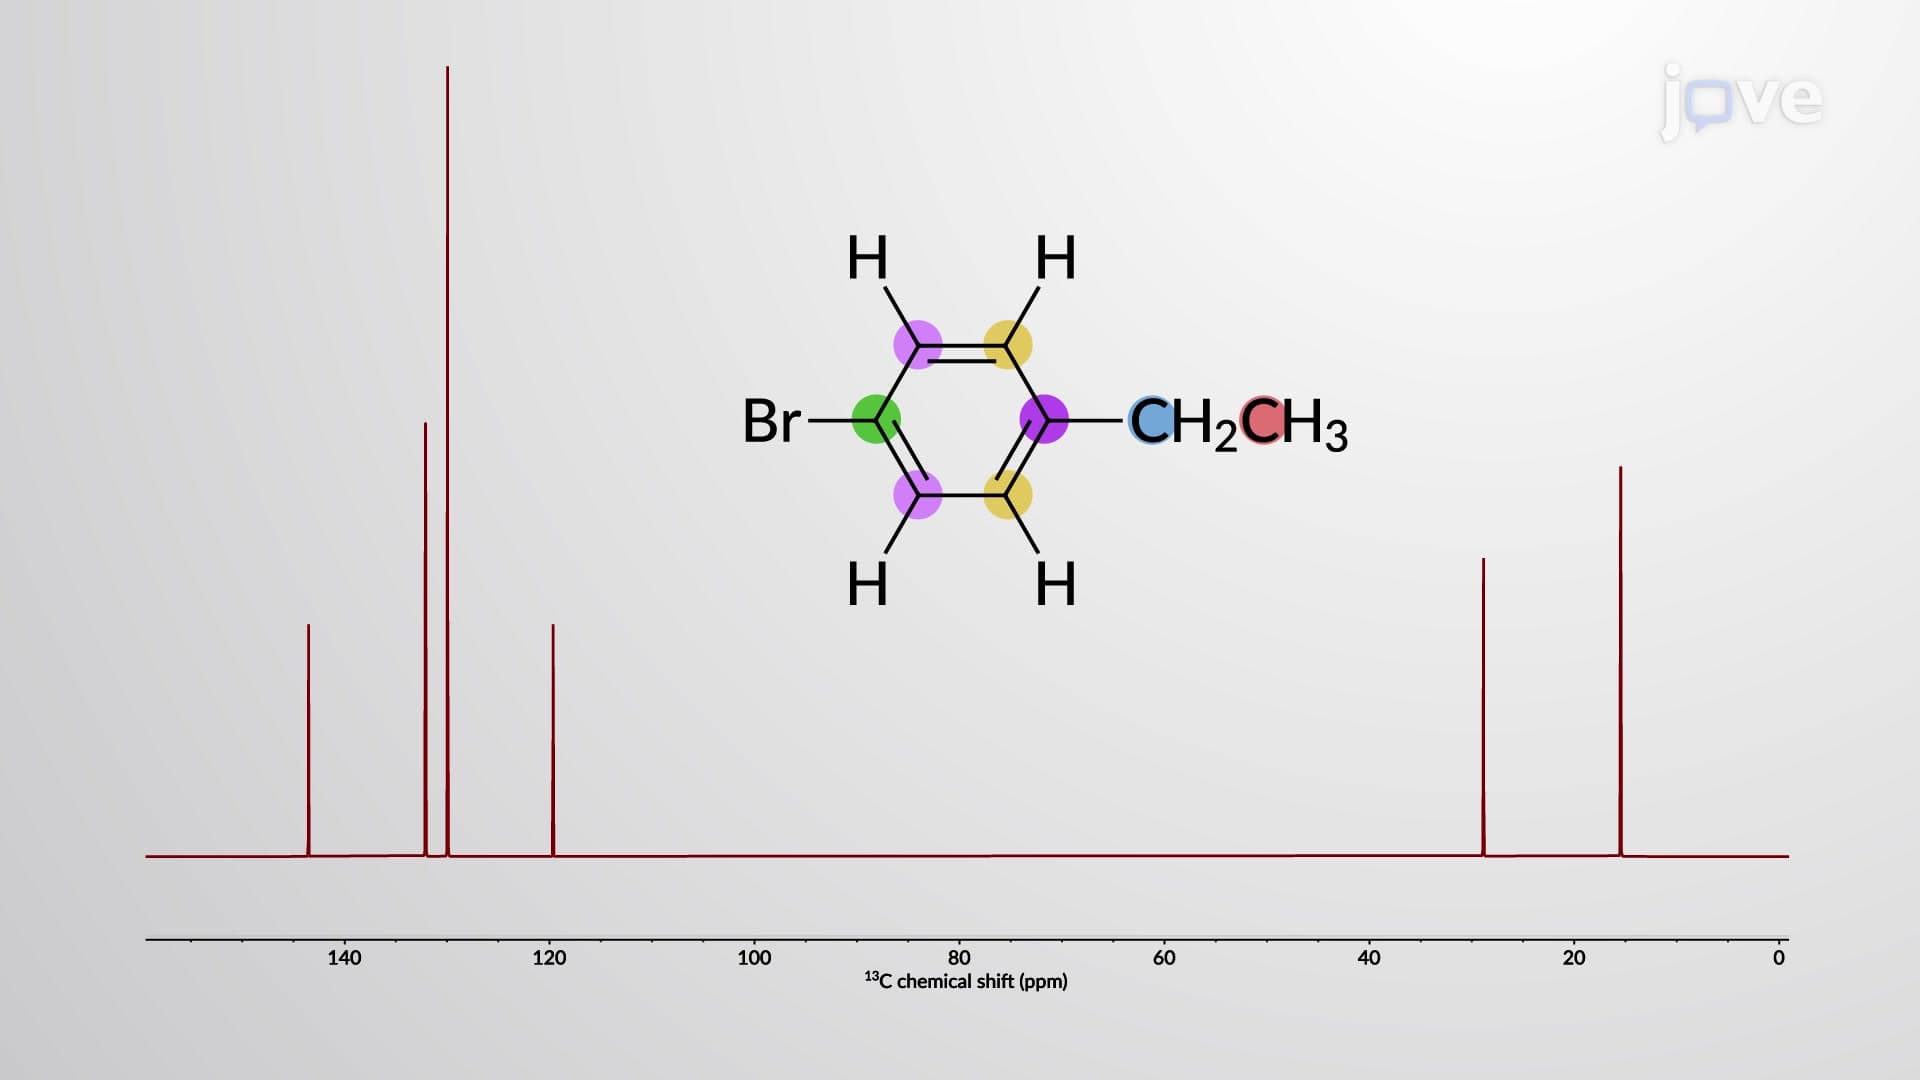
Simple unsubstituted benzene has six aromatic protons, all chemically equivalent. Therefore, benzene exhibits only a singlet peak at δ 7.3 ppm in the 1H NMR spectrum. The observed shift is far downfield because the aromatic ring current strongly deshields the protons. Any substitution on the benzene ring makes the aromatic protons nonequivalent, and the protons split each other. The peak is, therefore, no longer a singlet and the splitting pattern and their associated coupling constants depend...