Related Experiment Video
Updated: May 30, 2026
Modeling Ligands into Maps Derived from Electron Cryomicroscopy
Published on: July 19, 2024
Free Energy Perturbation Hamiltonian Replica-Exchange Molecular Dynamics (FEP/H-REMD) for Absolute Ligand Binding
1Biosciences Division, Argonne National Laboratory, 9700 South Cass Avenue, Building 221, Argonne, Illinois 60439.

Abstract:
Free Energy Perturbation with Replica Exchange Molecular Dynamics (FEP/REMD) offers a powerful strategy to improve the convergence of free energy computations. In particular, it has been shown previously that a FEP/REMD scheme allowing random moves within an extended replica ensemble of thermodynamic coupling parameters "lambda" can improve the statistical convergence in calculations of absolute binding free energy of ligands to proteins [J. Chem. Theory Comput. 2009, 5, 2583]. In the present study, FEP/REMD is extended and combined with an accelerated MD simulations method based on Hamiltonian replica-exchange MD (H-REMD) to overcome the additional problems arising from the existence of kinetically trapped conformations within the protein receptor. In the combined strategy, each system with a given thermodynamic coupling factor lambda in the extended ensemble is further coupled with a set of replicas evolving on a biased energy surface with boosting potentials used to accelerate the inter-conversion among different rotameric states of the side chains in the neighborhood of the binding site. Exchanges are allowed to occur alternatively along the axes corresponding to the thermodynamic coupling parameter lambda and the boosting potential, in an extended dual array of coupled lambda- and H-REMD simulations. The method is implemented on the basis of new extensions to the REPDSTR module of the biomolecular simulation program CHARMM. As an illustrative example, the absolute binding free energy of p-xylene to the nonpolar cavity of the L99A mutant of T4 lysozyme was calculated. The tests demonstrate that the dual lambda-REMD and H-REMD simulation scheme greatly accelerates the configurational sampling of the rotameric states of the side chains around the binding pocket, thereby improving the convergence of the FEP computations.
More Related Videos
 15:05
Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 02:18
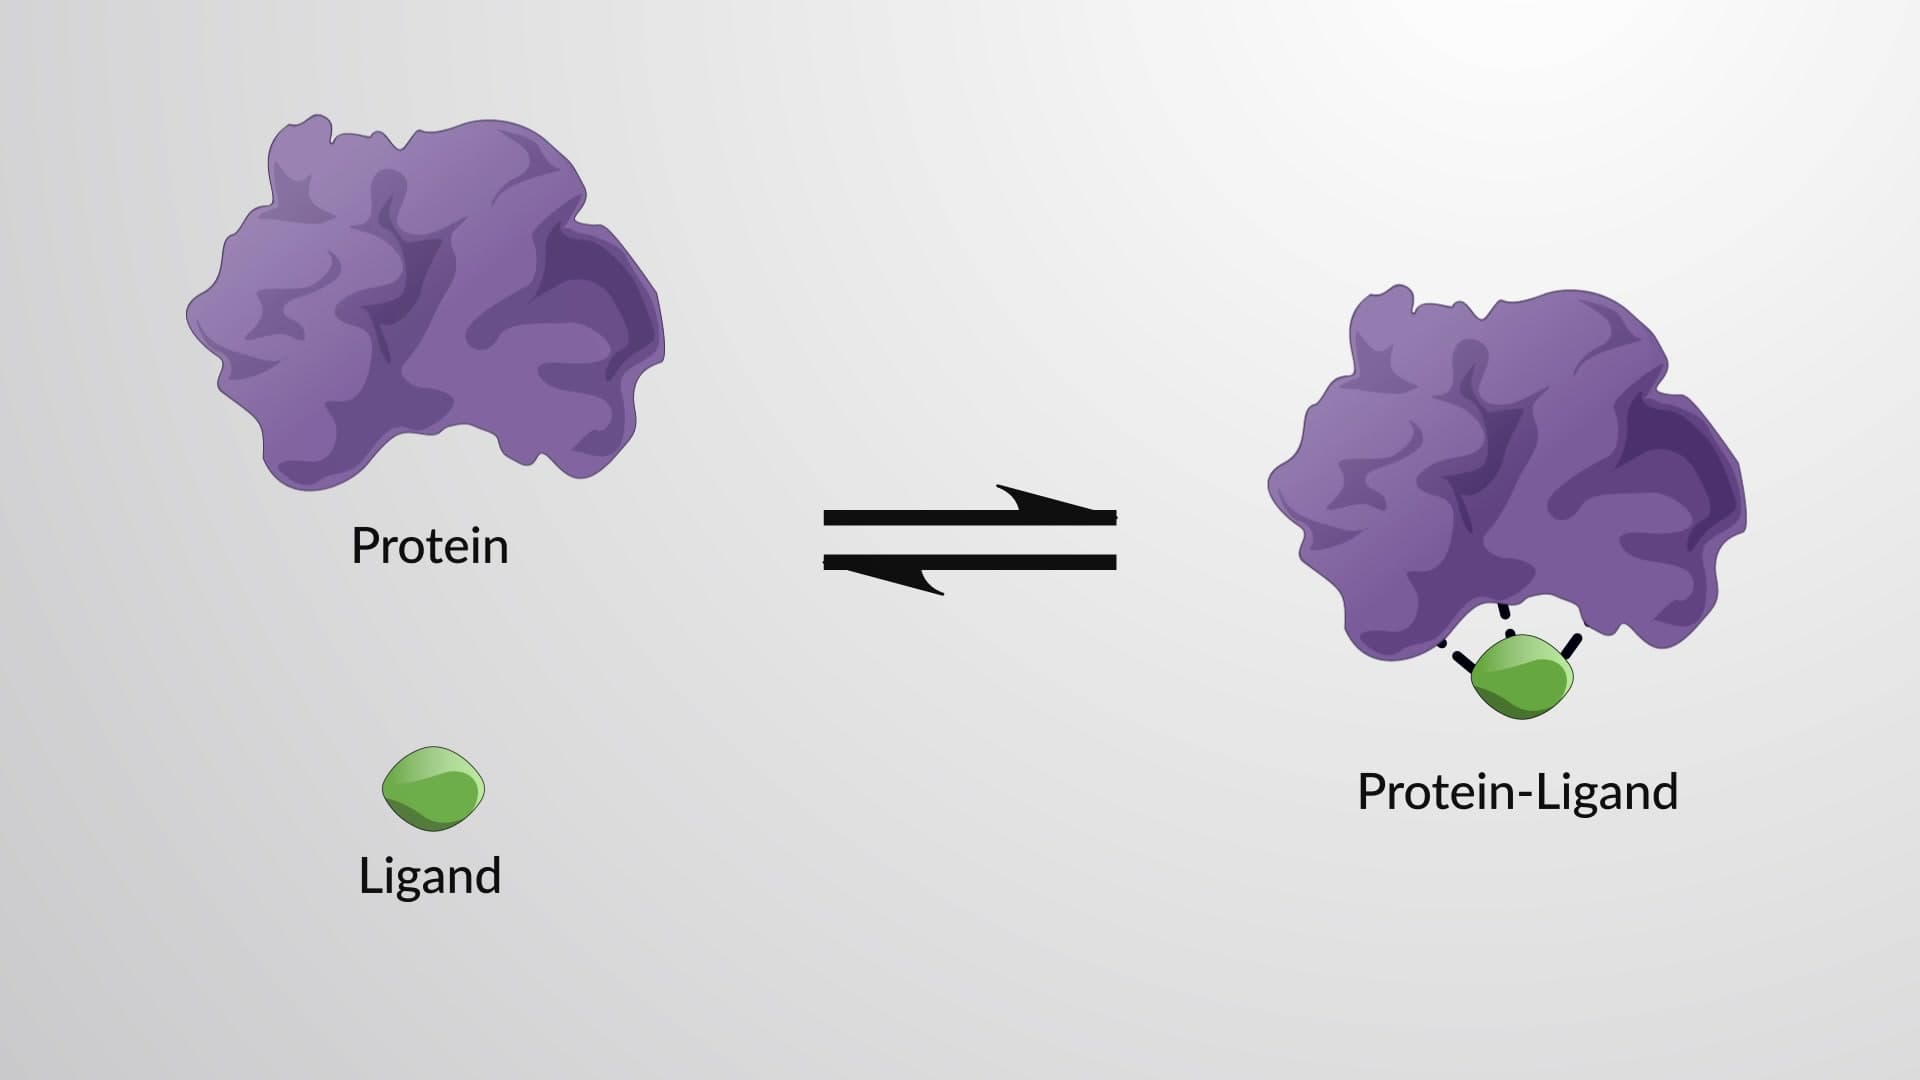
The Equilibrium Binding Constant and Binding Strength
 02:39
Gibbs Free Energy
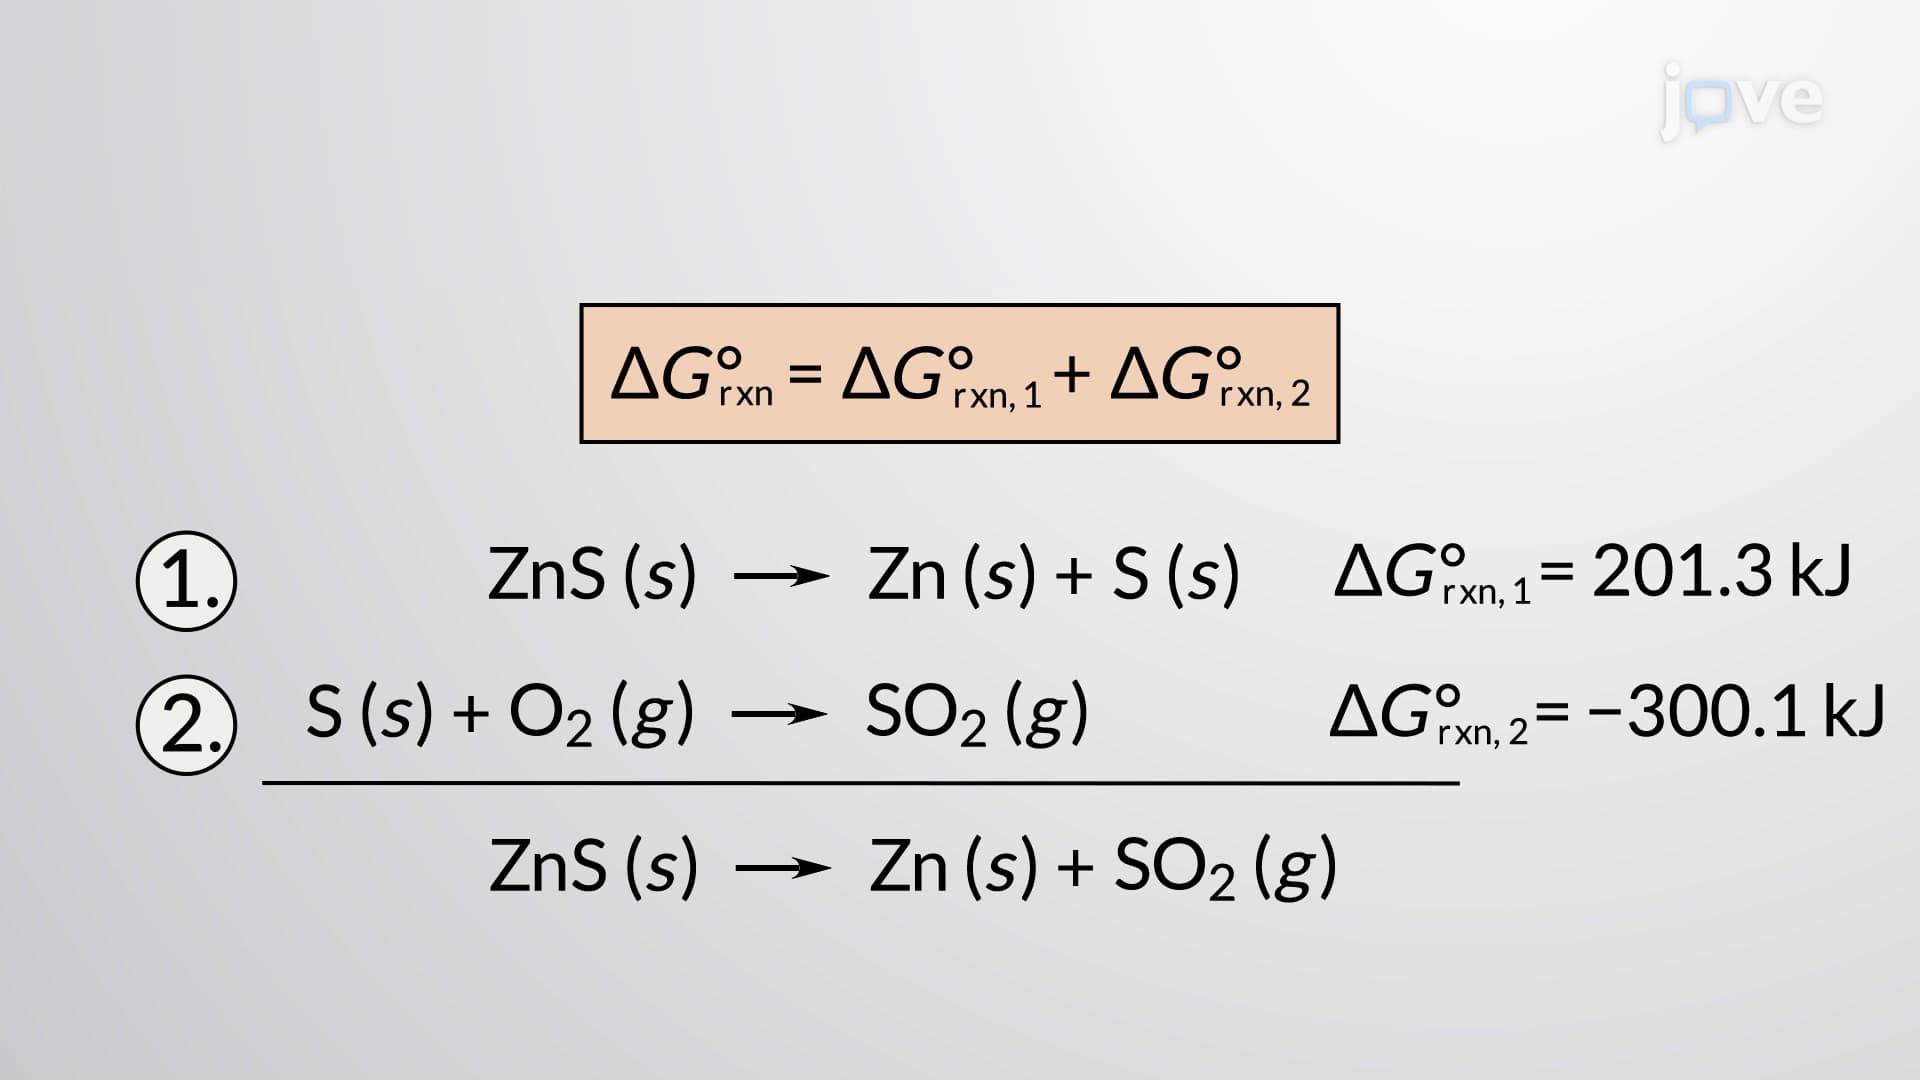 02:49
Calculating Standard Free Energy Changes
 01:05
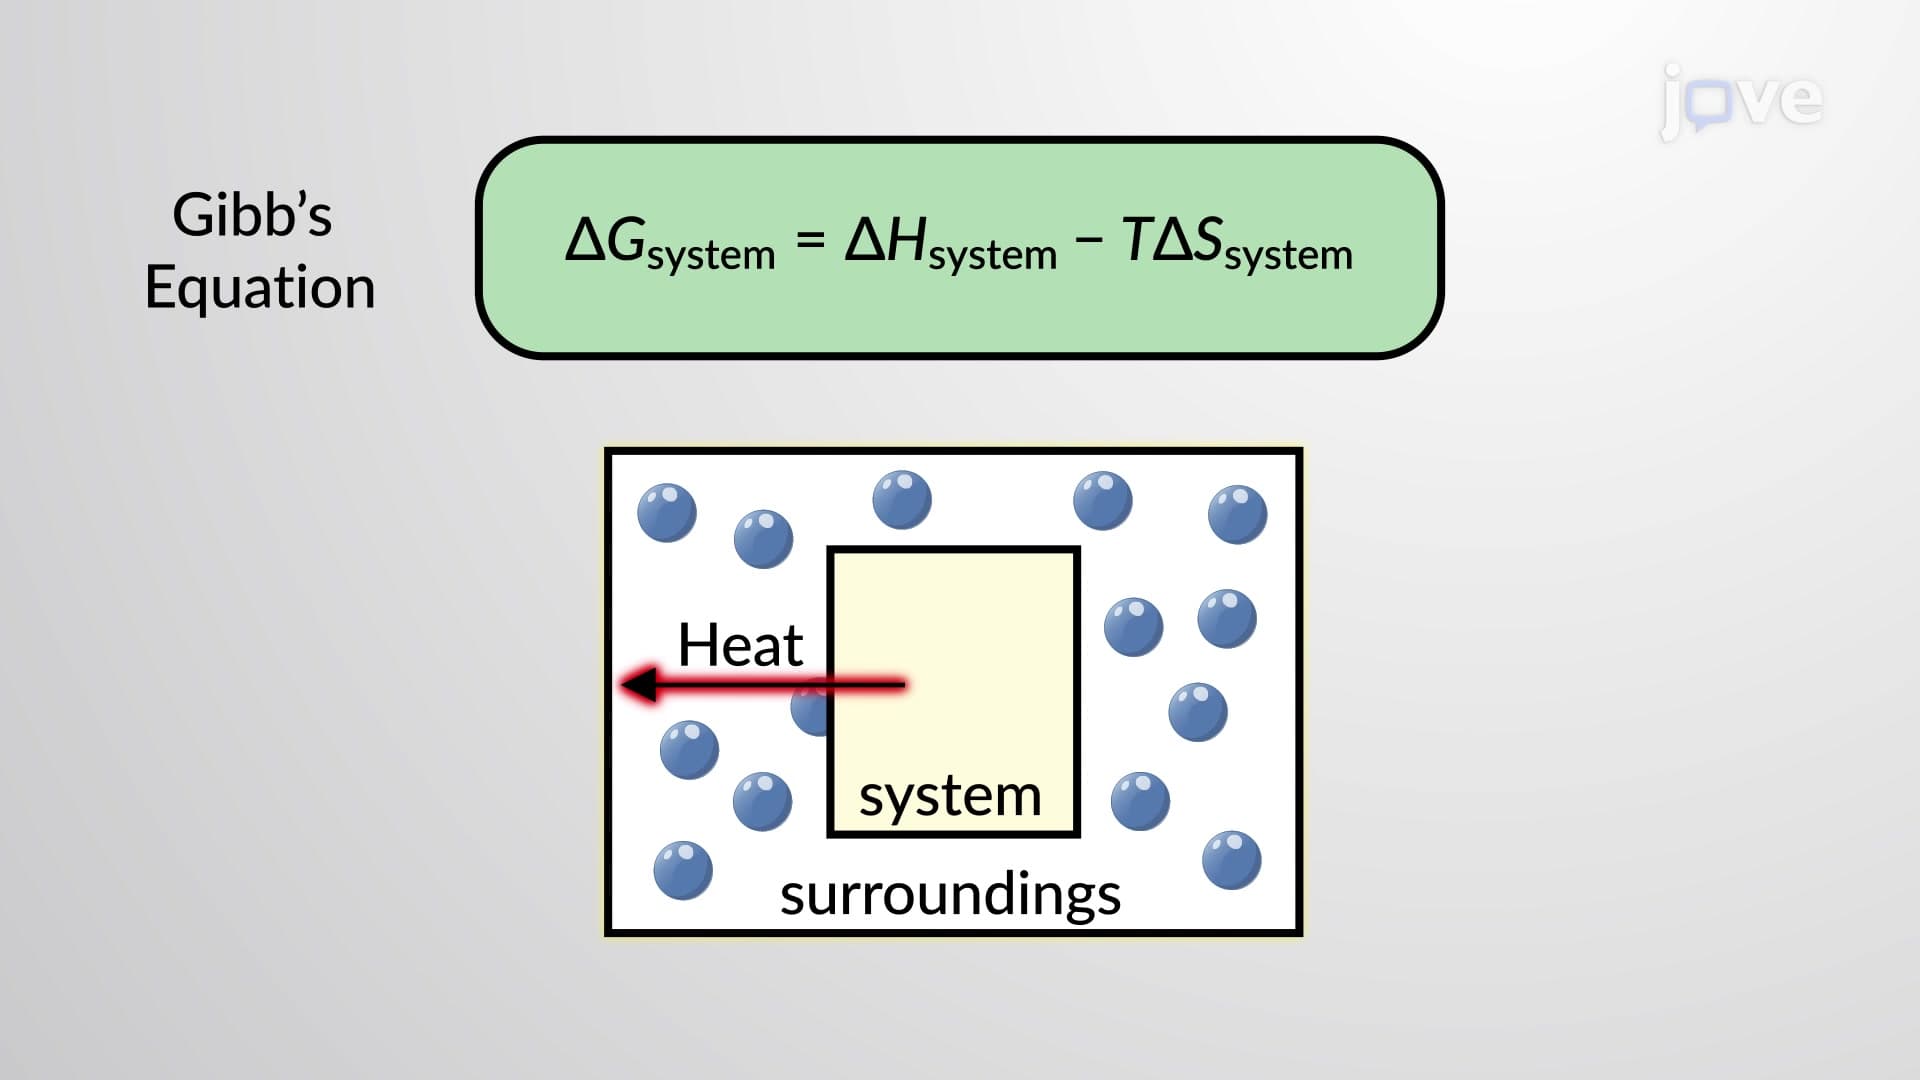
An Introduction to Free Energy
 01:13
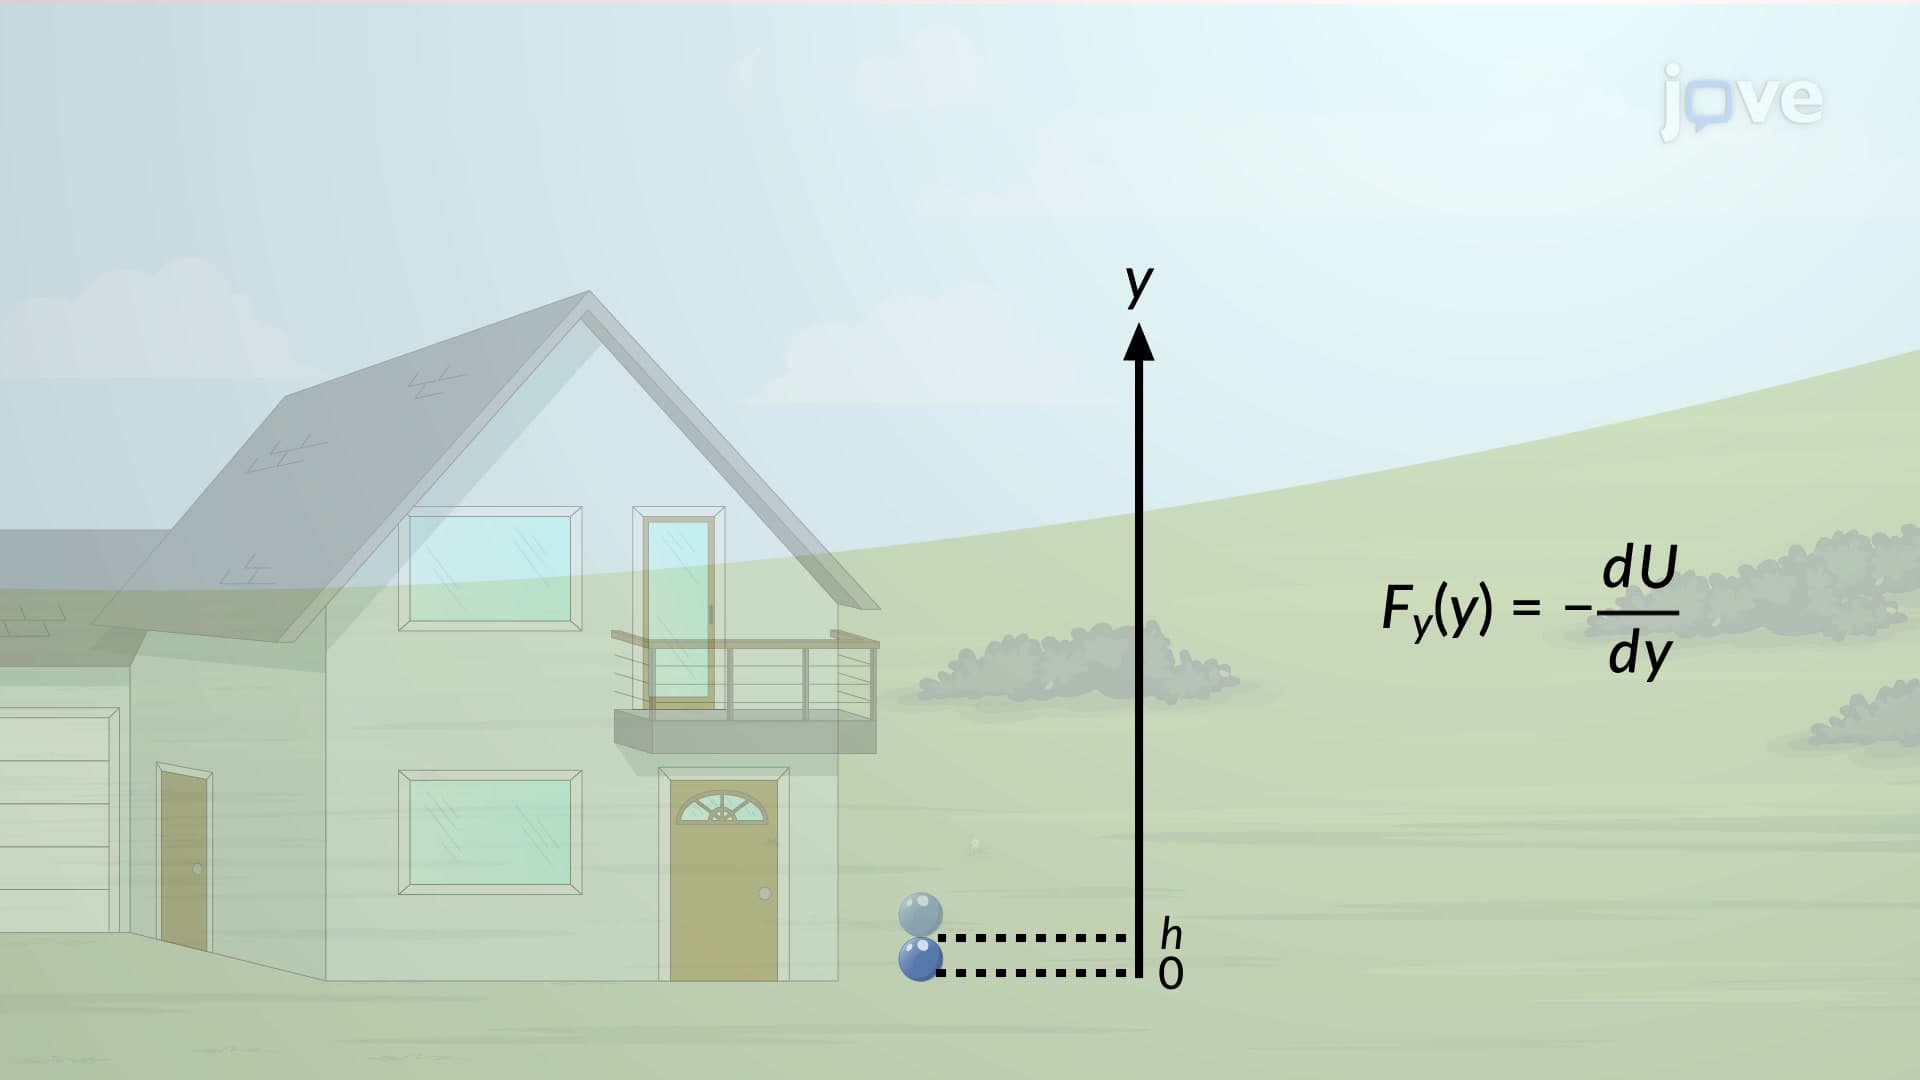
Force and Potential Energy in One Dimension
 02:56
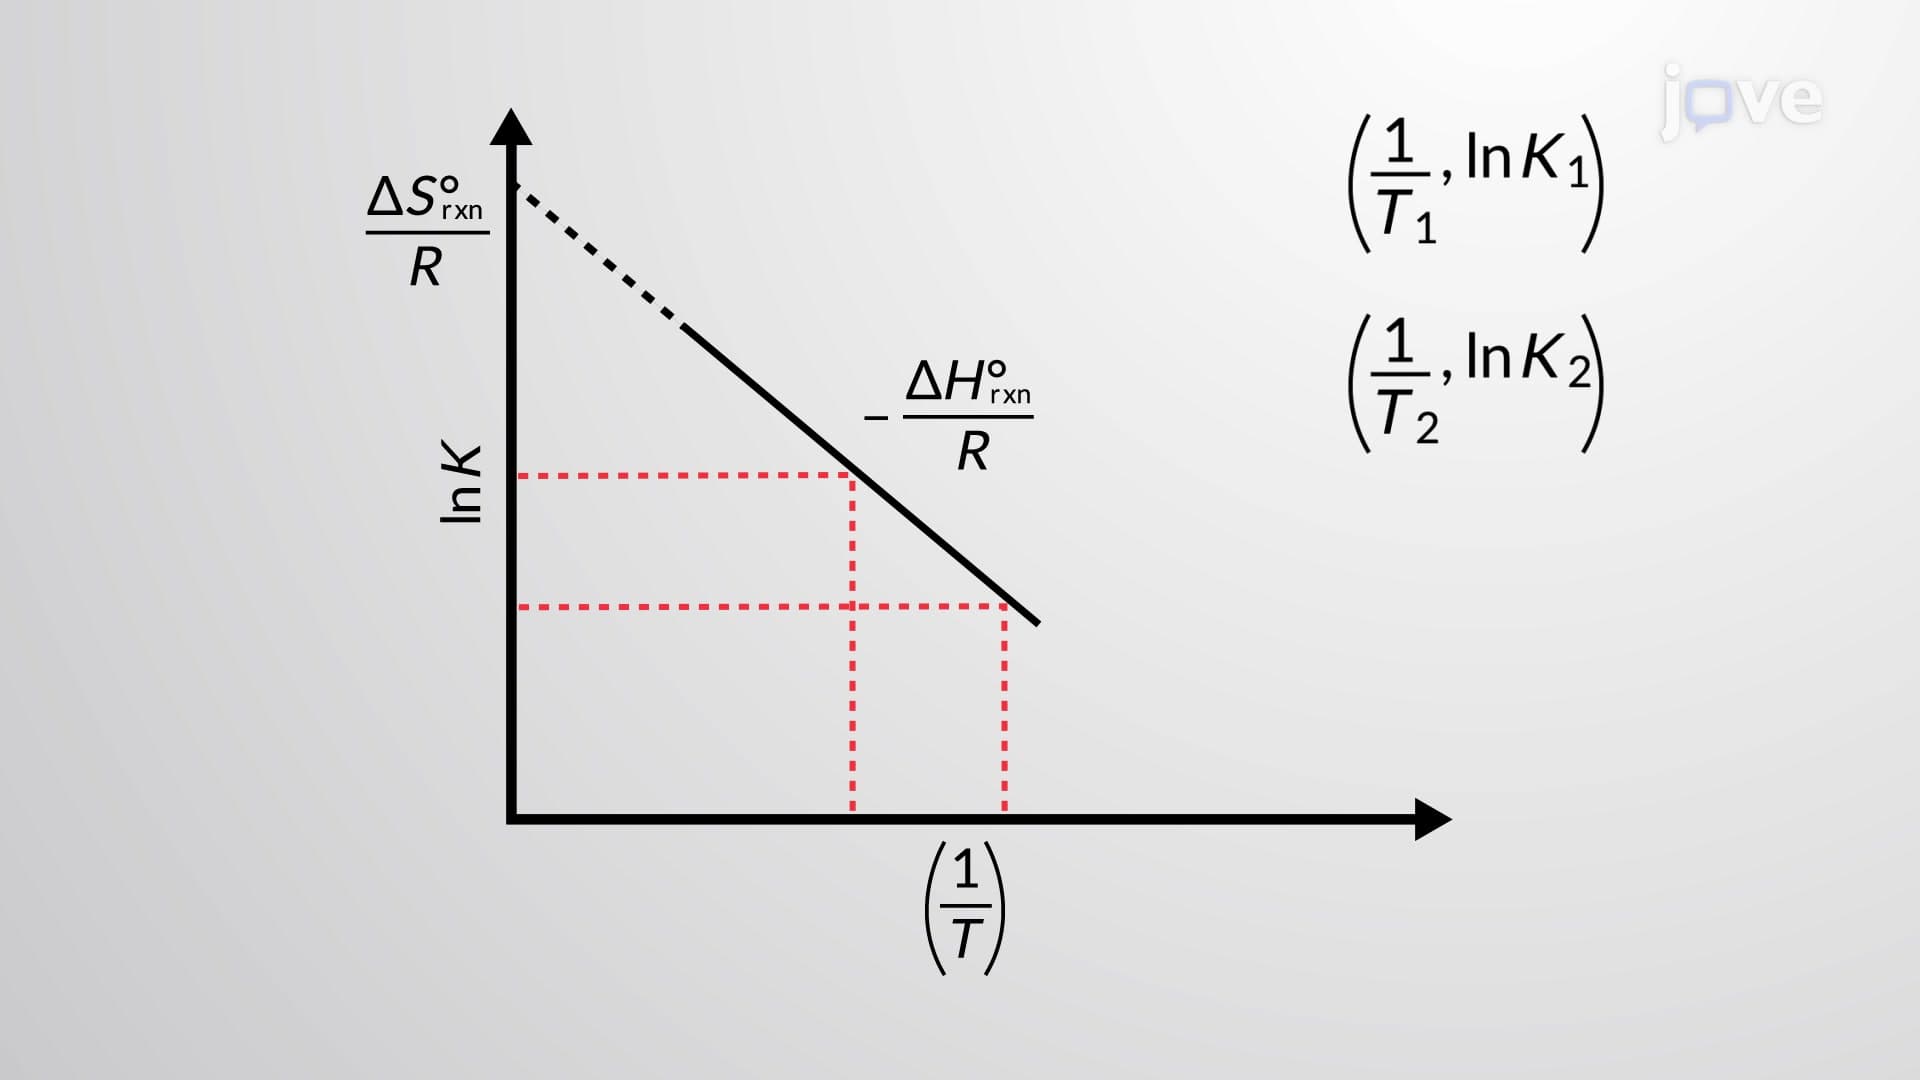
Free Energy and Equilibrium
Recall that Q is the numerical value of the mass action expression...