Predicting the open conformations of protein kinases using molecular dynamics simulations
Related Experiment Video
Updated: May 30, 2026
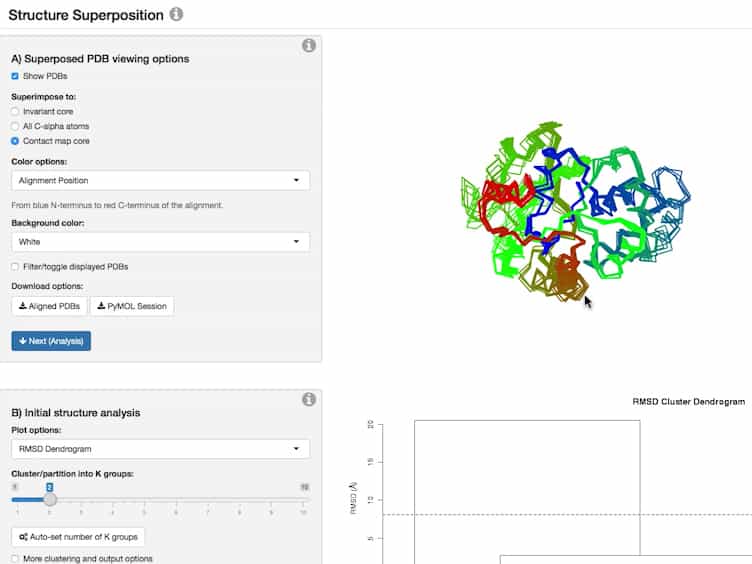
Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Una Bjarnadottir1, Jens Erik Nielsen
1UCD Conway Institute, University College Dublin, Belfield, Dublin 4, Ireland.

Abstract:
Protein kinases (PK) control phosphorylation in eukaryotic cells, and thereby regulate metabolic pathways, cell cycle progression, apoptosis, and transcription. Consequently, there is significant interest in manipulating PK activity and treat diseases by using small-molecule drugs. All PK catalytic domains undergo large conformational changes as a result of substrate binding and phosphorylation. The "closed" state of a PK catalytic domain is the only state able to phosphorylate the target substrate, which makes the two other observed states (the "open" and the "intermediate" states) interesting drug targets. We investigate whether molecular dynamics (MD) simulations starting from the closed state of the catalytic domain of protein kinase A (C-PKA) can be used to produce realistic structures representing the intermediate and/or open conformation of C-PKA, because this would allow for drug docking calculations and drug design using MD snapshots. We perform 36 ten-nanosecond MD simulations starting from the closed conformation [PDB ID: ATP] of C-PKA in various liganded and phosphorylated states. The results show that MD simulations are capable of reproducing the open conformation of C-PKA with good accuracy within 1 ns of simulation as measured by Cα root mean square deviations (RMSDs) and RMSDs of atoms defining the ATP-binding pocket. Importantly, we are able to show that even without knowledge of the structure of the open form of C-PKA, we can identify the MD snapshots resembling the open conformation most using the open structure of a different PK displaying only 23% sequence identity to C-PKA.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:58
Cooperative Allosteric Transitions
 02:54
Protein Kinases and Phosphatases
Protein kinases
Many proteins in the cell are regulated by phosphorylation, the addition of a phosphate group. A family of enzymes called kinases...
 02:04
Protein-protein Interfaces