Related Experiment Video
Updated: May 29, 2026

Simultaneous Affinity Enrichment of Two Post-Translational Modifications for Quantification and Site Localization
Published on: February 27, 2020
An improved method for the construction of decoy peptide MS/MS spectra suitable for the accurate estimation of false
Erik Ahrné1, Yuki Ohta, Frederic Nikitin
1Proteome Informatics Group, Swiss Institute of Bioinformatics, Geneva, Switzerland. erik.ahrne@isb-sib.ch

Abstract:
The relevance of libraries of annotated MS/MS spectra is growing with the amount of proteomic data generated in high-throughput experiments. These reference libraries provide a fast and accurate way to identify newly acquired MS/MS spectra. In the context of multiple hypotheses testing, the control of the number of false-positive identifications expected in the final result list by means of the calculation of the false discovery rate (FDR). In a classical sequence search where experimental MS/MS spectra are compared with the theoretical peptide spectra calculated from a sequence database, the FDR is estimated by searching randomized or decoy sequence databases. Despite on-going discussion on how exactly the FDR has to be calculated, this method is widely accepted in the proteomic community. Recently, similar approaches to control the FDR of spectrum library searches were discussed. We present in this paper a detailed analysis of the similarity between spectra of distinct peptides to set the basis of our own solution for decoy library creation (DeLiberator). It differs from the previously published results in some key points, mainly in implementing new methods that prevent decoy spectra from being too similar to the original library spectra while keeping important features of real MS/MS spectra. Using different proteomic data sets and library creation methods, we evaluate our approach and compare it with alternative methods.
Related Concept Videos
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:21
Tandem Mass Spectrometry
 01:20
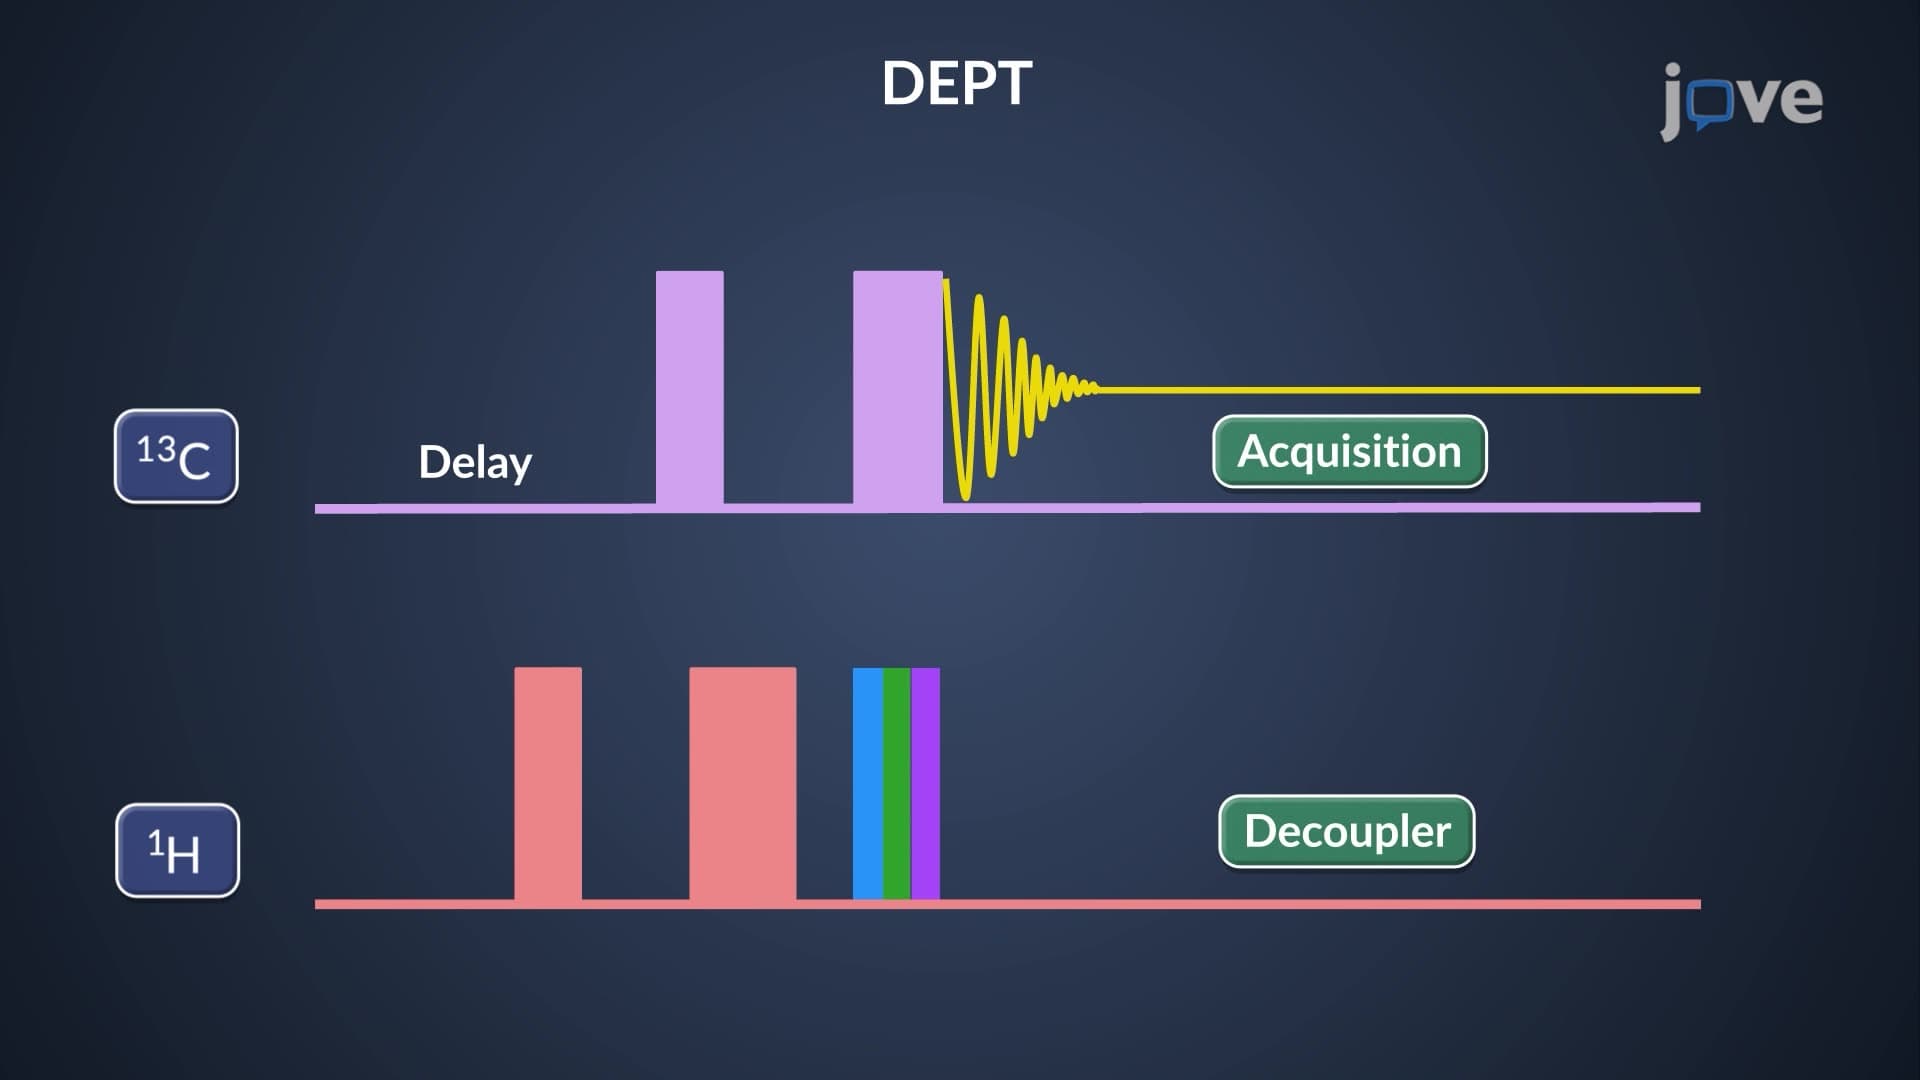
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:19
MALDI-TOF Mass Spectrometry