Related Experiment Video
Updated: May 29, 2026
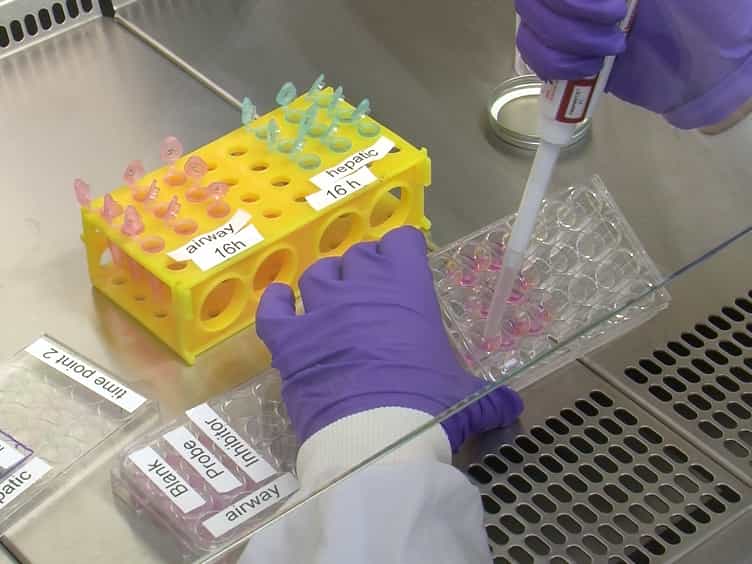
Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Determining the time course of CYP3A inhibition by potent reversible and irreversible CYP3A inhibitors using A
S Katzenmaier1, C Markert, K-D Riedel
1Department of Clinical Pharmacology and Pharmacoepidemiology, Heidelberg University, Heidelberg, Germany.

Abstract:
We established a new limited sampling strategy to assess CYP3A activity and evaluated the time course of reversible (voriconazole) and irreversible (ritonavir) CYP3A inhibition. In this randomized trial, two groups, each with eight healthy participants, received CYP3A inhibitors voriconazole or ritonavir orally for 9 days, with 3 mg midazolam (MDZ) administered before the inhibitor treatment, on days 1, 2, 3, 5, 8, and 9 during inhibitor treatment, and on days 10, 11, and 12 (3 days) after discontinuation. Plasma MDZ area under the curve (AUC) between 2 and 4 h after oral administration in the form of a solution strongly correlated with MDZ clearance. Using this parameter, maximum inhibition of voriconazole and ritonavir was calculated to have occurred only 48 h after starting of the inhibitor (percentage of baseline MDZ clearance, voriconazole: 10.6%; ritonavir: 8.4%). Recovery of CYP3A activity occurred with a half-life of 24 h after voriconazole, whereas ritonavir inhibition was still strong 3 days after discontinuation. These findings underscore the substantial and gradual alterations in dose requirements in the first days of and after such combination therapies.
Insights
A new limited sampling strategy effectively assessed CYP3A inhibition by voriconazole and ritonavir. Maximum inhibition occurred within 48 hours, with varying recovery times impacting drug dosage adjustments.
Area of Science:
- Pharmacology
- Drug Metabolism
- Clinical Trials
Background:
- Cytochrome P450 3A (CYP3A) is a key enzyme in drug metabolism.
- Understanding CYP3A inhibition kinetics is crucial for safe drug combinations.
- Voriconazole and ritonavir are known CYP3A inhibitors with different mechanisms.
Purpose of the Study:
- To establish and validate a limited sampling strategy for assessing CYP3A activity.
- To evaluate the time course of reversible (voriconazole) and irreversible (ritonavir) CYP3A inhibition.
- To correlate plasma midazolam (MDZ) area under the curve (AUC) with MDZ clearance.
Main Methods:
- A randomized trial involving 16 healthy participants (2 groups of 8).
- Administration of oral voriconazole or ritonavir for 9 days.
- Timed oral administration of midazolam (MDZ) at multiple time points relative to inhibitor treatment.
Main Results:
- Plasma MDZ AUC (2-4h) strongly correlated with MDZ clearance, validating the limited sampling strategy.
- Maximum CYP3A inhibition by both voriconazole and ritonavir occurred within 48 hours.
- Voriconazole inhibition showed a 24-hour recovery half-life, while ritonavir inhibition persisted 3 days post-discontinuation.
Conclusions:
- The limited sampling strategy is effective for assessing CYP3A inhibition.
- Early and gradual alterations in CYP3A activity occur during and after inhibitor therapy.
- Findings highlight the need for careful dose adjustments in combination therapies involving CYP3A inhibitors.
Related Concept Videos
 01:14
Time Course of Drug Effect
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
 01:20
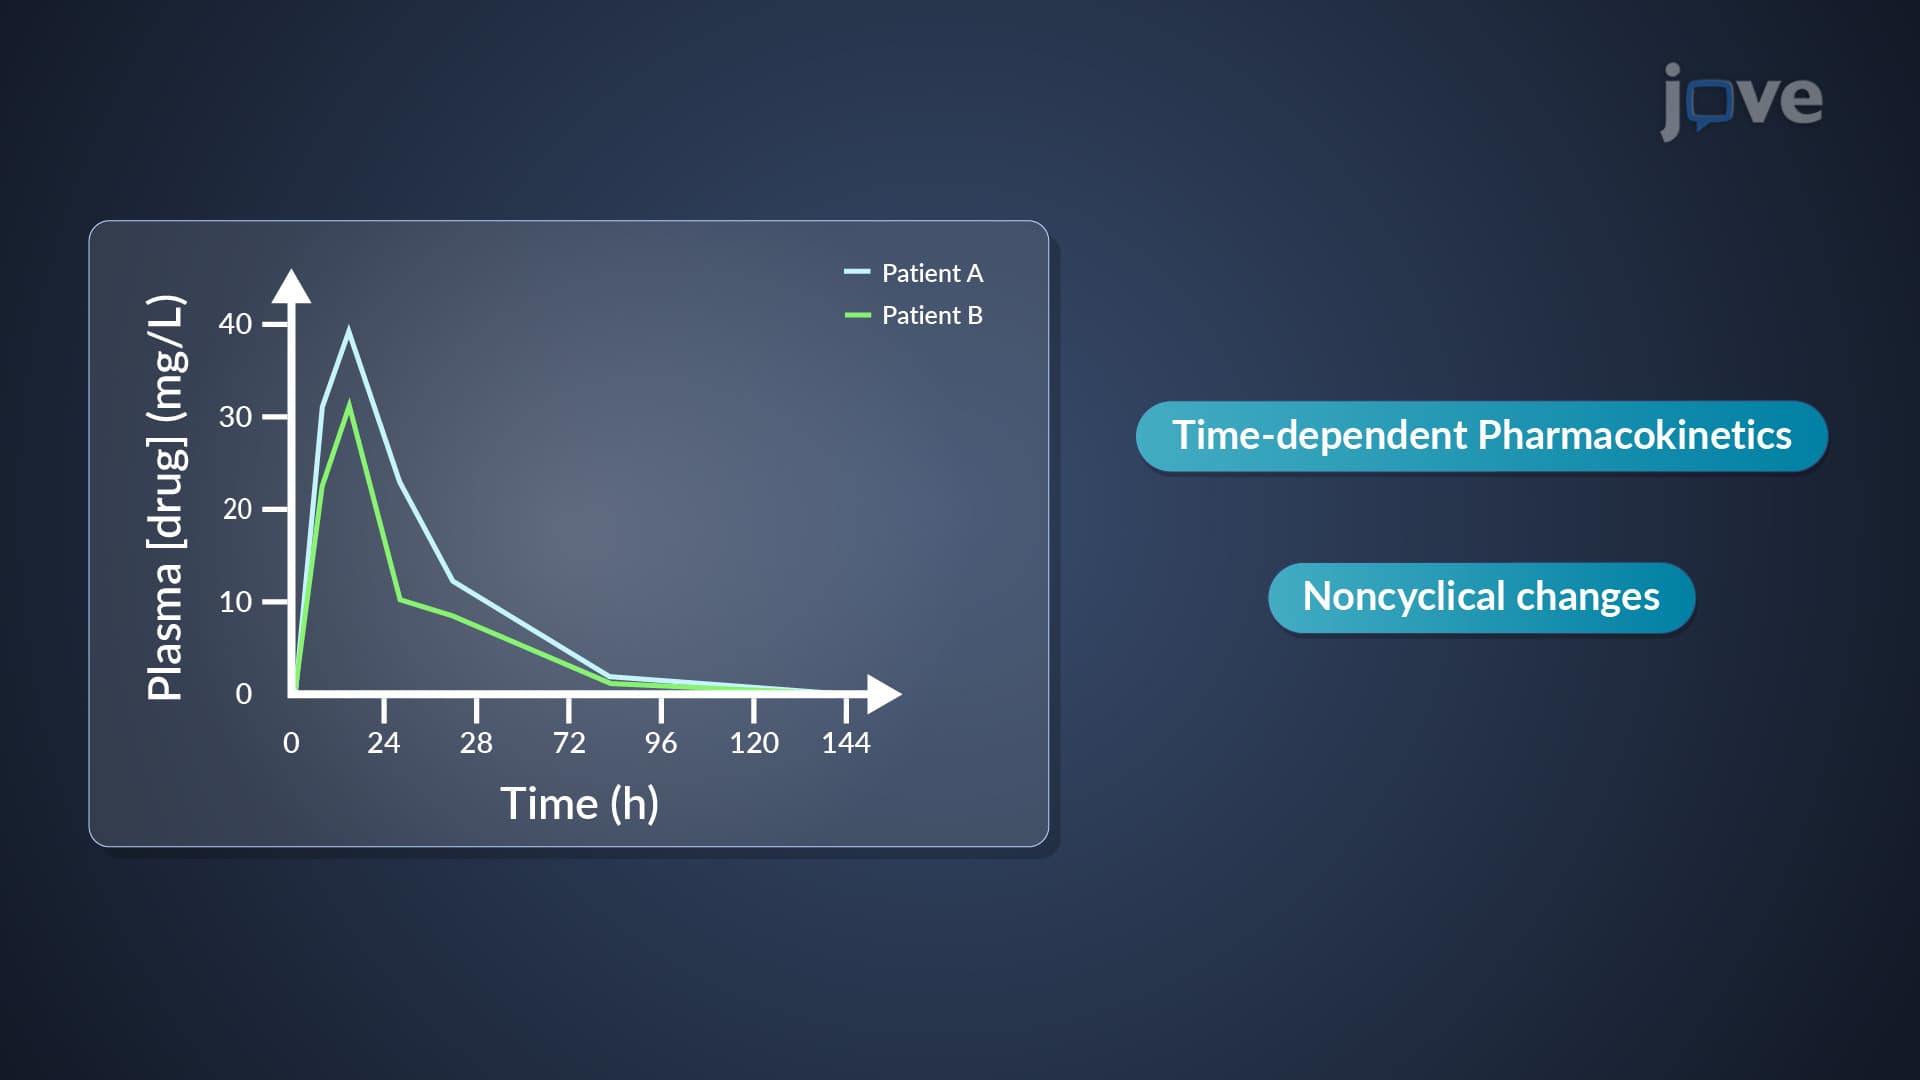
Chronopharmacokinetics: Time-Dependent Pharmacokinetics
Time-dependent pharmacokinetics refers to non-cyclical changes in drug rate processes over a period of time. It can lead to nonlinear pharmacokinetics, where the relationship between drug concentration and time is not proportional. Non-cyclical...
 01:11
Bioavailability Study Design: Single Versus Multiple Dose Studies
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the lowest drug...