
Analysis of electronic states and energy level structure of uranyl in compounds
1Chemical Sciences and Engineering Division, Argonne National Laboratory, Argonne, Illinois 60439, United States.

Abstract:
An effective operator Hamiltonian has been developed for evaluating the excited states of uranyl ion in compounds. Including free-ion and crystal-field interactions, the parametrized Hamiltonian is able to reproduce the energy level structure in good agreement with experimental measurements on uranyl in chloride compounds and nitride complexes. It is shown that the low-lying excited states belong to a nonbonding σf configuration (therefore, one Slater integral of G(3) is sufficient to account for the Coulomb electron exchange interaction) and that the spin-orbit coupling surpasses the electrostatic interactions in energy level splitting. In comparison with previously reported ab initio calculations, the present studies find that the Coulomb interactions in uranyl were excessively evaluated in the first-principle calculations.
Related Concept Videos
 01:18
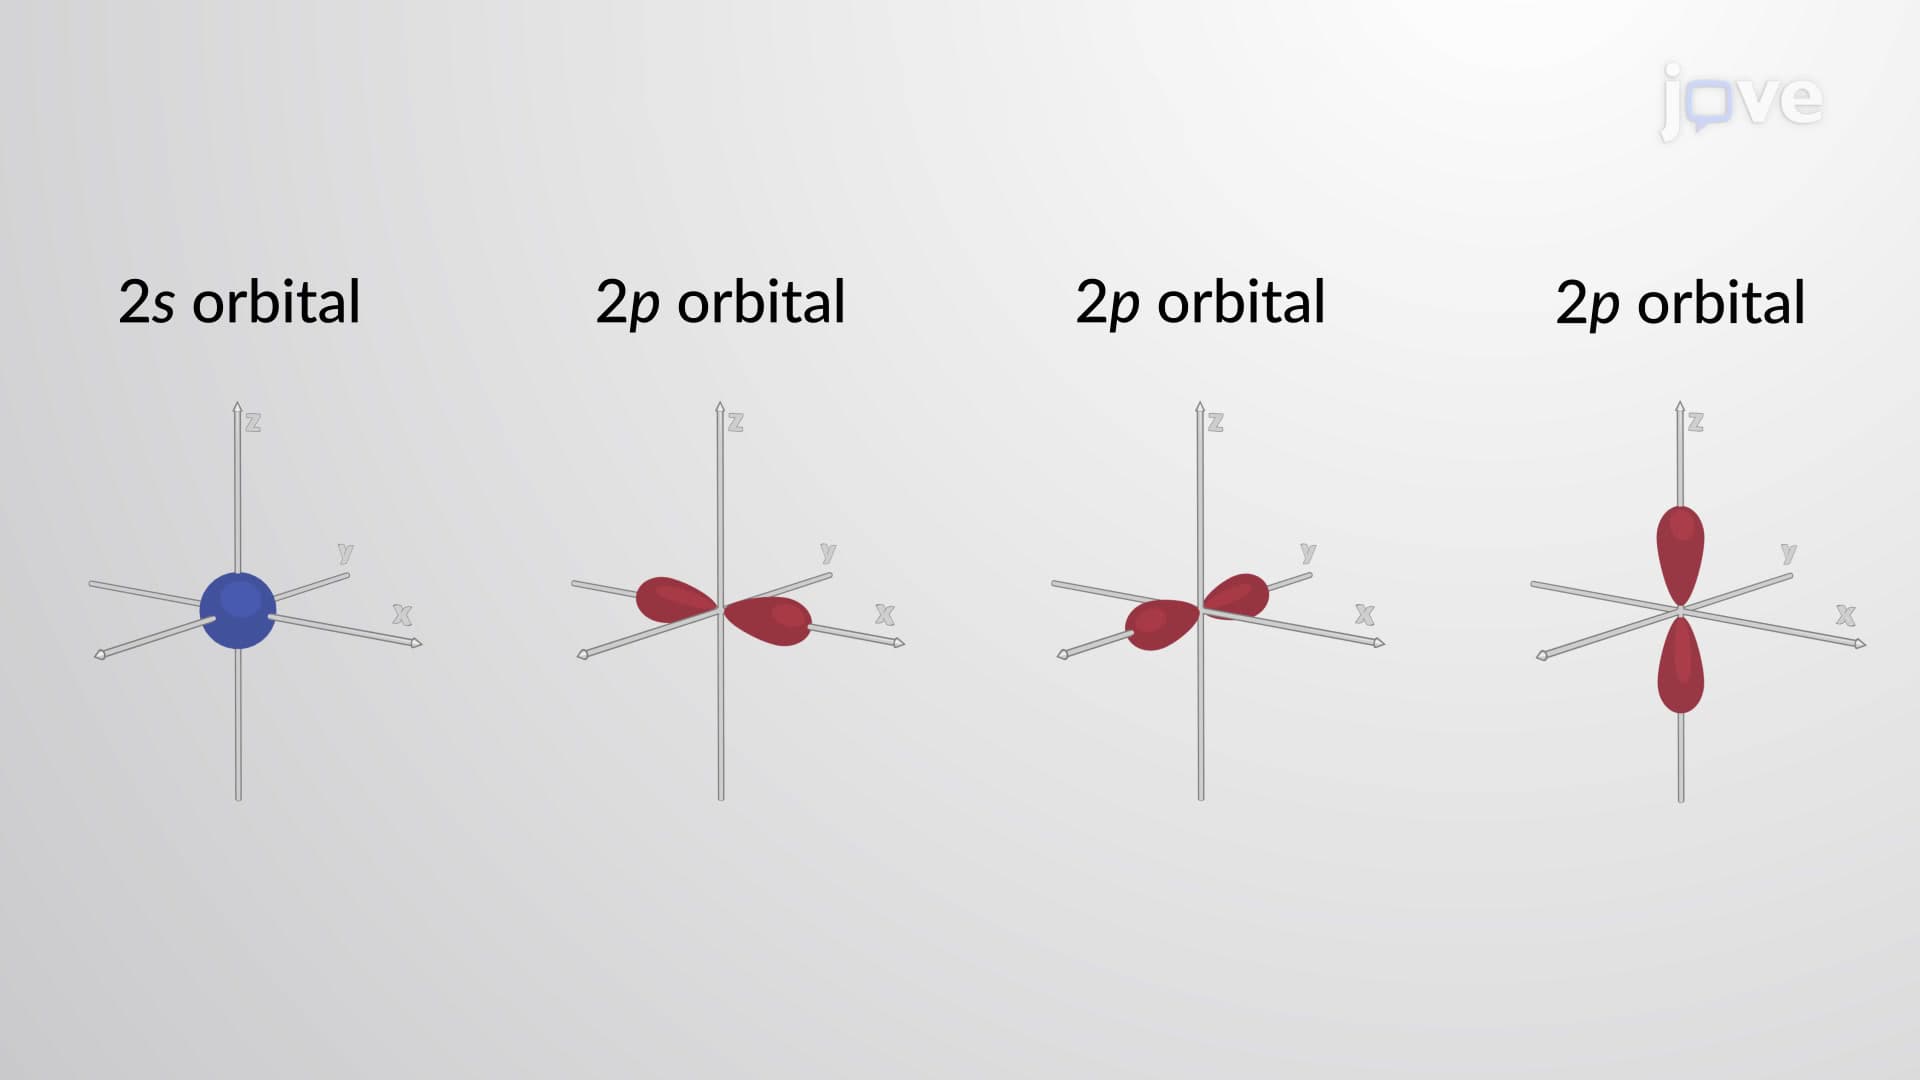
Electron Orbital Model
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p, 4s,...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 03:21
The Energies of Atomic Orbitals
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...