Related Experiment Video
Updated: May 29, 2026
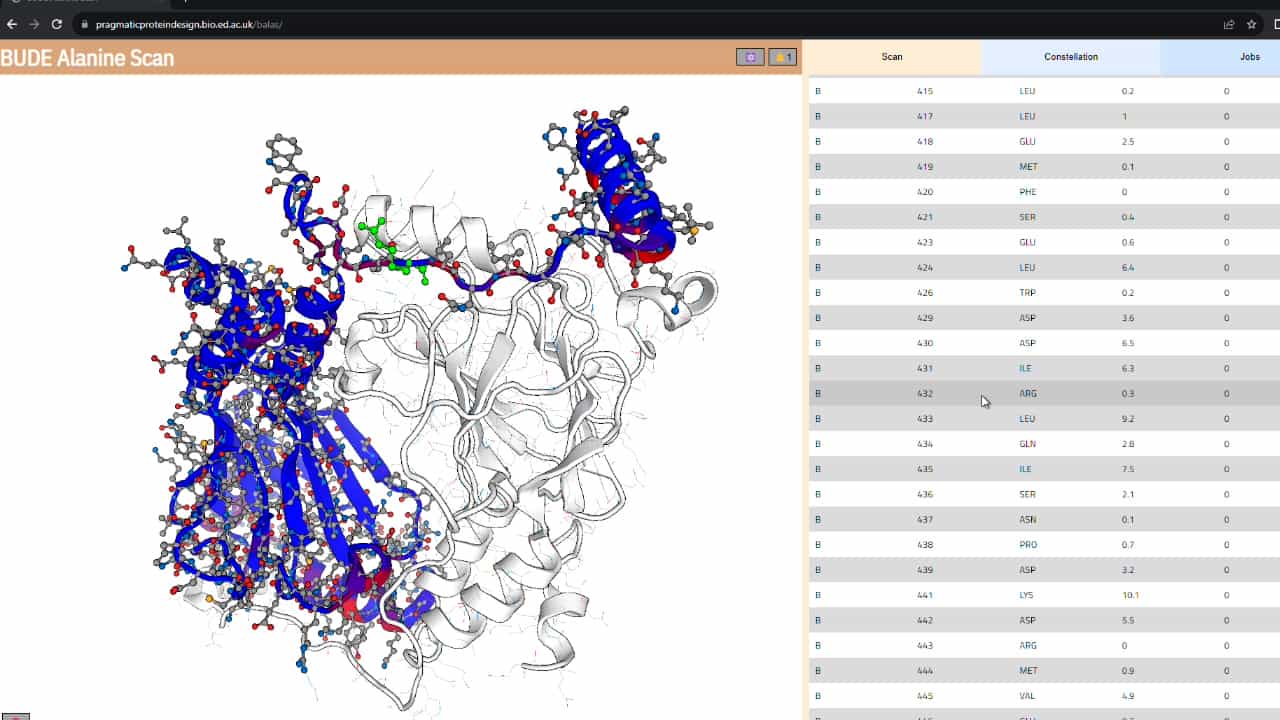
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
Structure-based characterization of the binding of peptide to the human endophilin-1 Src homology 3 domain using
Chunjiang Fu1, Gang Wu, Fenglin Lv
1Department of Cardiology, Daping Hospital, The Third Military Medical University, Chongqing, China.

Abstract:
Many protein-protein interactions are mediated by a peptide-recognizing domain, such as WW, PDZ, or SH3. In the present study, we describe a new method called position-dependent noncovalent potential analysis (PDNPA), which can accurately characterize the nonbonding profile between the human endophilin-1 Src homology 3 (hEndo1 SH3) domain and its peptide ligands and quantitatively predict the binding affinity of peptide to hEndo1 SH3. In this procedure, structure models of diverse peptides in complex with the hEndo1 SH3 domain are constructed by molecular dynamics simulation and a virtual mutagenesis protocol. Subsequently, three noncovalent interactions associated with each position of the peptide ligand in the complexed state are analyzed using empirical potential functions, and the resulting potential descriptors are then correlated with the experimentally measured affinity on the basis of 1997 hEndo1 SH3-binding peptides with known activities, using linear partial least squares regression (PLS) and the nonlinear support vector machine (SVM). The results suggest that: (i) the electrostatics appears to be more important than steric properties and hydrophobicity in the formation of the hEndo1 SH3-peptide complex; (ii) P(-4) of the core decapeptide ligand with the sequence pattern P(-6)P(-5)P(-4)P(-3)P(-2)P(-1)P(0)P(1)P(2)P(3) is the most important position in terms of determining both the stability and specificity of the architecture of the complex, and; (iii) nonlinear SVM appears to be more effective than linear PLS for accurately predicting the binding affinity of a peptide ligand to hEndo1 SH3, whereas PLS models are straightforward and easy to interpret as compared to those built by SVM.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 02:18
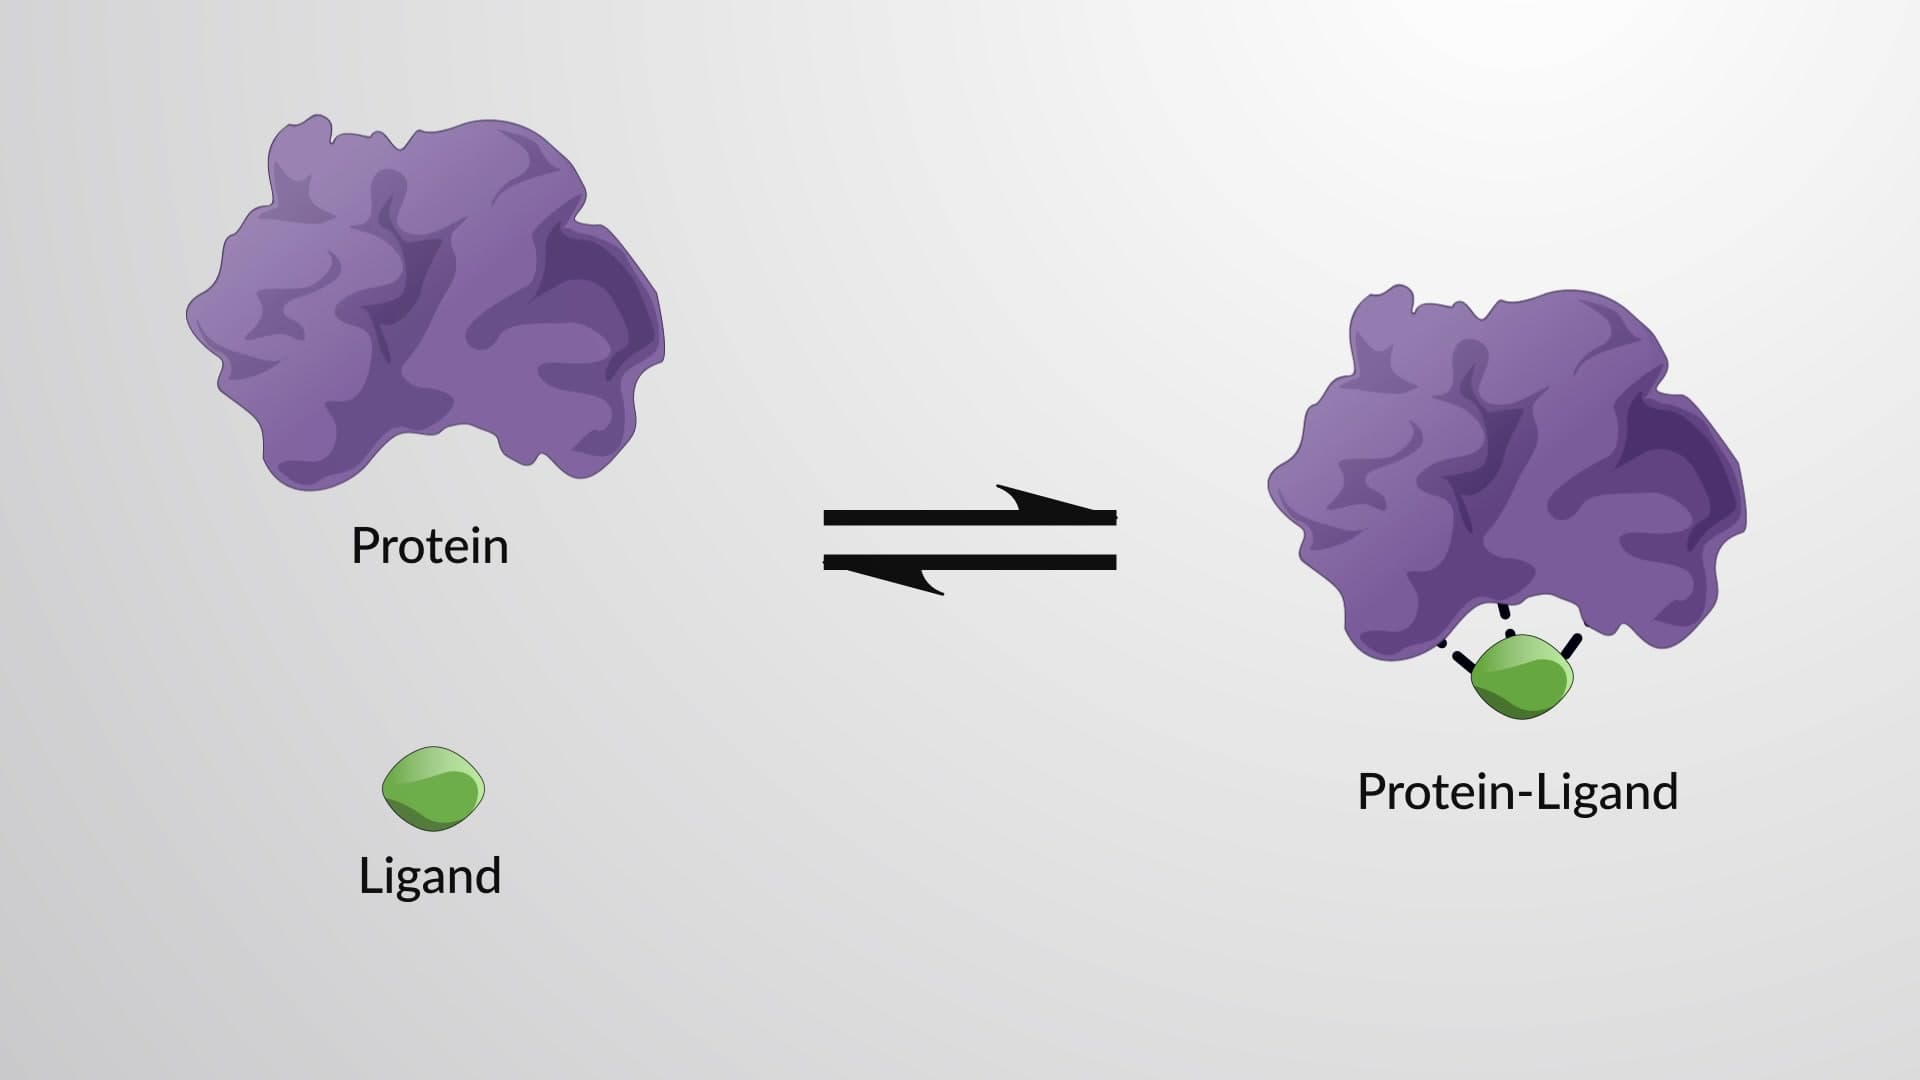
The Equilibrium Binding Constant and Binding Strength