Related Experiment Video
Updated: May 28, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
First principles calculations on structure, bonding, and vibrational frequencies of SiP2
F Bachhuber1, J Rothballer, F Pielnhofer
1Universität Regensburg, Institute of Inorganic Chemistry, Universitätsstr. 31, 93040 Regensburg, Germany.

Abstract:
Pyrite type SiP(2) is reinvestigated by first principles calculations on various levels of functionals including local density approximation, generalized gradient approximation, Becke-Lee-Yang-Parr hybrid functional, and the Hartree-Fock method. SiP(2) is seen as a model compound with molecular [P-P] entities and [SiP(6)] octahedra. Structure and bonding are addressed by electronic structure calculations. Special attention is spent on P-P and Si-P bonds in terms of bond lengths and respective stretching modes from simulated Raman spectra. The electronic structure is analyzed in both direct and momentum space by the electron localization function and site projected density of states. The main goals of this work are to understand the nature of chemical bonding in SiP(2) and to compare and contrast the different methods of calculation.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 02:52
VSEPR Theory and the Basic Shapes
 03:51
Molecular Orbital Theory II
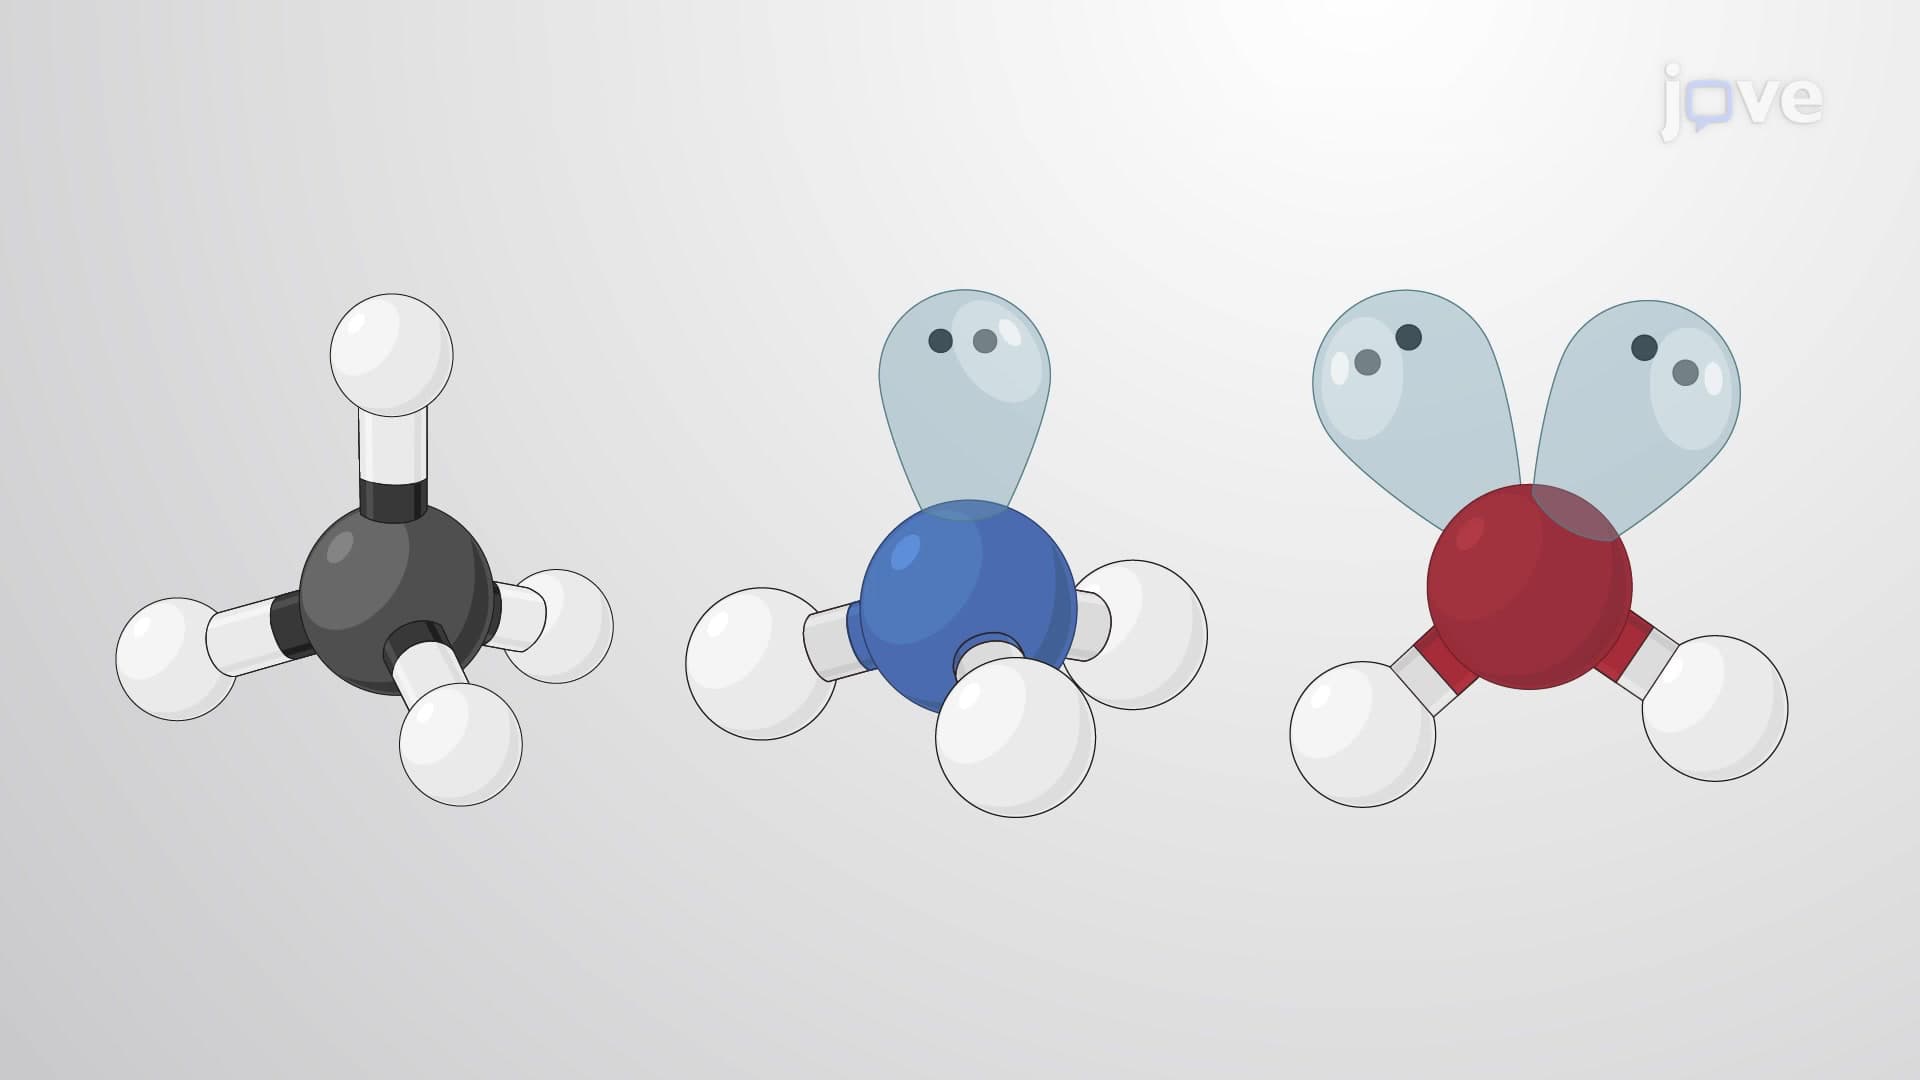 02:37
VSEPR Theory
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:24
Hybridization of Atomic Orbitals I
 02:40
MO Theory and Covalent Bonding