Related Experiment Video
Updated: May 28, 2026

Rapid Point-of-Care Assay of Enoxaparin Anticoagulant Efficacy in Whole Blood
Published on: October 12, 2012
Pharmacokinetics of the low molecular weight heparin dalteparin in cats
Reinhard Mischke1, Judith Schmitt, Sonja Wolken
1Small Animal Clinic, University of Veterinary Medicine Hannover, Bünteweg 9, D-30559 Hannover, Germany. Reinhard.Mischke@tiho-hannover.de

Low molecular weight heparin (LMWH) is used as an anticoagulant in cats although only limited pharmacokinetic data are available in this species. The aim of the present study was to establish the pharmacokinetics of dalteparin in cats based on anti-FXa heparin activities. Groups of clinically healthy cats (six animals per treatment) received individual LMWH injections at three different doses intravenously (IV) (25, 50, 100 anti-factor Xa international units [IU anti-FXa]/kg) or subcutaneously (SC) (50, 100, 200 IU anti-FXa/kg). Blood samples were collected before and at various times after injection. Anti-FXa activity was measured with a chromogenic substrate test. Following IV injection, maximum plasma heparin activities (C(max)) were 0.67 ± 0.14, 1.44 ± 0.22 and 2.87 ± 0.38 IU anti-FXa/mL, respectively. The calculated mean half-life (t(½)) was between 39 and 57 min and was not significantly dose-dependent (P=0.139). The volume of distribution (35-39 mL/kg) was almost equivalent to the plasma volume. After SC injection, C(max) values of 0.41 ± 0.10, 0.86 ± 0.17 or 1.91 ± 0.16 IU anti-FXa/mL, respectively, were calculated at 91-110 min post-injection. The t(½) values were between 106 and 122 min and were not significantly influenced by dose (P=0.784). The bioavailability after SC injection was approximately 100%. The high bioavailability of the SC administered LMWH dalteparin in cats was consistent with other species and indicated predictable blood levels. However, the comparatively short t(½) may indicate the necessity of multiple daily injections, which should be verified in clinical trials.
Low molecular weight heparin (LMWH) is used as an anticoagulant in cats although only limited pharmacokinetic data are available in this species. The aim of the present study was to establish the pharmacokinetics of dalteparin in cats based on anti-FXa heparin activities. Groups of clinically healthy cats (six animals per treatment) received individual LMWH injections at three different doses intravenously (IV) (25, 50, 100 anti-factor Xa international units [IU anti-FXa]/kg) or subcutaneously (SC) (50, 100, 200 IU anti-FXa/kg). Blood samples were collected before and at various times after injection. Anti-FXa activity was measured with a chromogenic substrate test. Following IV injection, maximum plasma heparin activities (C(max)) were 0.67 ± 0.14, 1.44 ± 0.22 and 2.87 ± 0.38 IU anti-FXa/mL, respectively. The calculated mean half-life (t(½)) was between 39 and 57 min and was not significantly dose-dependent (P=0.139). The volume of distribution (35-39 mL/kg) was almost equivalent to the plasma volume. After SC injection, C(max) values of 0.41 ± 0.10, 0.86 ± 0.17 or 1.91 ± 0.16 IU anti-FXa/mL, respectively, were calculated at 91-110 min post-injection. The t(½) values were between 106 and 122 min and were not significantly influenced by dose (P=0.784). The bioavailability after SC injection was approximately 100%. The high bioavailability of the SC administered LMWH dalteparin in cats was consistent with other species and indicated predictable blood levels. However, the comparatively short t(½) may indicate the necessity of multiple daily injections, which should be verified in clinical trials.
More Related Videos
Related Concept Videos
 01:30
Anticoagulant Drugs: Low-Molecular-Weight Heparins
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Duration of Dose-Effect Relationship
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 00:59
Nonlinear Pharmacokinetics: Drug Elimination for IV Bolus Injection
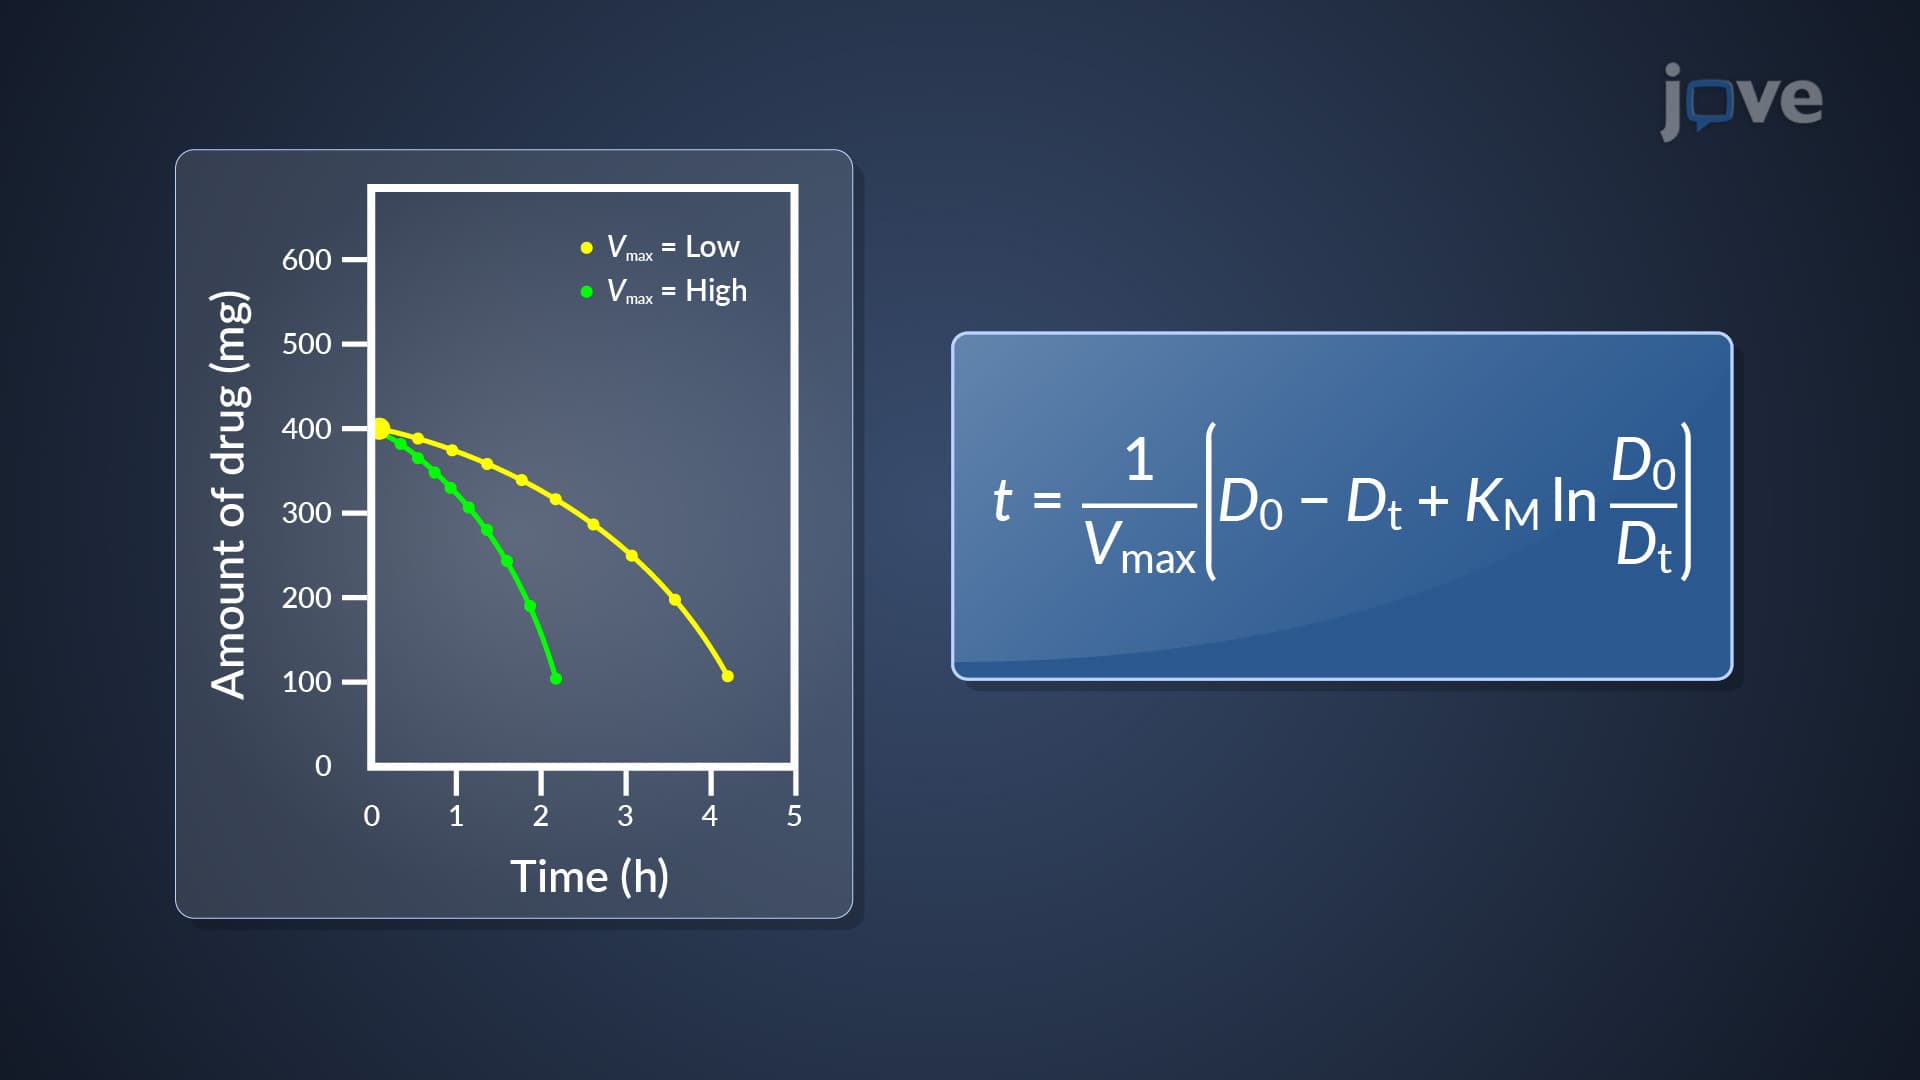
Following the administration of a single intravenous (IV) bolus injection, we can determine the concentration of the drug in the plasma at any given time. This calculation is achieved using a specific equation that integrates the values of Vmax and KM.
We can also...
 01:01
Dosage Regimens: Partial Pharmacokinetic Parameters
 01:14
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug